Related Experiment Video
Updated: Jun 16, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Ab initio molecular dynamics simulations of the adsorption of H2 on palladium surfaces
1Institut für Theoretische Chemie, Universität Ulm, D-89069 Ulm, Germany. axel.gross@uni-ulm.de

Abstract:
The interaction of hydrogen with palladium surfaces represents a model system for the study of the adsorption and absorption at metal surfaces. Theoretical gas-surface dynamics studies have usually concentrated on the adsorption dynamics on clean surfaces. Only recently has it become possible, based on advances in electronic structure codes and improvements in computer power, to address the much more complex problem of the adsorption dynamics on precovered surfaces. Herein, recent ab initio molecular dynamics studies are discussed that address the adsorption dynamics of hydrogen molecules on hydrogen- and sulfur-precovered Pd surfaces. In addition, the relaxation dynamics of the hydrogen atoms after the dissociation on clean Pd(100) are presented.
More Related Videos
Related Concept Videos
 01:28
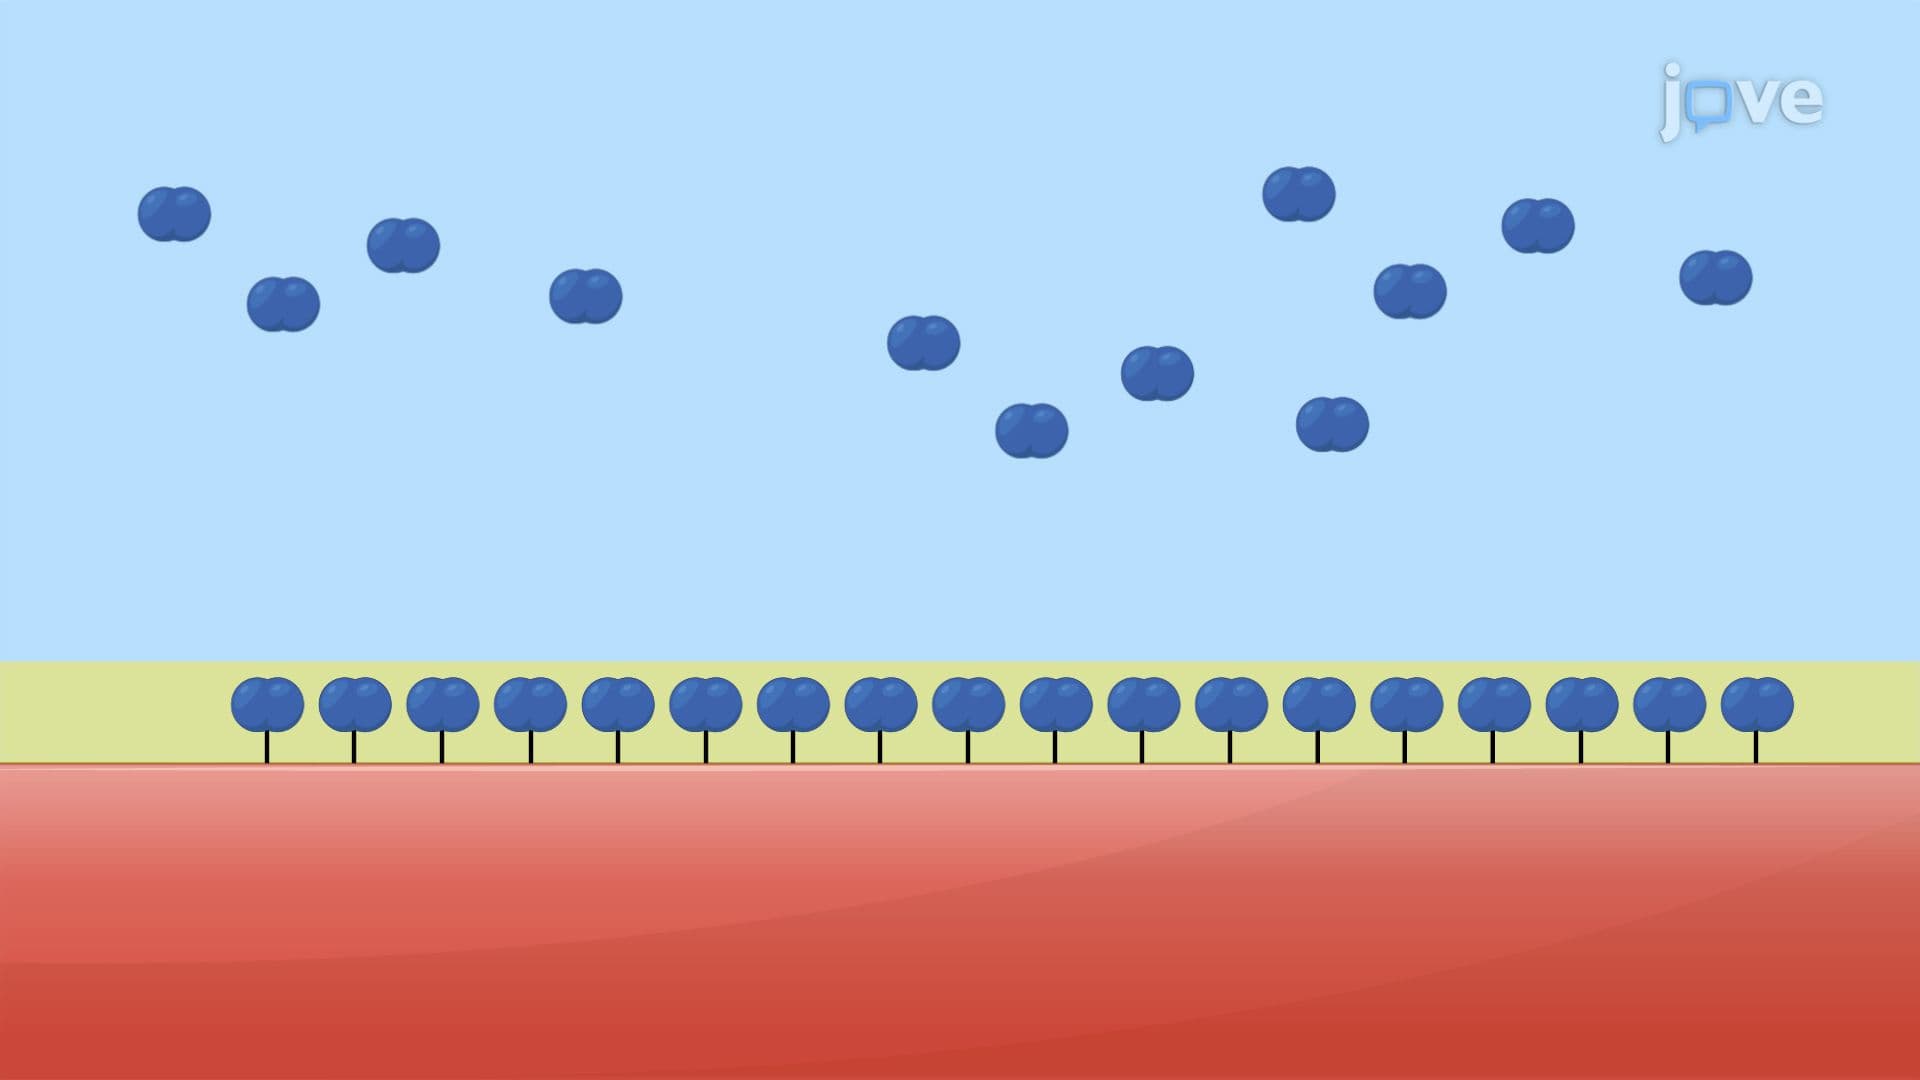
Adsorption of Gases on Solids
 01:22
Heterogeneous Catalysis
 01:29
Adsorption Isotherms I
 01:25
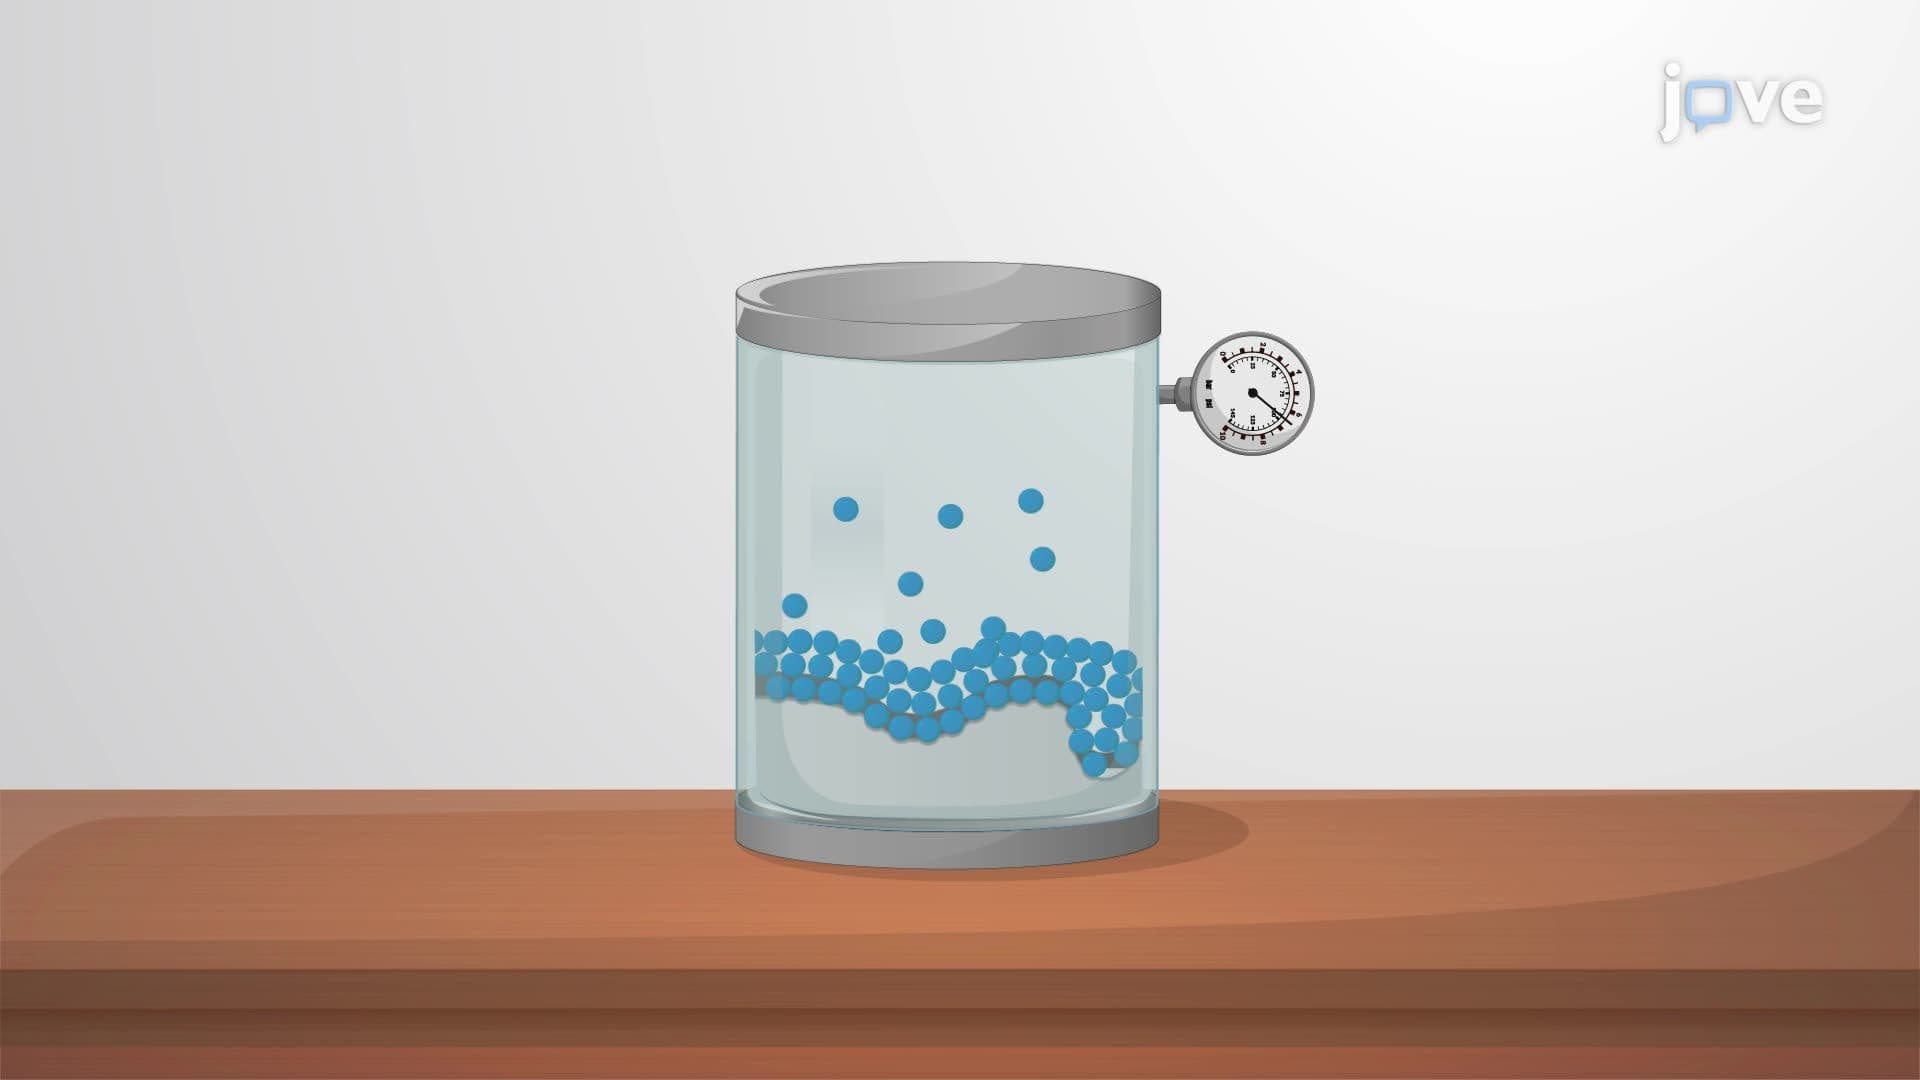
Adsorption Isotherms II
 03:51
Molecular Orbital Theory II
 02:13
Reduction of Alkenes: Catalytic Hydrogenation
Metals like palladium, platinum, and nickel are commonly used in their solid forms — fine powder on an inert surface. As these catalysts remain insoluble in the reaction mixture, they are referred to as heterogeneous catalysts.
The hydrogenation process takes place on the surface of...