Related Experiment Video
Updated: Jun 15, 2026
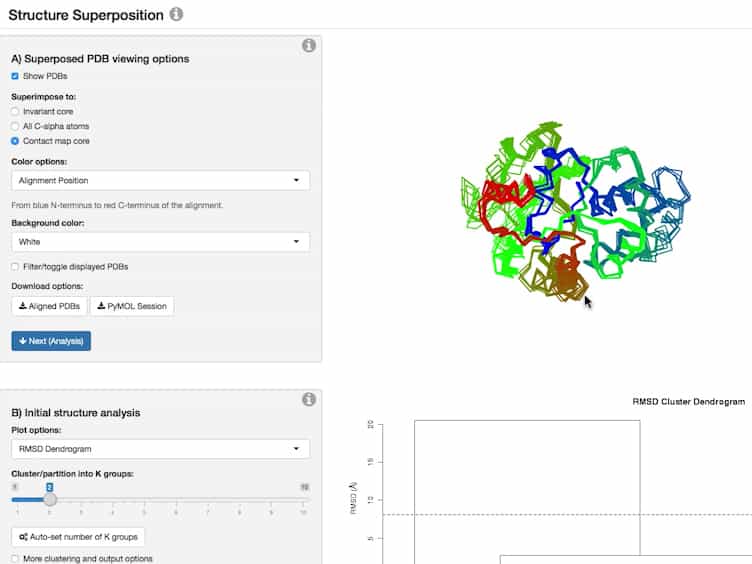

Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Atomic interaction networks in the core of protein domains and their native folds
Venkataramanan Soundararajan1, Rahul Raman, S Raguram
1Harvard-MIT Division of Health Sciences & Technology, and Department of Biological Engineering, Massachusetts Institute of Technology, Cambridge, Massachusetts, United States of America.

Abstract:
Vastly divergent sequences populate a majority of protein folds. In the quest to identify features that are conserved within protein domains belonging to the same fold, we set out to examine the entire protein universe on a fold-by-fold basis. We report that the atomic interaction network in the solvent-unexposed core of protein domains are fold-conserved, extraordinary sequence divergence notwithstanding. Further, we find that this feature, termed protein core atomic interaction network (or PCAIN) is significantly distinguishable across different folds, thus appearing to be "signature" of a domain's native fold. As part of this study, we computed the PCAINs for 8698 representative protein domains from families across the 1018 known protein folds to construct our seed database and an automated framework was developed for PCAIN-based characterization of the protein fold universe. A test set of randomly selected domains that are not in the seed database was classified with over 97% accuracy, independent of sequence divergence. As an application of this novel fold signature, a PCAIN-based scoring scheme was developed for comparative (homology-based) structure prediction, with 1-2 angstroms (mean 1.61A) C(alpha) RMSD generally observed between computed structures and reference crystal structures. Our results are consistent across the full spectrum of test domains including those from recent CASP experiments and most notably in the 'twilight' and 'midnight' zones wherein <30% and <10% target-template sequence identity prevails (mean twilight RMSD of 1.69A). We further demonstrate the utility of the PCAIN protocol to derive biological insight into protein structure-function relationships, by modeling the structure of the YopM effector novel E3 ligase (NEL) domain from plague-causative bacterium Yersinia Pestis and discussing its implications for host adaptive and innate immune modulation by the pathogen. Considering the several high-throughput, sequence-identity-independent applications demonstrated in this work, we suggest that the PCAIN is a fundamental fold feature that could be a valuable addition to the arsenal of protein modeling and analysis tools.

Related Concept Videos
 01:22
Protein Folding
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
01:22Protein Folding
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
02:26Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
 01:24
Protein Organization
The primary structure of a protein is its amino acid sequence.