Related Experiment Video
Updated: Jun 14, 2026

Study of Short Peptide Adsorption on Solution Dispersed Inorganic Nanoparticles Using Depletion Method
Published on: April 11, 2020
Adsorption of proline and glycine on the TiO2(110) surface: a density functional theory study
1Centre for Theoretical Chemistry and Physics, New Zealand Institute for Advanced Study, Massey University Albany, Private Bag 102904, North Shore City, 0745 Auckland, New Zealand. r.e.tonner@massey.ac.nz

Abstract:
The optimal adsorption modes for the amino acids glycine and proline on the ideal TiO(2)(110) surface are investigated by using density functional theory (PBE) applying periodic boundary conditions. Binding modes with anionic acid moieties bridging two titanium atoms after transferring a proton to the surface are the most stable configurations for both molecules investigated-similar to previous results for carboxylic acids. In contrast to the latter compounds, amino acids can form hydrogen bonds via the amino group towards the surface-bound proton; this provides an additional stabilisation of 15-20 kJ mol(-1). Zwitterionic binding modes are less stable (by 10-20 kJ mol(-1)) and are less important for proline. Neutral modes are energetically even less favourable. Calculations of vibrational frequencies and core-level shifts complement the adsorption study and provide guidance for future experimental investigations. Control of the computational parameters is crucial for the derivation of accurate results. The layout and thickness of the slab model used are also shown to be decisive factors. Calculations with a different GGA-functional (PW91) provide very similar relative energies, although the absolute energies change by about 20 kJ mol(-1). Results derived with the hybrid functional PBE0 show an even greater stabilisation of the anionic binding modes with respect to the zwitterionic modes. A previously observed discrepancy between experimental and theoretical results for glycine could be solved, although the experimentally proposed free rotation of the C-C bond could not be reproduced.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 16:24
Controlling the Size, Shape and Stability of Supramolecular Polymers in Water
Published on: August 2, 2012
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:15
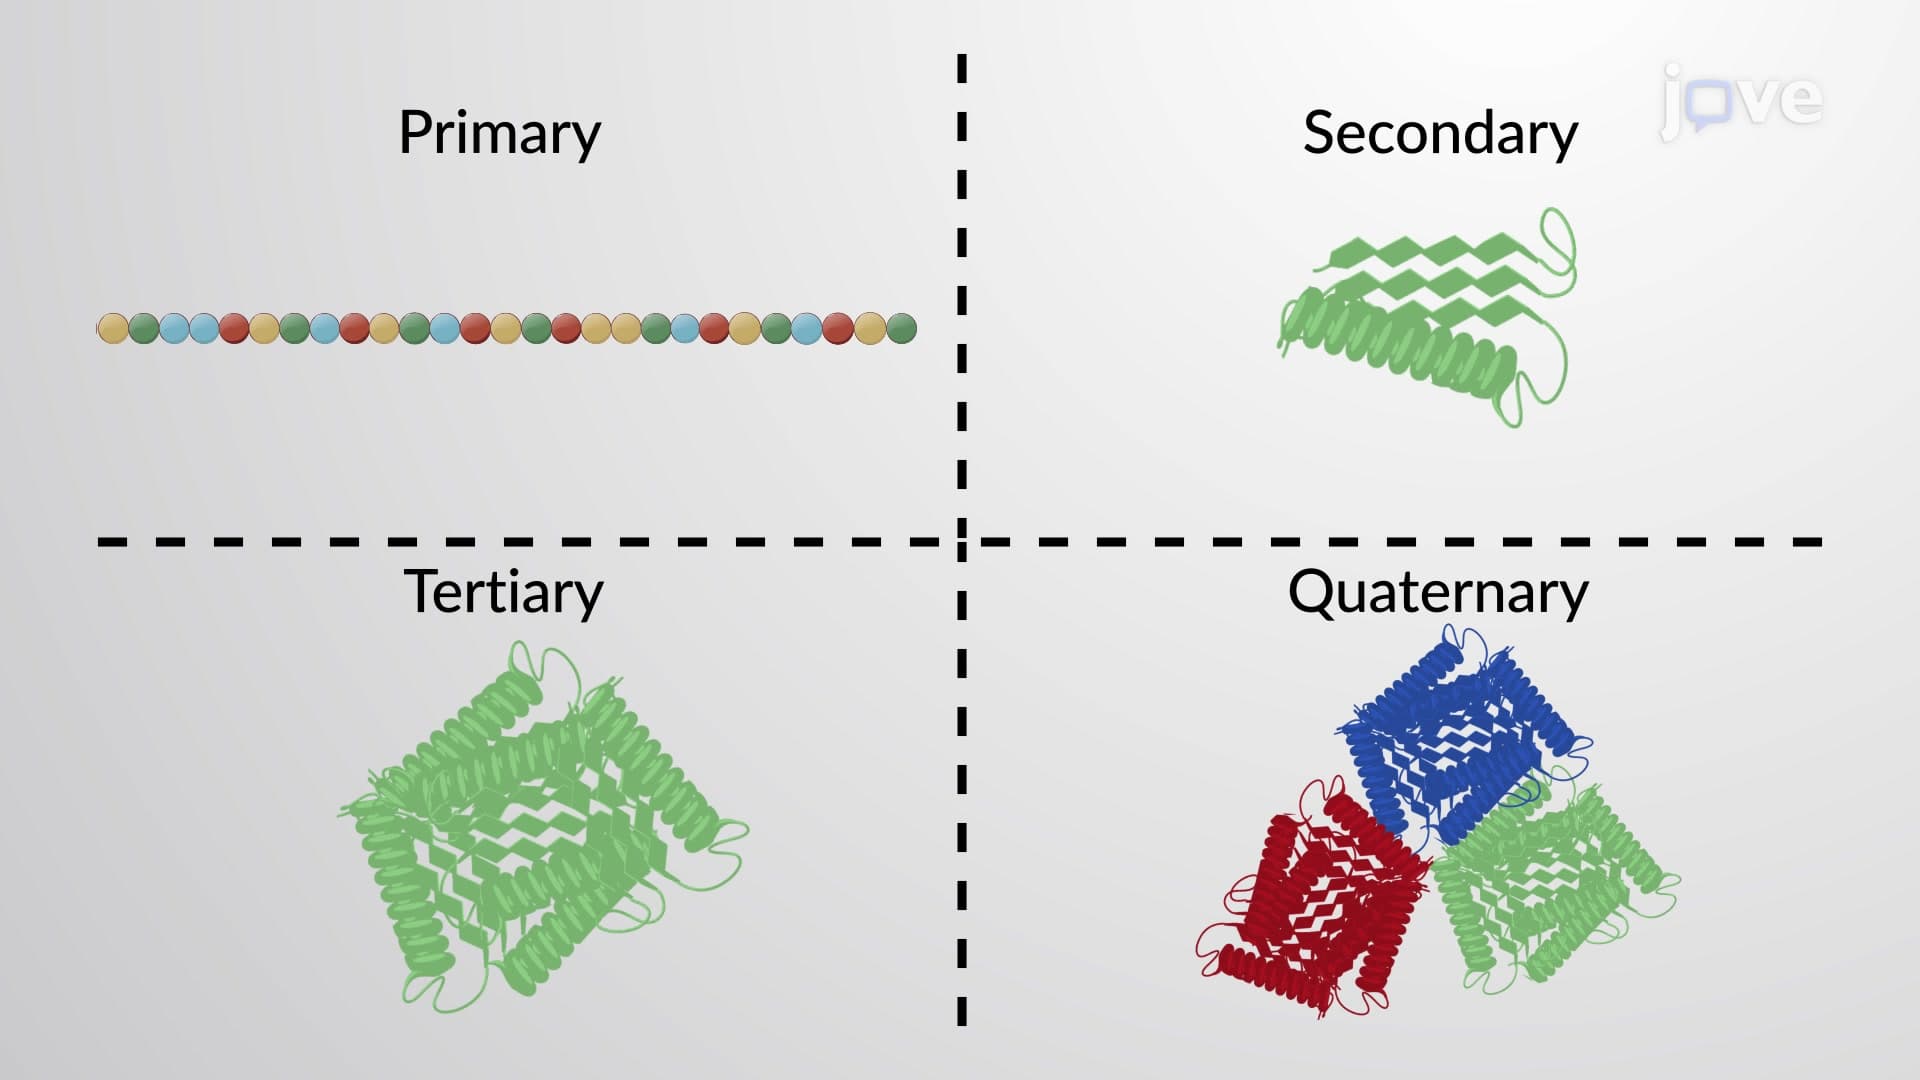
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme can...
 02:35
Noncovalent Attractions in Biomolecules
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
 01:28
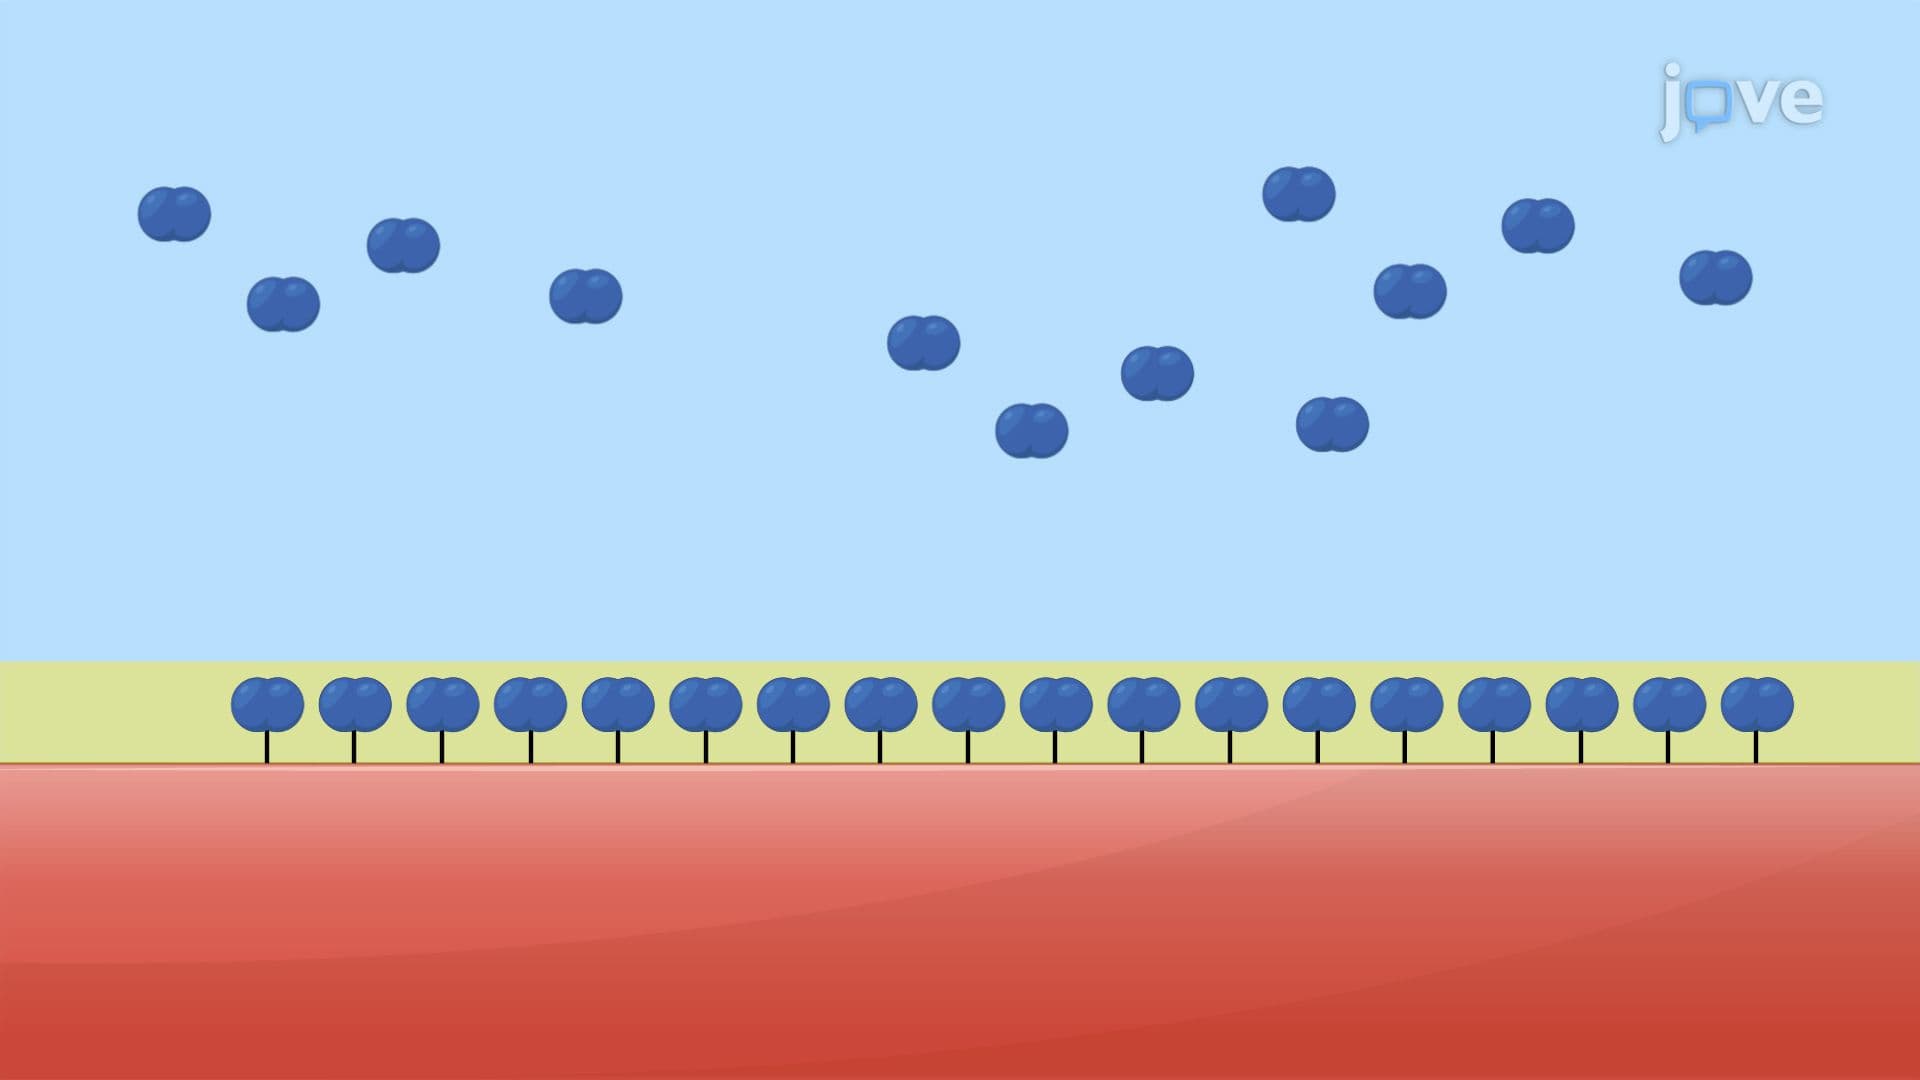
Adsorption of Gases on Solids
 02:04
Protein-protein Interfaces