Related Experiment Video
Updated: Jun 14, 2026

Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Toward the synthetic control of the HOMO-LUMO gap in binuclear systems: insights from density functional calculations
Alvaro Muñoz-Castro1, Desmond Mac-Leod Carey, Cesar Morales-Verdejo
1Departamento de Ciencias Quimicas, Universidad Andres Bello, Republica 275, Santiago, Chile.

Abstract:
Computational methods based on density functional theory have been applied to address the design of tailored HOMO-LUMO gap bimetallic complexes. We focus our attention on the [Cp*Fe-(L)-FeCp*] system, where two ferrocenyl units are linked through the dianion of fused ring ligands such as pentalene, s-indacene, dicyclopenta-[b,g]-naphthalene, dicyclopenta-[b,i]-anthracene and dicyclopenta-[b,l]-tetracene. Our DFT calculations on the title organometallic complexes suggest a controlled decrease in the HOMO-LUMO gap, which is desirable for studies on electron-transfer phenomena, as well as the design potential devices for molecular electronic purposes.
Related Concept Videos
 03:35
Hybridization of Atomic Orbitals II
 03:24
Hybridization of Atomic Orbitals I
 03:51
Molecular Orbital Theory II
 02:35
Molecular Orbital Theory I
 01:12
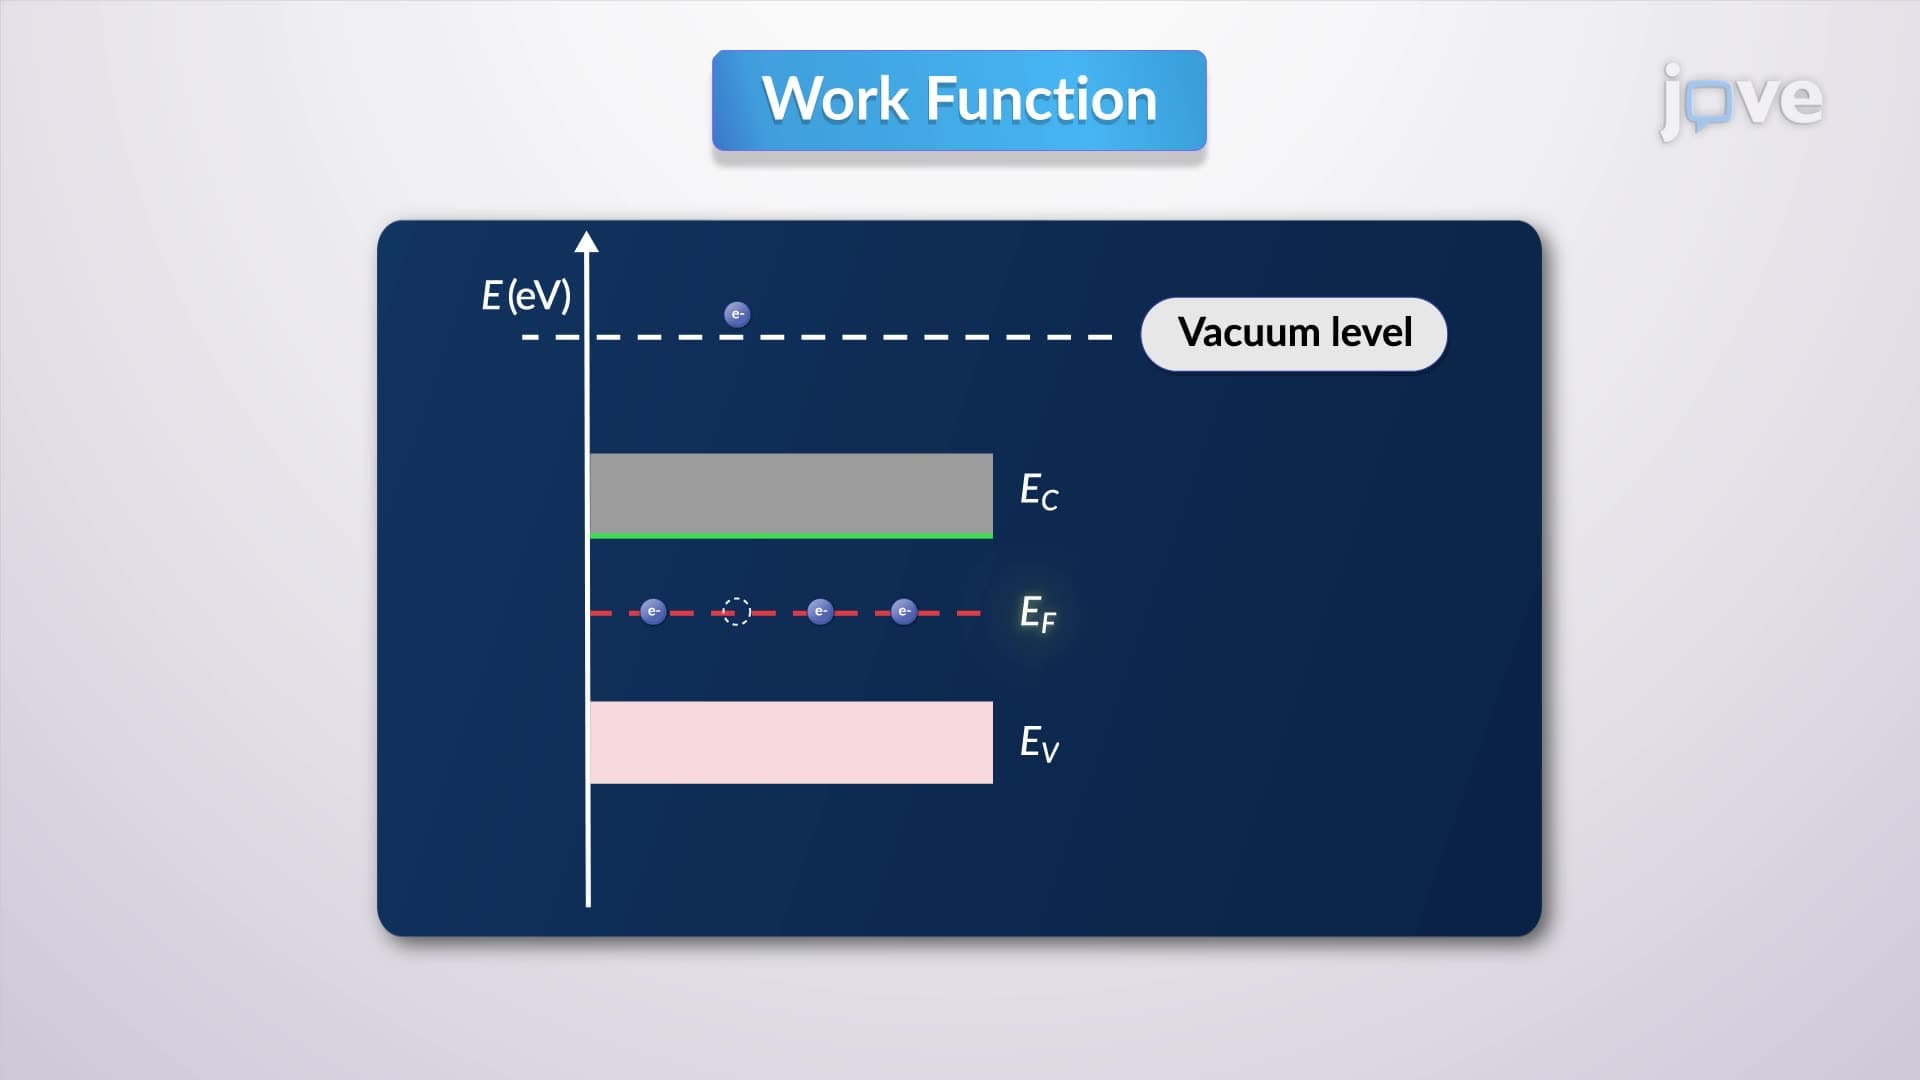
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 02:54
Trends in Lattice Energy: Ion Size and Charge