Related Experiment Video
Updated: Jun 13, 2026

From a 2DE-Gel Spot to Protein Function: Lesson Learned From HS1 in Chronic Lymphocytic Leukemia
Published on: October 19, 2014
Primary and secondary hemophagocytic lymphohistiocytosis: clinical features, pathogenesis and therapy
1Division of Haematology/Oncology, The Hospital for Sick Children, 555 University Avenue, Toronto, ON M5G 1X8, Canada. sumit.gupta@sickkids.ca

Hemophagocytic lymphohistiocytosis (HLH) is a potentially fatal hyperinflammatory syndrome with prolonged high fever, hepatosplenomegaly and characteristic laboratory findings. HLH may be inherited (primary) or may be secondary to any severe infection, malignancy or rheumatologic condition. The last several years have witnessed an explosion in our understanding of HLH. Of the inherited causes for which the underlying genetic cause is known, most involve abnormalities of proteins important in the exocytosis cytolytic pathway, whereby perforin and granzymes are delivered to a target cell to induce apoptosis. The exact mechanisms underlying this process remain unclear. However, when a known genetic defect is not present, the diagnosis of HLH is still made on a constellation of clinical features and good clinical judgment. Rapid diagnosis is crucial, as early therapy with immunosuppressive agents and/or proapoptotic chemotherapy can be life-saving. This article examines recent advances in our understanding of the pathophysiology, clinical features, diagnosis, etiology and treatment of HLH, as well as the challenges that lie ahead.
Hemophagocytic lymphohistiocytosis (HLH) is a potentially fatal hyperinflammatory syndrome with prolonged high fever, hepatosplenomegaly and characteristic laboratory findings. HLH may be inherited (primary) or may be secondary to any severe infection, malignancy or rheumatologic condition. The last several years have witnessed an explosion in our understanding of HLH. Of the inherited causes for which the underlying genetic cause is known, most involve abnormalities of proteins important in the exocytosis cytolytic pathway, whereby perforin and granzymes are delivered to a target cell to induce apoptosis. The exact mechanisms underlying this process remain unclear. However, when a known genetic defect is not present, the diagnosis of HLH is still made on a constellation of clinical features and good clinical judgment. Rapid diagnosis is crucial, as early therapy with immunosuppressive agents and/or proapoptotic chemotherapy can be life-saving. This article examines recent advances in our understanding of the pathophysiology, clinical features, diagnosis, etiology and treatment of HLH, as well as the challenges that lie ahead.
Related Concept Videos
 01:16
Primary Lymphoid Organs
The red bone marrow is a soft, spongy tissue nestled in the interior of long bones such as the humerus and femur. It is the site...
 01:23
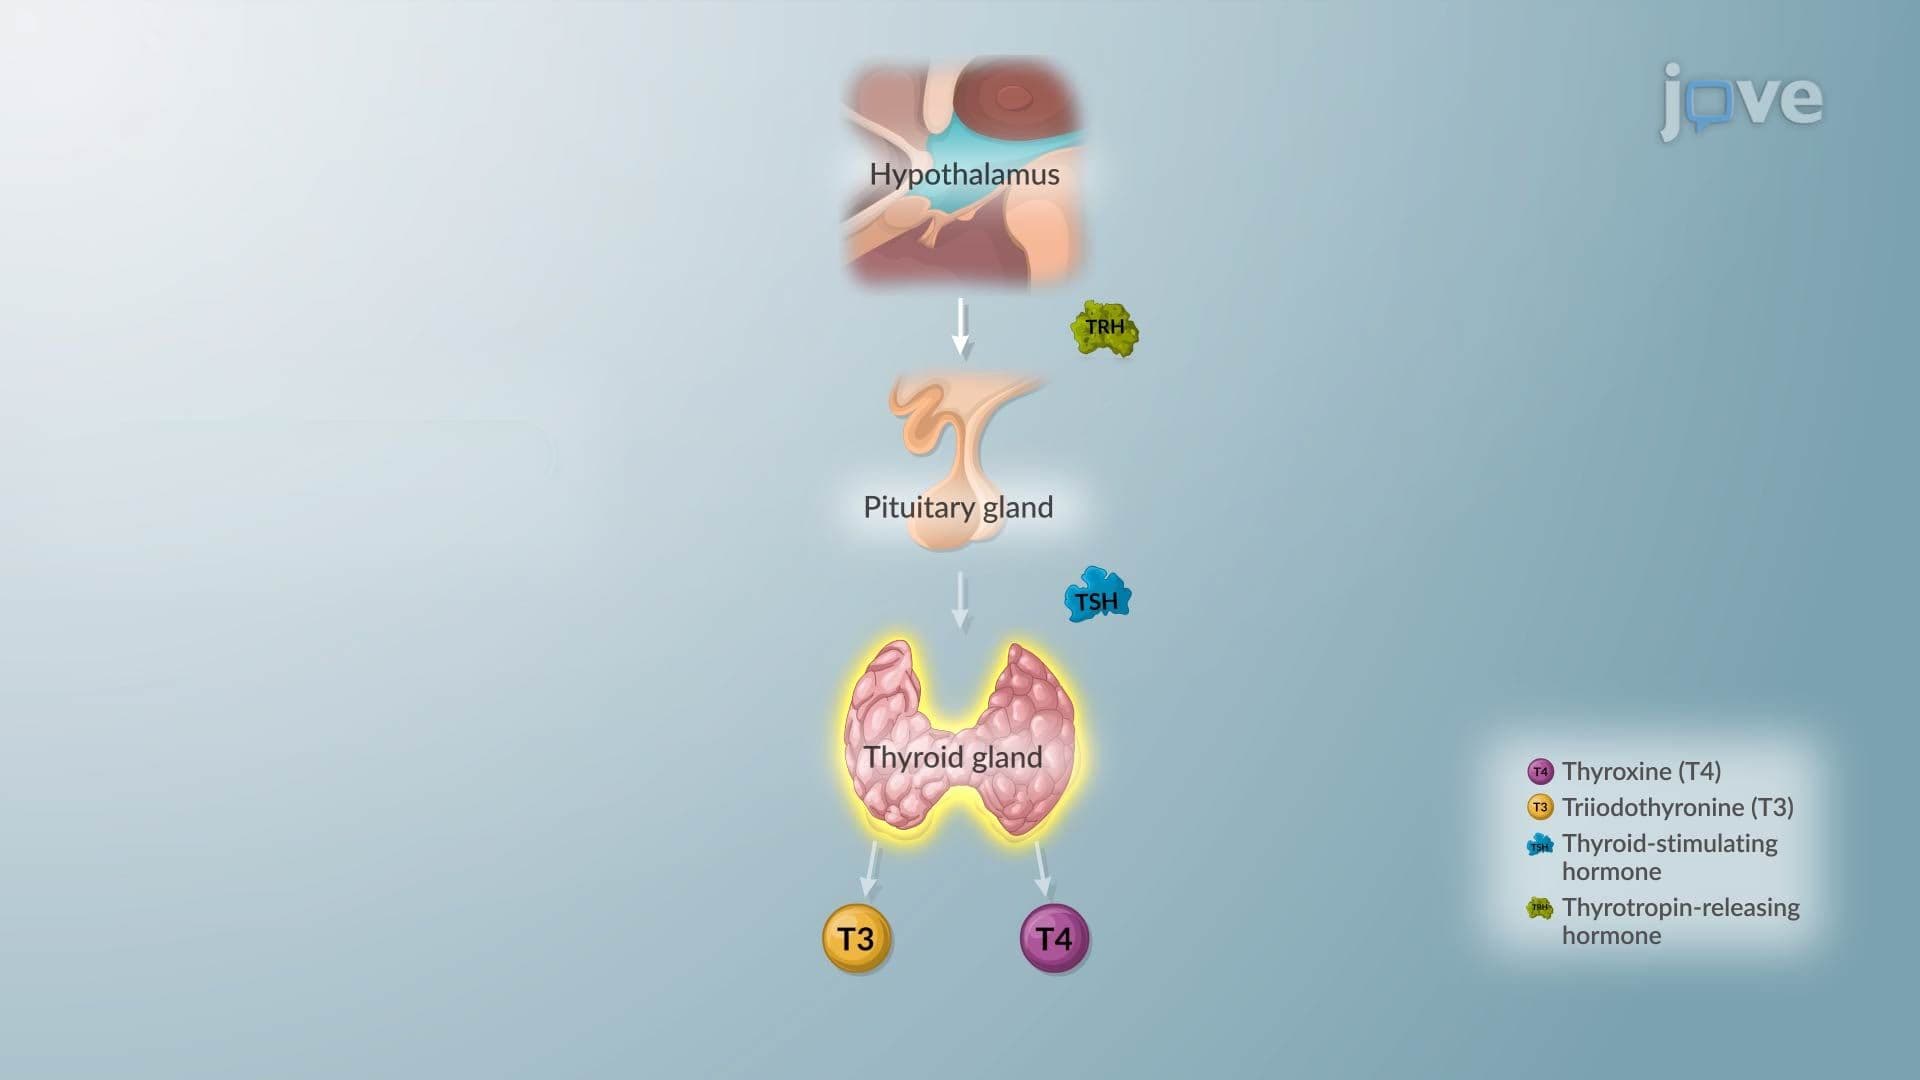
Hypothyroidism II: Pathophysiology
 01:01
Regulation of Hematopoietic Stem Cells
 01:15
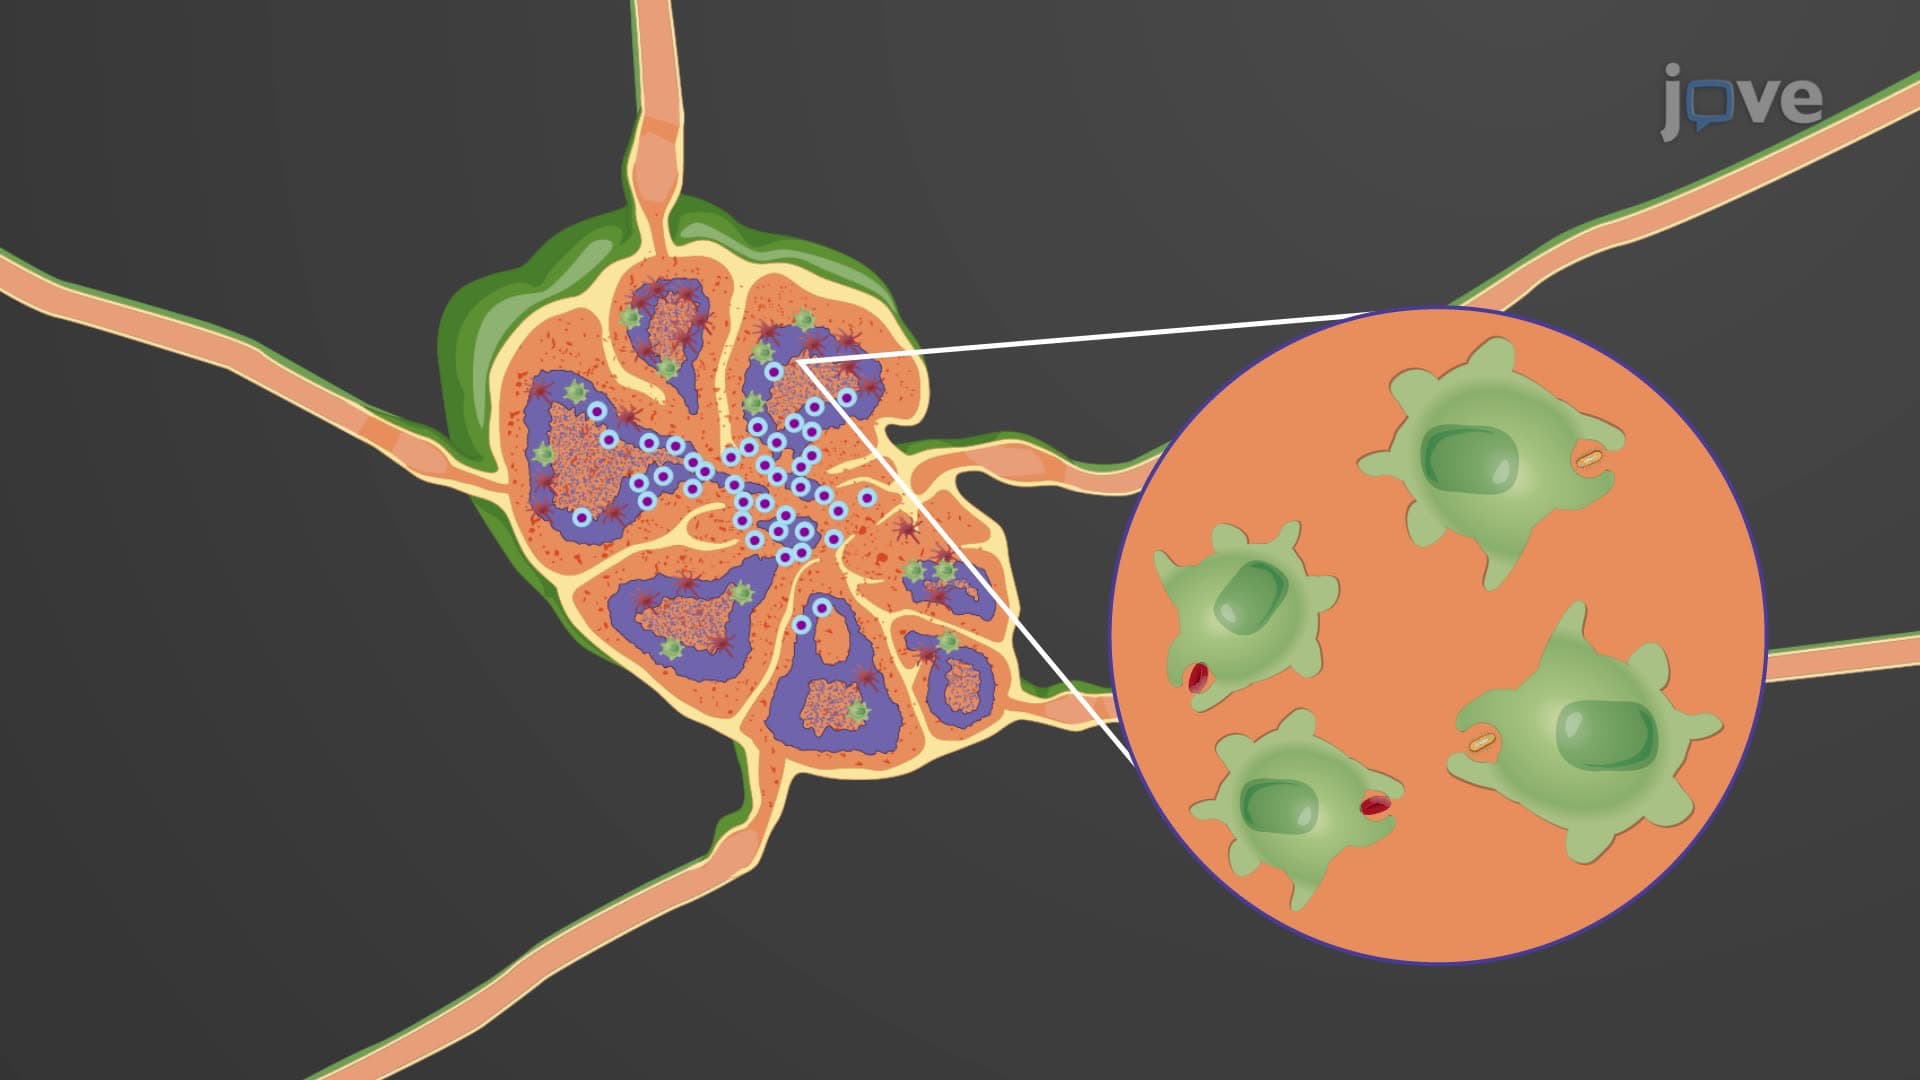
Secondary Lymphoid Organs
The spleen is a vital organ in the lymphatic system, nestled in the upper left side of the abdomen. It is composed of two primary regions: the red pulp and the white pulp, each having distinct functions. The red pulp performs a significant role in blood filtration. It efficiently purges the blood of old or damaged red blood cells and...
 01:24
Cirrhosis II: Pathophysiology
 01:01
Hypersensitivity Reactions: Cytolytic Reactions