Related Experiment Video
Updated: Jun 13, 2026
Surfactant Depletion Combined with Injurious Ventilation Results in a Reproducible Model of the Acute Respiratory Distress Syndrome (ARDS)
Published on: April 7, 2021
Surfactant protein A2 mutations associated with pulmonary fibrosis lead to protein instability and endoplasmic
Meenakshi Maitra1, Yongyu Wang, Robert D Gerard
1Eugene McDermott Center for Human Growth and Development, Dallas, Texas 75390, USA.

Abstract:
Rare heterozygous mutations in the gene encoding surfactant protein A2 (SP-A2, SFTPA2) are associated with adult-onset pulmonary fibrosis and adenocarcinoma of the lung. We have previously shown that two recombinant SP-A2 mutant proteins (G231V and F198S) remain within the endoplasmic reticulum (ER) of A549 cells and are not secreted into the culture medium. The pathogenic mechanism of the mutant proteins is unknown. Here we analyze all common and rare variants of the surfactant protein A2, SP-A2, in both A549 cells and in primary type II alveolar epithelial cells. We show that, in contrast with all other SP-A2 variants, the mutant proteins are not secreted into the medium with wild-type SP-A isoforms, form fewer intracellular dimer and trimer oligomers, are partially insoluble in 0.5% Nonidet P-40 lysates of transfected A549 cells, and demonstrate greater protein instability in chymotrypsin proteolytic digestions. Both the G231V and F198S mutant SP-A2 proteins are destroyed via the ER-association degradation pathway. Expression of the mutant proteins increases the transcription of a BiP-reporter construct, expression of BiP protein, and production of an ER stress-induced XBP-1 spliced product. Human bronchoalveolar wash samples from individuals who are heterozygous for the G231V mutation have similar levels of total SP-A as normal family members, which suggests that the mechanism of disease does not involve an overt lack of secreted SP-A but instead involves an increase in ER stress of resident type II alveolar epithelial cells.
Related Concept Videos
 01:05
Breathing
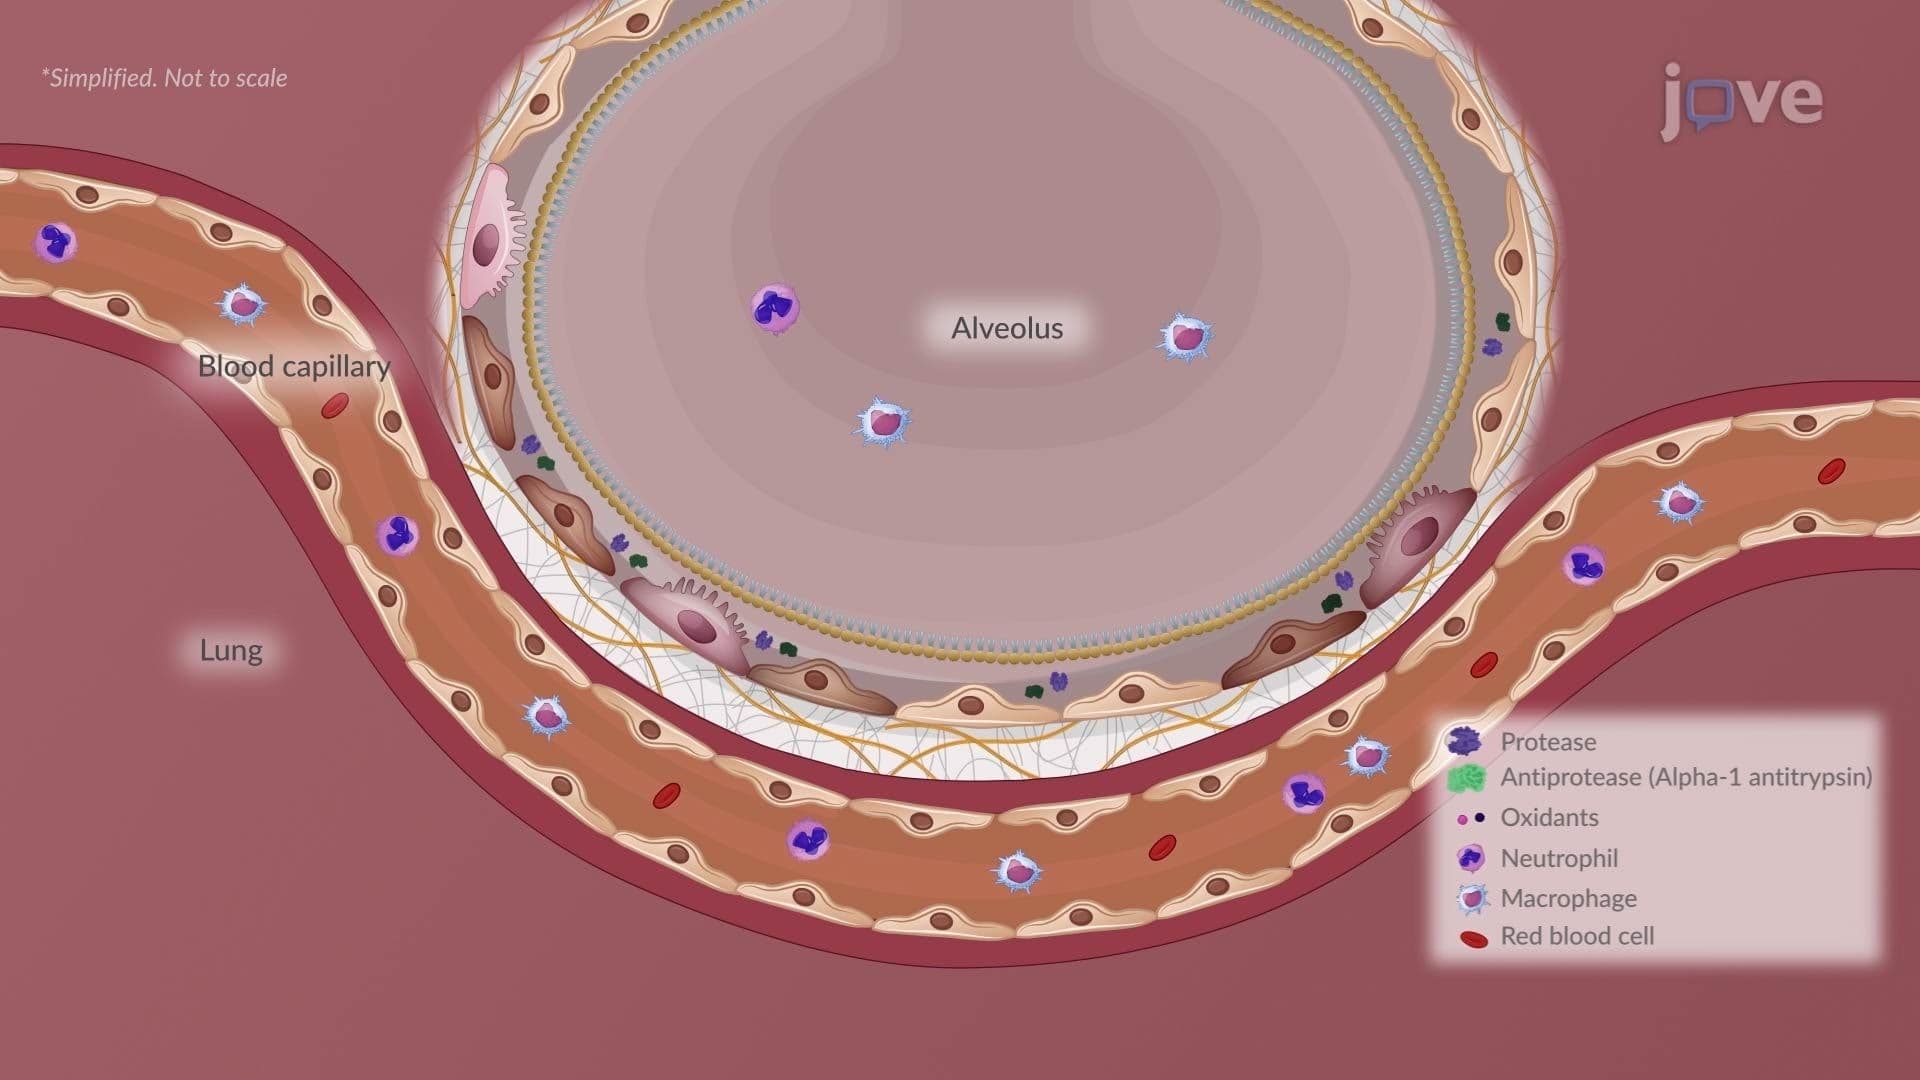 01:23
Chronic Obstructive Pulmonary Disease II: Emphysema
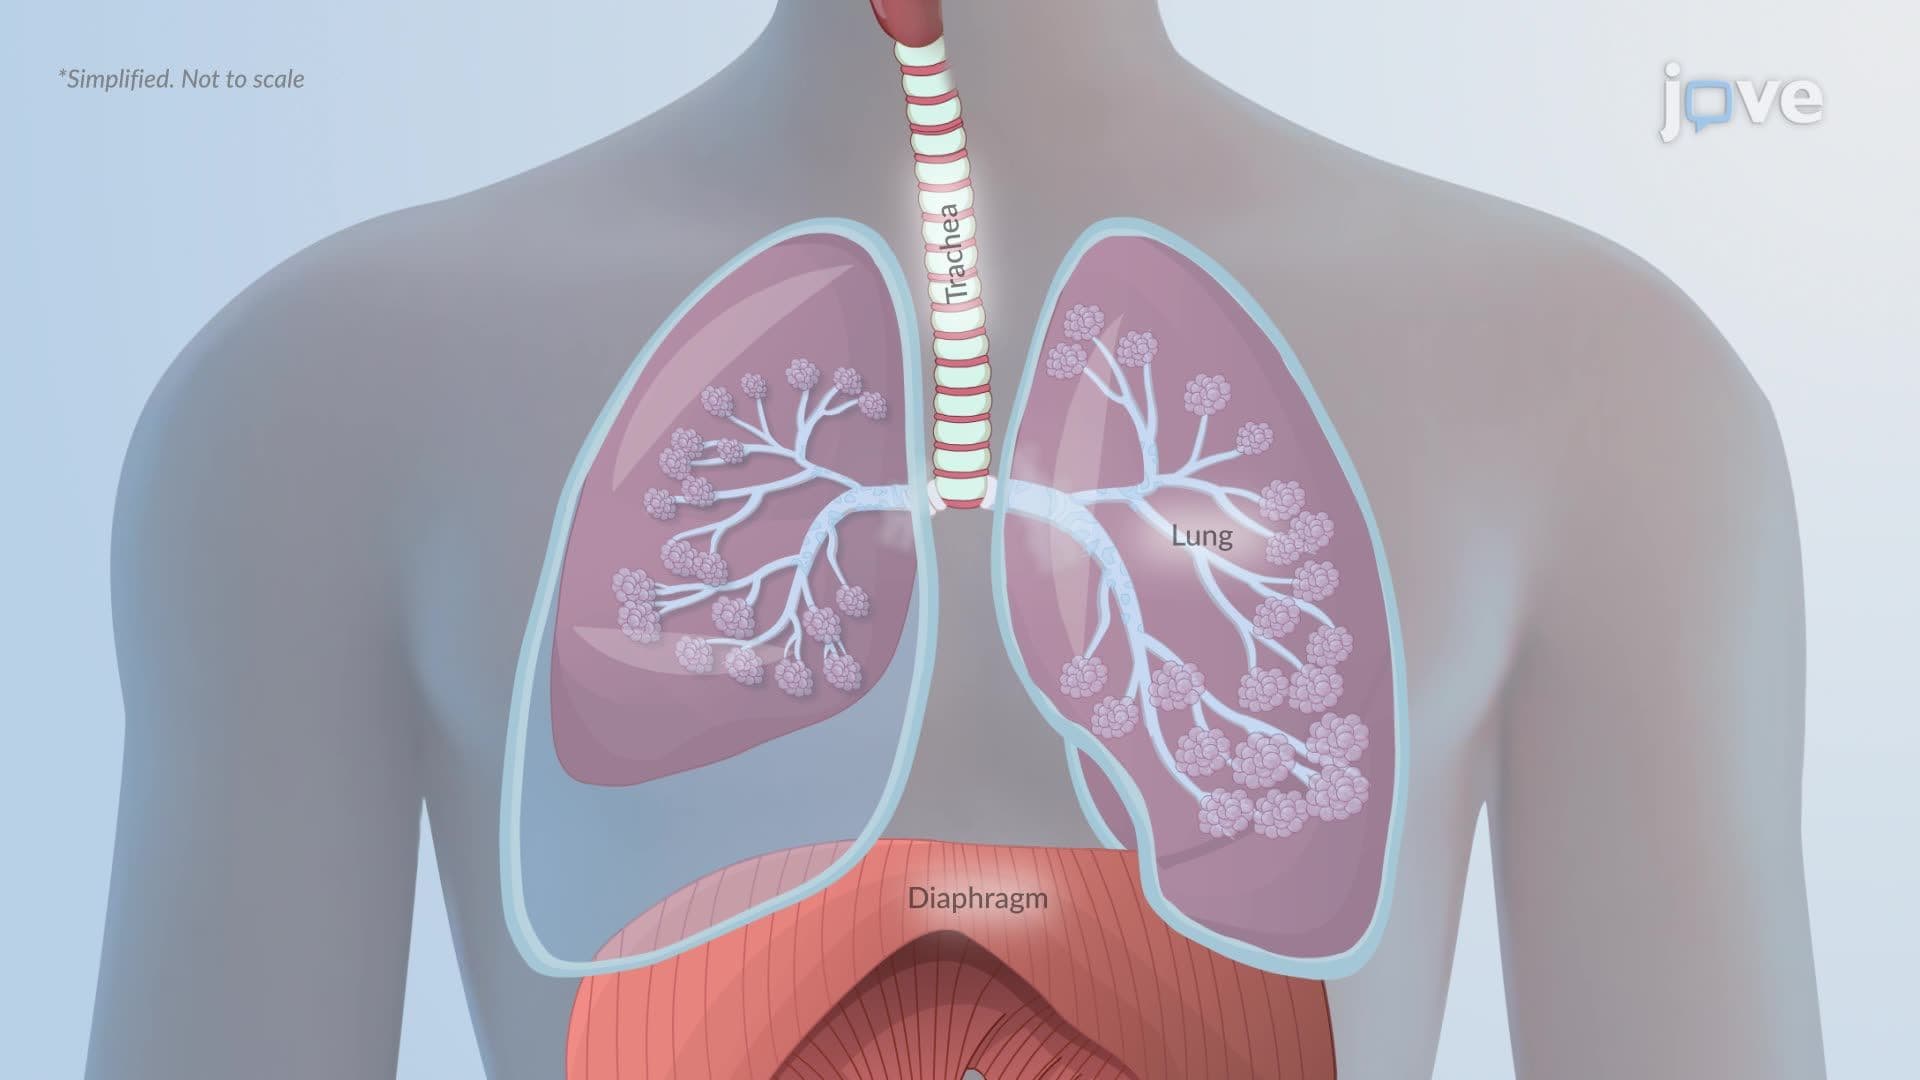 01:10
Atelectasis II: Pathophysiology
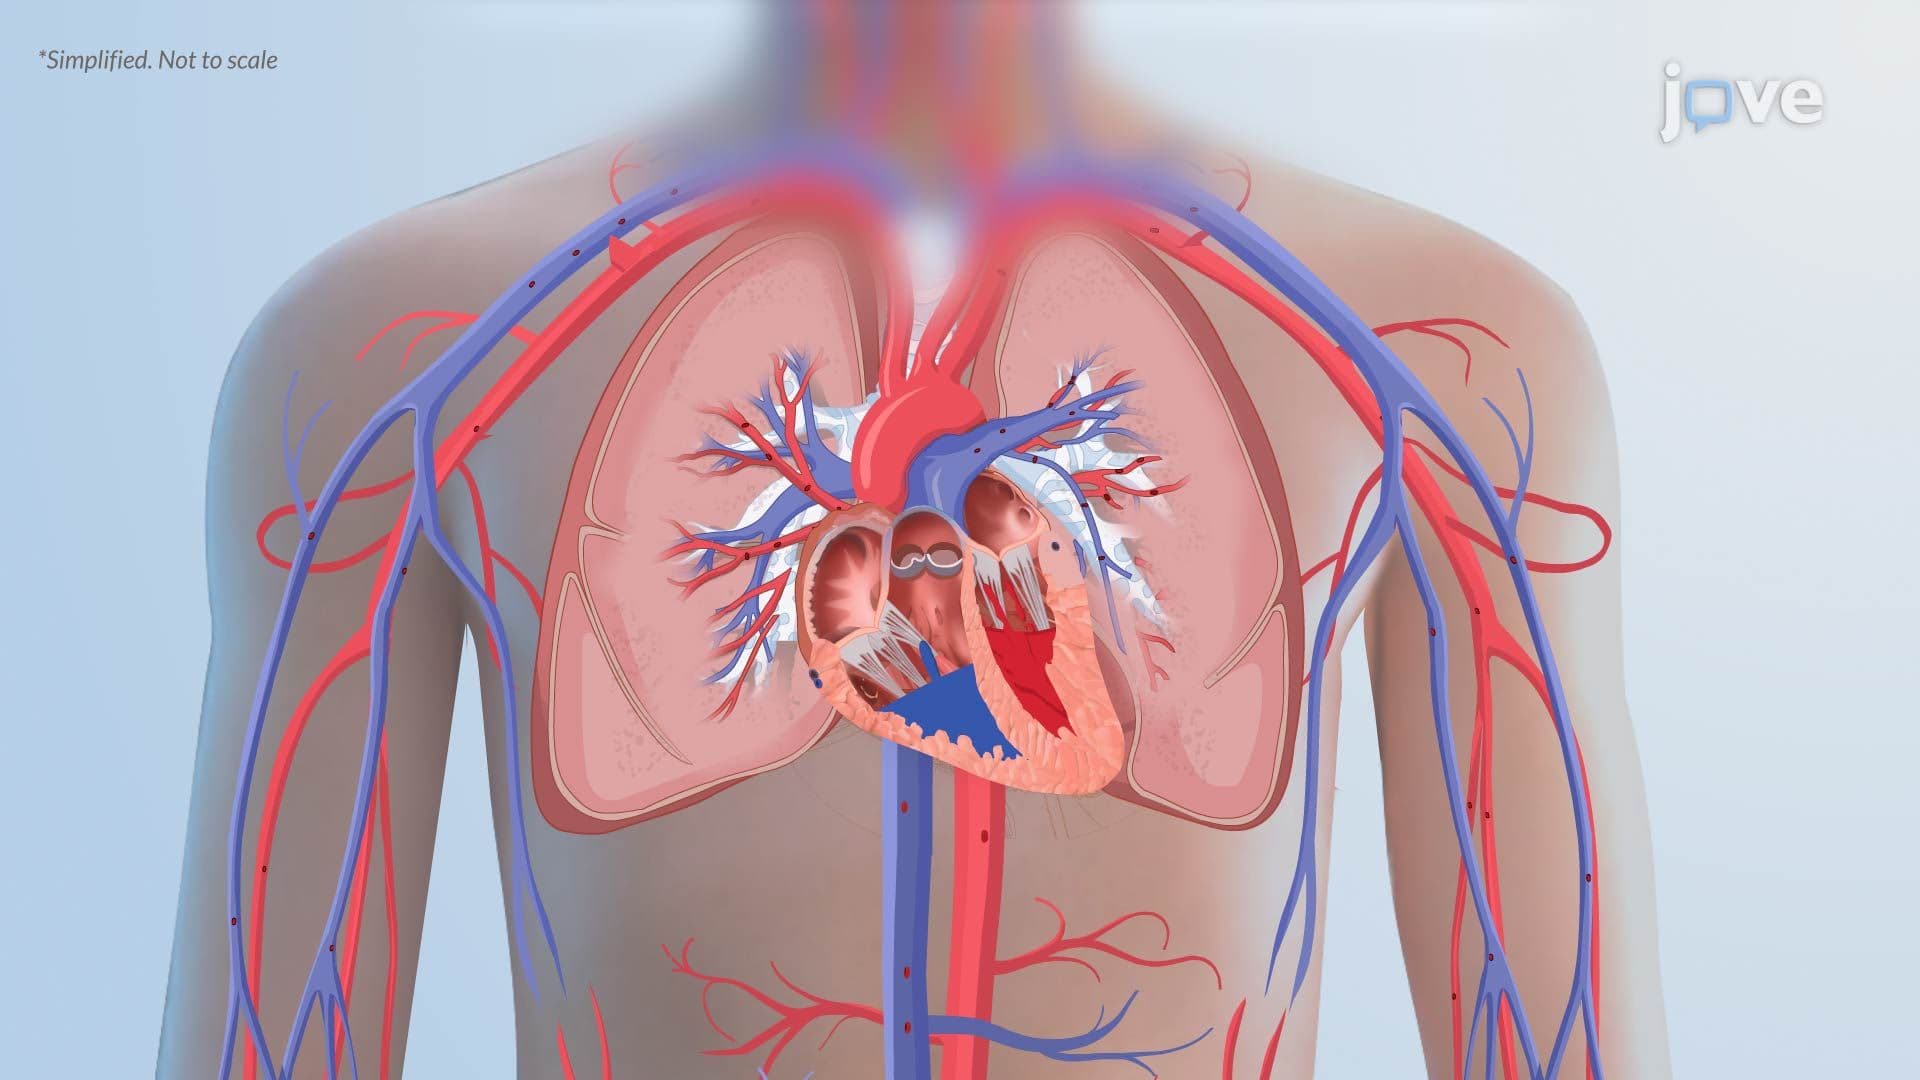 01:18
Pulmonary Edema II: Pathophysiology
 01:37
The Unfolded Protein Response
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...