Related Experiment Video
Updated: Jun 12, 2026

Microwave-assisted Intramolecular Dehydrogenative Diels-Alder Reactions for the Synthesis of Functionalized Naphthalenes/Solvatochromic Dyes
Published on: April 1, 2013
[Molecular geometries and theoretical electronic spectra of four 1,8-naphthyridine derivatives]
Shao-Ming Chi1, Li Li, Yong Chen
1Technical Institute of Physics and Chemistry, Chinese Academy of Sciences, Beijing 100190, China. chishaoming@mail.ipc.ac.cn

Abstract:
The molecular geometries of four 2,4-dimethyl-7-amino-1,8-naphthyridine derivatives were optimized with B3LYP/6-31G(d) method. The energies of their frontier molecular orbitals and the molecular structures were investigated theoretically. The theoretical electronic spectra were calculated with TD-DFT in gas phase, PCM-TD-B3LYP/6-31 + G(d) and semiempirical ZINDO in CH2 Cl2 solution. The influences of solvent model and calculation methods on the electronic absorption spectra were also probed. The calculated results show that delocalized pi bonds exist in the four 1,8-naphthyridine derivatives, and their energy gaps (deltaE) between HOMO and LUMO are relatively small. The variation in their deltaE values gives a consistent trend with that of their electronic absorption with lambda(max). Theoretical spectra achieved prove that their absorptions are red-shifted when the delocalization of pi electrons is enhanced or the capability to donate electron by a substituted group is increased. The maximum absorption peaks of the four derivatives originate from pi (HOMO) --> pi * (LUMO) transition. The spectra calculated at the PCM-B3LYP/6-31 + G(d) level have little difference from experimental results: the differences in wavelength are 2.6, 10.3, 5.3 and 6.9 nm, whereas those in energies are 0.03, 0.09, 0.04 and 0.08 eV, respectively. The obtained results suggest that electronic spectra calculated by TD-DFT on the bases of geometries optimized with B3LYP/6-31(d) are in agreement with experimental ones, and can account for the different spectroscopic properties of the four 1,8-naphthyridine derivatives.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 10:51
The Synthesis, Characterization and Reactivity of a Series of Ruthenium N-triphosPh Complexes
Published on: April 10, 2015
Related Concept Videos
 01:37
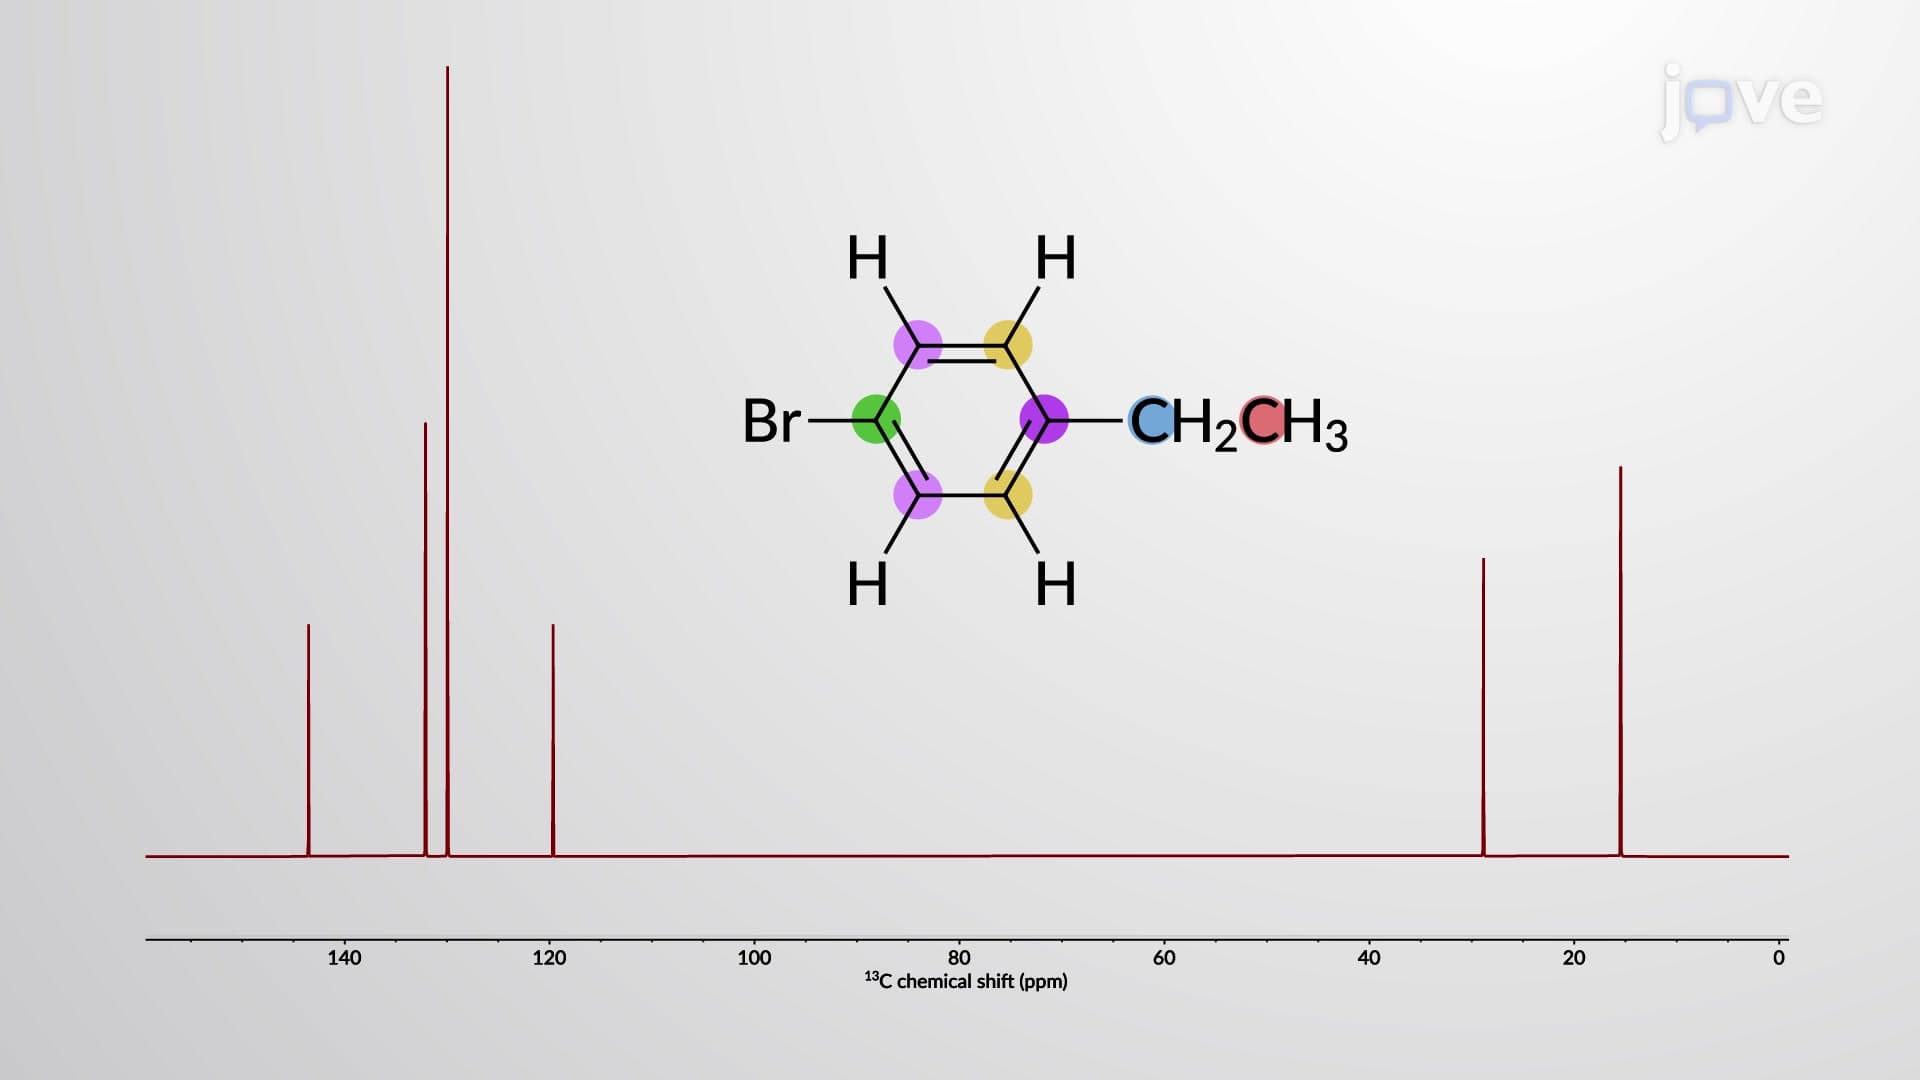
NMR Spectroscopy of Benzene Derivatives
 01:13
Five-Membered Heterocyclic Aromatic Compounds: Overview
 01:18
Frost Circles for Different Conjugated Systems
 01:20
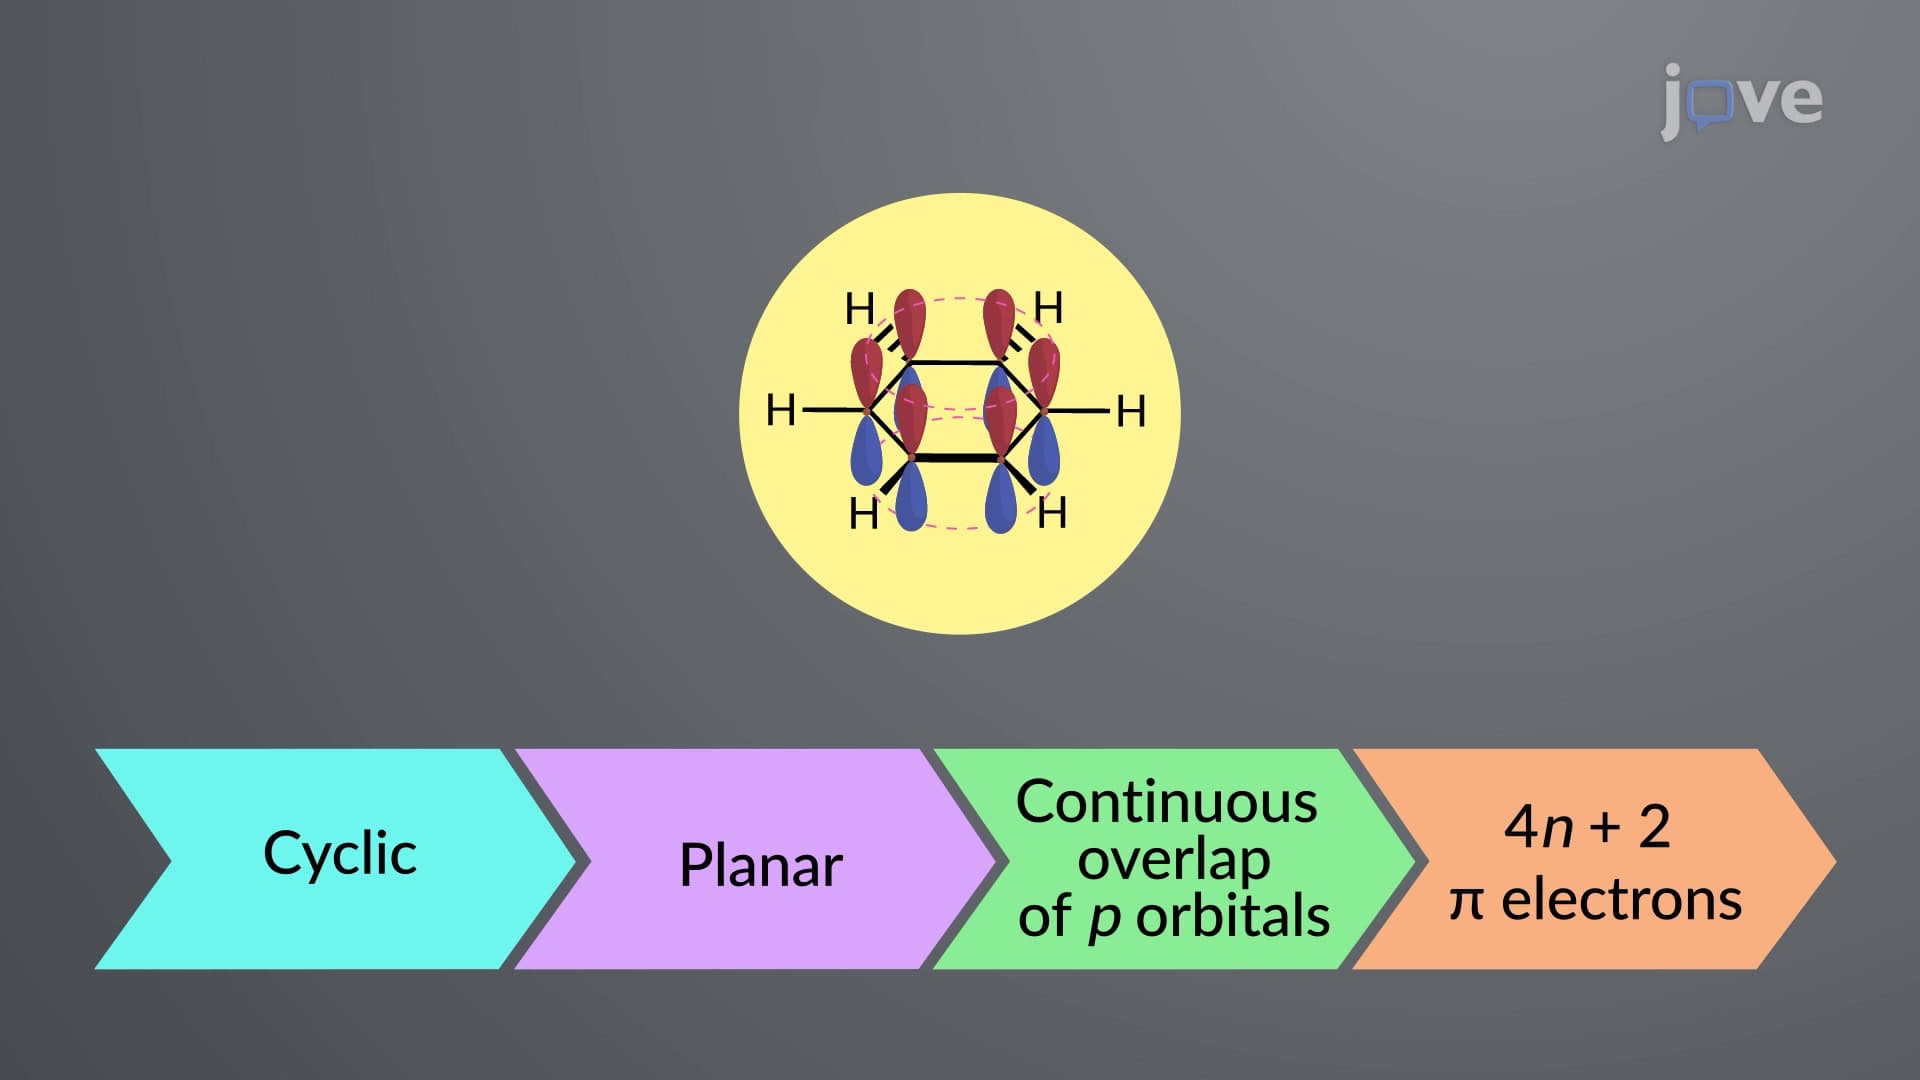
Criteria for Aromaticity and the Hückel 4n + 2 Rule
For the first time, Eric Hückel, a German chemical physicist, derived a set of structural features for a compound to be classified as aromatic. This is now known as Hückel’s rule or the 4n + 2 rule.
 02:37
VSEPR Theory
 04:01
VSEPR Theory and the Effect of Lone Pairs