Related Experiment Video
Updated: Sep 30, 2026

Efficient Construction of Drug-like Bispirocyclic Scaffolds Via Organocatalytic Cycloadditions of α-Imino γ-Lactones and Alkylidene Pyrazolones
Published on: February 7, 2019
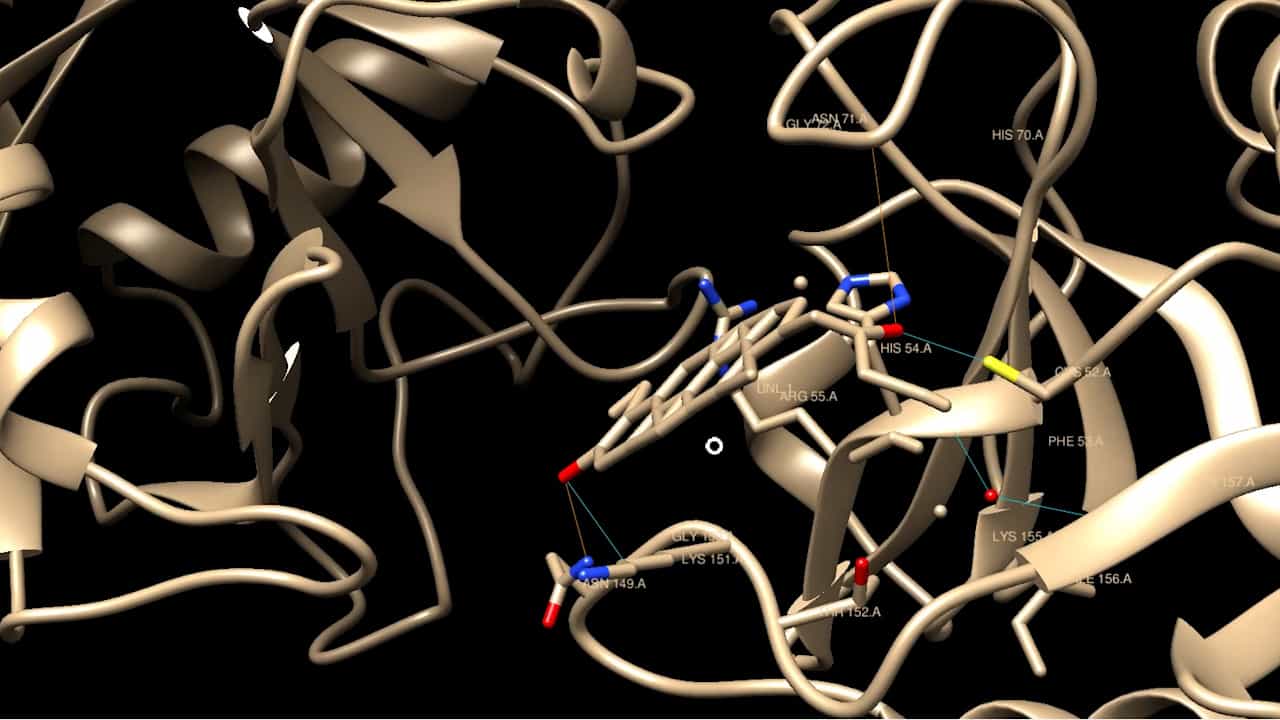
Optimization of a bis-imine scaffold as a glyoxalase I inhibitor: synthesis, SAR, and docking studies
Buthina A Al-Oudat1, Farah B Al-Omari2, Suaad A Audat3
1Department of Medicinal Chemistry and Pharmacognosy, Faculty of Pharmacy, University of Science and Technology, P.O. Box 3030, Irbid, 22110, Jordan. baoudat@just.edu.jo.

Abstract:
Glyoxalase I (Glo-I) enzyme is a validated anticancer target due to its role in methylglyoxal detoxification and its association with tumor progression and drug resistance. Inhibition of Glo-I promotes the accumulation of toxic metabolites and can trigger apoptosis in cancer cells. Building on our previously reported bis-imine Glo-I inhibitor N3 (IC50 = 0.89 ± 0.06 µM), which contains two pyrogallol moieties linked via para-positioned imines on a central benzene linker, the present study introduces a focused scaffold-optimization strategy aimed at defining the structural elements required for stronger Glo-I inhibition. Structural simplification of N3, through removal of the central benzene linker, generated compound 8a, a simplified bis-imine analogue with two-fold improved potency (IC50 = 0.46 ± 0.10 µM). This enhancement was attributed to preserved hydrogen bonding capacity and improved Zn²⁺ chelation within Glo-I active site. A comprehensive structure-activity relationship (SAR) investigation of compound 8a and its analogues demonstrated that the number, position, and free phenolic character of the hydroxyl groups are critical determinants of activity, mainly by supporting hydrogen bonding and bidentate Zn²⁺ coordination. Masking the hydroxyl groups abolished activity, whereas selected alternative chelating groups preserved moderate inhibition, highlighting the importance and flexibility of metal-binding interactions. Importantly, replacement of the bis-imine with amide-imine linker, or reduction to mono-imine analogues, reduced potency, confirming that the bis-imine linker is a key pharmacophoric feature for optimal Glo-I inhibition. Molecular docking further rationalized these findings by identifying the essential interactions with Zn²⁺, polar residues at the active site entrance, and the hydrophobic pocket. Overall, this study provides a clear innovative-driven SAR model for bis-imine-based Glo-I inhibitors and identified compound 8a as an optimized lead for the future design of anticancer candidates targeting Glo-I enzyme.Clinical trial number: not applicable. The image presents a chemical structure at the top, connected by arrows to three groups of related chemical structures labeled 8a-p, 11a-f, and 14a-c. The top structure includes two benzene rings each with hydroxyl groups, linked by a diazene group and a phenyl ring. The three groups below show variations of this structure with different substituents labeled as R. The top right includes a molecular surface model showing a ligand bound within a protein pocket, illustrating the interaction between the molecule and the protein.