Related Experiment Video
Updated: Jun 12, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Ab-initio-based global double many-body expansion potential energy surface for the electronic ground state of the
1Departamento de Química, Universidade de Coimbra 3004-535 Coimbra, Portugal.

Abstract:
We report a single-sheeted global double many-body expansion potential energy surface for the ground electronic state of NH(3) that has been calibrated from newly calculated ab initio energies. It employs realistic double many-body expansion functions previously reported from accurate ab initio data for the triatomic fragments plus four-body energy terms that have been calibrated from multireference configuration calculations carried out in the present work for the title system using the full valence complete active space wave function as reference and a triple-zeta basis set of the correlation consistent type. It provides therefore a reliable description of both short-range and long-range forces while dissociating correctly at all asymptotic limits. The major attributes of the NH(3) double many-body expansion potential energy surface have been characterized and found to be in good agreement both with the calculated ones from the raw ab initio energies and theoretical results available in the literature. It can then be recommended both as a reliable functional form on which dynamics calculations can be performed and as a model for improvement that is open to refinement through further accurate ab initio calculations, vibrational calculations, or both.

Related Concept Videos
 03:25
Free Energy Changes for Nonstandard States
 04:01
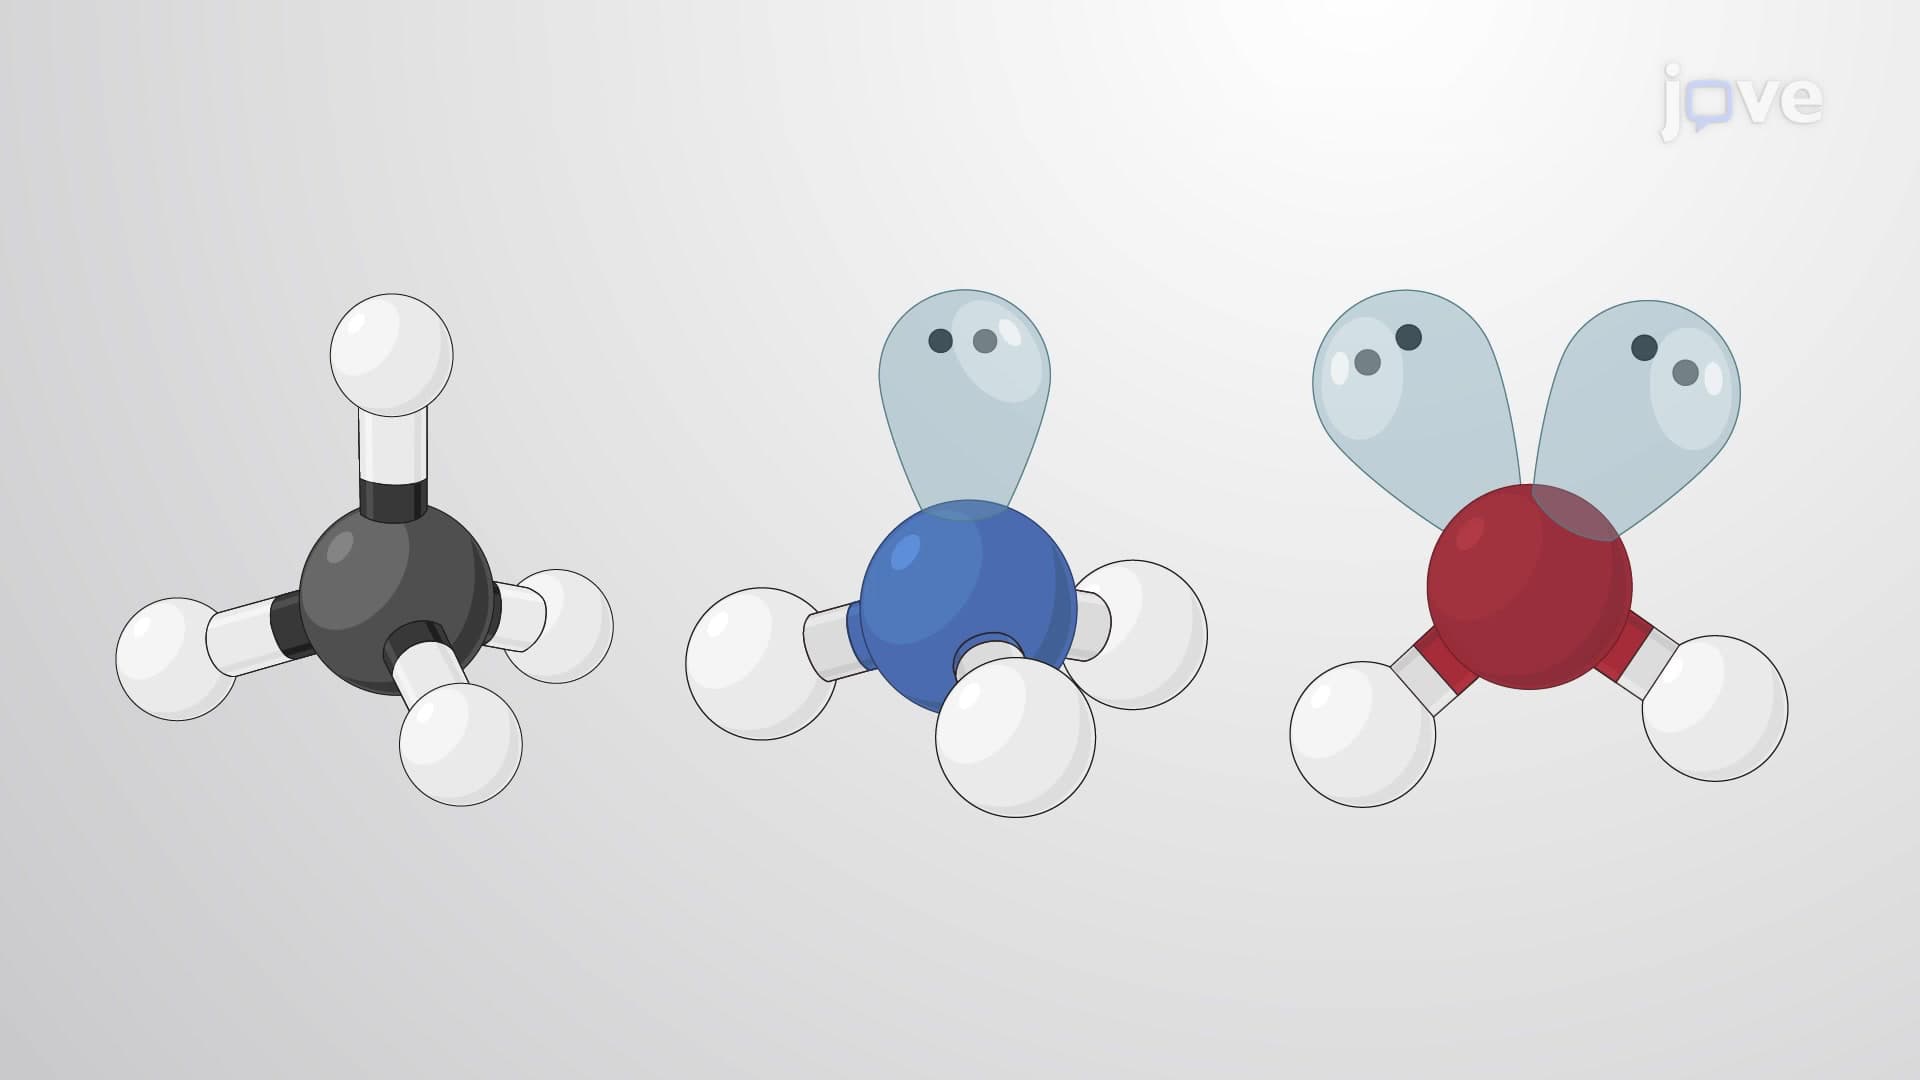
VSEPR Theory and the Effect of Lone Pairs
 01:19
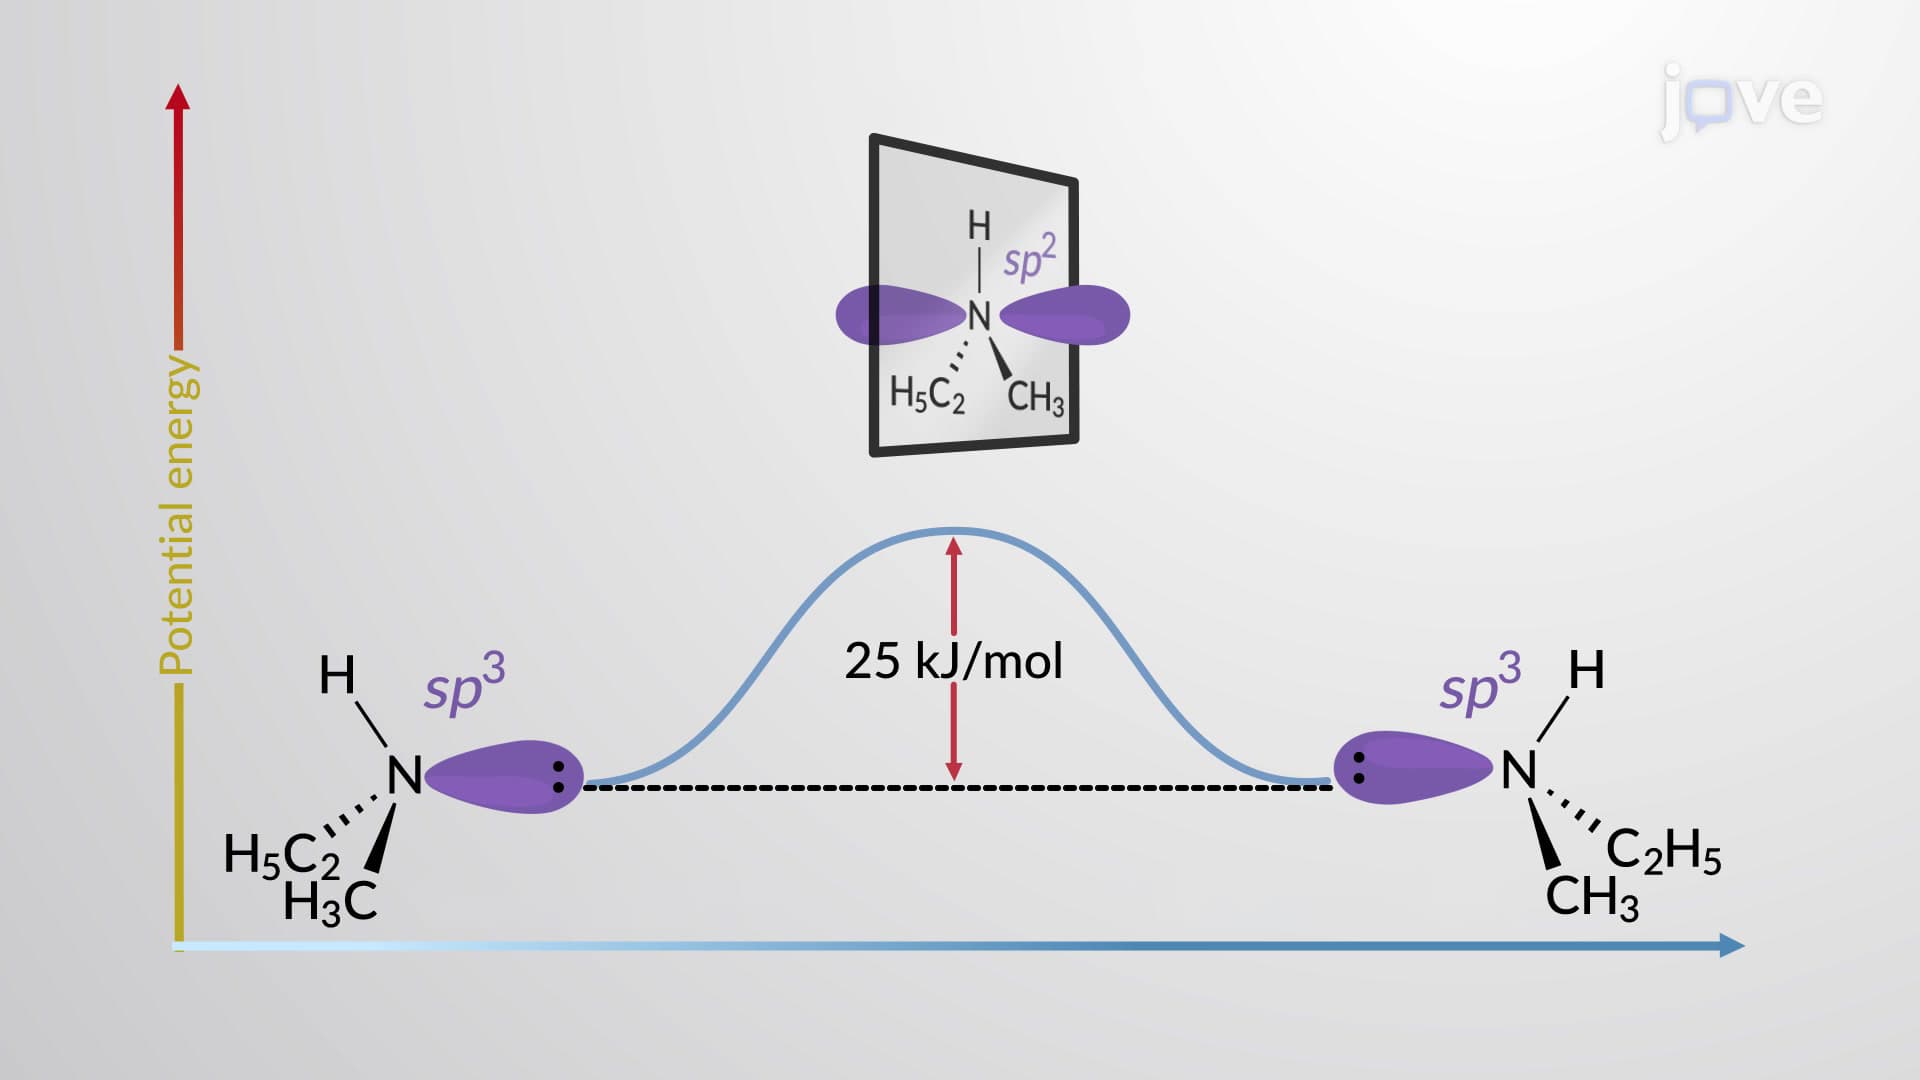
Structure of Amines
 01:19
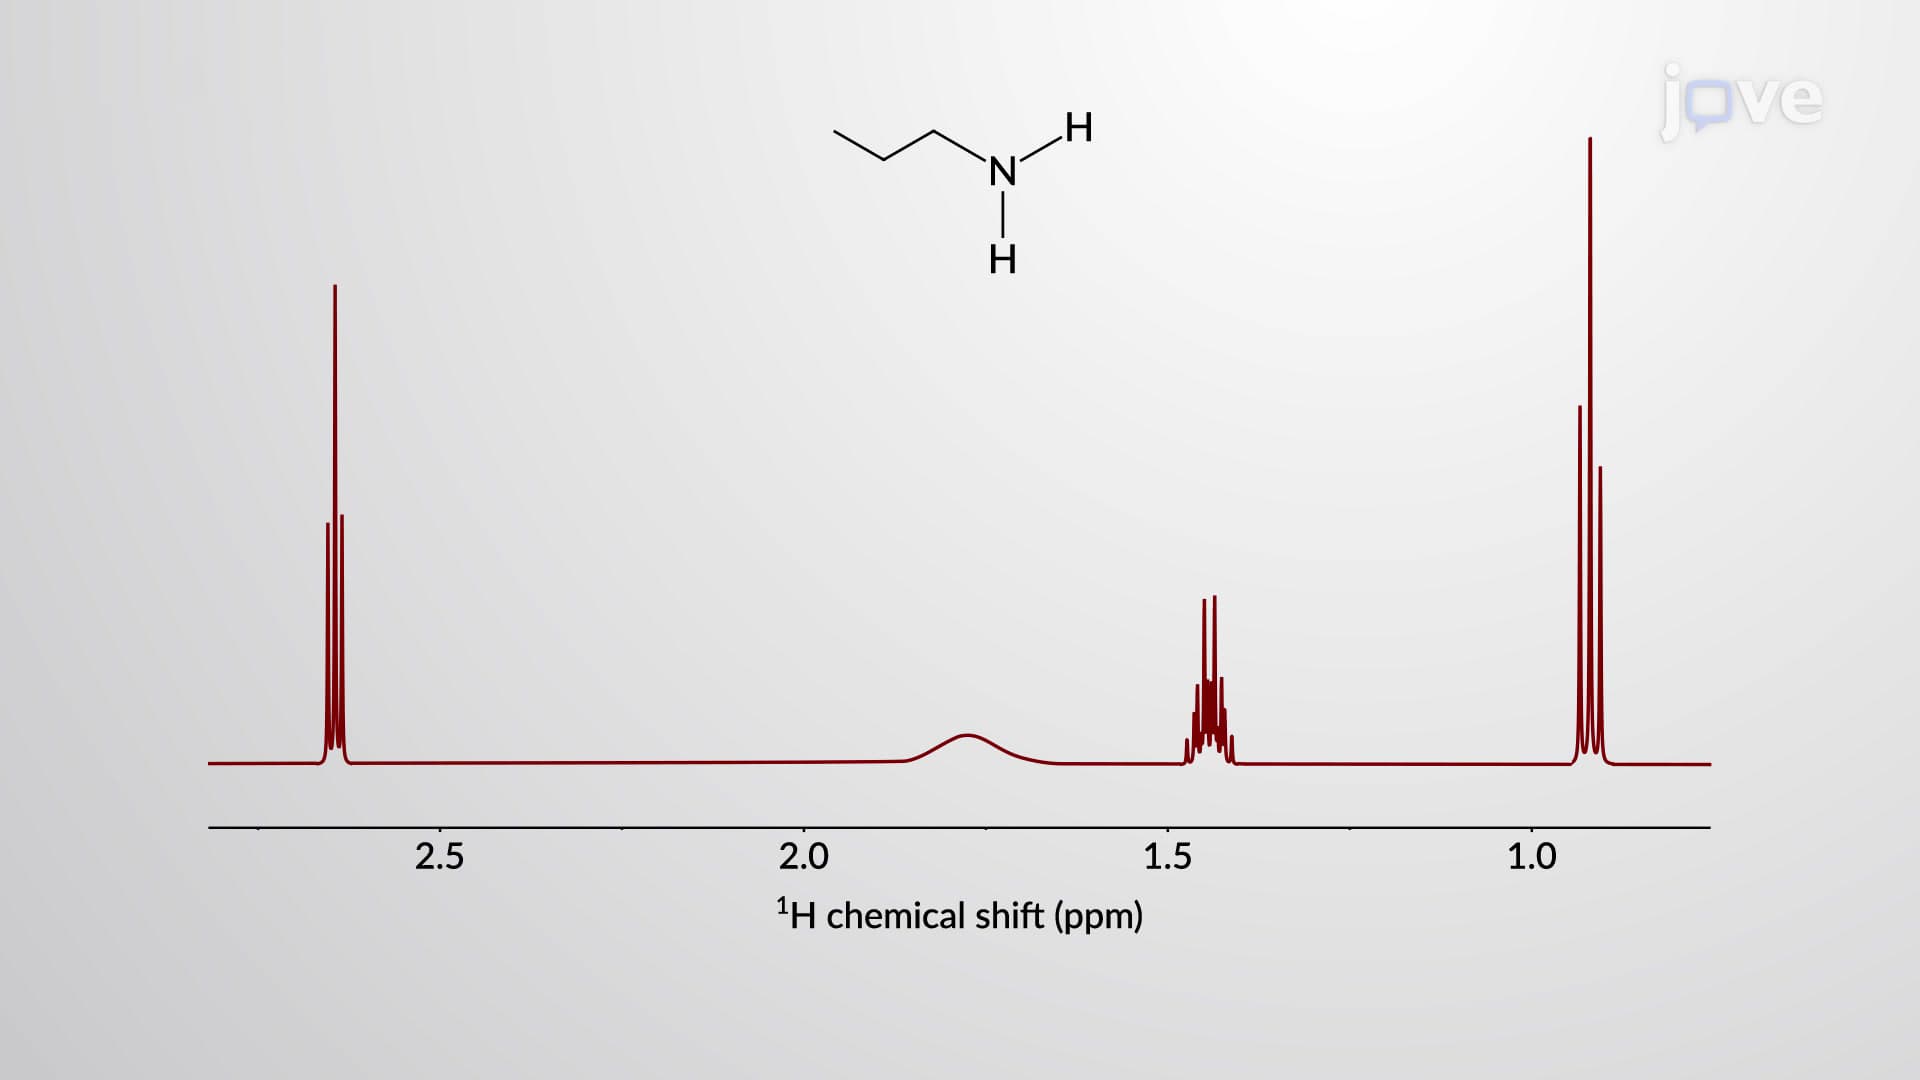
NMR Spectroscopy Of Amines
 02:37
VSEPR Theory
 01:15
Mass Spectrometry of Amines