Related Experiment Video
Updated: Jun 10, 2026


Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Quantum chemical modeling of benzene ethylation over H-ZSM-5 approaching chemical accuracy: a hybrid MP2:DFT study
Niels Hansen1, Torsten Kerber, Joachim Sauer
1Department of Chemical Engineering, Hamburg University of Technology, D-21073 Hamburg, Germany. n.hansen@tuhh.de

Abstract:
The alkylation of benzene by ethene over H-ZSM-5 is analyzed by means of a hybrid MP2:DFT scheme. Density functional calculations applying periodic boundary conditions (PBE functional) are combined with MP2 energy calculations on a series of cluster models of increasing size which allows extrapolation to the periodic MP2 limit. Basis set truncation errors are estimated by extrapolation of the MP2 energy to the complete basis set limit. Contributions from higher-order correlation effects are accounted for by CCSD(T) coupled cluster calculations. The sum of all contributions provides the "final estimates" for adsorption energies and energy barriers. Dispersion contributes significantly to the potential energy surface. As a result, the MP2:DFT potential energy profile is shifted downward compared to the PBE profile. More importantly, this shift is not the same for reactants and transition structures due to different self-interaction correction errors. The final enthalpies for ethene, benzene, and ethylbenzene adsorption on the Brønsted acid site at 298 K are -46, -78, and -110 kJ/mol, respectively. The intrinsic enthalpy barriers at 653 K are 117 and 119/94 kJ/mol for the one- and two-step alkylation, respectively. Intrinsic rate coefficients calculated by means of transition state theory are converted to apparent Arrhenius parameters by means of the multicomponent adsorption equilibrium. The simulated apparent activation energy (66 kJ/mol) agrees with experimental data (58-76 kJ/mol) within the uncertainty limit of the calculations. Adsorption energies obtained by adding a damped dispersion term to the PBE energies (PBE+D), agree within +/-7 kJ/mol, with the "final estimates", except for physisorption (pi-complex formation) and chemisorption of ethene (ethoxide formation) for which the PBE+D energies are 12.4 and 26.0 kJ/mol, respectively larger than the "final estimates". For intrinsic energy barriers, the PBE+D approach does not improve pure PBE results.
More Related Videos


Related Concept Videos
 01:18
Structure of Benzene: Molecular Orbital Model
 01:07
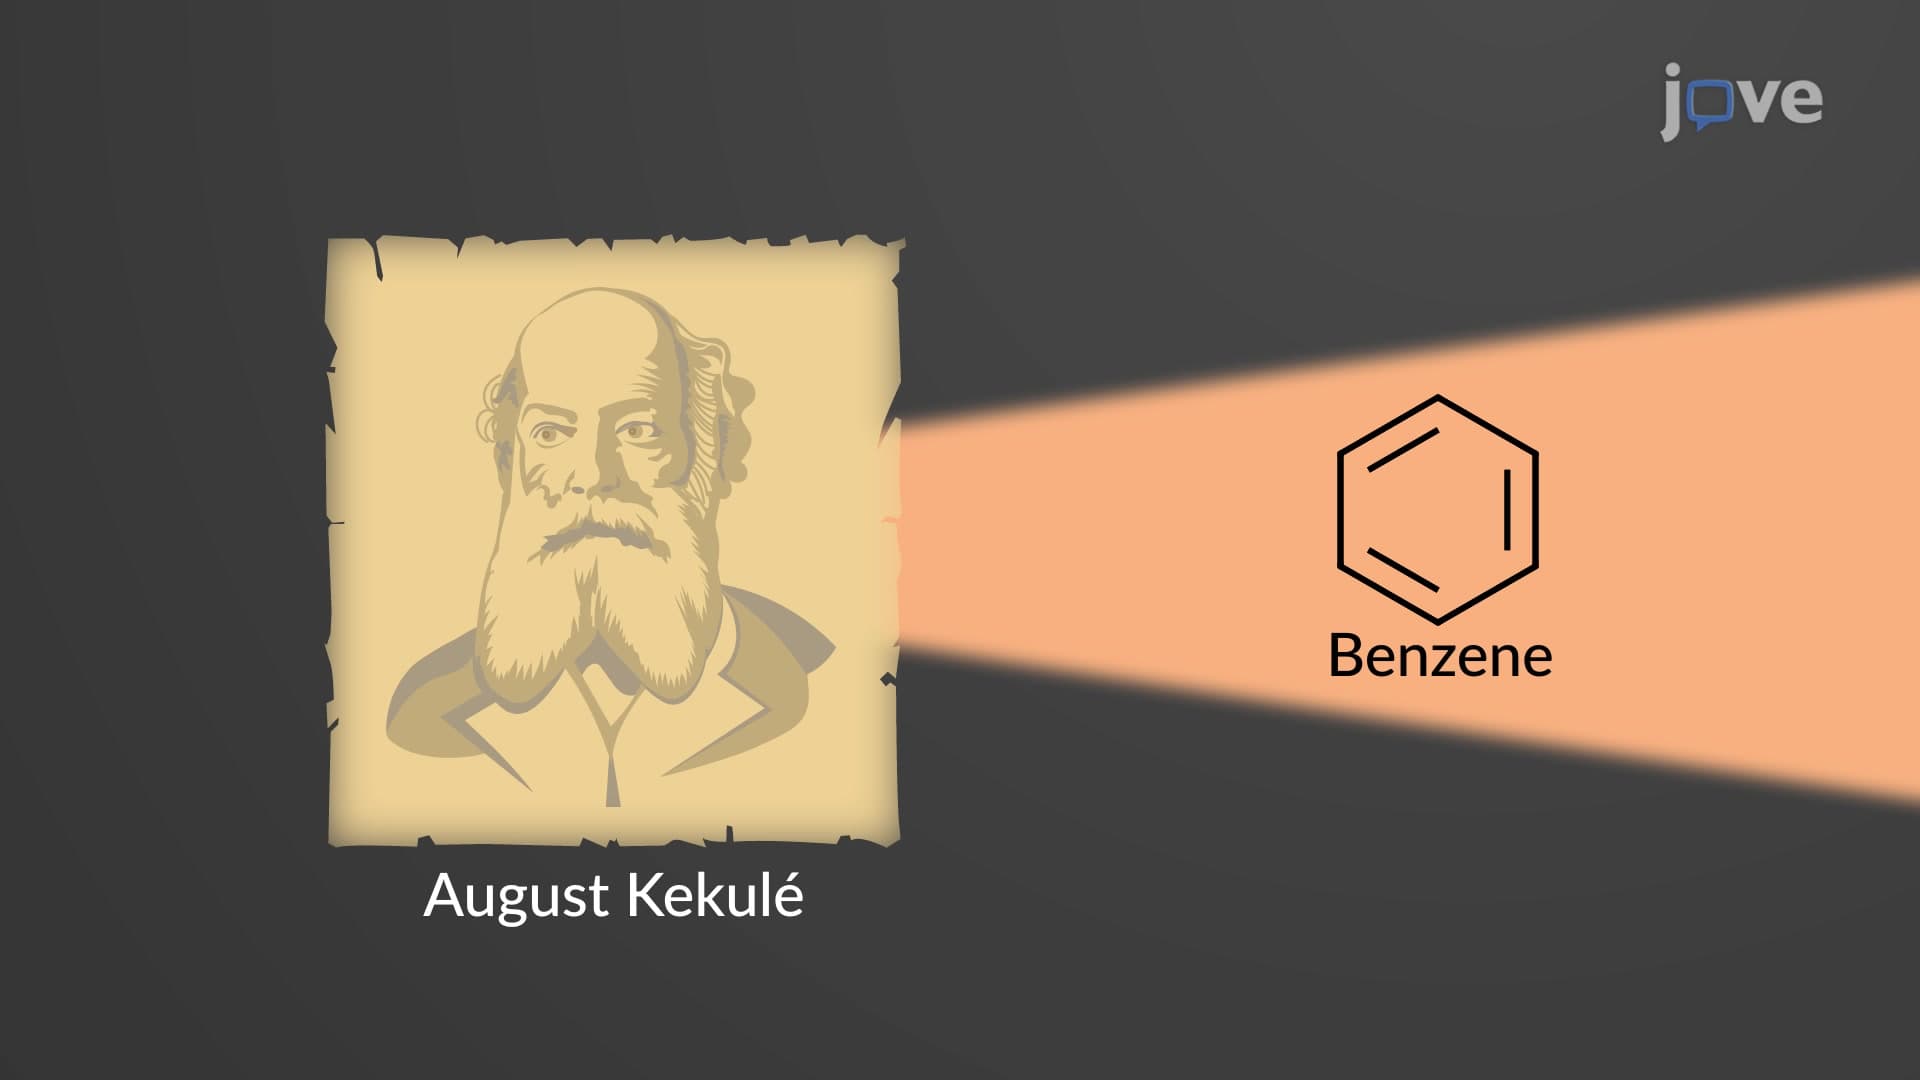
Structure of Benzene: Kekulé Model
He proposed that benzene has a cyclic structure of six carbon atoms attached to one hydrogen atom each, with three alternating pi bonds.
 01:37
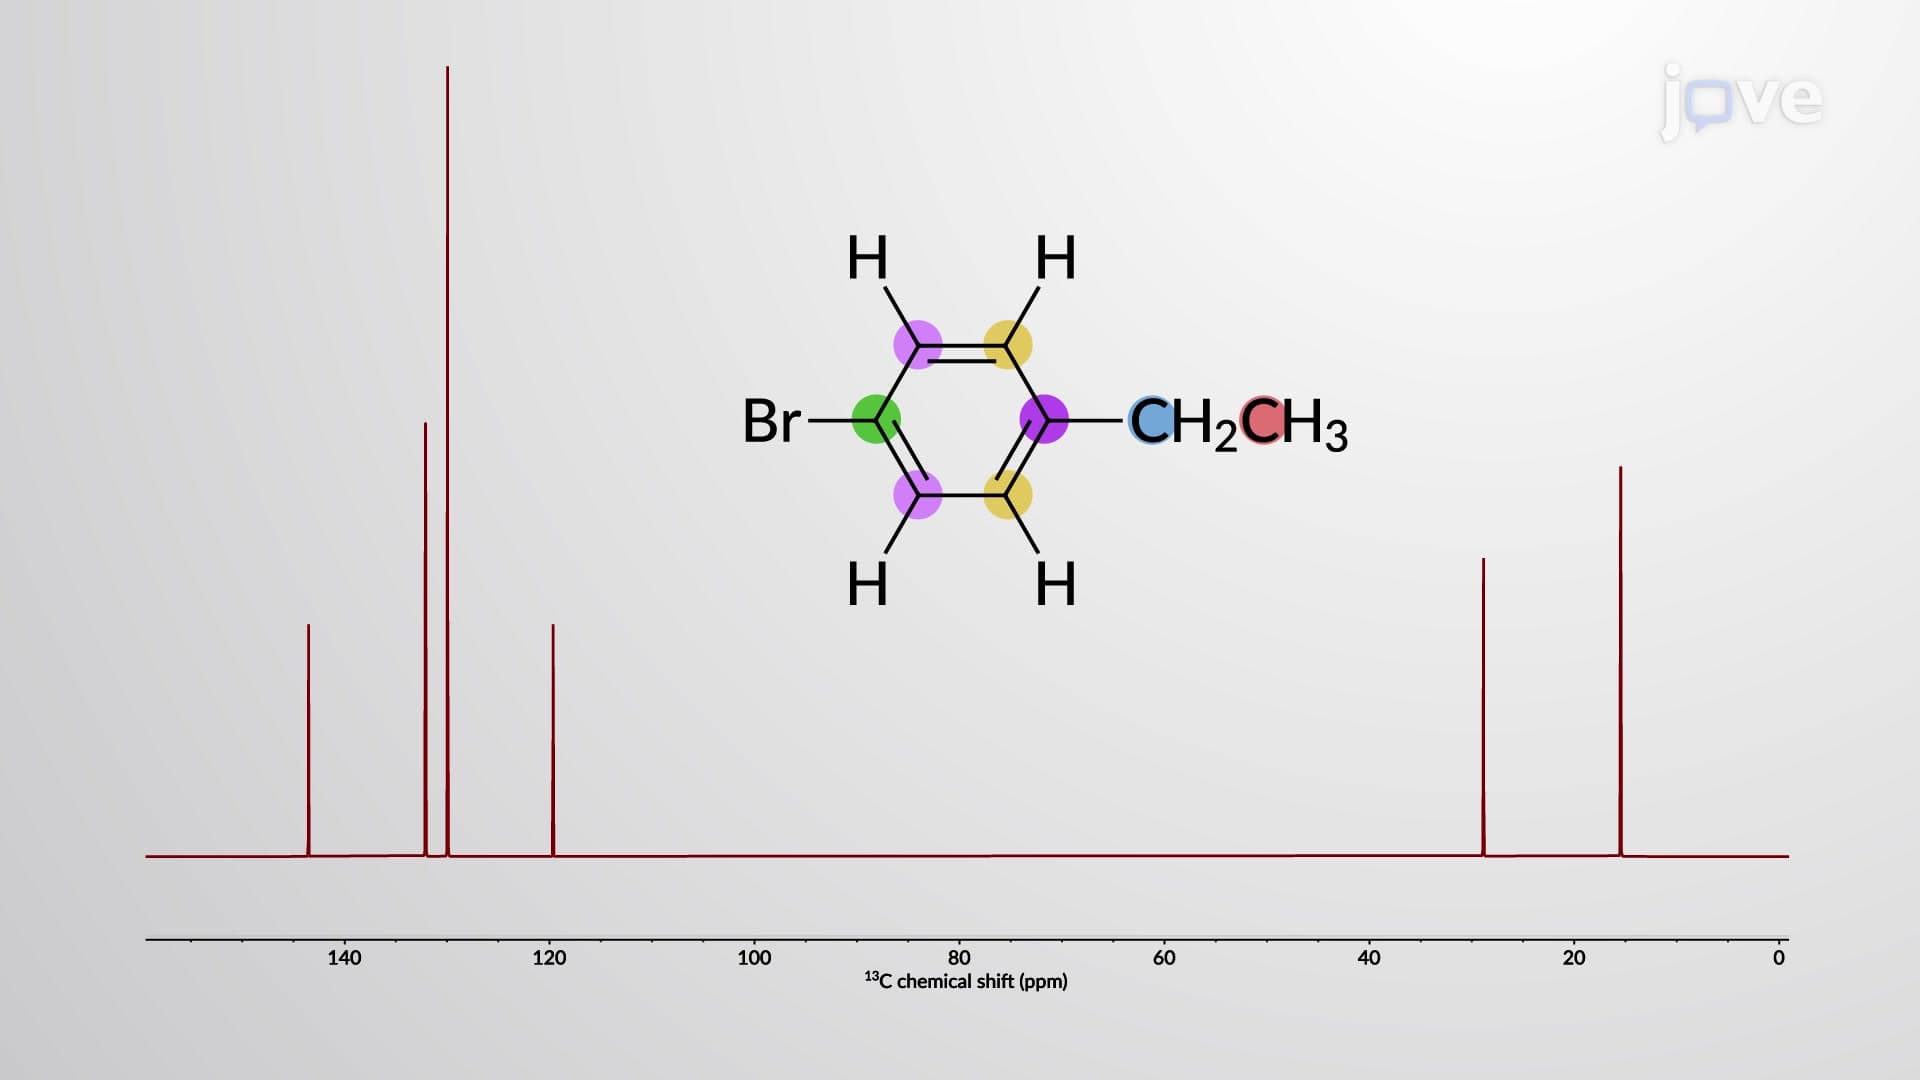
NMR Spectroscopy of Benzene Derivatives
 03:51
Molecular Orbital Theory II
 01:18
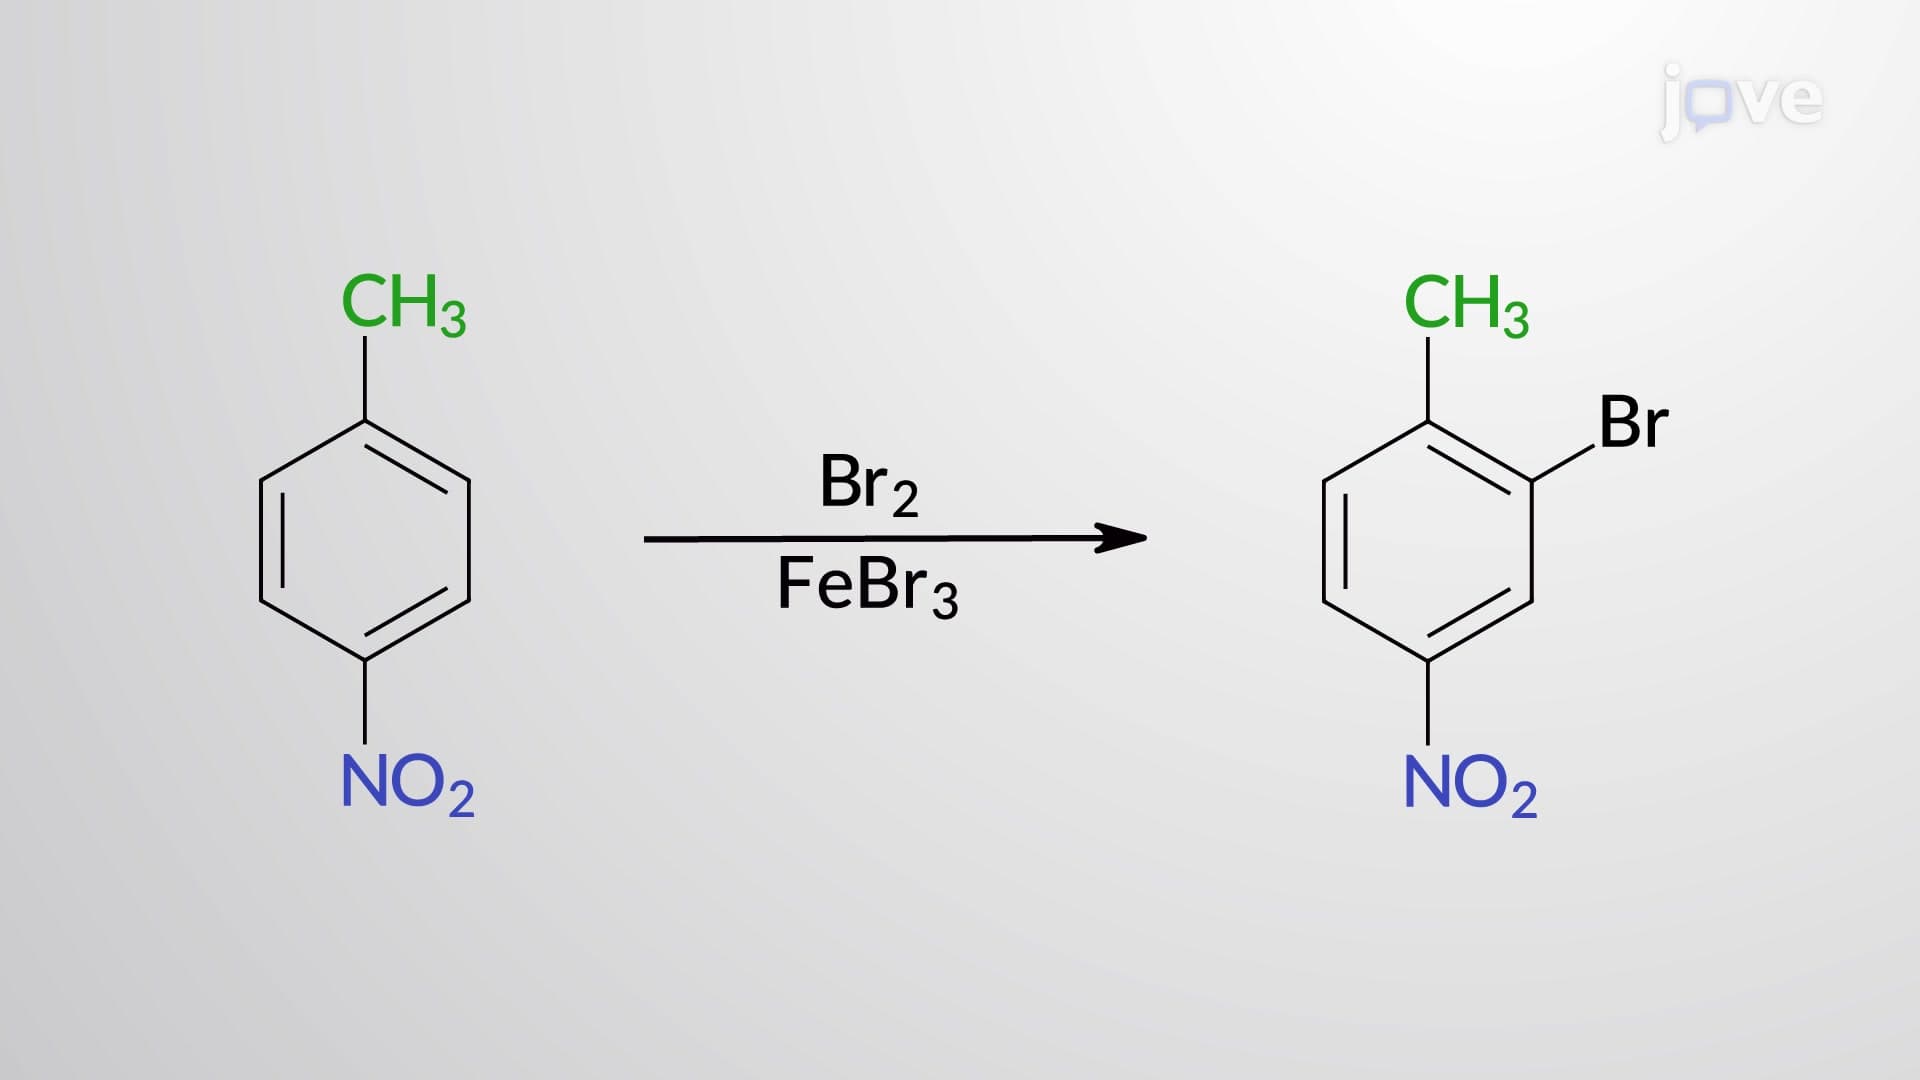
Directing and Steric Effects in Disubstituted Benzene Derivatives
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...