Related Experiment Video
Updated: Jun 7, 2026

High-Contrast and Fast Photorheological Switching of a Twist-Bend Nematic Liquid Crystal
Published on: October 31, 2019
Conformational stability, r0 structural parameters, and vibrational assignments of mono-substituted cyclobutanes:
Arindam Ganguly1, Joshua J Klaassen, Gamil A Guirgis
1Department of Chemistry, University of Missouri-Kansas City, Kansas City, MO 64110, USA.

Abstract:
Variable temperature (-55 to -100°C) studies of the infrared spectra (3500-400 cm(-1)) of fluorocyclobutane, c-C(4)H(7)F, dissolved in liquid xenon have been carried out as well as the infrared spectra of the gas. By utilizing eight pairs of conformers at 10 different temperatures, the enthalpy difference between the more stable equatorial conformer and the axial form has been determined to be 496±40 cm(-1) (5.93±0.48 kJ/mol). The percentage of the axial conformer present at ambient temperature is estimated to be 8±1%. The ab initio MP2(full) average predicted energy difference from a variety of basis sets is 732±47 cm(-1) (9.04±0.44 kJ/mol) and the average value of 602±20 cm(-1) from density functional theory predictions by the B3LYP method are significantly larger than the experimentally determined enthalpy value. By utilizing previously reported microwave rotational constants for the equatorial and axial conformers combined with ab initio MP2(full)/6-311+G(d,p) predicted structural values, adjusted r(0) parameters have been obtained. The determined heavy atom structural parameters for the equatorial [axial] conformer are: distances (Å) C-F=1.383(3) [1.407(3)], C(α)-C(β)=1.543(3) [1.546(3)], C(β)-C(γ)=1.554(3) [1.554(3)] and angles (°) ∠C(α)C(β)C(γ)=85.0(5) [89.2(5)], ∠C(β)C(α)C(β)=89.3(5) [89.2(5)], ∠F-(C(β)C(α)C(β))=117.4(5) [109.2(5)] and a puckering angle of 37.4(5) [20.7(5)]. The conformational stabilities, harmonic force fields, infrared intensities, Raman activities, depolarization ratios and vibrational frequencies have been obtained for both conformers from MP2(full)/6-31G(d) ab initio calculations and compared to experimental values where available. The results are discussed and compared to the corresponding properties of some other monosubstituted cyclobutanes with halogen and pseudo-halogen substituents.
Related Concept Videos
 02:02
Chair Conformation of Cyclohexane
The hydrogen atoms linked to carbons are arranged in two different axial and equatorial orientations to achieve this staggered...
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 02:29
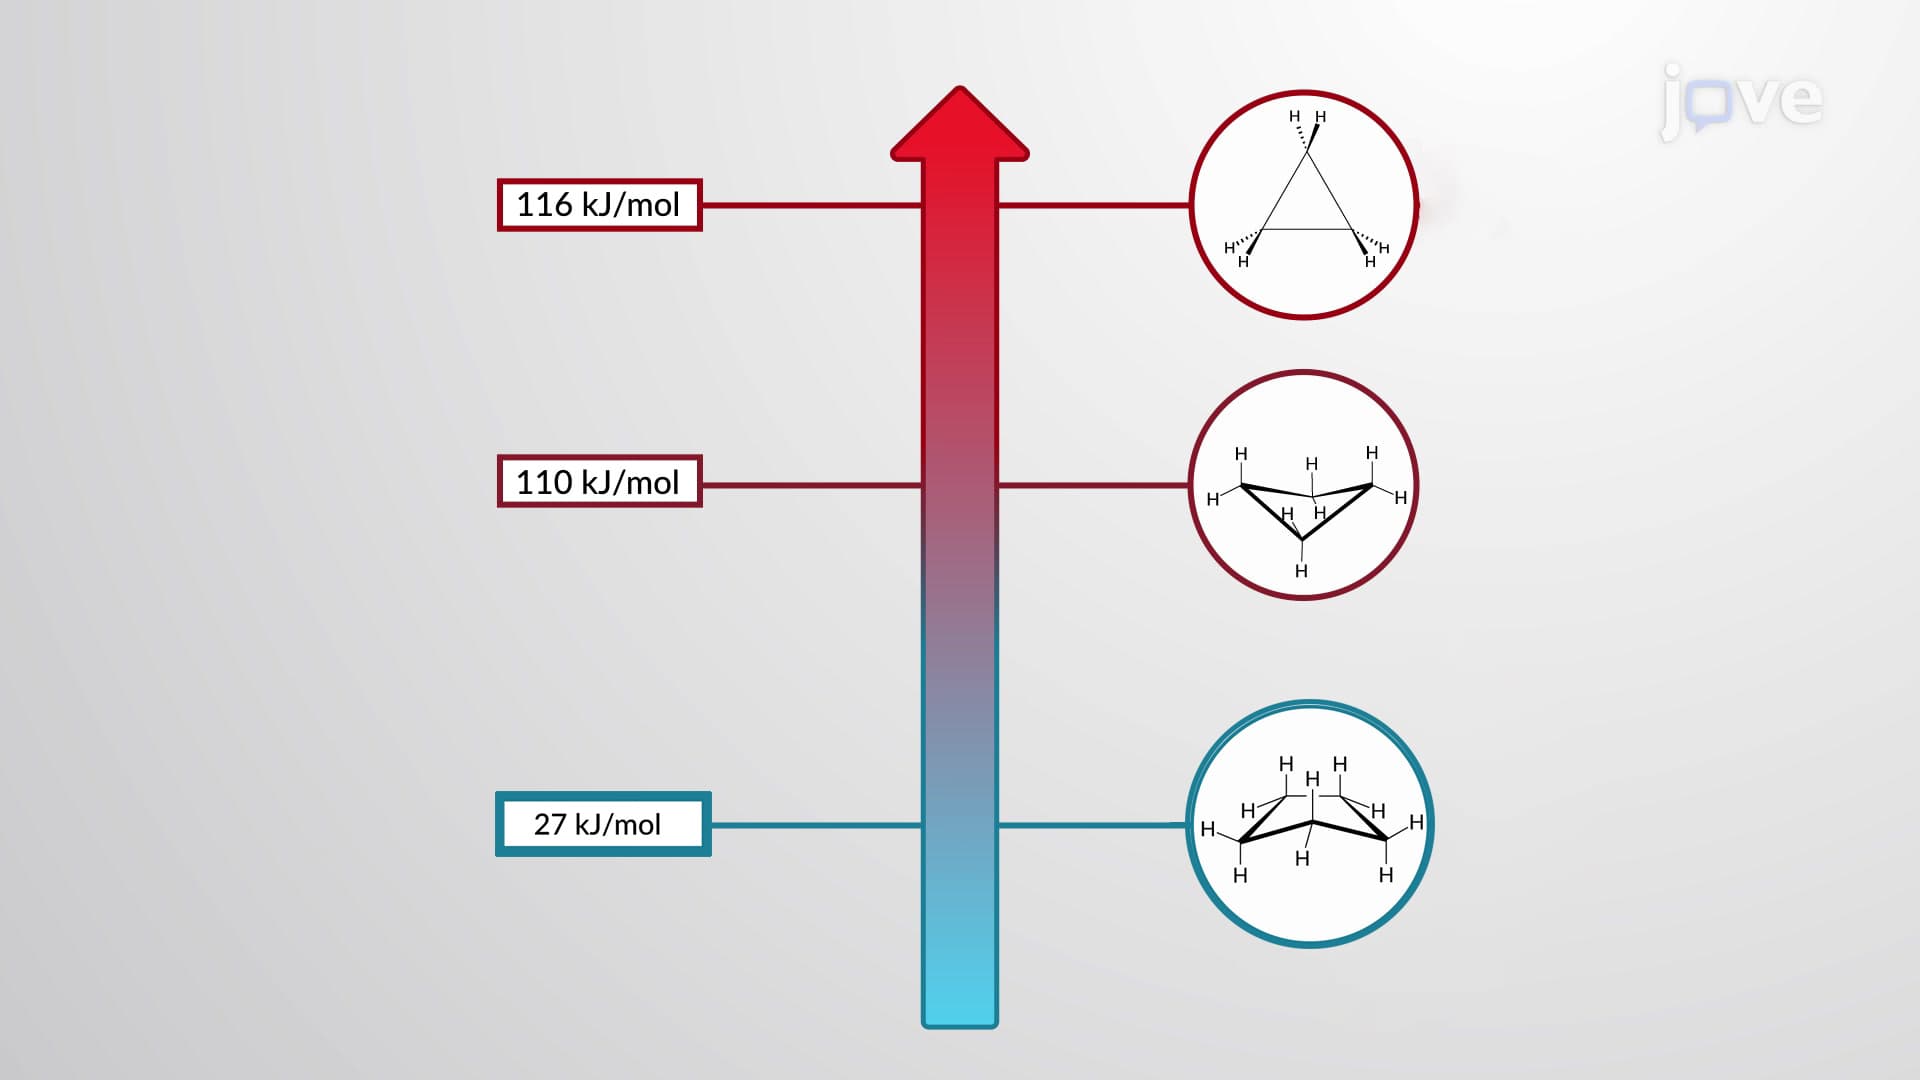
Conformations of Cycloalkanes
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal tetrahedral value,...
 01:18
Frost Circles for Different Conjugated Systems
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution