Related Experiment Video
Updated: Jun 7, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Comparative analysis of QSAR models for predicting pK(a) of organic oxygen acids and nitrogen bases from molecular
Haiying Yu1, Ralph Kühne, Ralf-Uwe Ebert
1UFZ Department of Ecological Chemistry, Helmholtz Centre for Environmental Research, Permoserstrasse 15, D-04318 Leipzig, Germany.

Abstract:
For 1143 organic compounds comprising 580 oxygen acids and 563 nitrogen bases that cover more than 17 orders of experimental pK(a) (from -5.00 to 12.23), the pK(a) prediction performances of ACD, SPARC, and two calibrations of a semiempirical quantum chemical (QC) AM1 approach have been analyzed. The overall root-mean-square errors (rms) for the acids are 0.41, 0.58 (0.42 without ortho-substituted phenols with intramolecular H-bonding), and 0.55 and for the bases are 0.65, 0.70, 1.17, and 1.27 for ACD, SPARC, and both QC methods, respectively. Method-specific performances are discussed in detail for six acid subsets (phenols and aromatic and aliphatic carboxylic acids with different substitution patterns) and nine base subsets (anilines, primary, secondary and tertiary amines, meta/para-substituted and ortho-substituted pyridines, pyrimidines, imidazoles, and quinolines). The results demonstrate an overall better performance for acids than for bases but also a substantial variation across subsets. For the overall best-performing ACD, rms ranges from 0.12 to 1.11 and 0.40 to 1.21 pK(a) units for the acid and base subsets, respectively. With regard to the squared correlation coefficient r², the results are 0.86 to 0.96 (acids) and 0.79 to 0.95 (bases) for ACD, 0.77 to 0.95 (acids) and 0.85 to 0.97 (bases) for SPARC, and 0.64 to 0.87 (acids) and 0.43 to 0.83 (bases) for the QC methods, respectively. Attention is paid to structural and method-specific causes for observed pitfalls. The significant subset dependence of the prediction performances suggests a consensus modeling approach.
More Related Videos
 05:47
In Silico Modeling Method for Computational Aquatic Toxicology of Endocrine Disruptors: A Software-Based Approach Using QSAR Toolbox
Published on: August 28, 2019
 06:50
O-cresol Concentration Online Measurement Based On Near Infrared Spectroscopy Via Partial Least Square Regression
Published on: November 8, 2019
Related Concept Videos
 02:00
Molecular Models
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 02:35
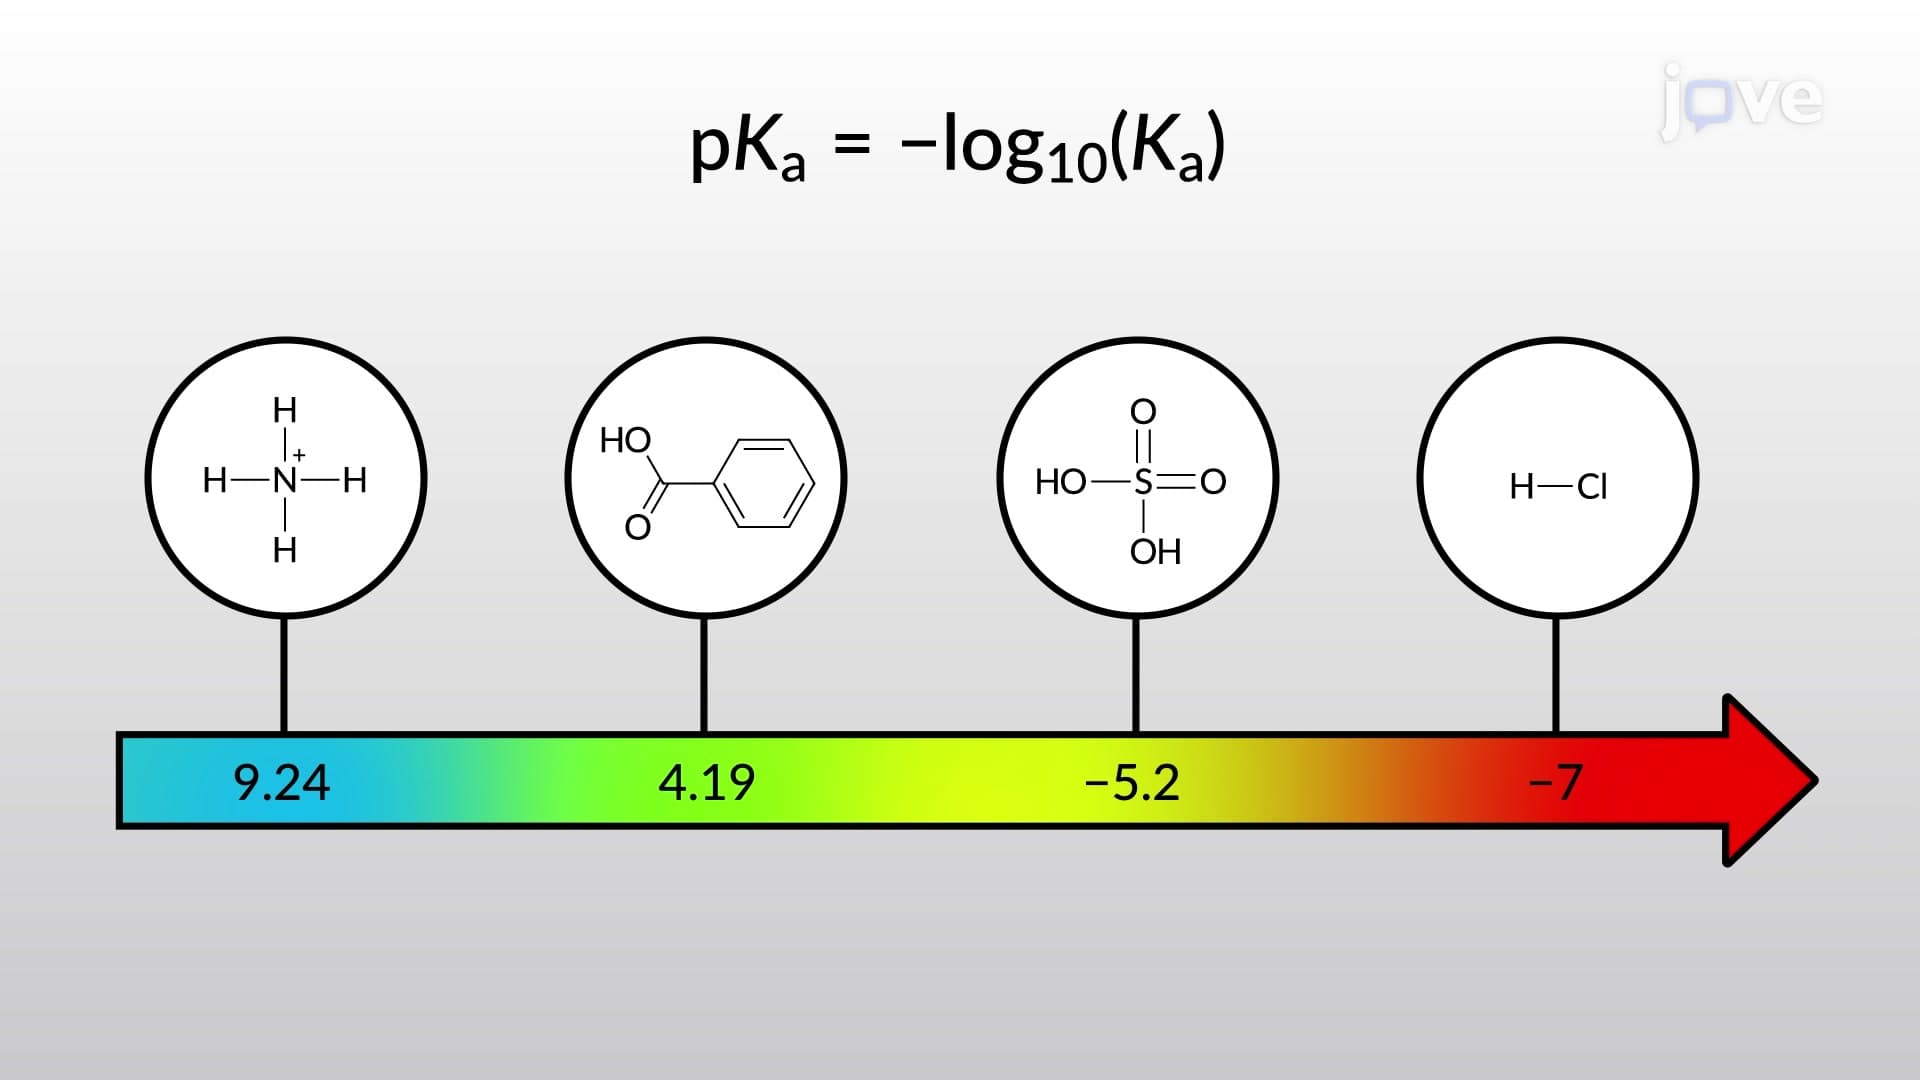
Acid and Bases: Ka, pKa, and Relative Strengths
 01:20
Pharmacokinetic Models: Overview
There are three primary types of models: empirical, compartment, and physiological. Empirical models, with minimal assumptions,...
 01:26
Pharmacokinetic Models: Comparison and Selection Criterion
Physiological models take a detailed approach by considering specific molecular processes. They can predict drug distribution, metabolism, and elimination changes, providing a comprehensive understanding of how drugs interact with the body.
 01:25
Basicity of Heterocyclic Aromatic Amines