Related Experiment Video
Updated: Jun 5, 2026

Using Live Cell STED Imaging to Visualize Mitochondrial Inner Membrane Ultrastructure in Neuronal Cell Models
Published on: June 30, 2023
Abnormal mitochondrial dynamics, mitochondrial loss and mutant huntingtin oligomers in Huntington's disease:
Ulziibat Shirendeb1, Arubala P Reddy, Maria Manczak
1Neurogenetics Laboratory, Division of Neuroscience, Oregon National Primate Research Center, Oregon Health & Science University, 505 NW 185th Avenue, Beaverton, OR 97006, USA.

Abstract:
The purpose of our study was to determine the relationship between mutant huntingtin (Htt) and mitochondrial dynamics in the progression of Huntington's disease (HD). We measured the mRNA levels of electron transport chain genes, and mitochondrial structural genes, Drp1 (dynamin-related protein 1), Fis1 (fission 1), Mfn1 (mitofusin 1), Mfn2 (mitofusin 2), Opa1 (optric atrophy 1), Tomm40 (translocase of outermembrane 40) and CypD (cyclophilin D) in grade III and grade IV HD patients and controls. The mutant Htt oligomers and the mitochondrial structural proteins were quantified in the striatum and frontal cortex of HD patients. Changes in expressions of the electron transport chain genes were found in HD patients and may represent a compensatory response to mitochondrial damage caused by mutant Htt. Increased expression of Drp1 and Fis1 and decreased expression of Mfn1, Mfn2, Opa1 and Tomm40 were found in HD patients relative to the controls. CypD was upregulated in HD patients, and this upregulation increased as HD progressed. Significantly increased immunoreactivity of 8-hydroxy-guanosine was found in the cortical specimens from stage III and IV HD patients relative to controls, suggesting increased oxidative DNA damage in HD patients. In contrast, significantly decreased immunoreactivities of cytochrome oxidase 1 and cytochrome b were found in HD patients relative to controls, indicating a loss of mitochondrial function in HD patients. Immunoblotting analysis revealed 15, 25 and 50 kDa mutant Htt oligomers in the brain specimens of HD patients. All oligomeric forms of mutant Htt were significantly increased in the cortical tissues of HD patients, and mutant Htt oligomers were found in the nucleus and in mitochondria. The increase in Drp1, Fis1 and CypD and the decrease in Mfn1 and Mfn2 may be responsible for abnormal mitochondrial dynamics that we found in the cortex of HD patients, and may contribute to neuronal damage in HD patients. The presence of mutant Htt oligomers in the nucleus of HD neurons and in mitochondria may disrupt neuronal functions. Based on these findings, we propose that mutant Htt in association with mitochondria imbalance and mitochondrial dynamics impairs axonal transport of mitochondria, decreases mitochondrial function and damages neurons in affected brain regions of HD patients.
Related Concept Videos
 01:21
Huntington Disease l: Introduction
 01:24
Parkinson Disease ll: Pathophysiology
 01:23
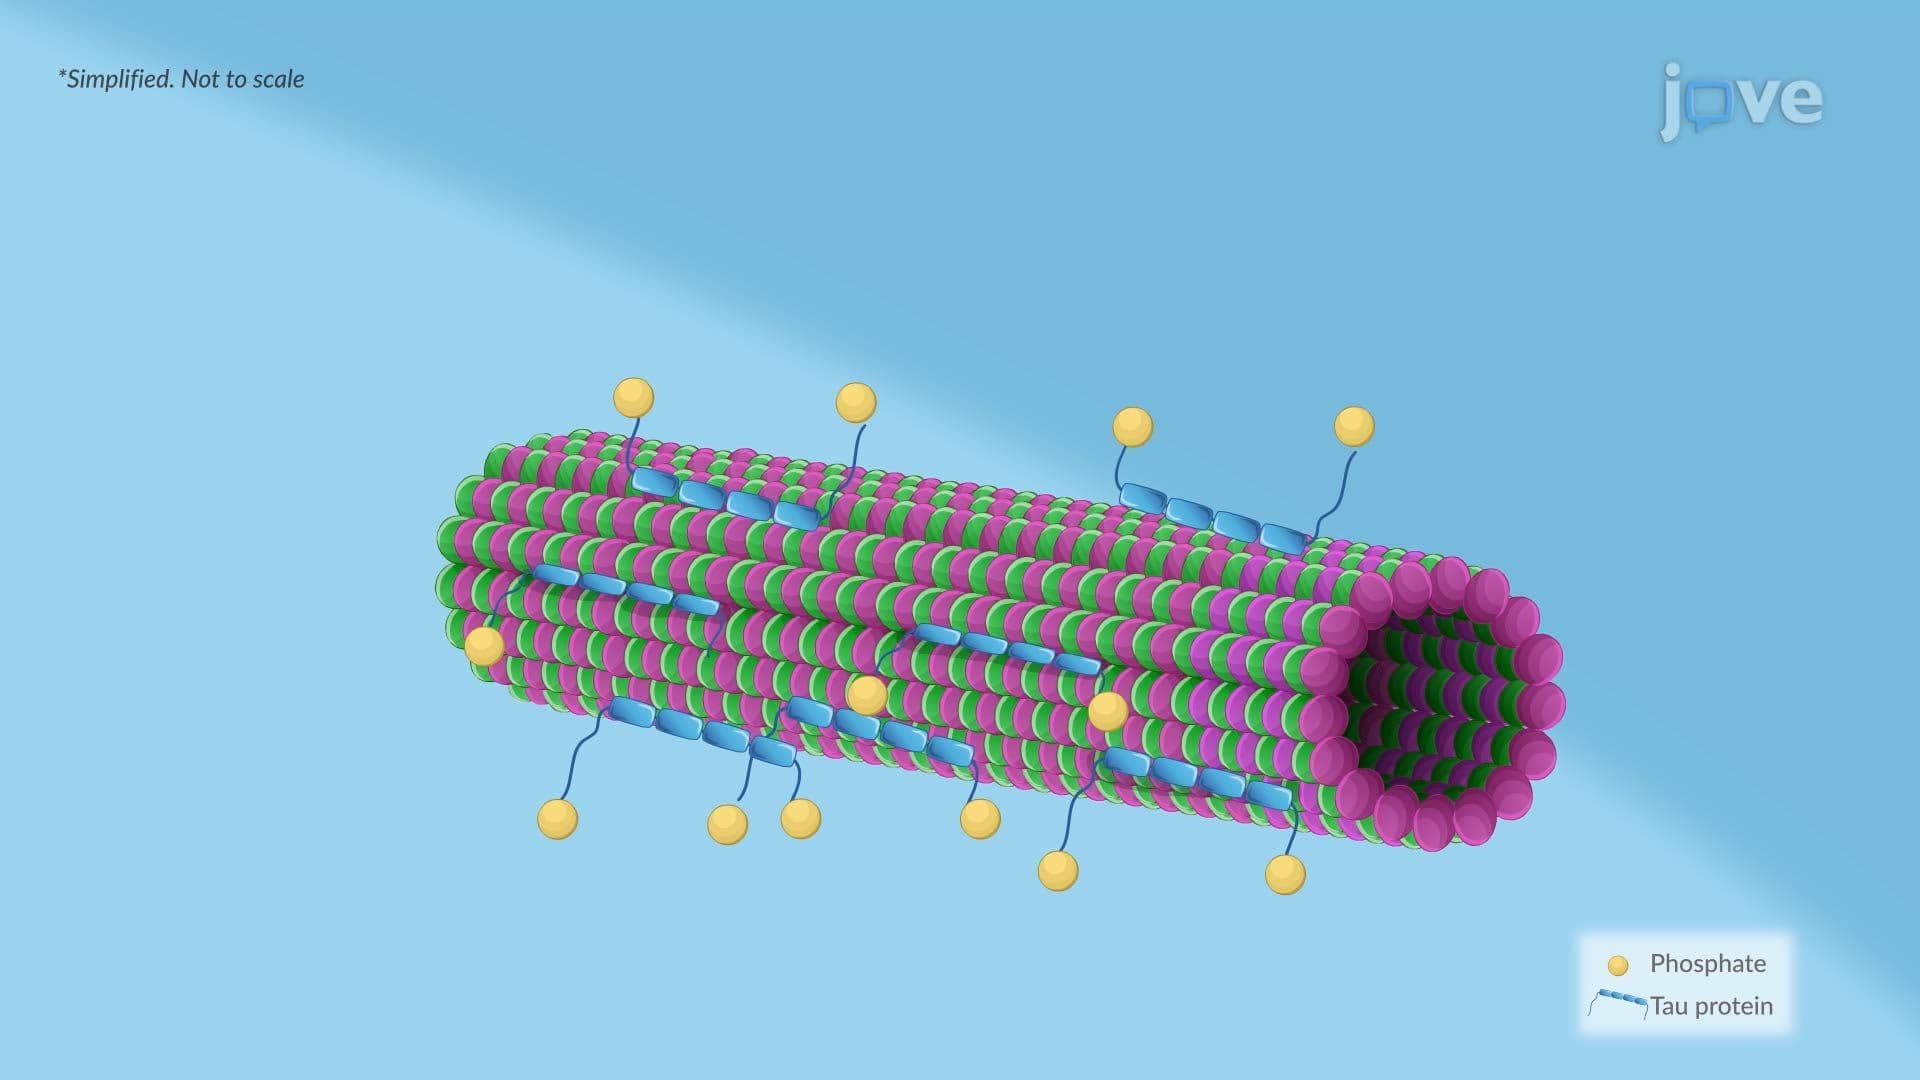
Alzheimer Disease ll: Pathophysiology
 01:22
Lysosomal Hydrolases