Related Experiment Video
Updated: Jun 4, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Assessment of the ΔSCF density functional theory approach for electronic excitations in organic dyes
Tim Kowalczyk1, Shane R Yost, Troy Van Voorhis
1Department of Chemistry, Massachusetts Institute of Technology, Cambridge, Massachusetts 02139-4307, USA.

Abstract:
This paper assesses the accuracy of the ΔSCF method for computing low-lying HOMO→LUMO transitions in organic dye molecules. For a test set of vertical excitation energies of 16 chromophores, surprisingly similar accuracy is observed for time-dependent density functional theory and for ΔSCF density functional theory. In light of this performance, we reconsider the ad hoc ΔSCF prescription and demonstrate that it formally obtains the exact stationary density within the adiabatic approximation, partially justifying its use. The relative merits and future prospects of ΔSCF for simulating individual excited states are discussed.
More Related Videos
 11:44
Using Cyclic Voltammetry, UV-Vis-NIR, and EPR Spectroelectrochemistry to Analyze Organic Compounds
Published on: October 18, 2018
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:29
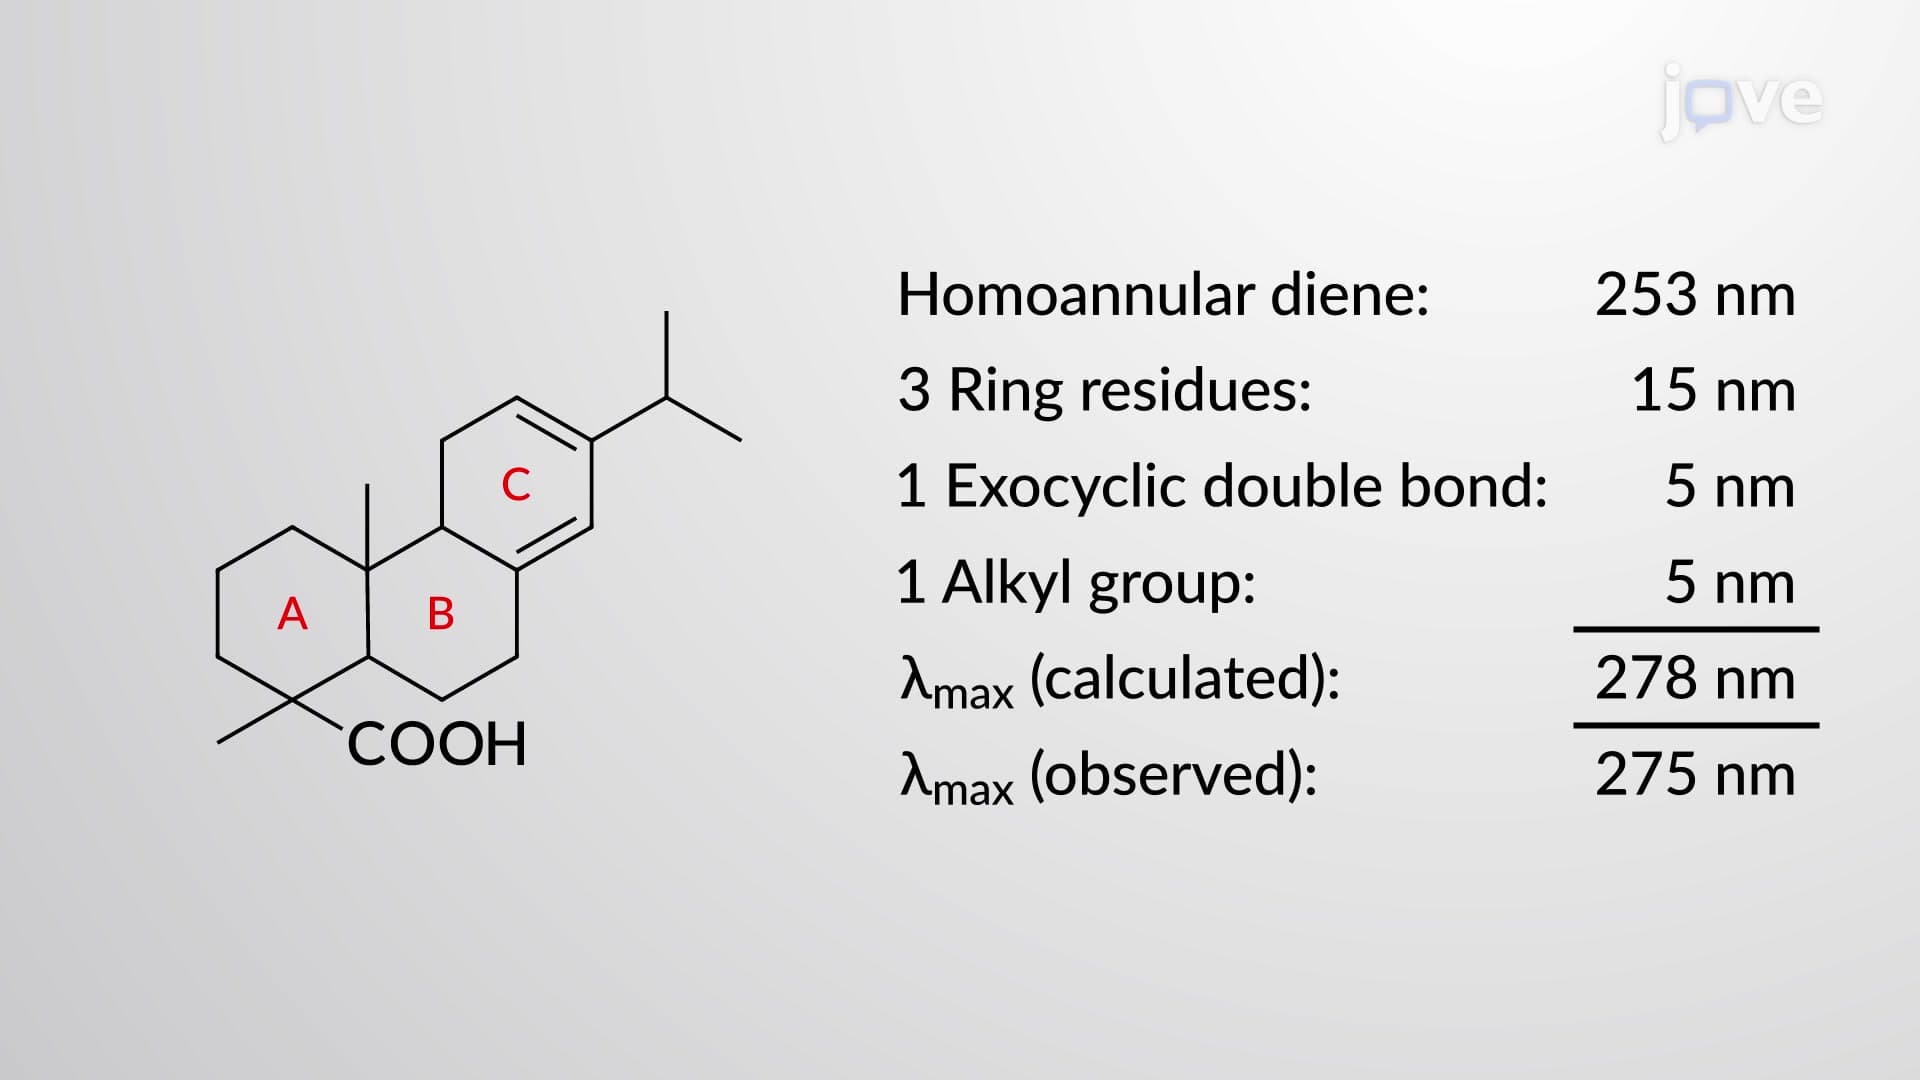
UV–Vis Spectroscopy: Woodward–Fieser Rules
 02:13
Energy Diagrams, Transition States, and Intermediates
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
Selection Rules: Photochemical Activation
 01:27
π Electron Effects on Chemical Shift: Overview