
Relaxation pathways of excited N-(triphenylmethyl)salicylidenimine in solutions
Renata Karpicz1, Vidmantas Gulbinas, Aleksandra Lewanowicz
1Institute of Physics, Center for Physical Sciences and Technology, Savanoriu Ave 231, LT-02300 Vilnius, Lithuania. renata@ar.fi.lt

Abstract:
Excited state relaxation of N-(triphenylmethyl)-salicylidenimine (MS1) in protic and aprotic solvents has been investigated by means of absorption pump-probe spectroscopy with femtosecond time resolution and fluorescence spectroscopy with picosecond time resolution. Short-lived excited states and long-lived photoproducts have been identified from the differential absorption spectra. Excited states and photoproducts were different under excitation of enol-closed and cis-keto tautomers. As a result, the commonly accepted excited state relaxation model of aromatic anils, which assumes an ultrafast transformation of excited enol-closed tautomers into cis-keto tautomers, has been modified. Performed quantum chemical calculations suggest that hydrogen-bonded ethanol molecules facilitate formation of cis-keto tautomers and are responsible for their different relaxation pathways in comparison with relaxation of excited enol-closed tauromers. Fluorescence decay on a nanosecond time scale was attributed to aggregated MS1 molecules.
More Related Videos
Related Concept Videos
 01:28
Atomic Nuclei: Types of Nuclear Relaxation
In spin–lattice or longitudinal relaxation, the excited spins exchange energy with the surrounding lattice as they return to the lower energy level. Among several mechanisms that contribute to spin–lattice relaxation, magnetic dipolar interactions are significant. Here, the excited nucleus transfers energy to a nearby...
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
Selection Rules: Photochemical Activation
 01:17
Thermal Electrocyclic Reactions: Stereochemistry
Selection Rules: Thermal Activation
Conjugated systems containing an even number of π-electron pairs undergo a conrotatory ring closure. For example, thermal electrocyclization of (2E,4E)-2,4-hexadiene, a conjugated diene containing two π-electron pairs, gives trans-3,4-dimethylcyclobutene.
 01:30
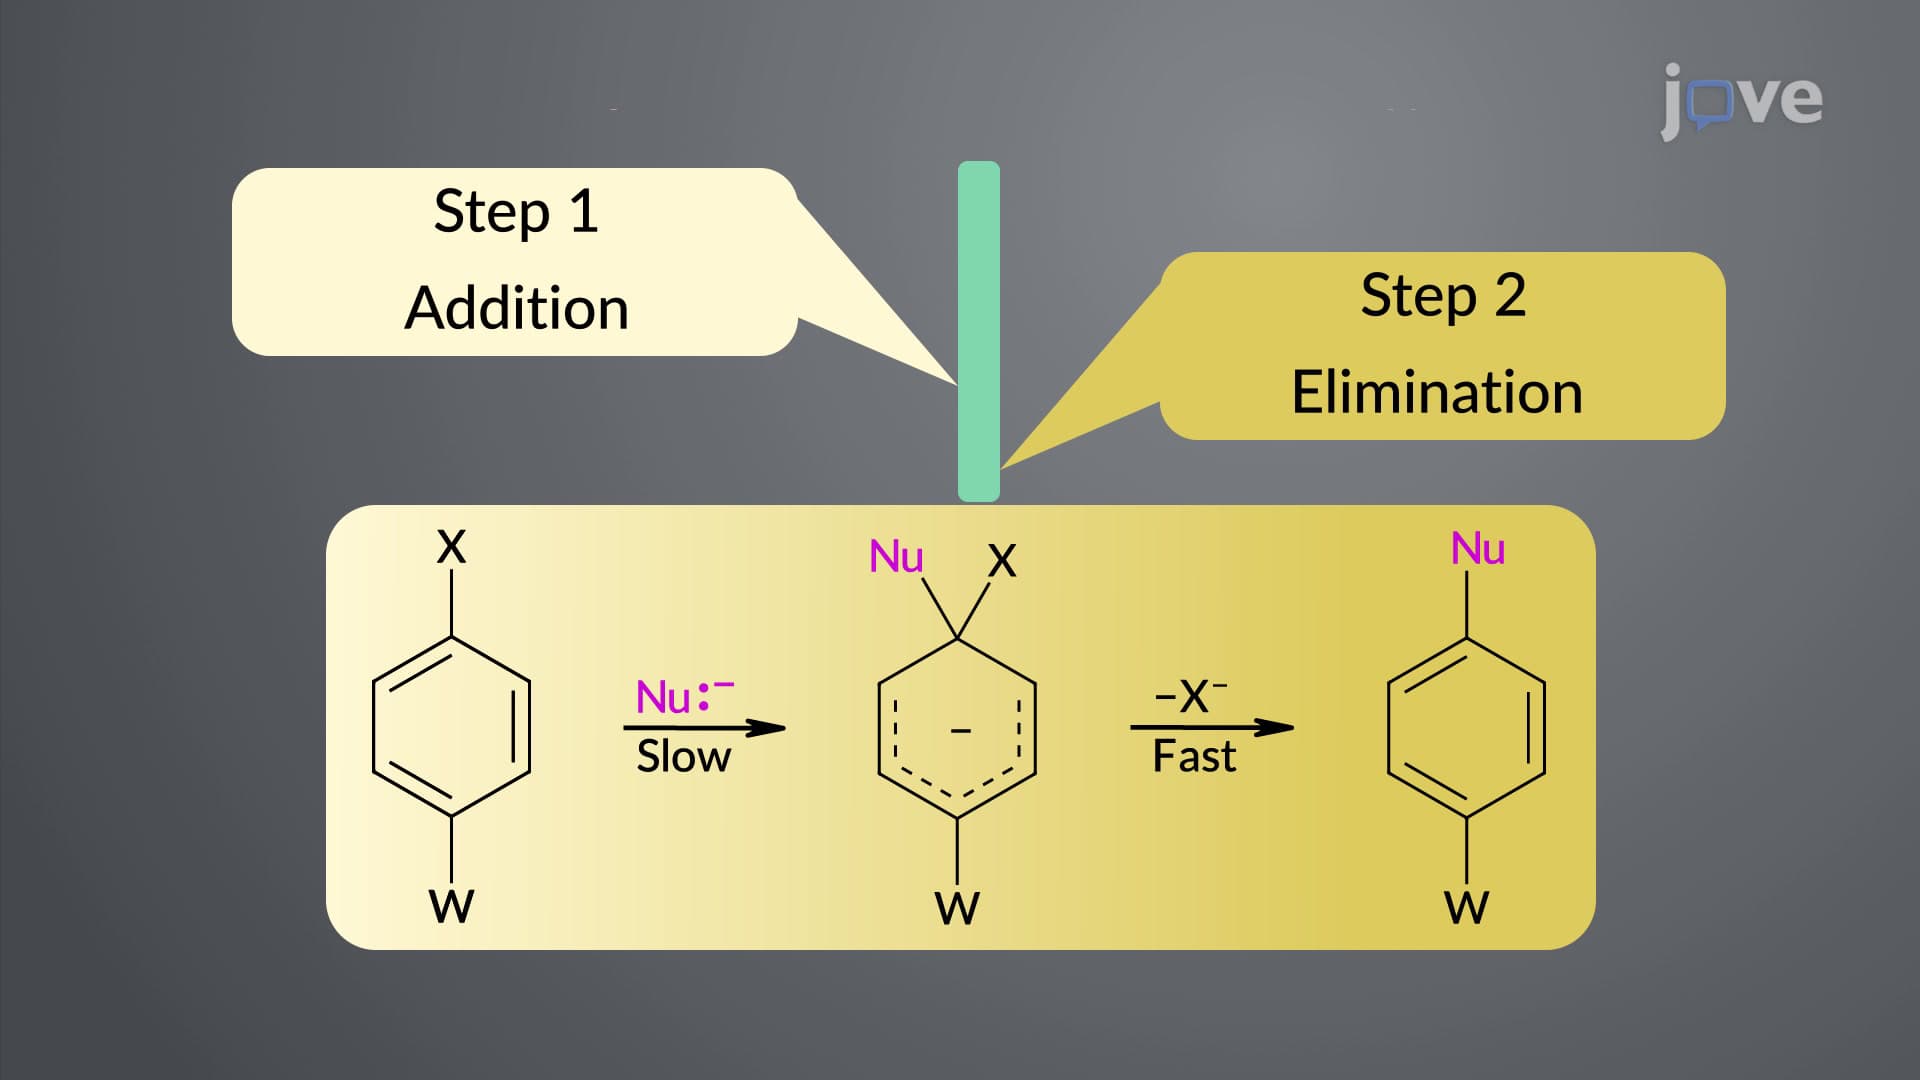
Nucleophilic Aromatic Substitution: Addition–Elimination (SNAr)
The reaction begins with an attack of the nucleophile on the carbon that holds the leaving group. This results in the delocalization of the π electrons over the ring carbons. The resonance interaction between the...
 01:11
Nucleophilic Aromatic Substitution: Elimination–Addition
 01:14
Nucleophilic Aromatic Substitution of Aryldiazonium Salts: Aromatic SN1
In the Sandmeyer reaction, for example, the diazonio group is replaced by a chloro, bromo, or cyano...