Related Experiment Video
Updated: Jun 3, 2026

High-throughput Screening for Protein-based Inheritance in S. cerevisiae
Published on: August 8, 2017
Prions
David W Colby1, Stanley B Prusiner
1Institute for Neurodegenerative Diseases, University of California, San Francisco, San Francisco, California 94143, USA.

Abstract:
The discovery of infectious proteins, denoted prions, was unexpected. After much debate over the chemical basis of heredity, resolution of this issue began with the discovery that DNA, not protein, from pneumococcus was capable of genetically transforming bacteria (Avery et al. 1944). Four decades later, the discovery that a protein could mimic viral and bacterial pathogens with respect to the transmission of some nervous system diseases (Prusiner 1982) met with great resistance. Overwhelming evidence now shows that Creutzfeldt-Jakob disease (CJD) and related disorders are caused by prions. The prion diseases are characterized by neurodegeneration and lethality. In mammals, prions reproduce by recruiting the normal, cellular isoform of the prion protein (PrP(C)) and stimulating its conversion into the disease-causing isoform (PrP(Sc)). PrP(C) and PrP(Sc) have distinct conformations: PrP(C) is rich in α-helical content and has little β-sheet structure, whereas PrP(Sc) has less α-helical content and is rich in β-sheet structure (Pan et al. 1993). The conformational conversion of PrP(C) to PrP(Sc) is the fundamental event underlying prion diseases. In this article, we provide an introduction to prions and the diseases they cause.
Insights
Prions are infectious proteins that cause fatal neurodegenerative diseases like Creutzfeldt-Jakob disease (CJD). They convert normal prion proteins (PrP(C)) into disease-causing forms (PrP(Sc)) through conformational changes.
Area of Science:
- Neurobiology
- Infectious Diseases
- Molecular Biology
Background:
- The discovery of prions, infectious proteins, challenged established biological principles.
- Prion diseases, such as Creutzfeldt-Jakob disease (CJD), are characterized by neurodegeneration and lethality.
- Mammalian prions propagate by converting normal cellular prion proteins (PrP(C)) into abnormal, disease-associated isoforms (PrP(Sc)).
Purpose of the Study:
- To introduce the concept of prions and the diseases they cause.
- To explain the mechanism of prion replication and disease pathogenesis.
- To highlight the structural differences between PrP(C) and PrP(Sc).
Main Methods:
- Review of historical discoveries in genetics and infectious agents.
- Analysis of structural differences between normal and abnormal prion protein isoforms.
- Explanation of the conformational conversion process.
Main Results:
- Prions, unlike DNA or conventional pathogens, are proteinaceous infectious agents.
- PrP(Sc) isoforms possess distinct secondary structures (rich in beta-sheet) compared to PrP(C) (rich in alpha-helix).
- The conformational conversion of PrP(C) to PrP(Sc) is the central event in prion disease development.
Conclusions:
- Prion diseases represent a unique class of infectious disorders.
- Understanding prion structure and conversion is key to addressing these neurodegenerative conditions.
- This article provides a foundational overview of prions for researchers and clinicians.
Related Concept Videos
 01:51
Prokaryotic cells
 01:28
Prokaryotic Cells
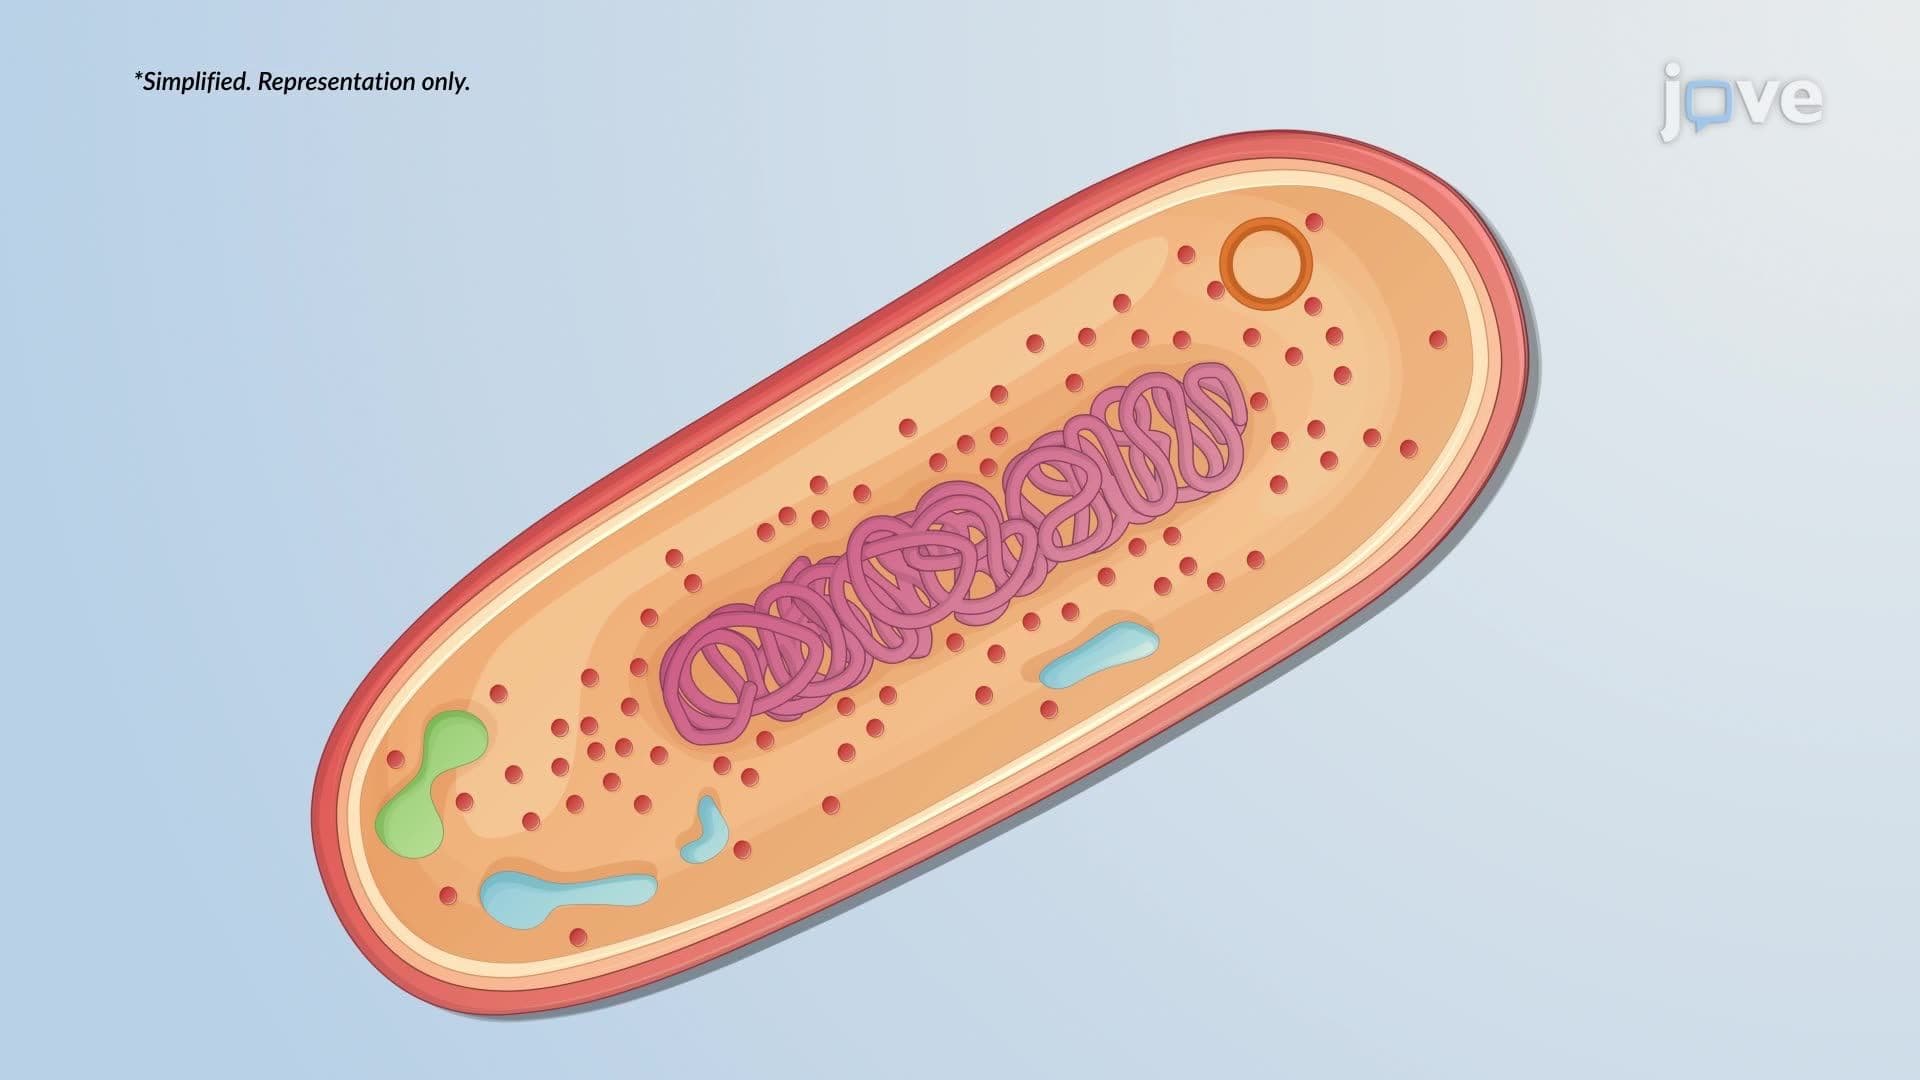
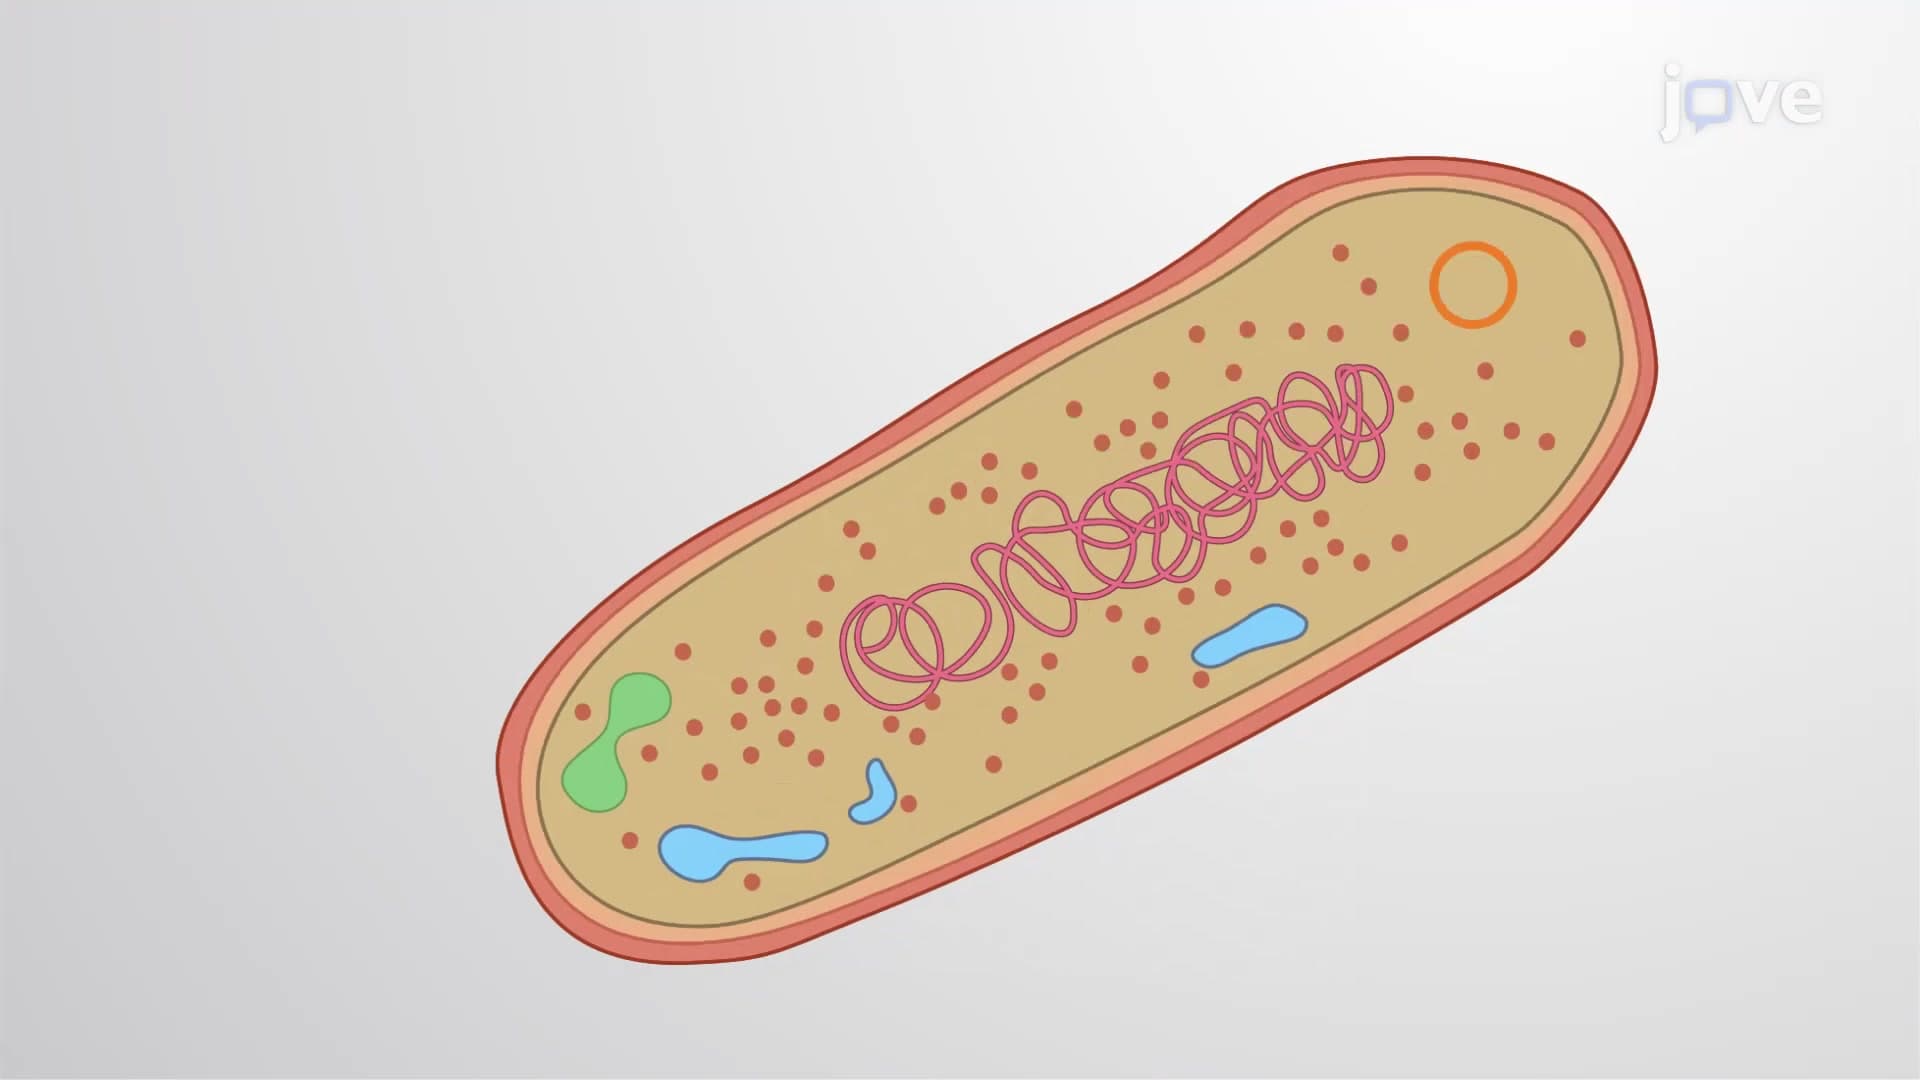
Like eukaryotic cells, all prokaryotic cells are surrounded by a plasma membrane, have genetic material in the form of single, circular DNA, a cytoplasm that fills the interior of the cell, and ribosomes that synthesize proteins.
01:51Prokaryotic Cells
 01:21
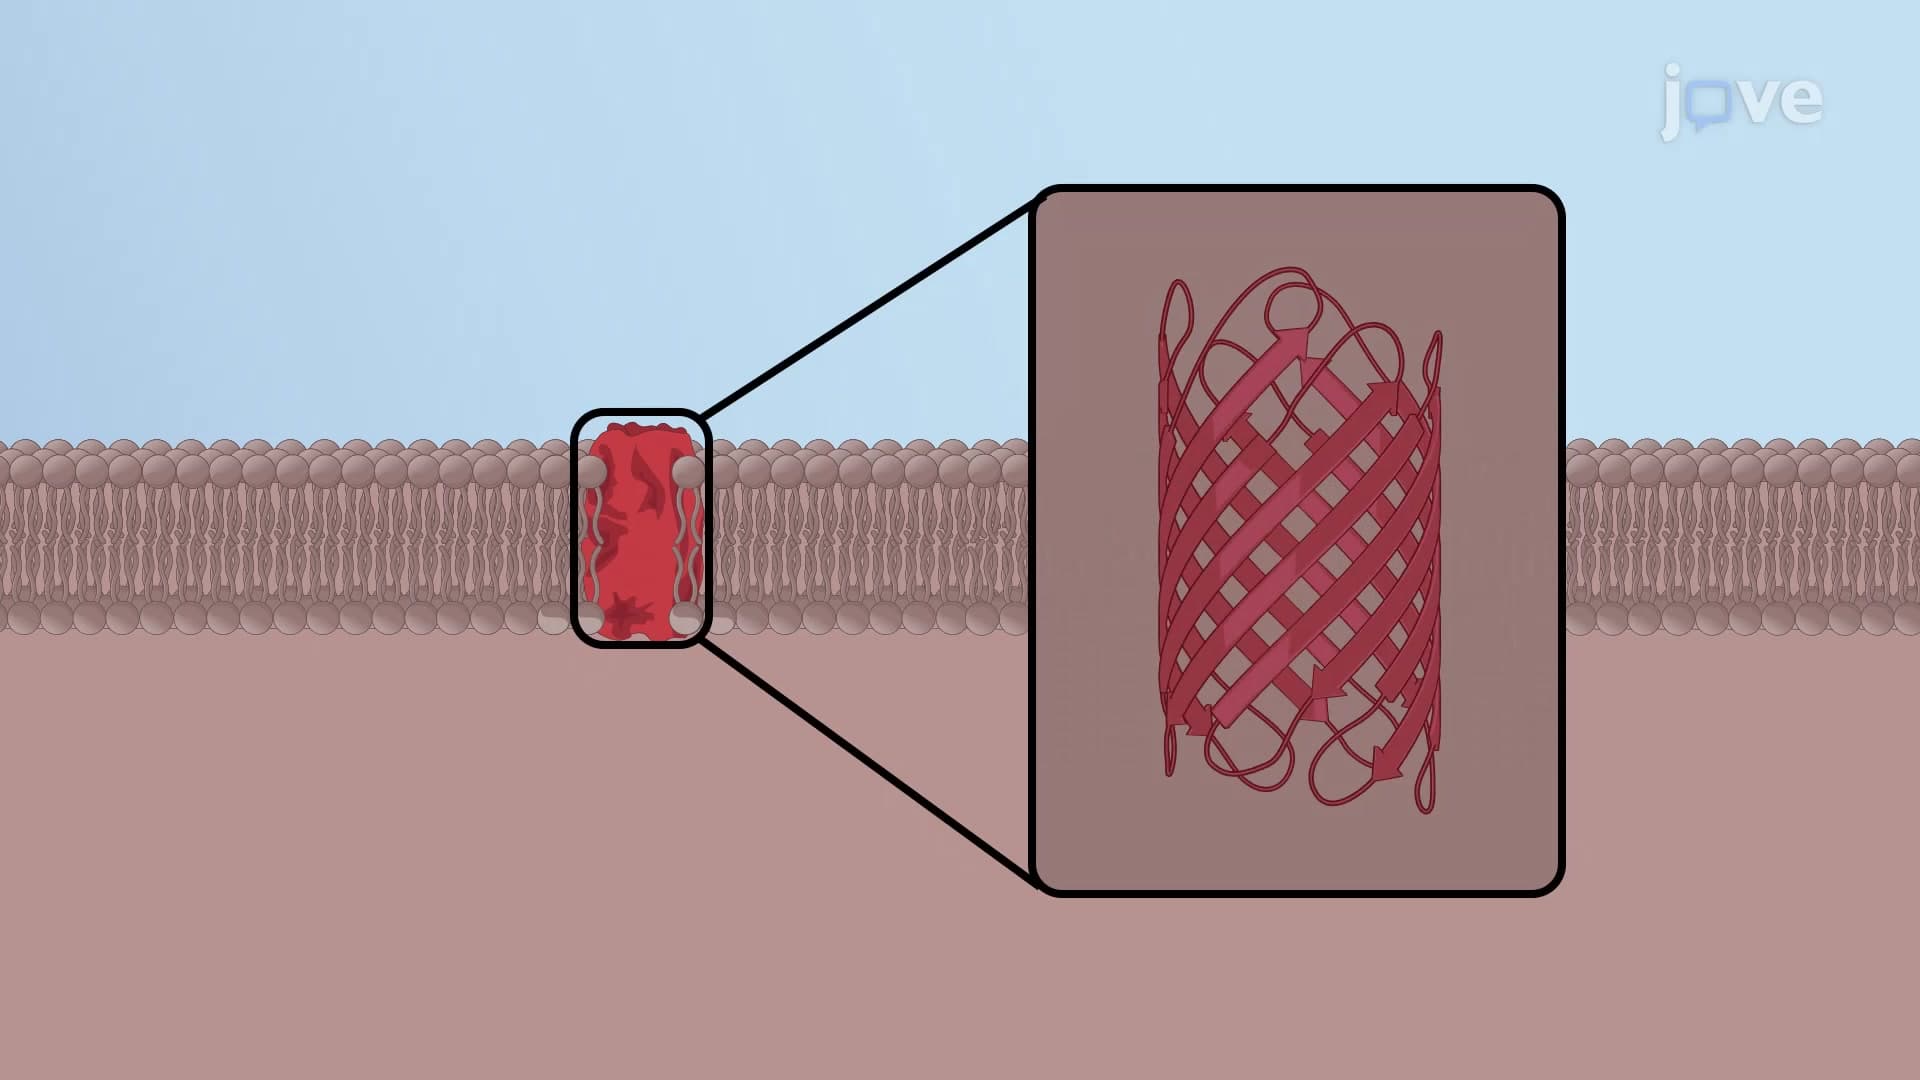
Structure of Porins
 02:57
piRNA - Piwi-interacting RNAs
 01:29
Translation in Prokaryotes