
Molecular dynamics simulations and free energy calculations on the enzyme 4-hydroxyphenylpyruvate dioxygenase
Stephanie B A De Beer1, Alice Glättli, Johannes Hutzler
1Leiden-Amsterdam Center for Drug Research, Section of Molecular Toxicology, Department of Chemistry and Pharmaceutical Sciences, Vrije Universiteit, De Boelelaan 1083, 1081 HV, Amsterdam, The Netherlands.

Abstract:
4-Hydroxyphenylpyruvate dioxygenase is a relevant target in both pharmaceutical and agricultural research. We report on molecular dynamics simulations and free energy calculations on this enzyme, in complex with 12 inhibitors for which experimental affinities were determined. We applied the thermodynamic integration approach and the more efficient one-step perturbation. Even though simulations seem well converged and both methods show excellent agreement between them, the correlation with the experimental values remains poor. We investigate the effect of slight modifications on the charge distribution of these highly conjugated systems and find that accurate models can be obtained when using improved force field parameters. This study gives insight into the applicability of free energy methods and current limitations in force field parameterization.
More Related Videos

Related Concept Videos
 01:19
Enzyme Kinetics
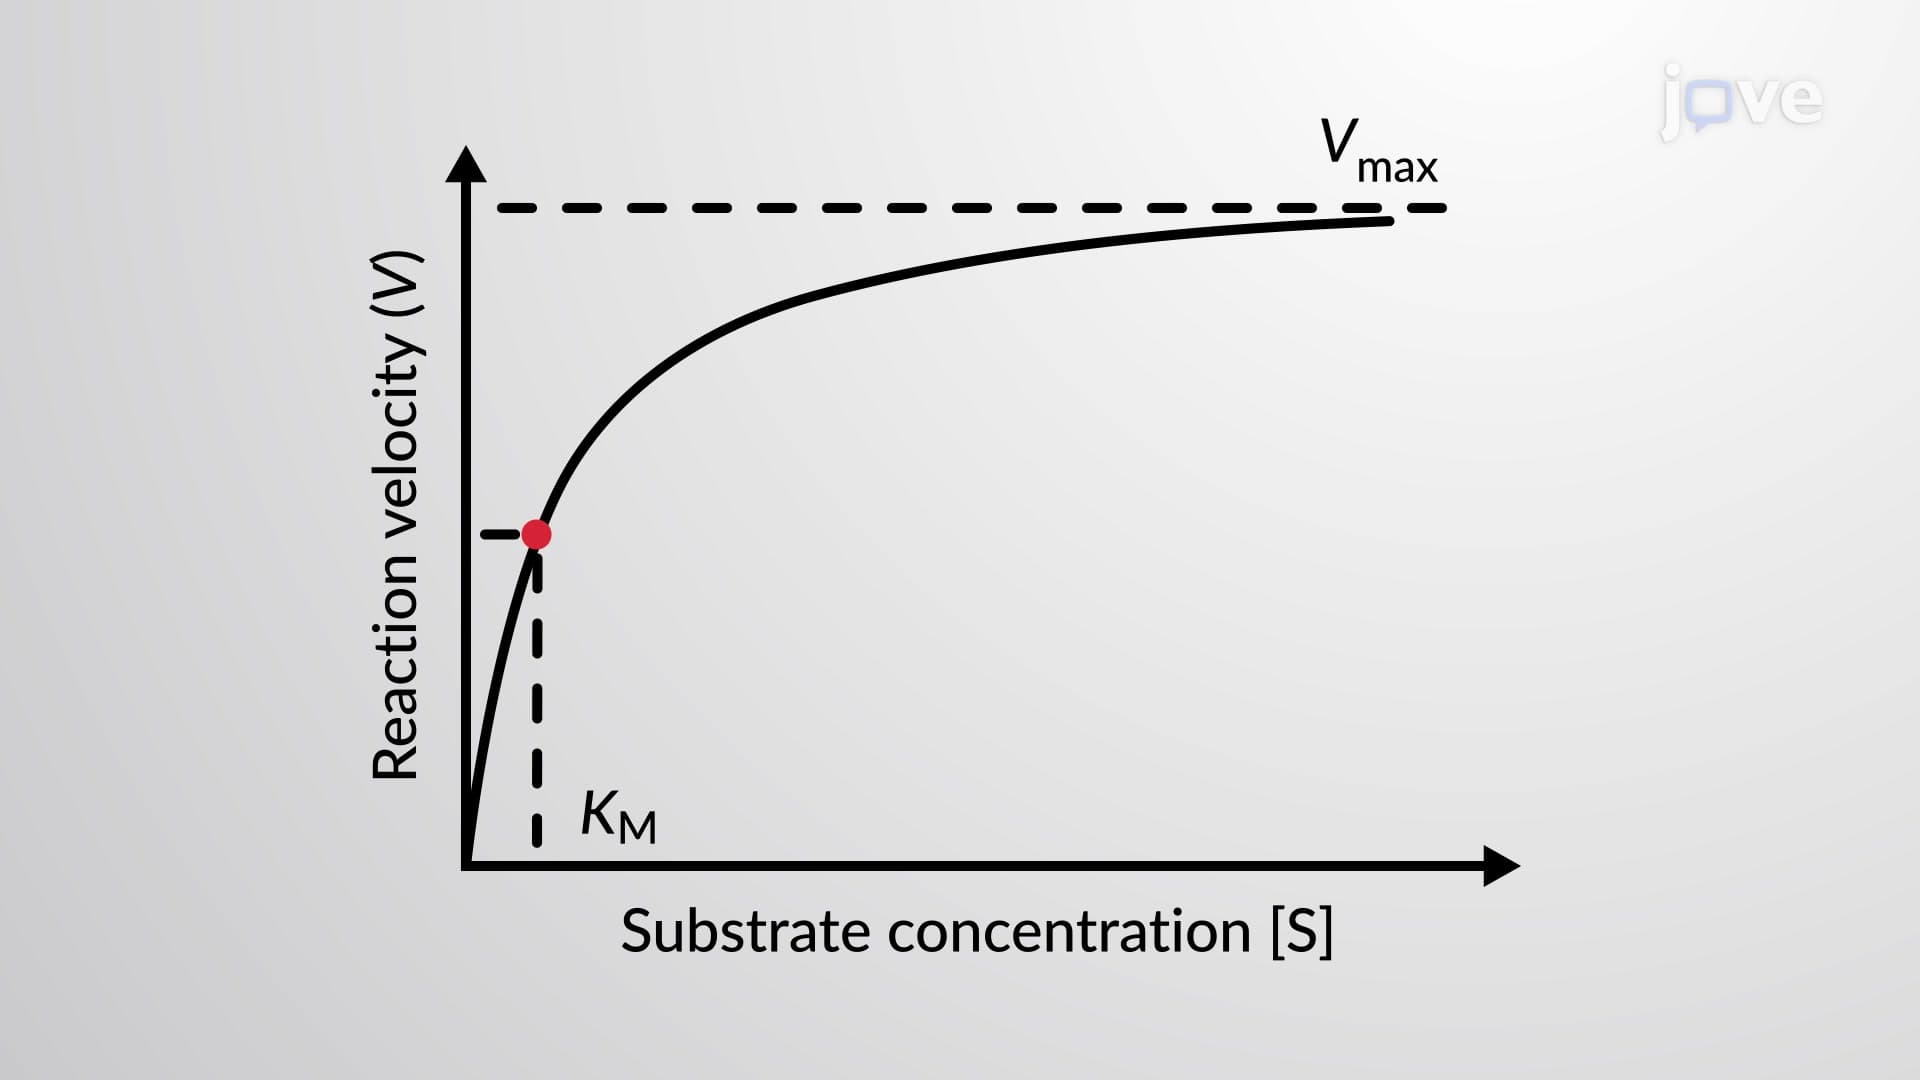
Scientists typically study enzyme kinetics with a fixed amount of enzyme in the controlled environment of a test tube. When more reactant, or substrate, is...
 01:19
Introduction to Enzyme Kinetics
The experimenter can then plot the initial reaction rate or velocity (Vo) of a given trial against the substrate concentration ([S]) to obtain a graph of the reaction properties. For many enzymatic reactions involving a...
 01:07
Catalytically Perfect Enzymes
 02:49
Calculating Standard Free Energy Changes
 01:21
Molecular Kinetic Energy
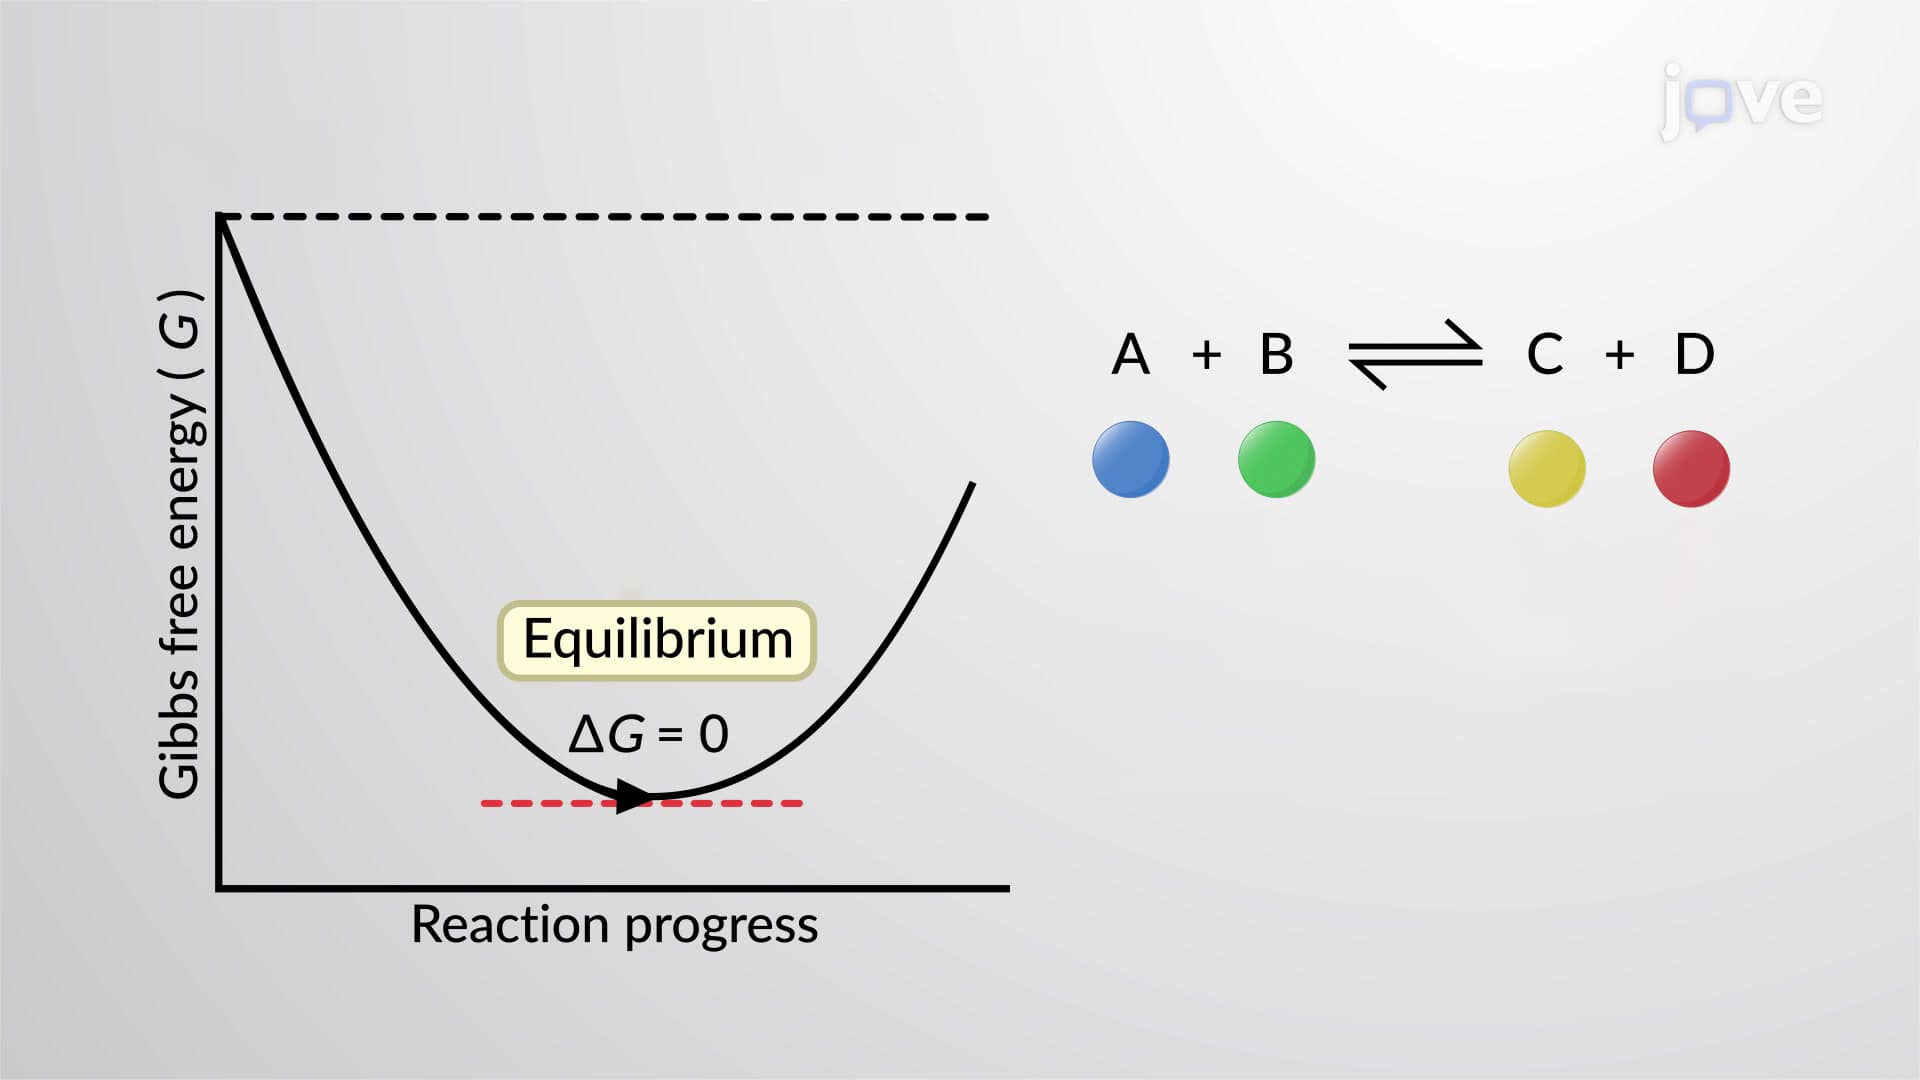 00:55
Free Energy and Equilibrium
The reaction quotient, Q, is a convenient measure of the status of an...