Related Experiment Video
Updated: Jun 2, 2026

Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
Density functional theory study of pyrophyllite and M-montmorillonites (M = Li, Na, K, Mg, and Ca): role of
Vamsee K Voora1, W A Al-Saidi, Kenneth D Jordan
1Department of Chemistry and Center for Molecular and Materials Simulations, University of Pittsburgh, Pittsburgh, Pennsylvania 15260, USA.

Abstract:
The stacking parameters, lattice constants, bond lengths, and bulk moduli of pyrophyllite and montmorillonites (MMTs), with alkali and alkali earth metal ions, are investigated using density functional theory with and without dispersion corrections. For pyrophyllite, it is found that the inclusion of the dispersion corrections significantly improves the agreement of the calculated values of the lattice parameters and bulk modulus with the experimental values. For the MMTs, the calculations predict that the interlayer spacing varies approximately linearly with the cation radius. The inclusion of dispersion corrections leads to sizable shifts of the interlayer spacings to shorter values. In Li-MMT, compaction of the interlayer distance triggers migration of the Li ion into the tetrahedral sheet and close coordination with basal oxygen atoms. Analysis of electron density distributions shows that the isomorphic octahedral Al(3+)/Mg(2+) substitution in MMT causes an increase of electron density on the basal oxygen atoms of the tetrahedral sheets.
Related Concept Videos
 01:24
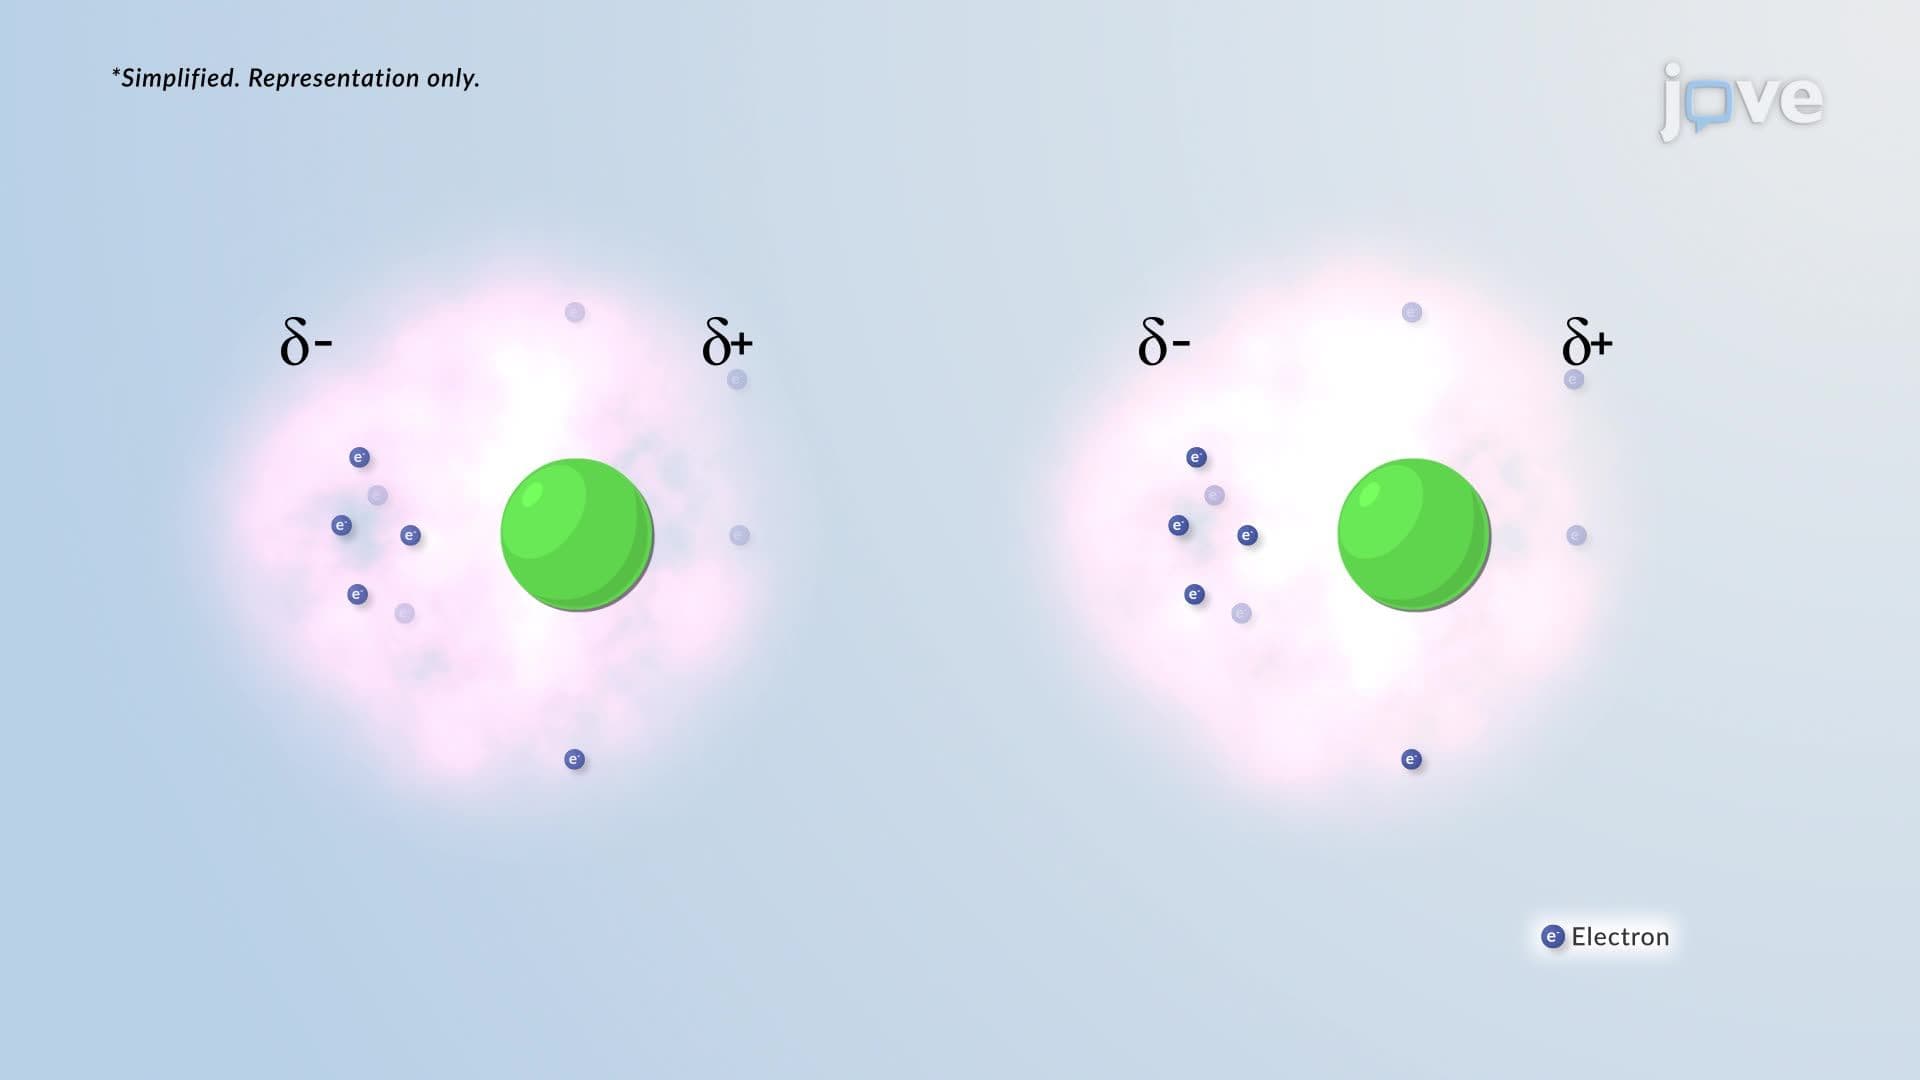
Van der Waals Interactions
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:56
Intermolecular Forces and Physical Properties
 01:29
The Colloidal State