Computational methods for transcriptome annotation and quantification using RNA-seq
Related Experiment Video
Updated: Jun 1, 2026
Rup (RNA-seq Usability Assessment Pipeline) - Quality Control for Bulk RNA-seq Experiments in Eukaryotes
Published on: November 7, 2025
Manuel Garber1, Manfred G Grabherr, Mitchell Guttman
1Broad Institute of Massachusetts Institute of Technology and Harvard, Cambridge, Massachusetts, USA. mgarber@broadinstitute.org

Abstract:
High-throughput RNA sequencing (RNA-seq) promises a comprehensive picture of the transcriptome, allowing for the complete annotation and quantification of all genes and their isoforms across samples. Realizing this promise requires increasingly complex computational methods. These computational challenges fall into three main categories: (i) read mapping, (ii) transcriptome reconstruction and (iii) expression quantification. Here we explain the major conceptual and practical challenges, and the general classes of solutions for each category. Finally, we highlight the interdependence between these categories and discuss the benefits for different biological applications.
Related Concept Videos
 03:21
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while microarray-based...
 02:24
Ribosome Profiling
Applications of ribosome profiling
Ribosome profiling has many applications, including in vivo monitoring of translation inside a particular organ or tissue type and quantifying new protein synthesis levels.
The technique helps...
 02:57
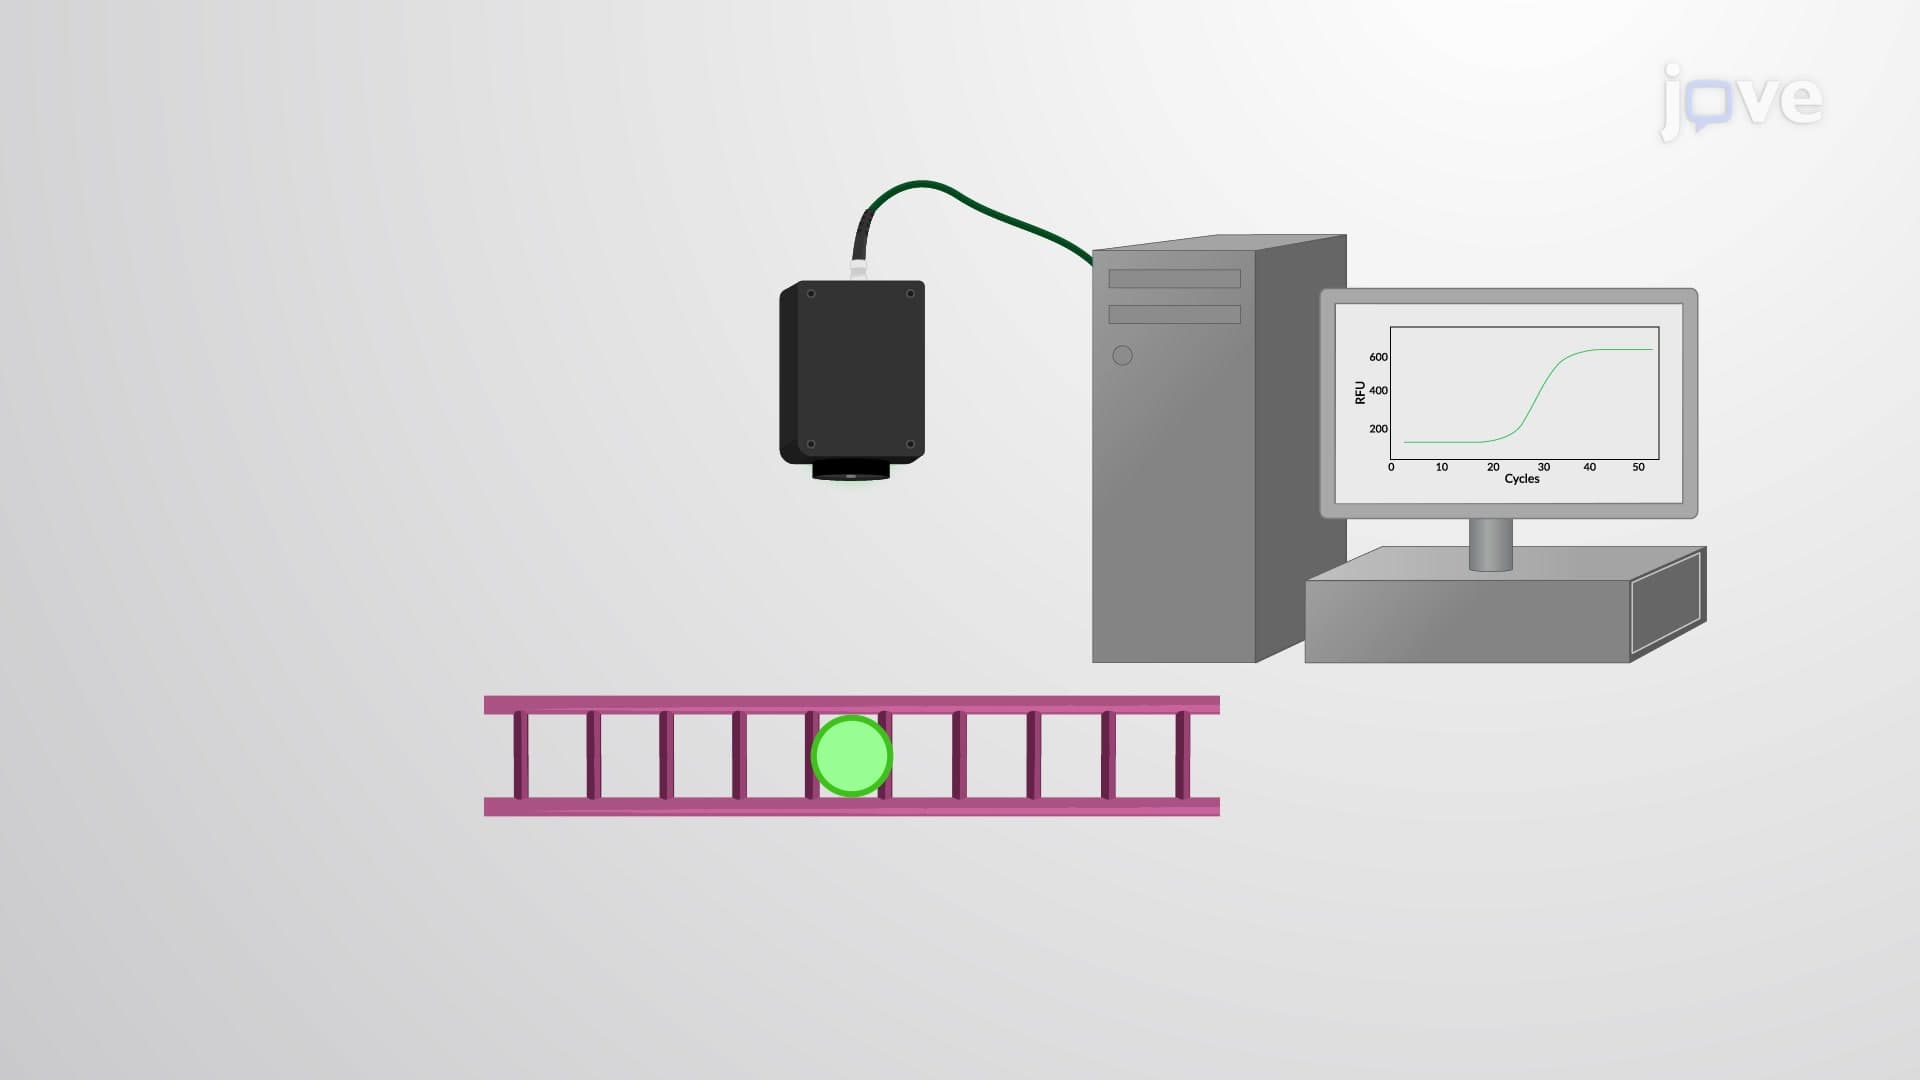
Real Time RT-PCR
The real-time quantification of the number of amplified products is...