Related Experiment Video
Updated: May 31, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Using free energy of binding calculations to improve the accuracy of virtual screening predictions
Robert D Malmstrom1, Stanley J Watowich
1Department of Biochemistry and Molecular Biology, Sealy Center for Structural Biology and Molecular Biophysics, University of Texas Medical Branch, Galveston, Texas 77555, USA.

Abstract:
Virtual screening of small molecule databases against macromolecular targets was used to identify binding ligands and predict their lowest energy bound conformation (i.e., pose). AutoDock4-generated poses were rescored using mean-field pathway decoupling free energy of binding calculations and evaluated if these calculations improved virtual screening discrimination between bound and nonbound ligands. Two small molecule databases were used to evaluate the effectiveness of the rescoring algorithm in correctly identifying binders of L99A T4 lysozyme. Self-dock calculations of a database containing compounds with known binding free energies and cocrystal structures largely reproduced experimental measurements, although the mean difference between calculated and experimental binding free energies increased as the predicted bound poses diverged from the experimental poses. In addition, free energy rescoring was more accurate than AutoDock4 scores in discriminating between known binders and nonbinders, suggesting free energy rescoring could be a useful approach to reduce false positive predictions in virtual screening experiments.

Related Concept Videos
 02:18
The Equilibrium Binding Constant and Binding Strength
02:18The Equilibrium Binding Constant and Binding Strength
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
01:49Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
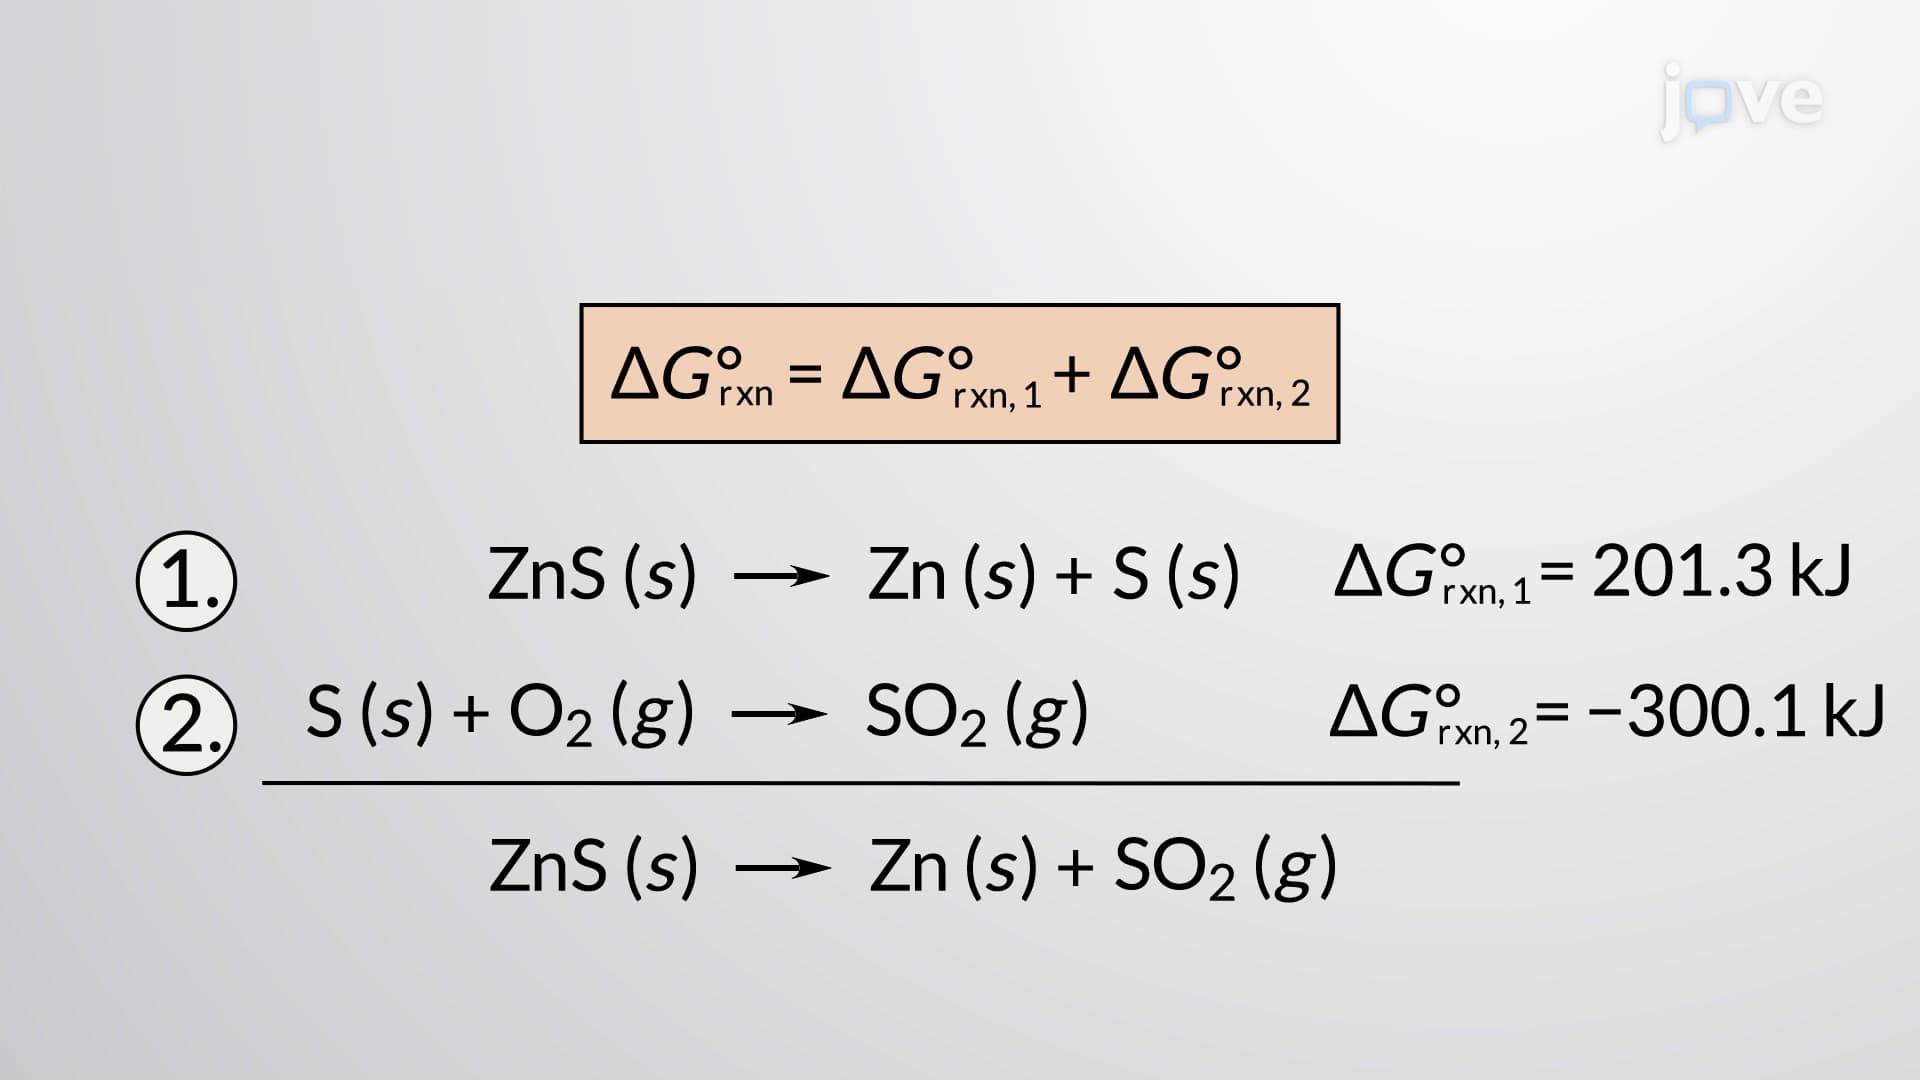 02:49
Calculating Standard Free Energy Changes