A characterization study on 2,6-dimethyl-4-nitropyridine N-oxide by density functional theory calculations
Related Experiment Video
Updated: May 31, 2026
Syntheses, Crystallization, and Spectroscopic Characterization of 3,5-Lutidine N-Oxide Dehydrate
Published on: April 24, 2018
G Yildirim1, Y Zalaoglu, C Kirilmis
1Abant Izzet Baysal University, Department of Physics, Bolu 14280, Turkey.

Abstract:
This study deals with the identification of a title compound, 2,6-dimethyl-4-nitropyridine N-oxide by means of theoretical calculations. The optimized molecular structures, vibrational frequencies, corresponding vibrational assignments, thermodynamic properties and atomic charges of the title compound in the ground state were evaluated using density functional theory (DFT) with the standard B3LYP/6-311G(d,p) method and basis set combination for the first time. Theoretical vibrational spectra were interpreted with the aid of normal coordinate analysis based on scaled density functional force field. The results show that the optimized geometric parameters (bond lengths and bond angles) and vibrational frequencies were observed to be in good agreement with the available experimental results. Based on the results of comparison between experimental results and theoretical data, the chosen calculation level is powerful approach for understanding the molecular structures and vibrational spectra of the 2,6-dimethyl-4-nitropyridine N-oxide. Moreover, we not only simulated frontier molecular orbitals (FMO) and molecular electrostatic potential (MEP) but also determined the transition state and energy band gap. Based on the investigations, the title compound is found to be useful to bond metallically and interact intermolecularly. Infrared intensities and Raman activities were also reported.
More Related Videos
Related Concept Videos
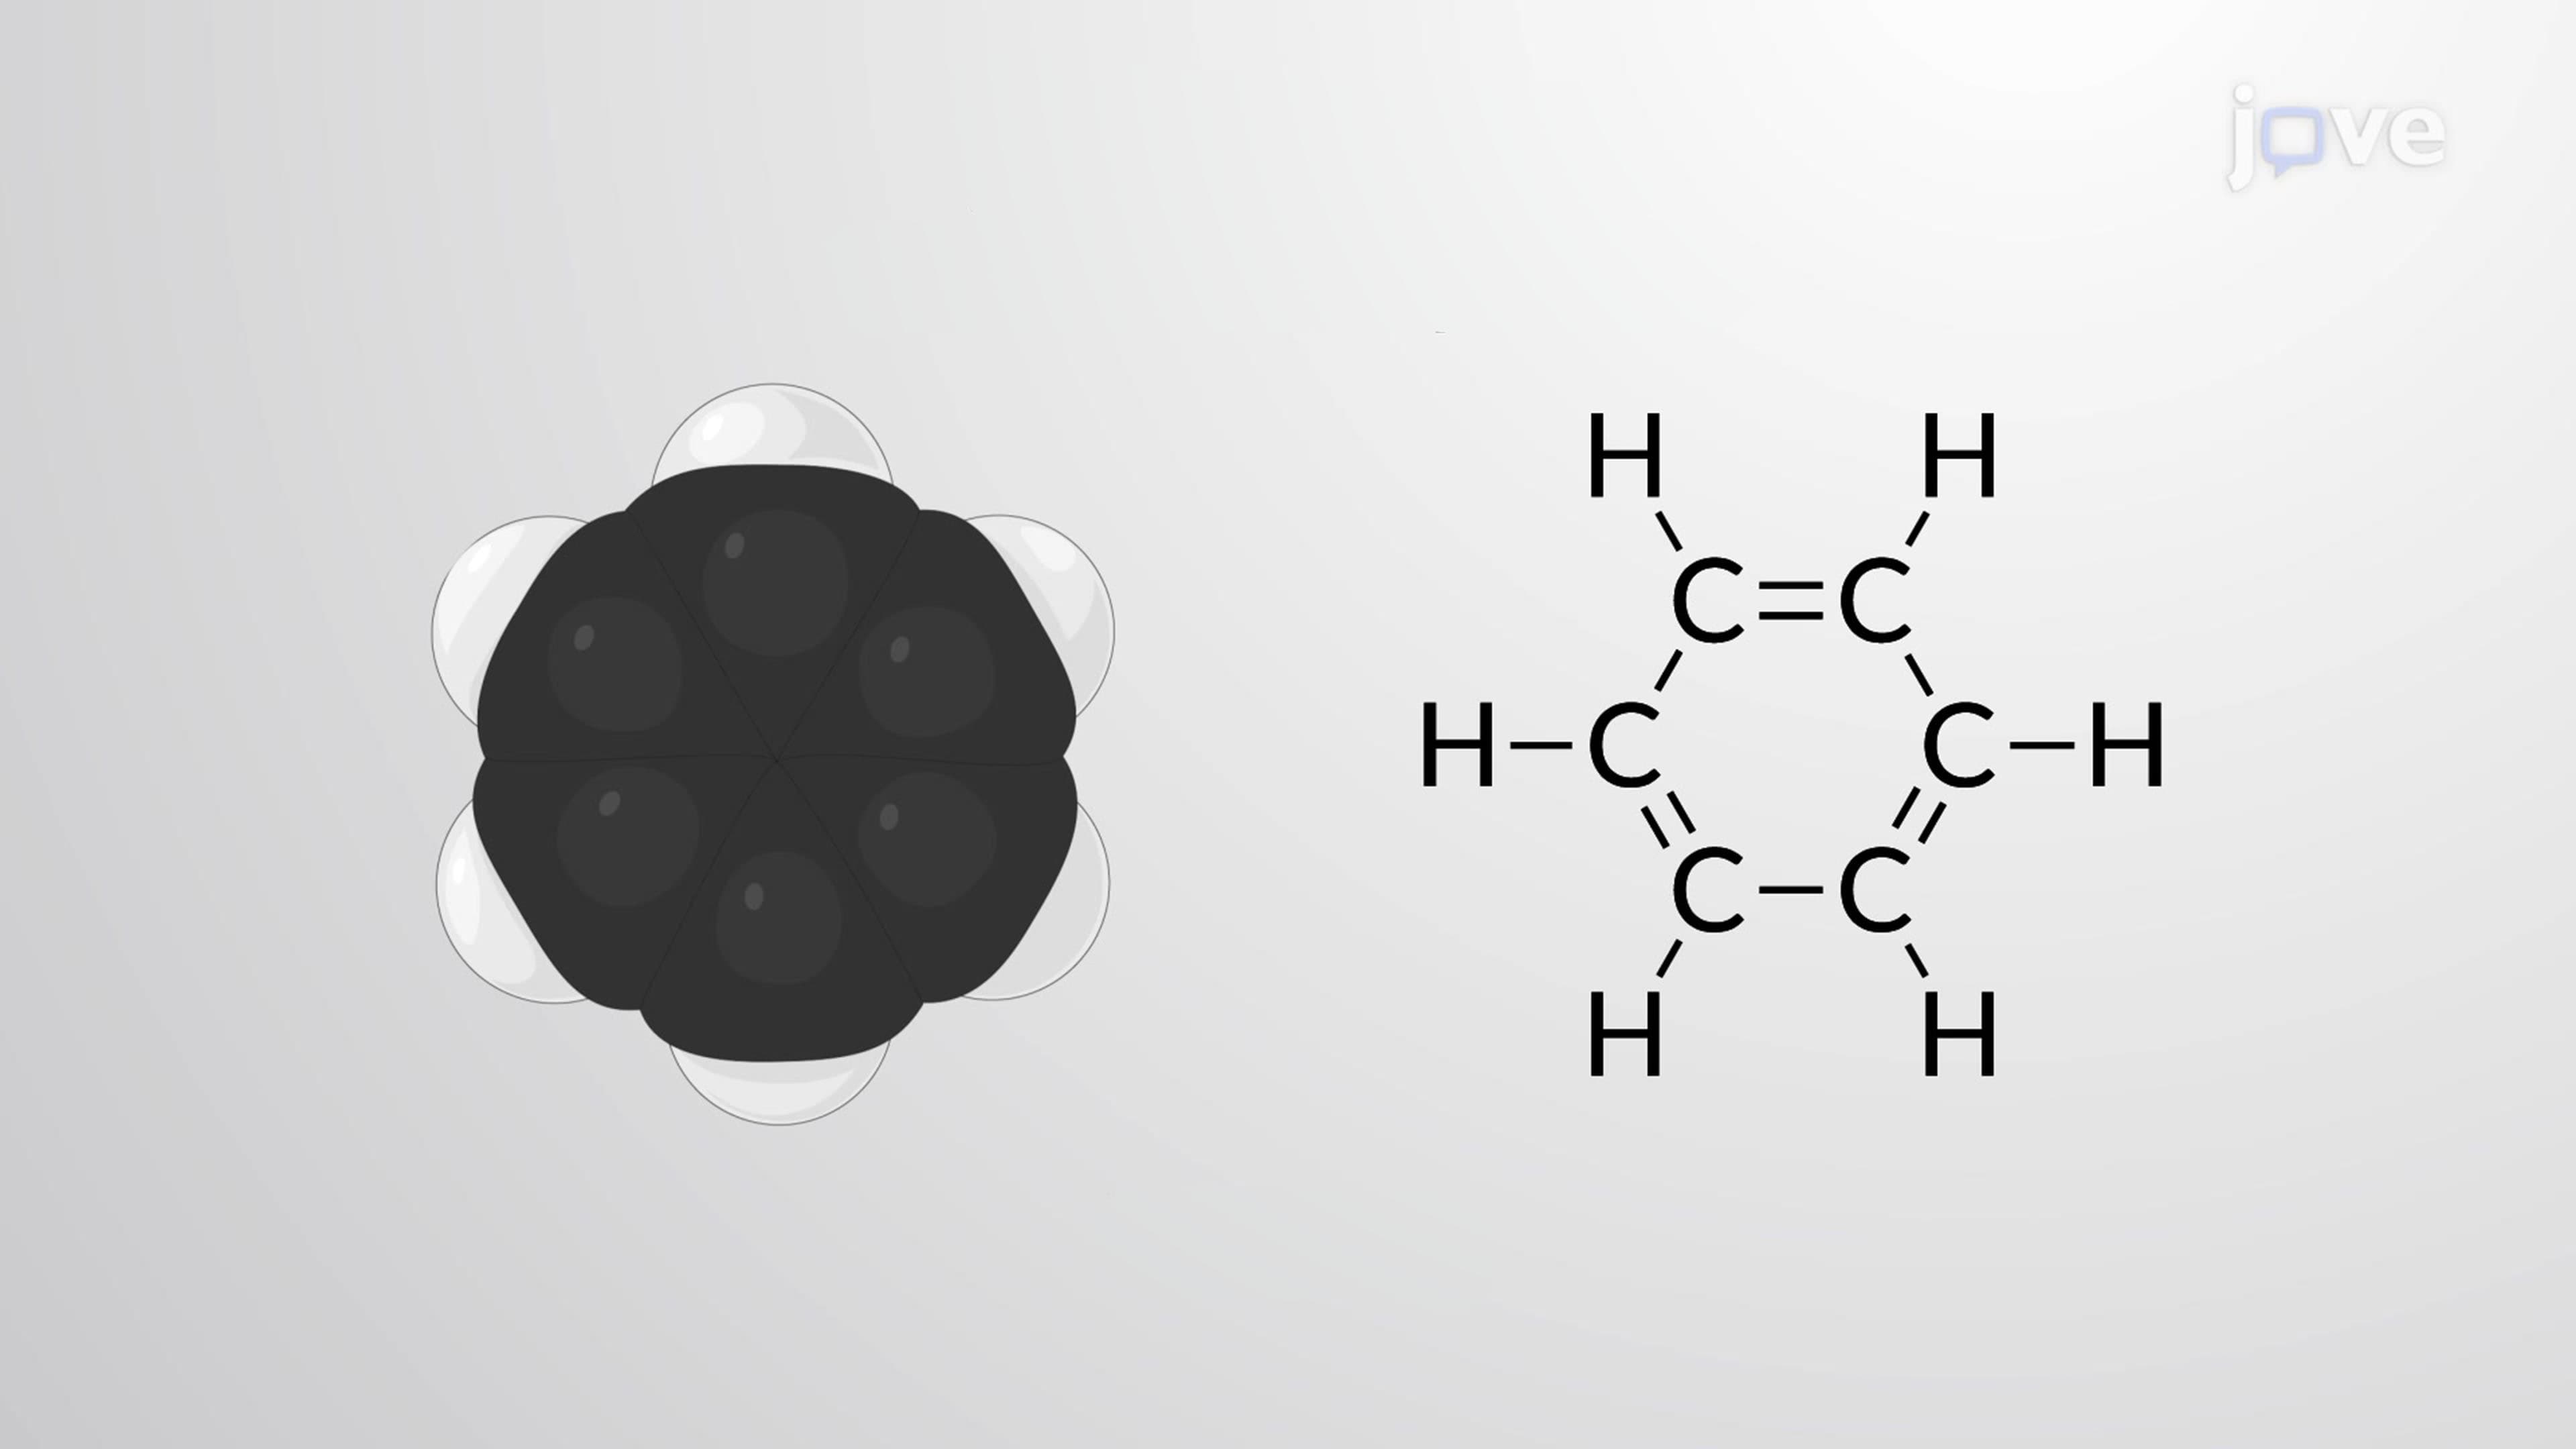 02:52
Resonance
 02:54
Lewis Structures of Molecular Compounds and Polyatomic Ions
 02:16
Resonance and Hybrid Structures
Resonance Structures and Resonance Hybrids
The Lewis structure of a nitrite anion (NO2−) may actually be drawn in two different ways, distinguished by the locations of the N–O and N=O bonds.
 02:40
MO Theory and Covalent Bonding
 03:51
Molecular Orbital Theory II
 02:42
Formal Charges