Related Experiment Video
Updated: May 30, 2026

Realistic Membrane Modeling Using Complex Lipid Mixtures in Simulation Studies
Published on: September 1, 2023
SAFT-γ force field for the simulation of molecular fluids. 1. A single-site coarse grained model of carbon dioxide
Carlos Avendaño1, Thomas Lafitte, Amparo Galindo
1Department of Chemical Engineering, Centre for Process Systems Engineering, Imperial College London, South Kensington Campus, London SW7 2AZ, UK.

Abstract:
An application of the "top-down" concept for the development of accurate coarse-grained intermolecular potentials of complex fluids is presented. With the more common "bottom-up" procedure, coarse-grained models are constructed from a suitable simplification of a detailed atomistic representation, and small adjustments to the intermolecular parameters are made by comparison with limited experimental data where necessary. In contrast, in the top-down approach, a molecular-based equation of state is used to obtain an effective coarse-grained intermolecular potential that reproduces the macroscopic experimental thermophysical properties over a wide range of conditions. These coarse-grained intermolecular potentials can then be used in conventional molecular simulation to obtain properties (such as structure or dynamics) that are not directly accessible from the equation of state or at extreme conditions where the theory is expected to fail. To demonstrate our procedure, a coarse-grained model for carbon dioxide (CO(2)) is obtained from a recent implementation of the statistical associating fluid theory of variable range (SAFT-VR) employing a Mie (generalized Lennard-Jones) potential; the parameters of this single-site Mie model of CO(2) are estimated by optimizing the equation of state's description of the experimental vapor-pressure and saturated liquid density data. This approach is only viable because of the excellent agreement of the SAFT-VR Mie EoS with simulation data. Our single-site SAFT-γ coarse-grained model for CO(2) is used in Monte Carlo molecular simulation to assess the adequacy of the description of the fluid-phase behavior and properties that were not used to develop the potential model such as the enthalpy of vaporization, interfacial tension, density profiles, supercritical densities, and second-derivative thermodynamic properties (thermal expansivity, isothermal compressibility, heat capacity, Joule-Thompson coefficient, and speed of sound). The accuracy of the description with the single-site SAFT-γ model of CO(2) is found to be of similar quality to that of more sophisticated intermolecular potentials such as a six-site (three LJ centers and three charged sites) all-atom model. The SAFT-γ top-down approach to coarse-graining resolves a key challenge with coarse-graining techniques: the provision of a direct robust link between the microscopic and macroscopic scales.
Related Concept Videos
 01:34
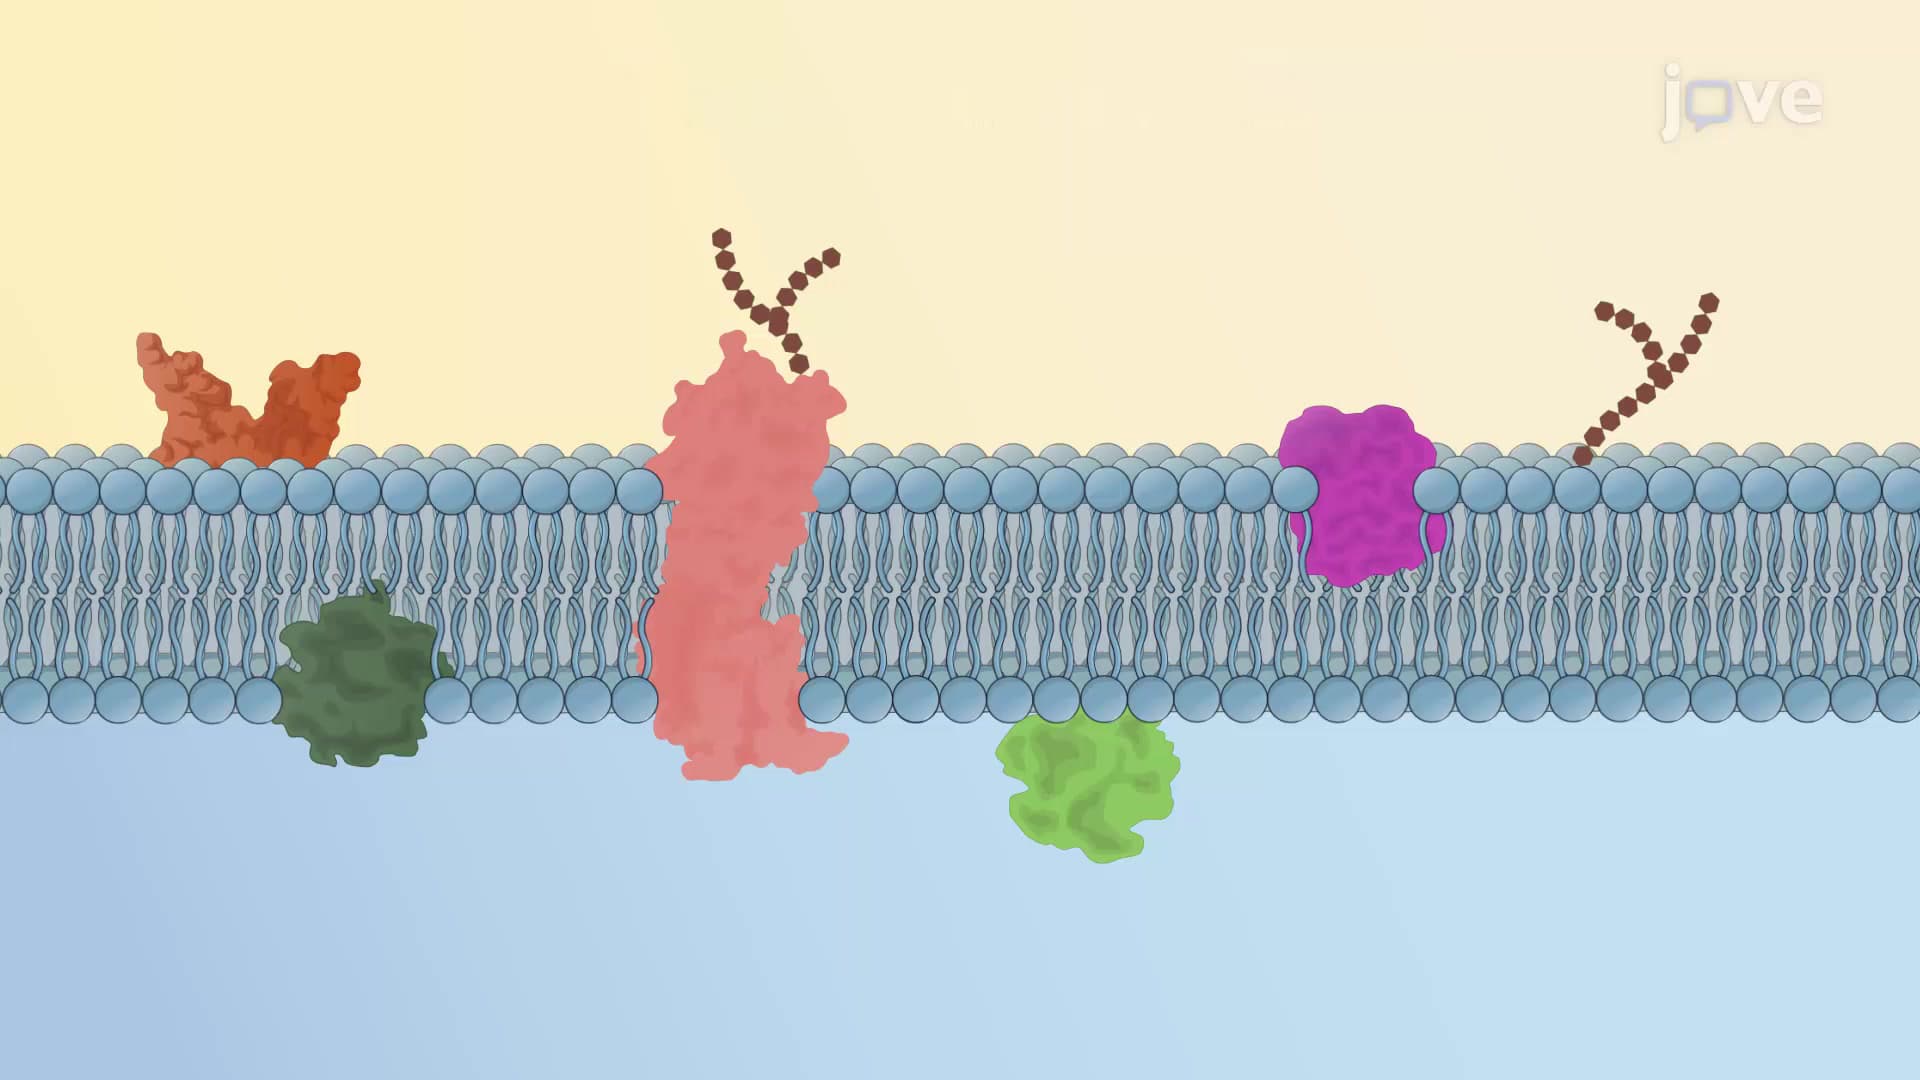
The Fluid Mosaic Model
 01:19
Fluid Mosaic Model
 02:28
Intermolecular Forces in Solutions
When the strengths of the intermolecular forces of attraction between solute and solvent species in a solution are no different than those present in the separated components, the solution is formed with no accompanying energy change. Such a solution is called an ideal solution. A mixture of ideal gases (or gases such as helium and argon,...
 03:13
Intermolecular Forces
03:13Intermolecular Forces
 02:26
Molecular Comparison of Gases, Liquids, and Solids