Related Experiment Video
Updated: May 30, 2026
Palladium N-Heterocyclic Carbene Complexes: Synthesis from Benzimidazolium Salts and Catalytic Activity in Carbon-carbon Bond-forming Reactions
Published on: July 30, 2017
Experimental and theoretical evidence of aromatic behavior in heterobenzene-like molecules with metal-metal multiple
Wenjuan Fang1, Qiao He, Zhong Fei Tan
1Department of Chemistry, Tongji University, Shanghai, 200092, P.R. China.

Abstract:
Binding two quadruply bonded dimolybdenum units [Mo(2)(DAniF)(3)](+) (DAniF=N,N'-di-p-anisylformamidinate) with two chalcogen atoms generated two molecules with a central core composed of a cyclic six-membered [Mo(2)](2)(μ-EH)(2) species (E=S in 1 and O in 3, and [Mo(2)] is a quadruple-bonded [Mo(2)(formamidinate)(3)] unit). Aerobic oxidation of 1 and 3 followed by concomitant deprotonation gave rise to the corresponding [Mo(2)](2)(μ-E)(2) compounds 2 and 4. The latter show a striking coplanarity and near-bond equalization of the Mo/E cluster. The oxidized species 2 and 4 are diamagnetic in the measured temperature range of 5 to 300 K, which is somewhat unexpected for molecules that have dimetal units with a σ(2)π(4)δ(1) electronic configuration. This suggests there are strong interactions between the dimolybdenum units through the E atoms. The large electronic delocalization of the δ electrons over the entire Mo/E core is supported by the exceptionally large potential separation for the two successive one-electron reductions of the linked Mo(2)(5+) units from the oxidized species (ΔE(½)=1.7 V for the sulfur analogue). This large electronic delocalization has an important effect on the NMR spectroscopic signals for the two sets of methine (N-(CH)-N) protons from the DAniF ligands. Those essentially parallel to the core, H(∥), and those essentially perpendicular to the core, H(⊥), exhibit downfield and upfield chemical shifts, respectively, that are separated by δ=1.32 ppm. The structural, electronic, magnetic, and chemical behaviors for 2 and 4 are consistent with aromaticity, with the [Mo(2)E(2)Mo(2)] cores that resemble the prototypical benzene molecule. Theoretical studies, including DFT calculations, natural bond orbital (NBO) analyses, and gauge-independent atomic orbital (GIAO) NMR spectroscopic calculations, are also consistent with the aromaticity of the [Mo(2)](2)(μ-E)(2) units being promoted by d(δ)(Mo(2))-p(π)(E) π conjugation. The cyclic π conjugation of the central moiety in 2 and 4 involves a total of six electrons with 2e from δ(Mo(2)) and 4e from p(π)(E) orbitals, thereby conforming to Hückel's rule when electrons in the MOs with δ character are considered part of the delocalized system.
Related Concept Videos
 01:18
Frost Circles for Different Conjugated Systems
 01:20
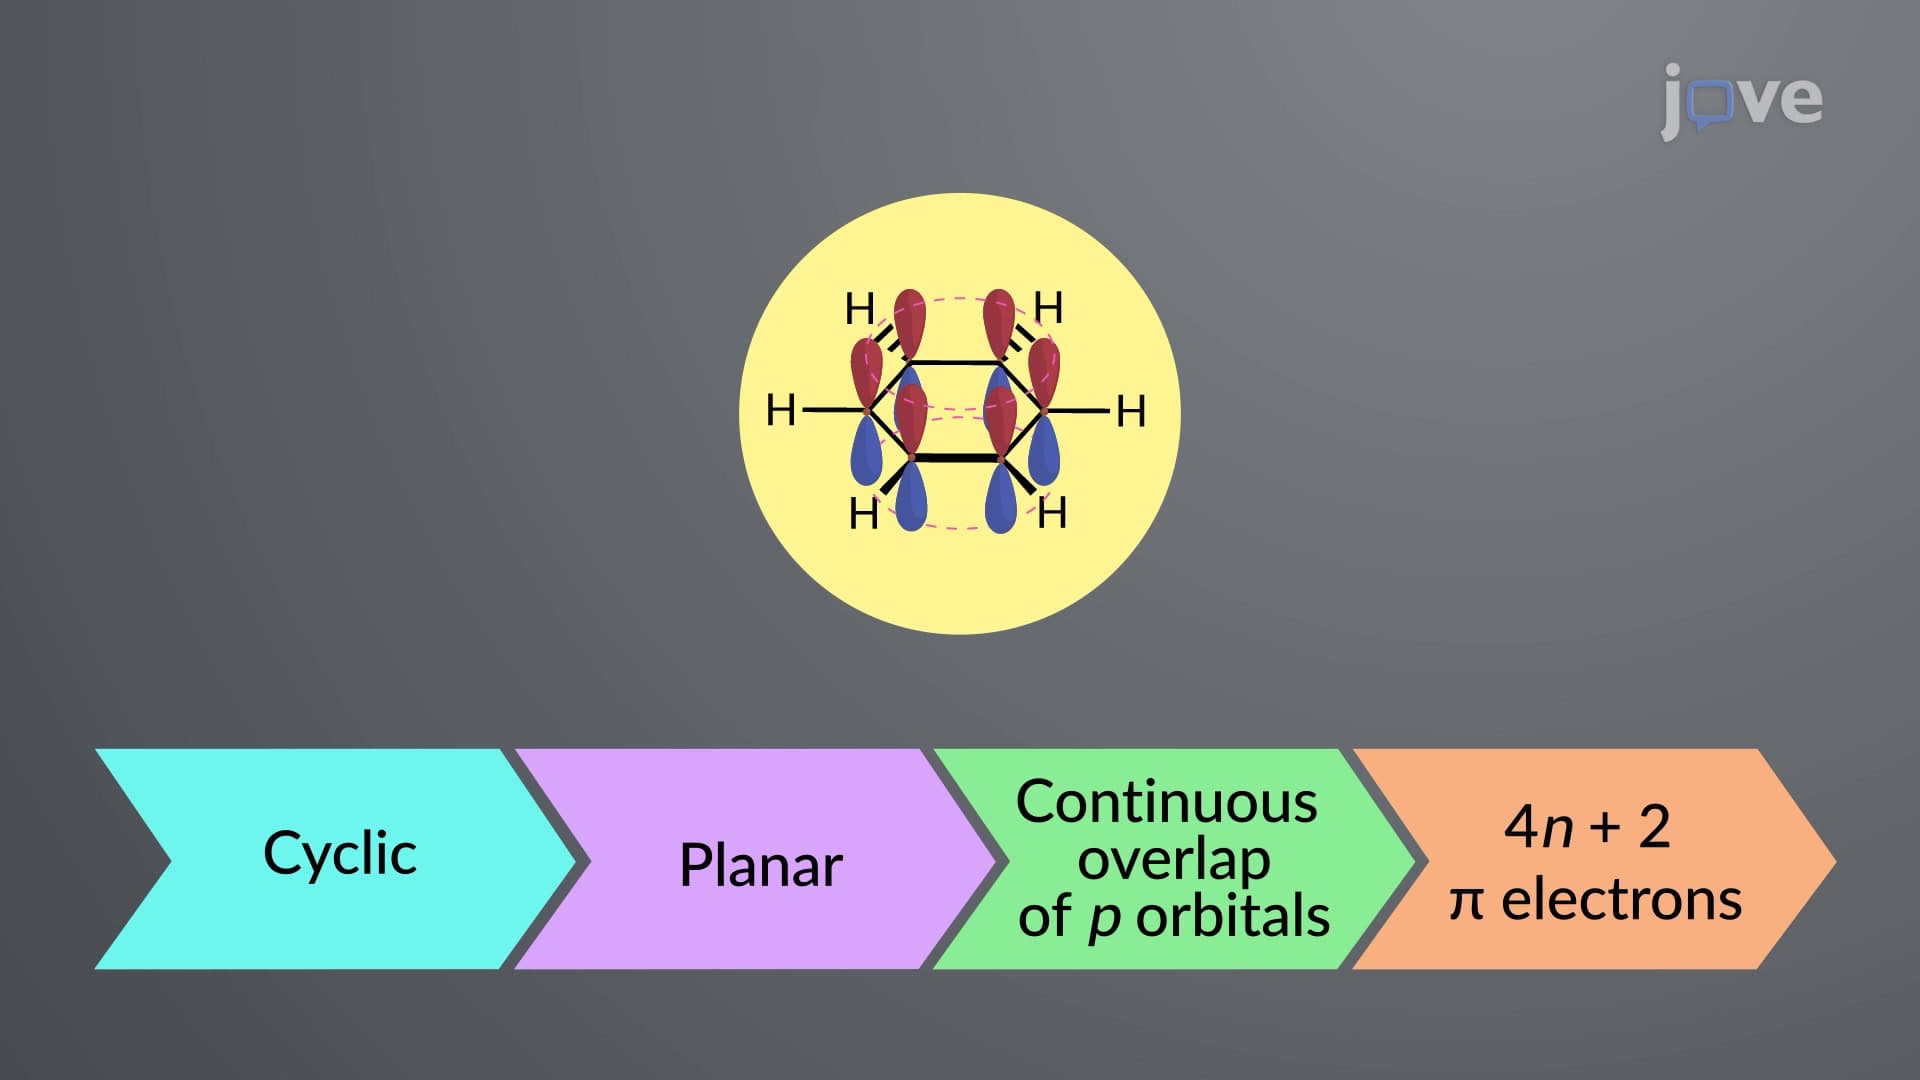
Criteria for Aromaticity and the Hückel 4n + 2 Rule
For the first time, Eric Hückel, a German chemical physicist, derived a set of structural features for a compound to be classified as aromatic. This is now known as Hückel’s rule or the 4n + 2 rule.
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:18
Structure of Benzene: Molecular Orbital Model
 01:37
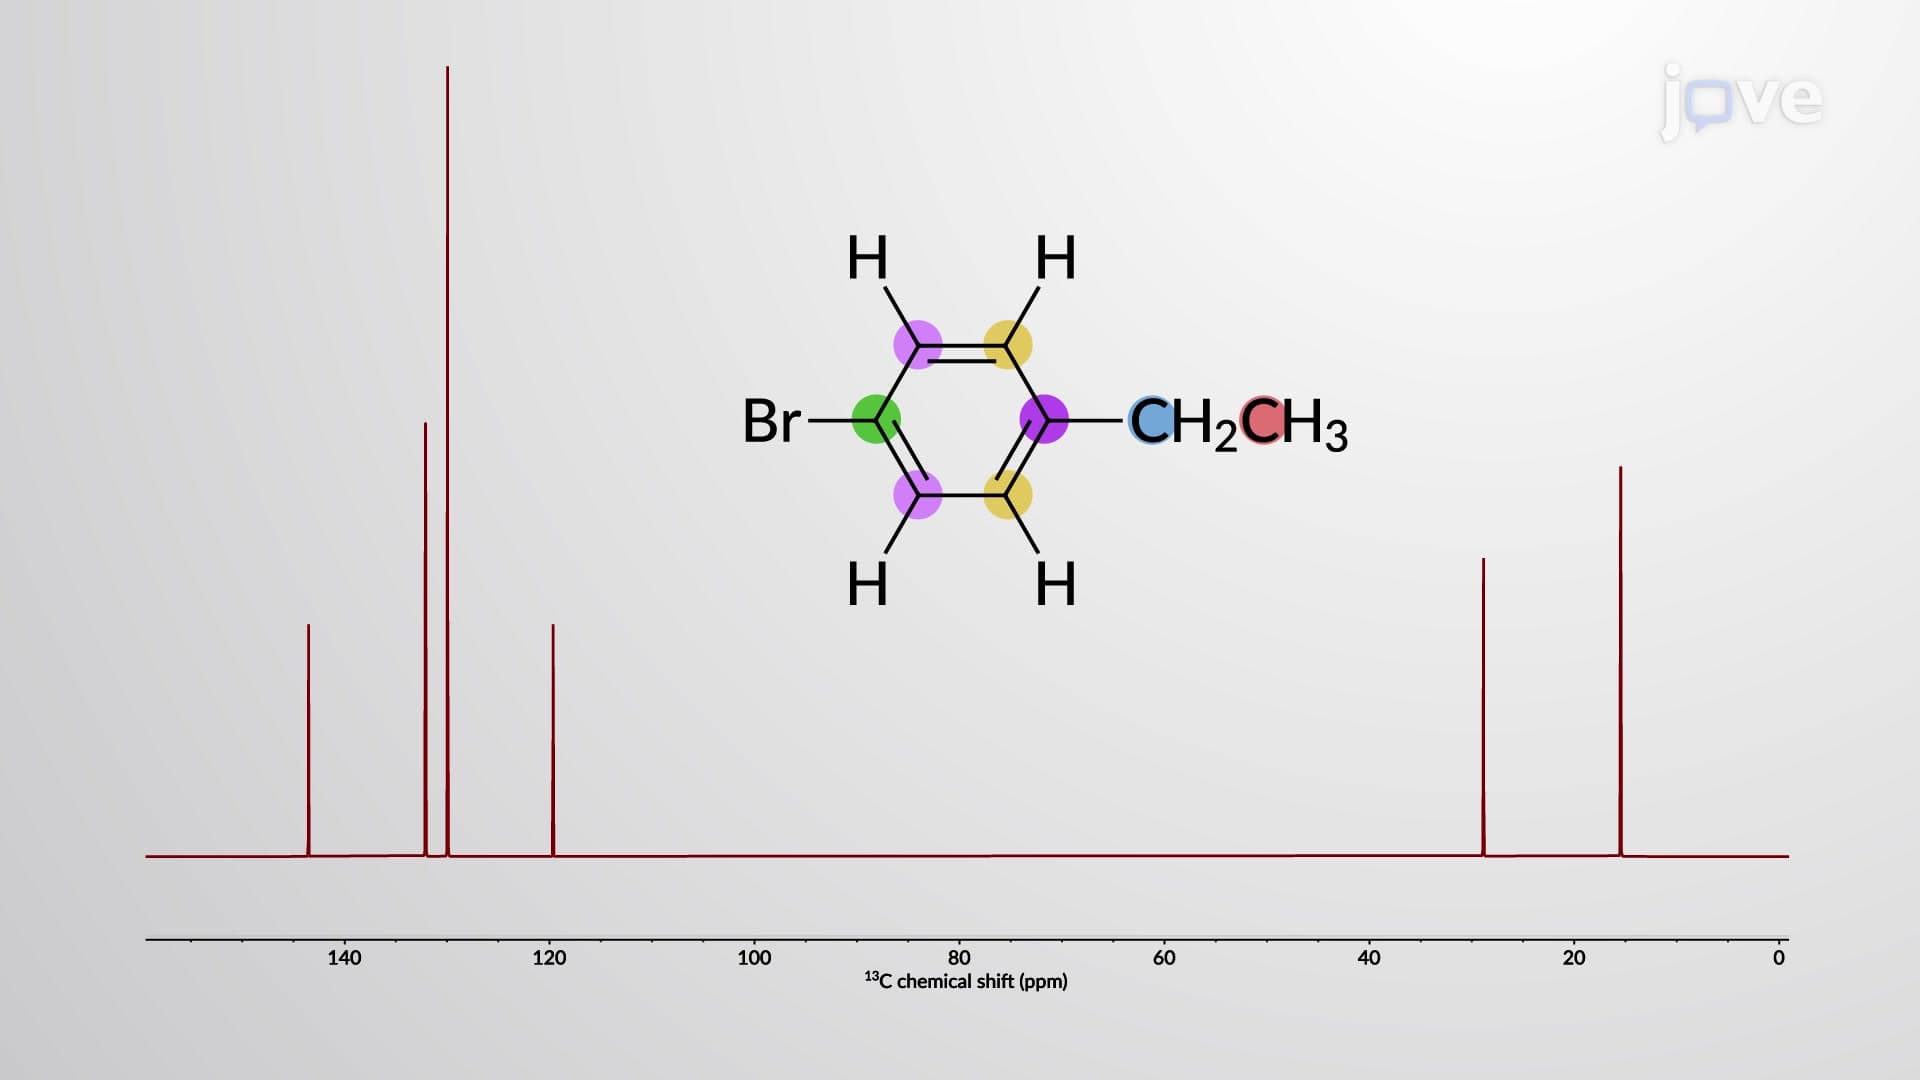
NMR Spectroscopy of Benzene Derivatives
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene π orbitals.