Related Experiment Video
Updated: May 28, 2026

Developing Photosensitizer-Cobaloxime Hybrids for Solar-Driven H2 Production in Aqueous Aerobic Conditions
Published on: October 5, 2019
Spectroscopic and electronic structure studies probing covalency contributions to C-H bond activation and
Joseph Sempombe1, Benjamin Stein, Martin L Kirk
1Department of Chemistry and Chemical Biology, The University of New Mexico, MSC03 2060, 1 University of New Mexico, Albuquerque, New Mexico 87131-0001, USA.

Abstract:
A detailed electron paramagnetic resonance (EPR) and computational study of a key paramagnetic form of xanthine oxidase (XO) has been performed and serves as a basis for developing a valence-bond description of C-H activation and transition-state (TS) stabilization along the reaction coordinate with aldehyde substrates. EPR spectra of aldehyde-inhibited XO have been analyzed in order to provide information regarding the relationship between the g, (95,97)Mo hyperfine (A(Mo)), and (13)C hyperfine (A(C)) tensors. Analysis of the EPR spectra has allowed for greater insight into the electronic origin of key delocalizations within the Mo-O(eq)-C fragment and how these contribute to C-H bond activation/cleavage and TS stabilization. A natural bond orbital analysis of the enzyme reaction coordinate with aldehyde substrates shows that both Mo═S π → C-H σ* (ΔE = 24.3 kcal mol(-1)) and C-H σ → Mo═S π* (ΔE = 20.0 kcal mol(-1)) back-donation are important in activating the substrate C-H bond for cleavage. Additional contributions to C-H activation derive from O(eq) lp → C-H σ* (lp = lone pair; ΔE = 8.2 kcal mol(-1)) and S lp → C-H σ* (ΔE = 13.2 kcal mol(-1)) stabilizing interactions. The O(eq)-donor ligand that derives from water is part of the Mo-O(eq)-C fragment probed in the EPR spectra of inhibited XO, and the observation of O(eq) lp → C-H σ* back-donation indicates a key role for O(eq) in activating the substrate C-H bond for cleavage. We also show that the O(eq) donor plays an even more important role in TS stabilization. We find that O(eq) → Mo + C charge transfer dominantly contributes to stabilization of the TS (ΔE = 89.5 kcal mol(-1)) and the Mo-O(eq)-C delocalization pathway reduces strong electronic repulsions that contribute to the classical TS energy barrier. The Mo-O(eq)-C delocalization at the TS allows for the TS to be described in valence-bond terms as a resonance hybrid of the reactant (R) and product (P) valence-bond wave functions.

Related Concept Videos
 01:19
Oxidation and Reduction of Organic Molecules
The removal of an electron from a molecule, results in a...
 01:15
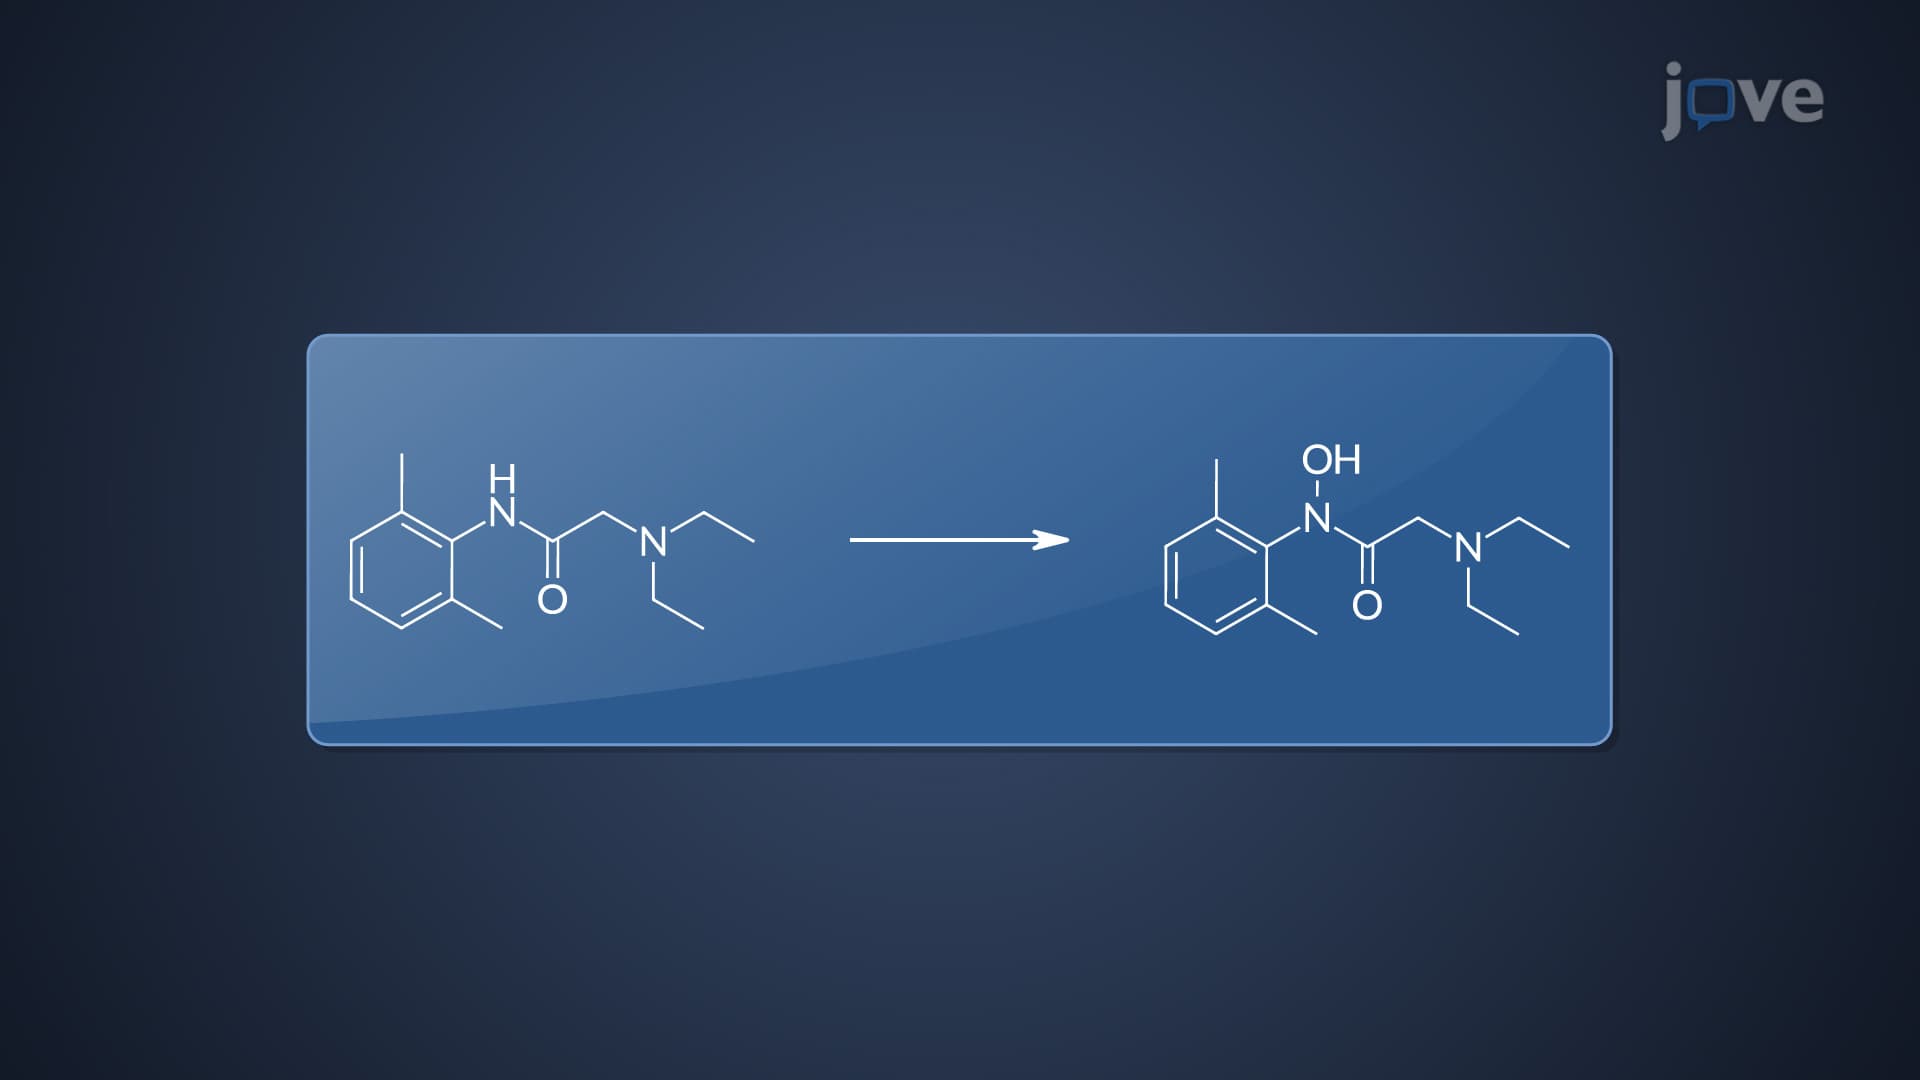
Phase I Reactions: Oxidation of Carbon-Heteroatom and Miscellaneous Systems
In carbon-nitrogen systems, aliphatic and aromatic amines can undergo oxidative reactions. Secondary and tertiary amines, like those found in tricyclic antidepressants, can undergo N-dealkylation, a process that involves the oxidation of the alkyl group. In addition, oxidative...
 01:46
Oxidative Cleavage of Alkenes: Ozonolysis
Ozone is a symmetrical bent molecule stabilized by a resonance structure.
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
Selection Rules: Photochemical Activation
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:19
Phase I Oxidative Reactions: Overview