CCL2 upregulation triggers hypoxic preconditioning-induced protection from stroke
Related Experiment Video
Updated: May 24, 2026
Quantification of Neurovascular Protection Following Repetitive Hypoxic Preconditioning and Transient Middle Cerebral Artery Occlusion in Mice
Published on: May 4, 2015
Ann M Stowe1, Bradley K Wacker, Petra D Cravens
1Department of Neurological Surgery, Washington University School of Medicine, 660 S, Euclid Ave,, Box 8057, St, Louis, MO 63110, USA.

Background:
A brief exposure to systemic hypoxia (i.e., hypoxic preconditioning; HPC) prior to transient middle cerebral artery occlusion (tMCAo) reduces infarct volume, blood-brain barrier disruption, and leukocyte migration. CCL2 (MCP-1), typically regarded as a leukocyte-derived pro-inflammatory chemokine, can also be directly upregulated by hypoxia-induced transcription. We hypothesized that such a hypoxia-induced upregulation of CCL2 is required for HPC-induced ischemic tolerance.
Methods:
Adult male SW/ND4, CCL2-null, and wild-type mice were used in these studies. Cortical CCL2/CCR2 message, protein, and cell-type specific immunoreactivity were determined following HPC (4 h, 8% O2) or room air control (21% O2) from 6 h through 2 weeks following HPC. Circulating leukocyte subsets were determined by multi-parameter flow cytometry in naïve mice and 12 h after HPC. CCL2-null and wild-type mice were exposed to HPC 2 days prior to tMCAo, with immunoneutralization of CCL2 during HPC achieved by a monoclonal CCL2 antibody.
Results:
Cortical CCL2 mRNA and protein expression peaked at 12 h after HPC (both p < 0.01), predominantly in cortical neurons, and returned to baseline by 2 days. A delayed cerebral endothelial CCL2 message expression (p < 0.05) occurred 2 days after HPC. The levels of circulating monocytes (p < 0.0001), T lymphocytes (p < 0.0001), and granulocytes were decreased 12 h after HPC, and those of B lymphocytes were increased (p < 0.0001), but the magnitude of these respective changes did not differ between wild-type and CCL2-null mice. HPC did decrease the number of circulating CCR2+ monocytes (p < 0.0001) in a CCL2-dependent manner, but immunohistochemical analyses at this 12 h timepoint indicated that this leukocyte subpopulation did not move into the CNS. While HPC reduced infarct volumes by 27% (p < 0.01) in wild-type mice, CCL2-null mice subjected to tMCAo were not protected by HPC. Moreover, administration of a CCL2 immunoneutralizing antibody prior to HPC completely blocked (p < 0.0001 vs. HPC-treated mice) the development of ischemic tolerance.
Conclusions:
The early expression of CCL2 in neurons, the delayed expression of CCL2 in cerebral endothelial cells, and CCL2-mediated actions on circulating CCR2+ monocytes, appear to be required to establish ischemic tolerance to focal stroke in response to HPC, and thus represent a novel role for this chemokine in endogenous neurovascular protection.
More Related Videos
 07:46
Bilateral Common Carotid Artery Occlusion as an Adequate Preconditioning Stimulus to Induce Early Ischemic Tolerance to Focal Cerebral Ischemia
Published on: May 9, 2013
 10:13
Experimental Models to Study the Neuroprotection of Acidic Postconditioning Against Cerebral Ischemia
Published on: July 31, 2017
Related Concept Videos
 01:15
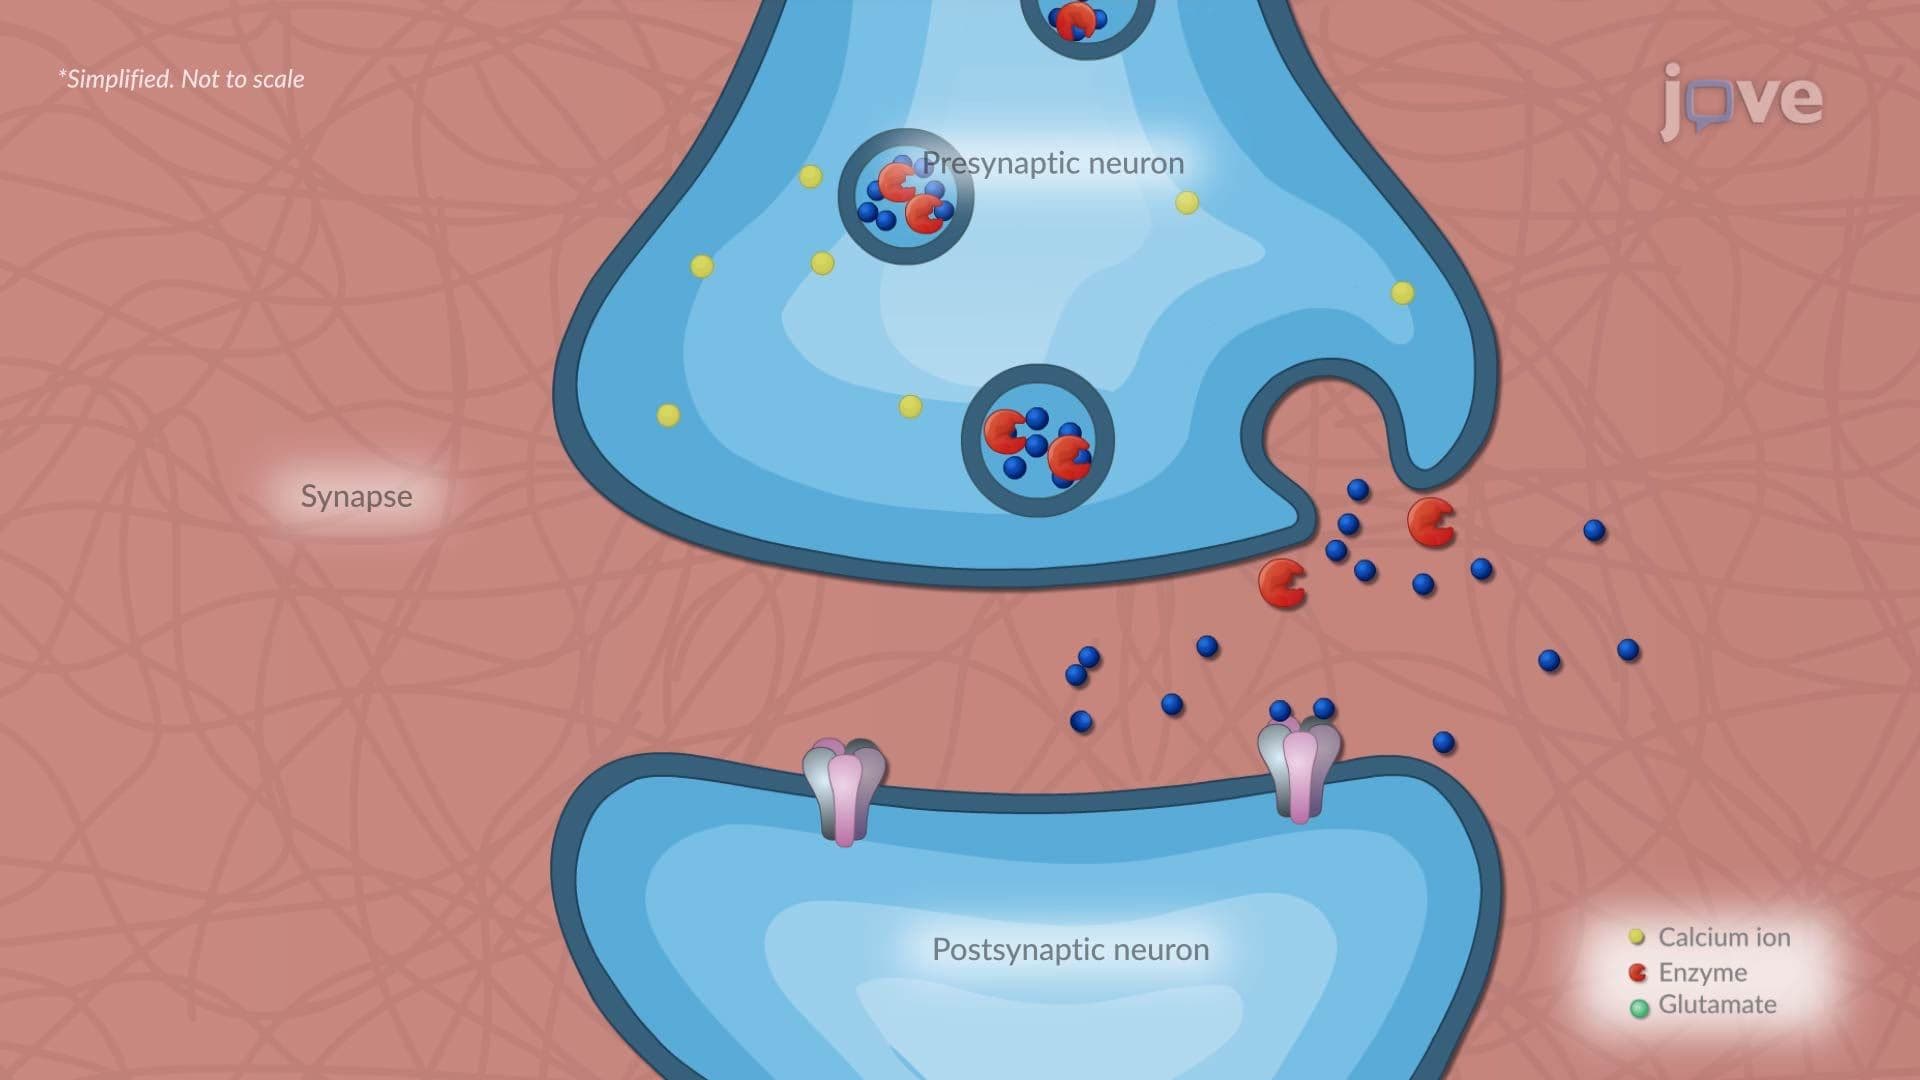
Ischemic Stroke ll: Pathophysiology
 01:15
Ischemic Stroke l: Introduction