

Electronic structure, molecular electrostatic potential and spectral characteristics of pillar[6]arene hosts and
Swarada R Peerannawar1, Shridhar P Gejji
1Department of Chemistry, University of Pune, Pune 411 007, India. spgejji@chem.unipune.ac.in.

Abstract:
Electronic structure, charge distribution and (1)H NMR in pillar[6]arene (P6) conformers, their diisobutoxy derivatives and their host-guest complexes have been investigated by employing the density functional theory. It has been shown that a P6 conformer obtained by flipping of alternate hydroquinone units turns out to be of lowest energy, owing to the hydrogen bonded network at both rims of the host. As opposed to this, a conformer void of hydrogen bonding interactions has largely been destabilized. The O-HO interactions are analyzed using molecular electrostatic potential topography as a tool. Modification of a P6 host by substituting a diisobutoxy group at reactive phenols (DIBP6) renders rigid pillar-shape architecture to the host in which electron-rich regions are localized within the cavity and near portals. Complexation of n-octyltriethylammonium ions (n-OTEA) with P6 and DIBP6 reveals qualitatively different binding patterns. It has been shown that the conformer in which n-OTEA penetrates from the lower rim of the host and partially encapsulates within the P6 cavity turns out to be 1.4 kJ mol(-1) lower in energy than the complex showing complete guest encapsulation. Host-guest binding patterns, viz. encapsulation or portal interactions, can be distinguished from (1)H NMR chemical shifts. The shielding of ethyl and n-octyl chain protons in an n-OTEA⊂DIBP6 complex points to encapsulation of the guest which has been rationalized from natural bond orbital analyses. These inferences are in consonance with (1)H NMR experiments.
More Related Videos
 06:35
Construction and Systematical Symmetric Studies of a Series of Supramolecular Clusters with Binary or Ternary Ammonium Triphenylacetates
Published on: February 15, 2016
 12:08
Catalytic Reactions at Amine-Stabilized and Ligand-Free Platinum Nanoparticles Supported on Titania During Hydrogenation of Alkenes and Aldehydes
Published on: June 24, 2022
Related Concept Videos
 01:19
Structure of Amines
 01:18
π Molecular Orbitals of the Allyl Cation and Anion
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:18
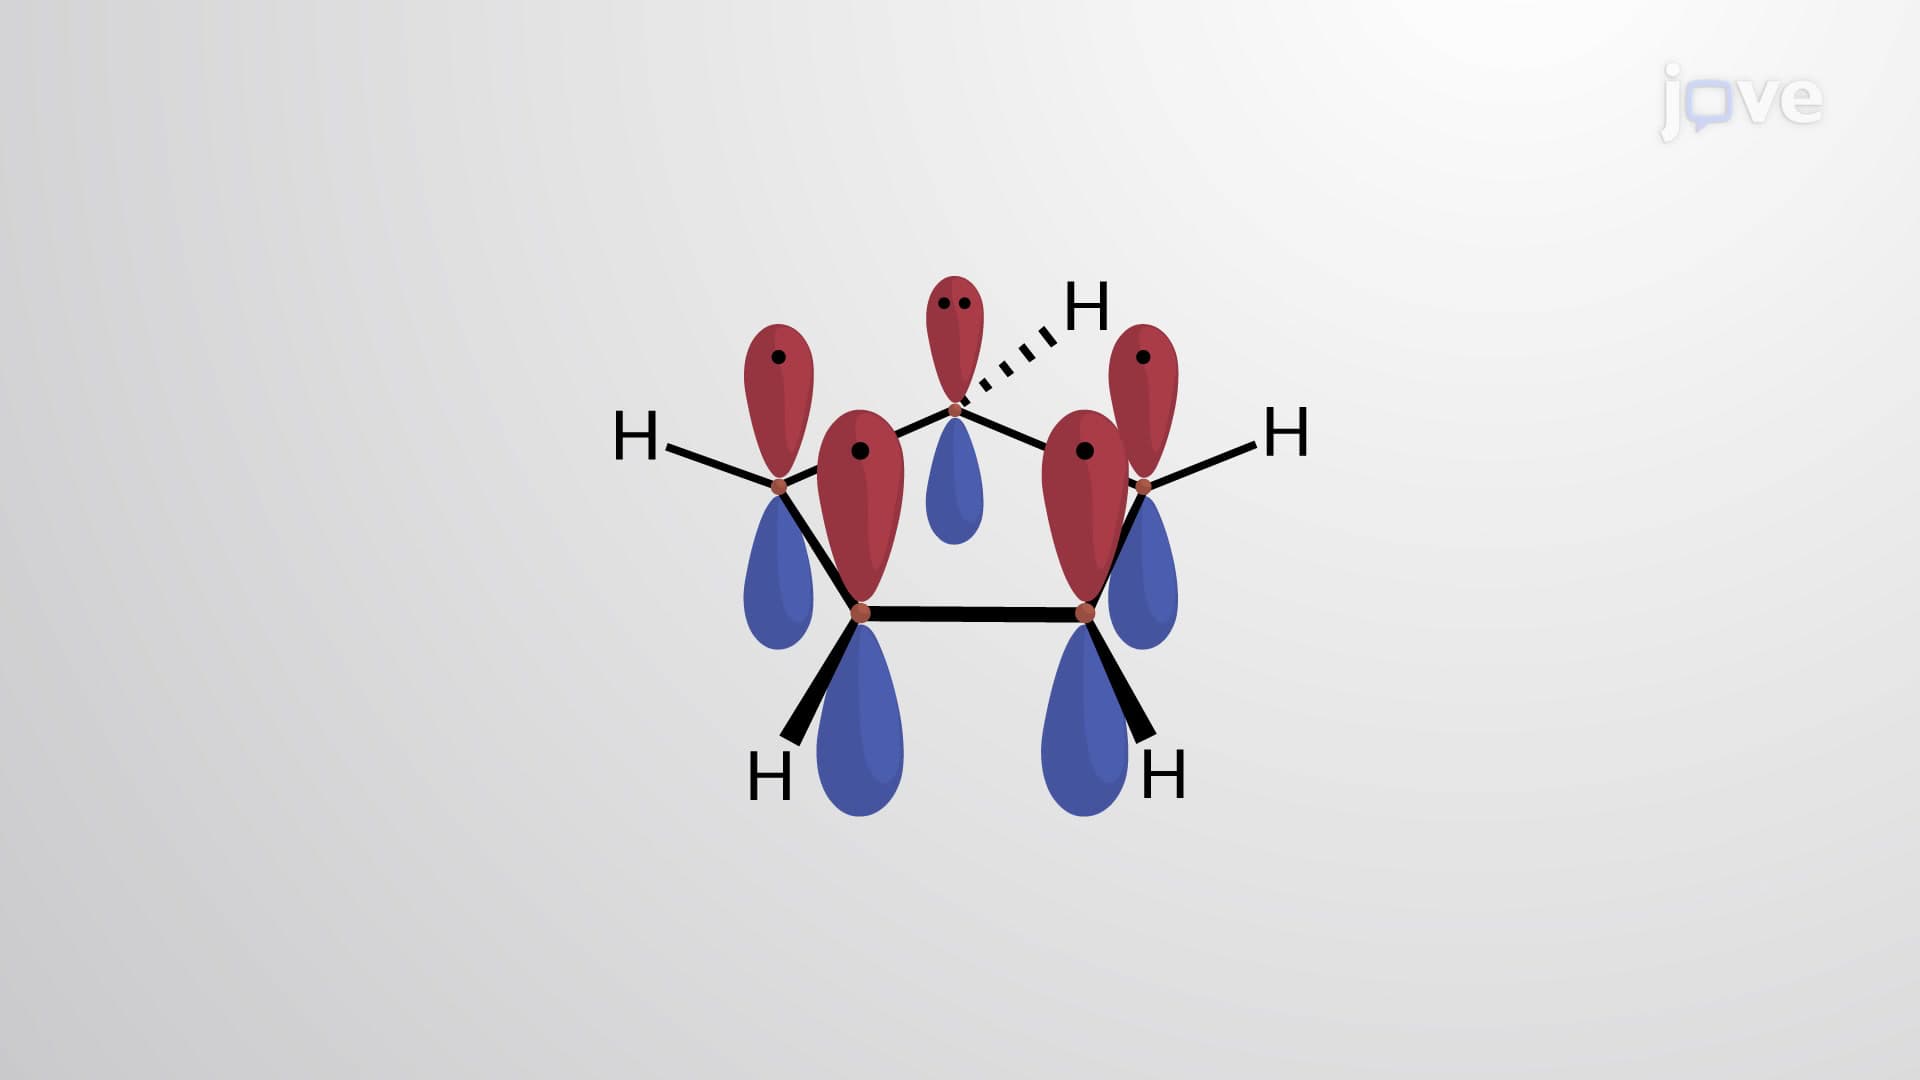
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous overlap of p...
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:27
π Electron Effects on Chemical Shift: Overview