Overcoming sequence misalignments with weighted structural superposition
Related Experiment Video
Updated: May 21, 2026
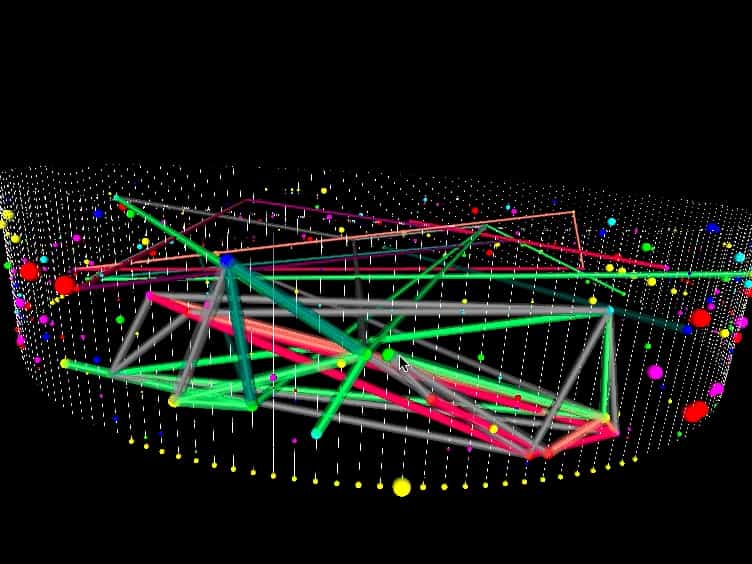
Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
Nickolay A Khazanov1, Kelly L Damm-Ganamet, Daniel X Quang
1Department of Computational Medicine and Bioinformatics, University of Michigan, Ann Arbor, MI 48109-2218, USA.

Abstract:
An appropriate structural superposition identifies similarities and differences between homologous proteins that are not evident from sequence alignments alone. We have coupled our Gaussian-weighted RMSD (wRMSD) tool with a sequence aligner and seed extension (SE) algorithm to create a robust technique for overlaying structures and aligning sequences of homologous proteins (HwRMSD). HwRMSD overcomes errors in the initial sequence alignment that would normally propagate into a standard RMSD overlay. SE can generate a corrected sequence alignment from the improved structural superposition obtained by wRMSD. HwRMSD's robust performance and its superiority over standard RMSD are demonstrated over a range of homologous proteins. Its better overlay results in corrected sequence alignments with good agreement to HOMSTRAD. Finally, HwRMSD is compared to established structural alignment methods: FATCAT, secondary-structure matching, combinatorial extension, and Dalilite. Most methods are comparable at placing residue pairs within 2 Å, but HwRMSD places many more residue pairs within 1 Å, providing a clear advantage. Such high accuracy is essential in drug design, where small distances can have a large impact on computational predictions. This level of accuracy is also needed to correct sequence alignments in an automated fashion, especially for omics-scale analysis. HwRMSD can align homologs with low-sequence identity and large conformational differences, cases where both sequence-based and structural-based methods may fail. The HwRMSD pipeline overcomes the dependency of structural overlays on initial sequence pairing and removes the need to determine the best sequence-alignment method, substitution matrix, and gap parameters for each unique pair of homologs.

Related Concept Videos
 01:20
Method of Superposition
When applying the method of superposition, each type of load—whether...
 02:04
Fixing Double-strand Breaks
02:04Fixing Double-strand Breaks
 02:07
Improving Translational Accuracy
02:07Improving Translational Accuracy
 01:28
Internal Loadings in Structural Members: Problem Solving
To illustrate this, let's consider a beam OC of 5 kN, inclined at an angle of 53.13° with the horizontal and supported at both ends. Determine the internal loadings...