Related Experiment Video
Updated: May 20, 2026


Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Assessment of the PW86+PBE+XDM density functional on van der Waals complexes at non-equilibrium geometries
1Department of Chemistry, Dalhousie University, 6274 Coburg Road, P.O. Box 15000, Halifax, Nova Scotia B3H 4R2, Canada.

Abstract:
The deficiency of conventional density-functional theory (DFT) in properly describing van der Waals (vdW) (especially dispersion-bound) complexes has been extensively addressed in the past decade. There are now several new methods published in the literature that are capable of accurately capturing weak dispersion interactions in complexes at equilibrium geometries. However, the performance of these new methods at non-equilibrium geometries remains to be assessed. We have previously published [F. O. Kannemann and A. D. Becke, J. Chem. Theory Comput. 6, 1081 (2010); A. D. Becke, A. A. Arabi, and F. O. Kannemann, Can. J. Chem. 88, 1057 (2010)] that the functional PW86+PBE+XDM for exchange + correlation + dispersion, respectively, is a highly accurate functional for general thermochemistry and vdW complexes at equilibrium geometries. Here, we show that this nonempirical, except for two parameters in the dispersion damping part, functional also performs well for vdW complexes at compressed and stretched intermonomer separations. The mean absolute relative error (MARE) is 9.4% overall for vdW complexes in the "S22×5" database incorporating compressed and stretched geometries [J. Rezac, K. E. Riley, and P. Hobza, J. Chem. Theory Comput. 7, 2427 (2011)]. Our largest MARE on the S22×5 database is 13.3% on the compressed geometry set.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:10
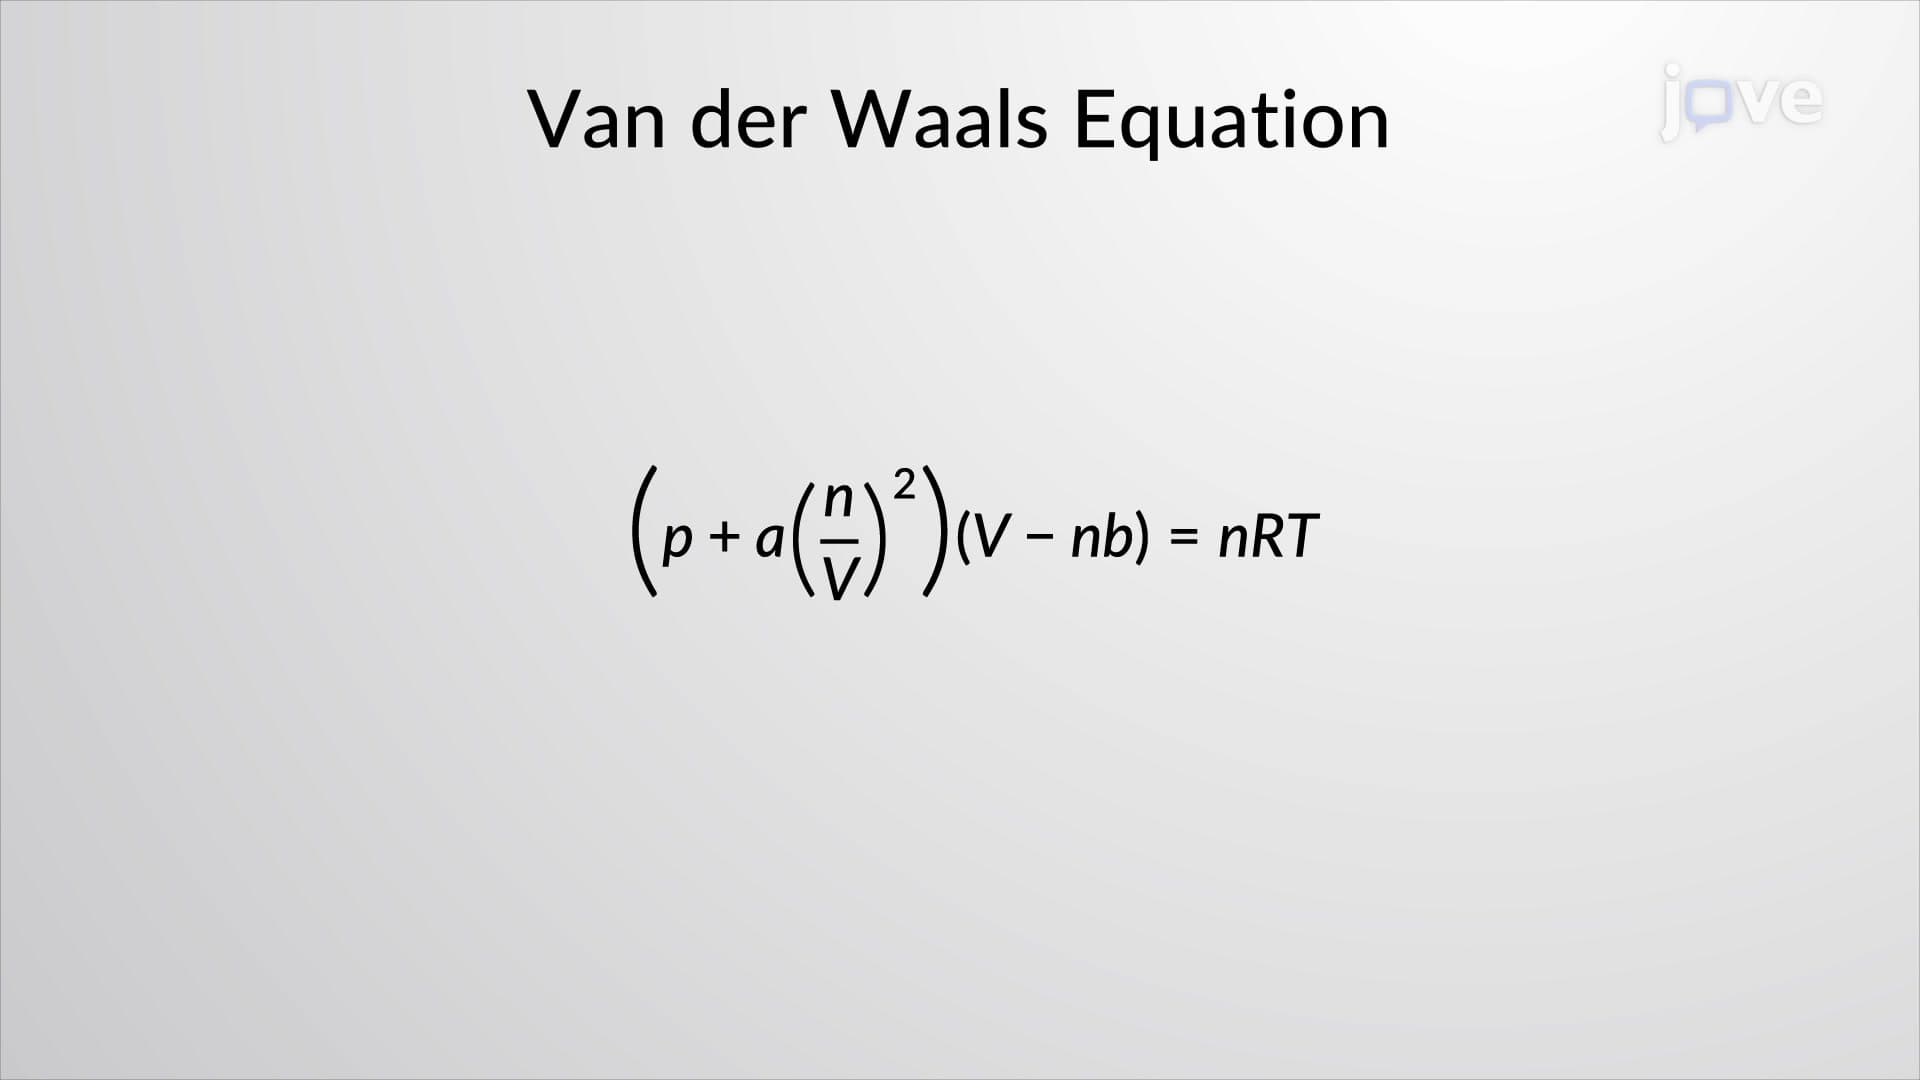
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:13
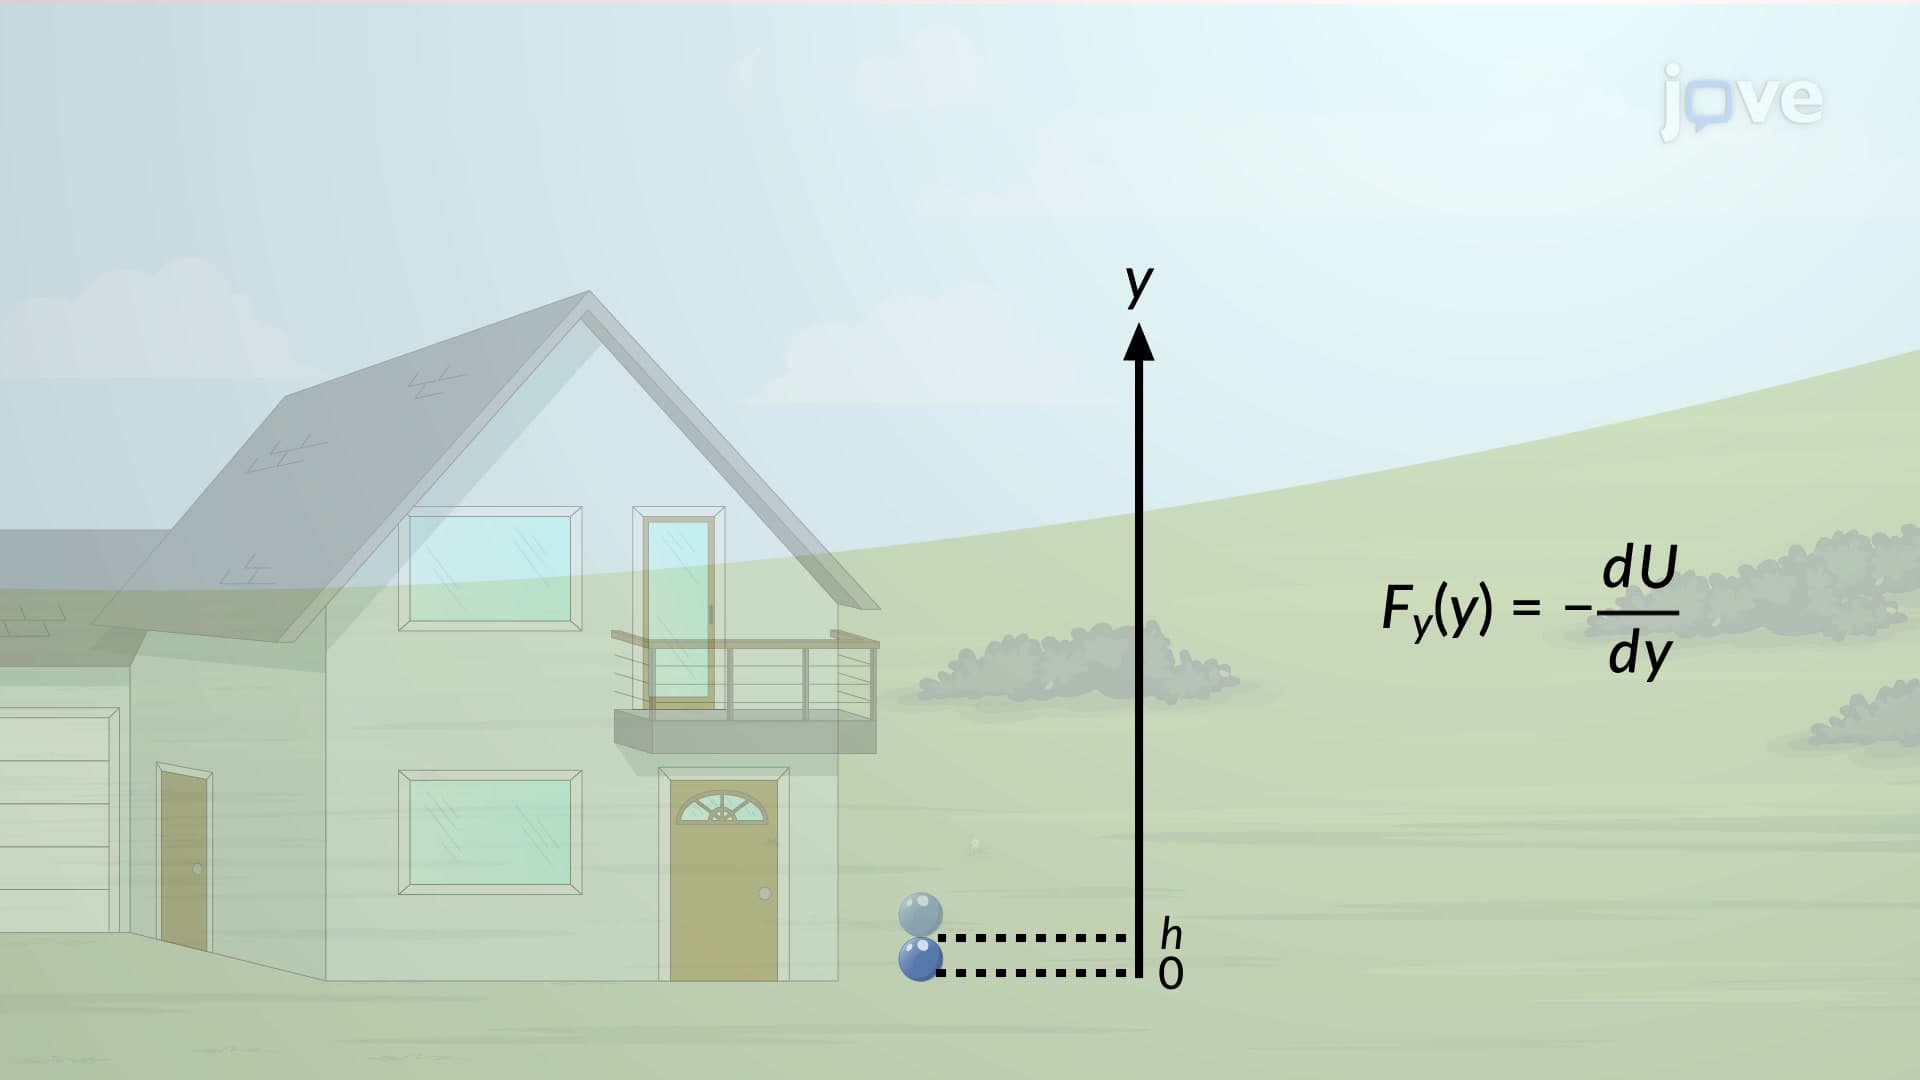
Force and Potential Energy in One Dimension
 01:24
Van der Waals Interactions
 01:26
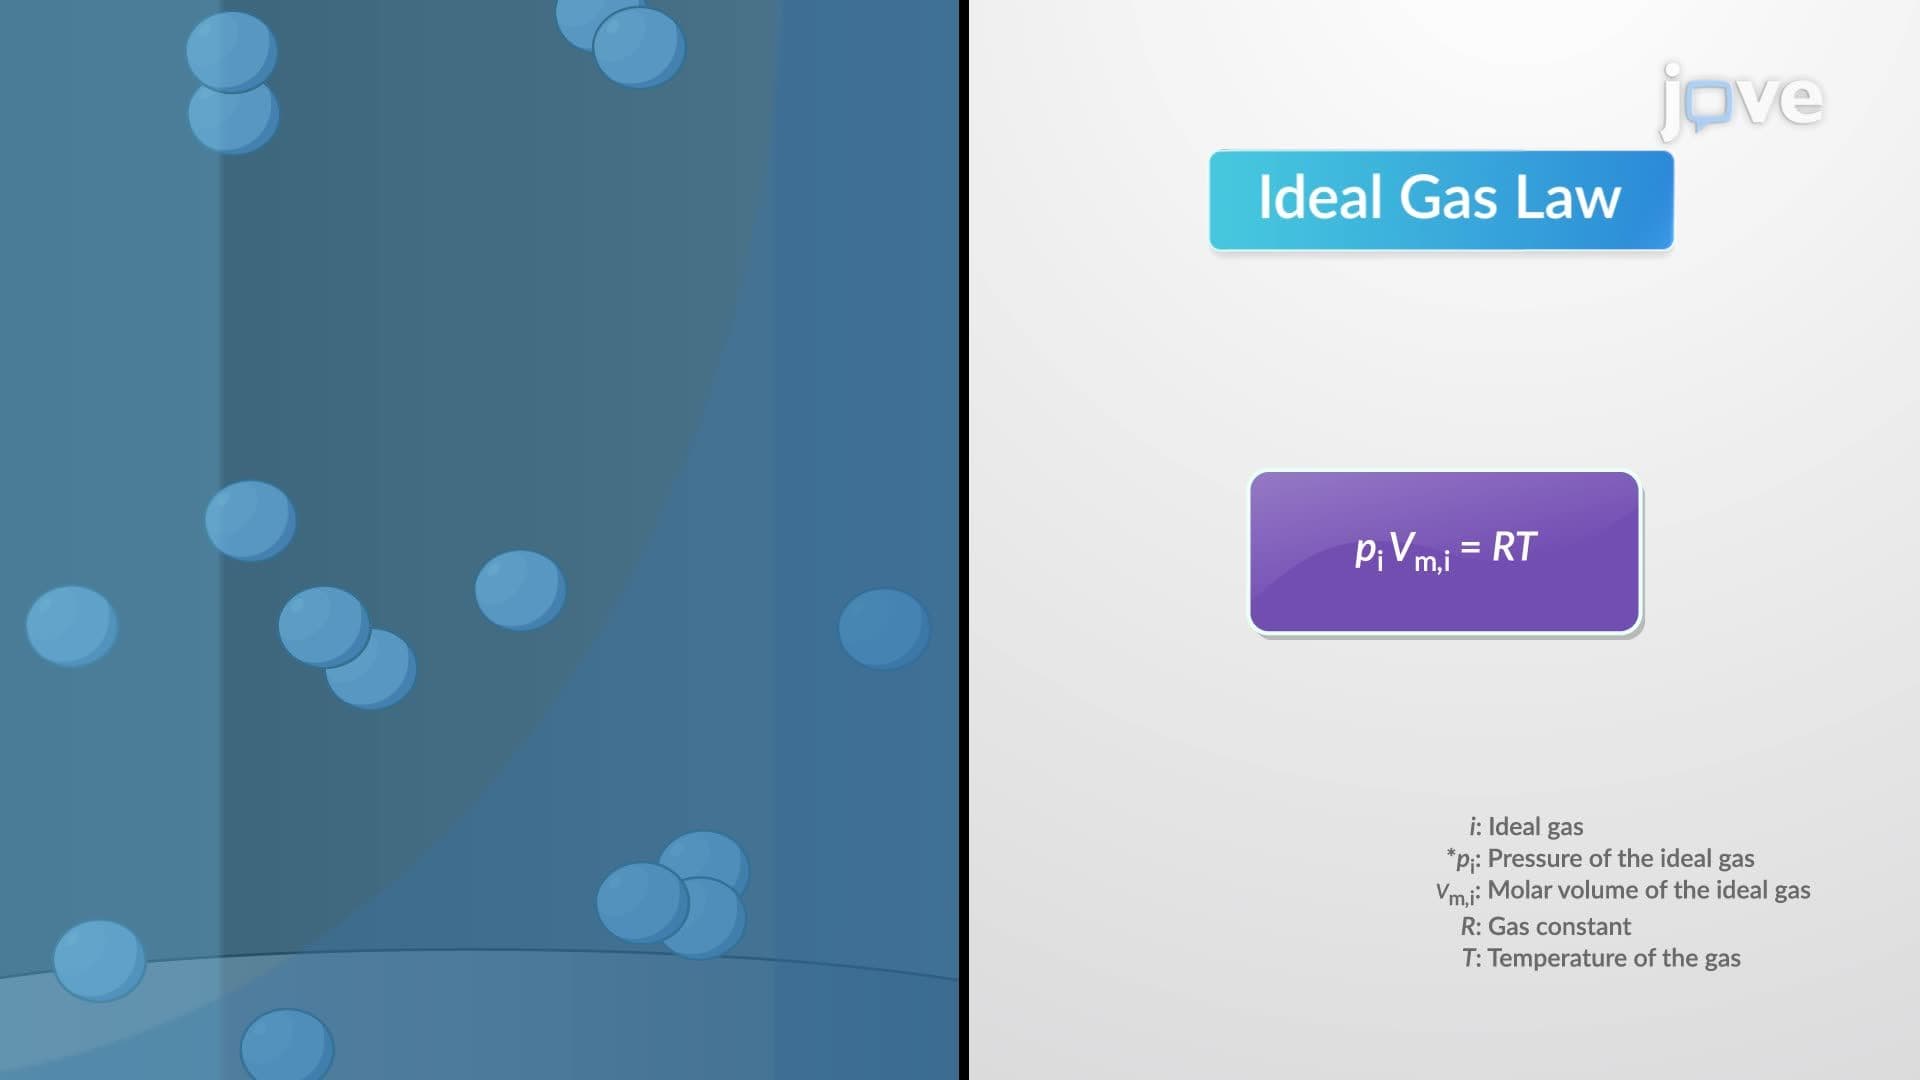
The Van der Waals Equation
 02:42
Valence Bond Theory