
Aromatic interactions as control elements in stereoselective organic reactions
Elizabeth H Krenske1, K N Houk
1School of Chemistry, University of Melbourne, VIC 3010, Australia. ekrenske@unimelb.edu.au

Abstract:
This Account describes how attractive interactions of aromatic rings with other groups can influence and control the stereoselectivity of many reactions. Recent developments in theory have improved the accuracy in the modeling of aromatic interactions. Quantum mechanical modeling can now provide insights into the roles of these interactions at a level of detail not previously accessible, both for ground-state species and for transition states of chemical reactions. In this Account, we show how transition-state modeling led to the discovery of the influence of aryl groups on the stereoselectivities of several types of organic reactions, including asymmetric dihydroxylations, transfer hydrogenations, hetero-Diels-Alder reactions, acyl transfers, and Claisen rearrangements. Our recent studies have also led to a novel mechanistic picture for two classes of (4 + 3) cycloadditions, both of which involve reactions of furans with oxyallyl intermediates. The first class of cycloadditions, developed by Hsung, features neutral oxyallyl intermediates that contain a chiral oxazolidinone auxiliary. Originally, it was thought that these cycloadditions relied on differential steric crowding of the two faces of a planar intermediate. Computations reveal a different picture and show that cycloaddition with furan takes place preferentially through the more crowded transition state: the furan adds on the same side as the Ph substituent of the oxazolidinone. The crowded transition state is stabilized by a CH-π interaction between furan and Ph worth approximately 2 kcal/mol. Attractive interactions with aromatic rings also control the stereoselectivity in a second class of (4+3) cycloadditions involving chiral alkoxy siloxyallyl cations. Alkoxy groups derived from chiral α-methylbenzyl alcohols favor crowded transition states, where a stabilizing CH-π interaction is present between the furan and the Ar group. The cationic cycloadditions are stepwise, while the Hsung cycloadditions are concerted. Our results suggest that this form of CH- π-directed stereocontrol is quite general and likely controls the stereoselectivities of other addition reactions in which one face of a planar intermediate bears a pendant aromatic substituent.
Related Concept Videos
 02:17
Regioselectivity of Electrophilic Additions to Alkenes: Markovnikov's Rule
The hydrohalogenation of an unsymmetrical alkene can yield two haloalkane products, depending on which vinylic carbon takes up the halogen. However, one product usually predominates, where hydrogen adds to the vinylic carbon bearing the...
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn...
 01:17
Thermal Electrocyclic Reactions: Stereochemistry
Selection Rules: Thermal Activation
Conjugated systems containing an even number of π-electron pairs undergo a conrotatory ring closure. For example, thermal electrocyclization of (2E,4E)-2,4-hexadiene, a conjugated diene containing two π-electron pairs, gives trans-3,4-dimethylcyclobutene.
 01:28
Diels–Alder Reaction Forming Cyclic Products: Stereochemistry
 01:11
ortho–para-Directing Activators: –CH3, –OH, –NH2, –OCH3
 01:30
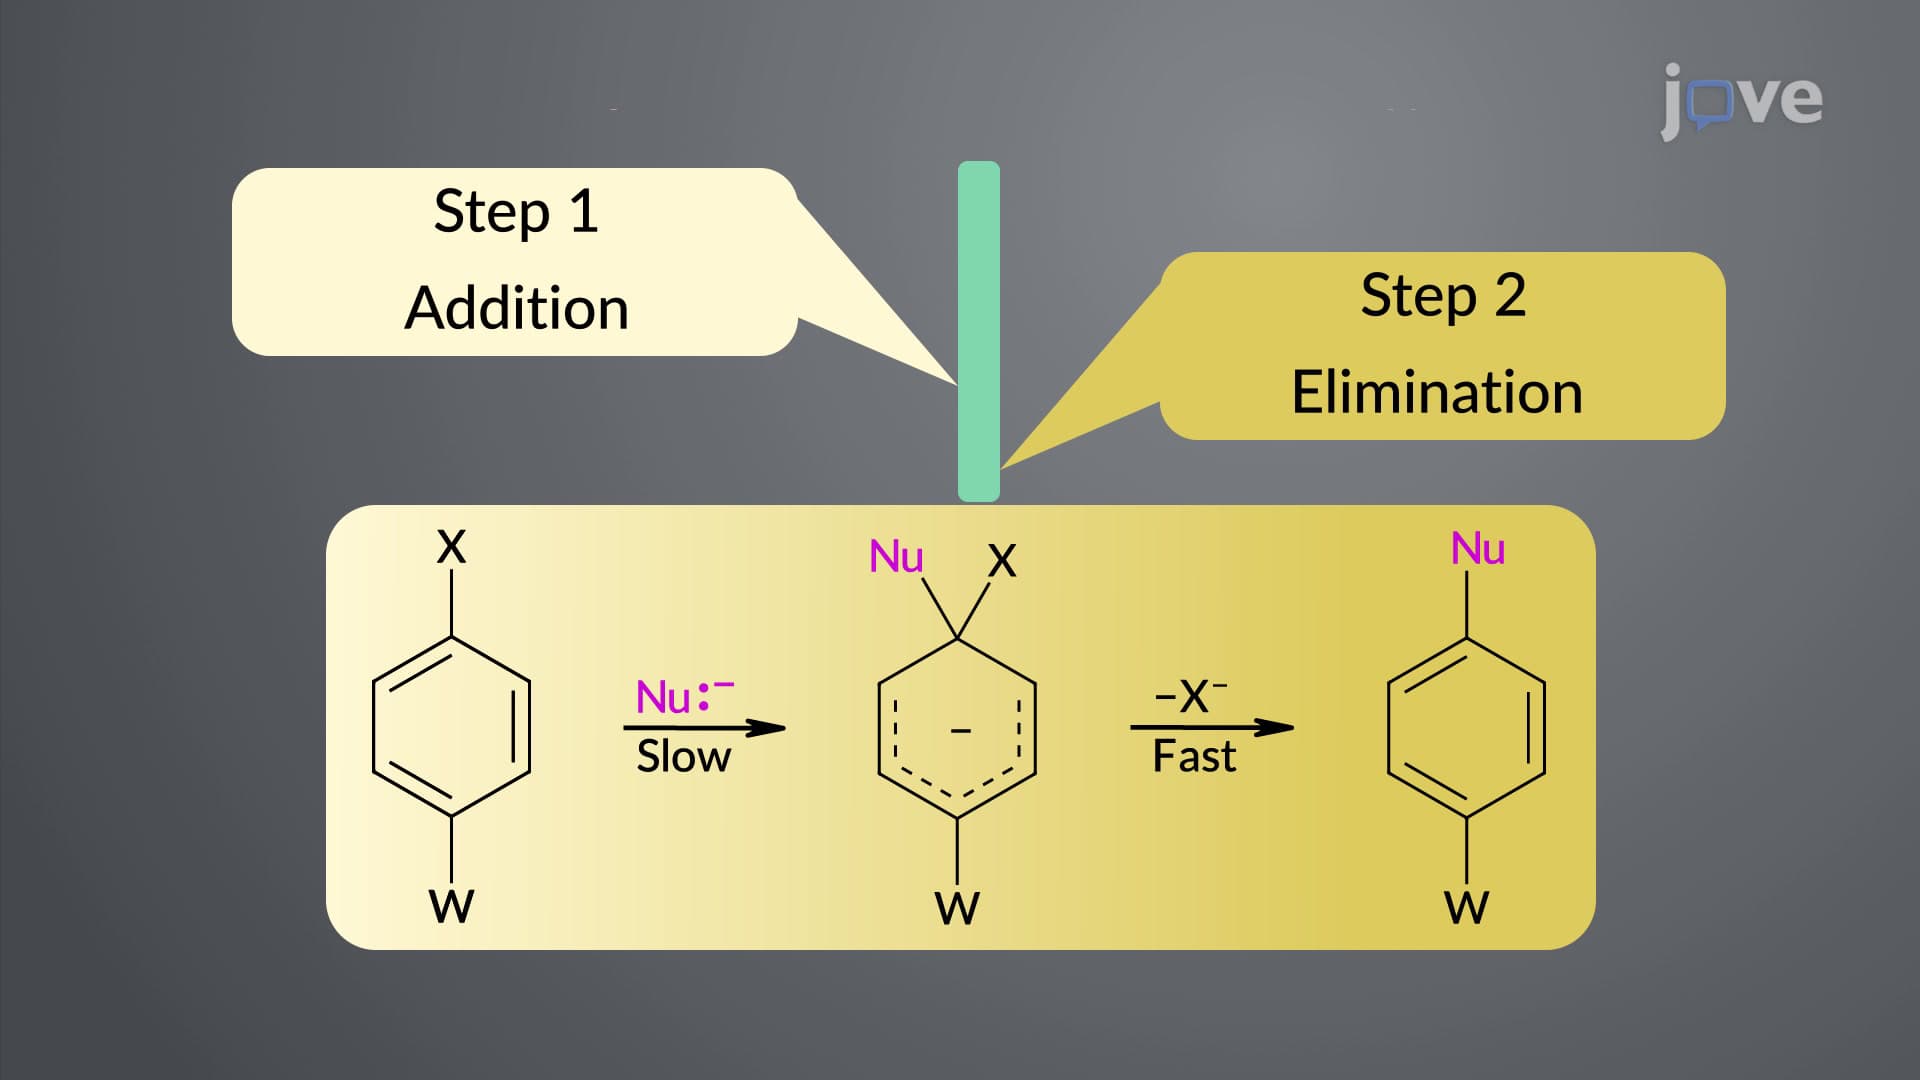
Nucleophilic Aromatic Substitution: Addition–Elimination (SNAr)
The reaction begins with an attack of the nucleophile on the carbon that holds the leaving group. This results in the delocalization of the π electrons over the ring carbons. The resonance interaction between...