Related Experiment Video
Updated: May 19, 2026


Realistic Membrane Modeling Using Complex Lipid Mixtures in Simulation Studies
Published on: September 1, 2023
Force field development for actinyl ions via quantum mechanical calculations: an approach to account for many body
Neeraj Rai1, Surya P Tiwari, Edward J Maginn
1Department of Chemical and Biomolecular Engineering, University of Notre Dame, 182 Fitzpatrick Hall, Notre Dame, Indiana 46556, United States.

Abstract:
Advances in computational algorithms and methodologies make it possible to use highly accurate quantum mechanical calculations to develop force fields (pair-wise additive intermolecular potentials) for condensed phase simulations. Despite these advances, this approach faces numerous hurdles for the case of actinyl ions, AcO2(n+) (high-oxidation-state actinide dioxo cations), mainly due to the complex electronic structure resulting from an interplay of s, p, d, and f valence orbitals. Traditional methods use a pair of molecules (“dimer”) to generate a potential energy surface (PES) for force field parametrization based on the assumption that many body polarization effects are negligible. We show that this is a poor approximation for aqueous phase uranyl ions and present an alternative approach for the development of actinyl ion force fields that includes important many body solvation effects. Force fields are developed for the UO2(2+) ion with the SPC/Fw, TIP3P, TIP4P, and TIP5P water models and are validated by carrying out detailed molecular simulations on the uranyl aqua ion, one of the most characterized actinide systems. It is shown that the force fields faithfully reproduce available experimental structural data and hydration free energies. Failure to account for solvation effects when generating PES leads to overbinding between UO2(2+) and water, resulting in incorrect hydration free energies and coordination numbers. A detailed analysis of arrangement of water molecules in the first and second solvation shell of UO2(2+) is presented. The use of a simple functional form involving the sum of Lennard-Jones + Coulomb potentials makes the new force field compatible with a large number of available molecular simulation engines and common force fields.
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:17
Factors Affecting Activity Coefficient
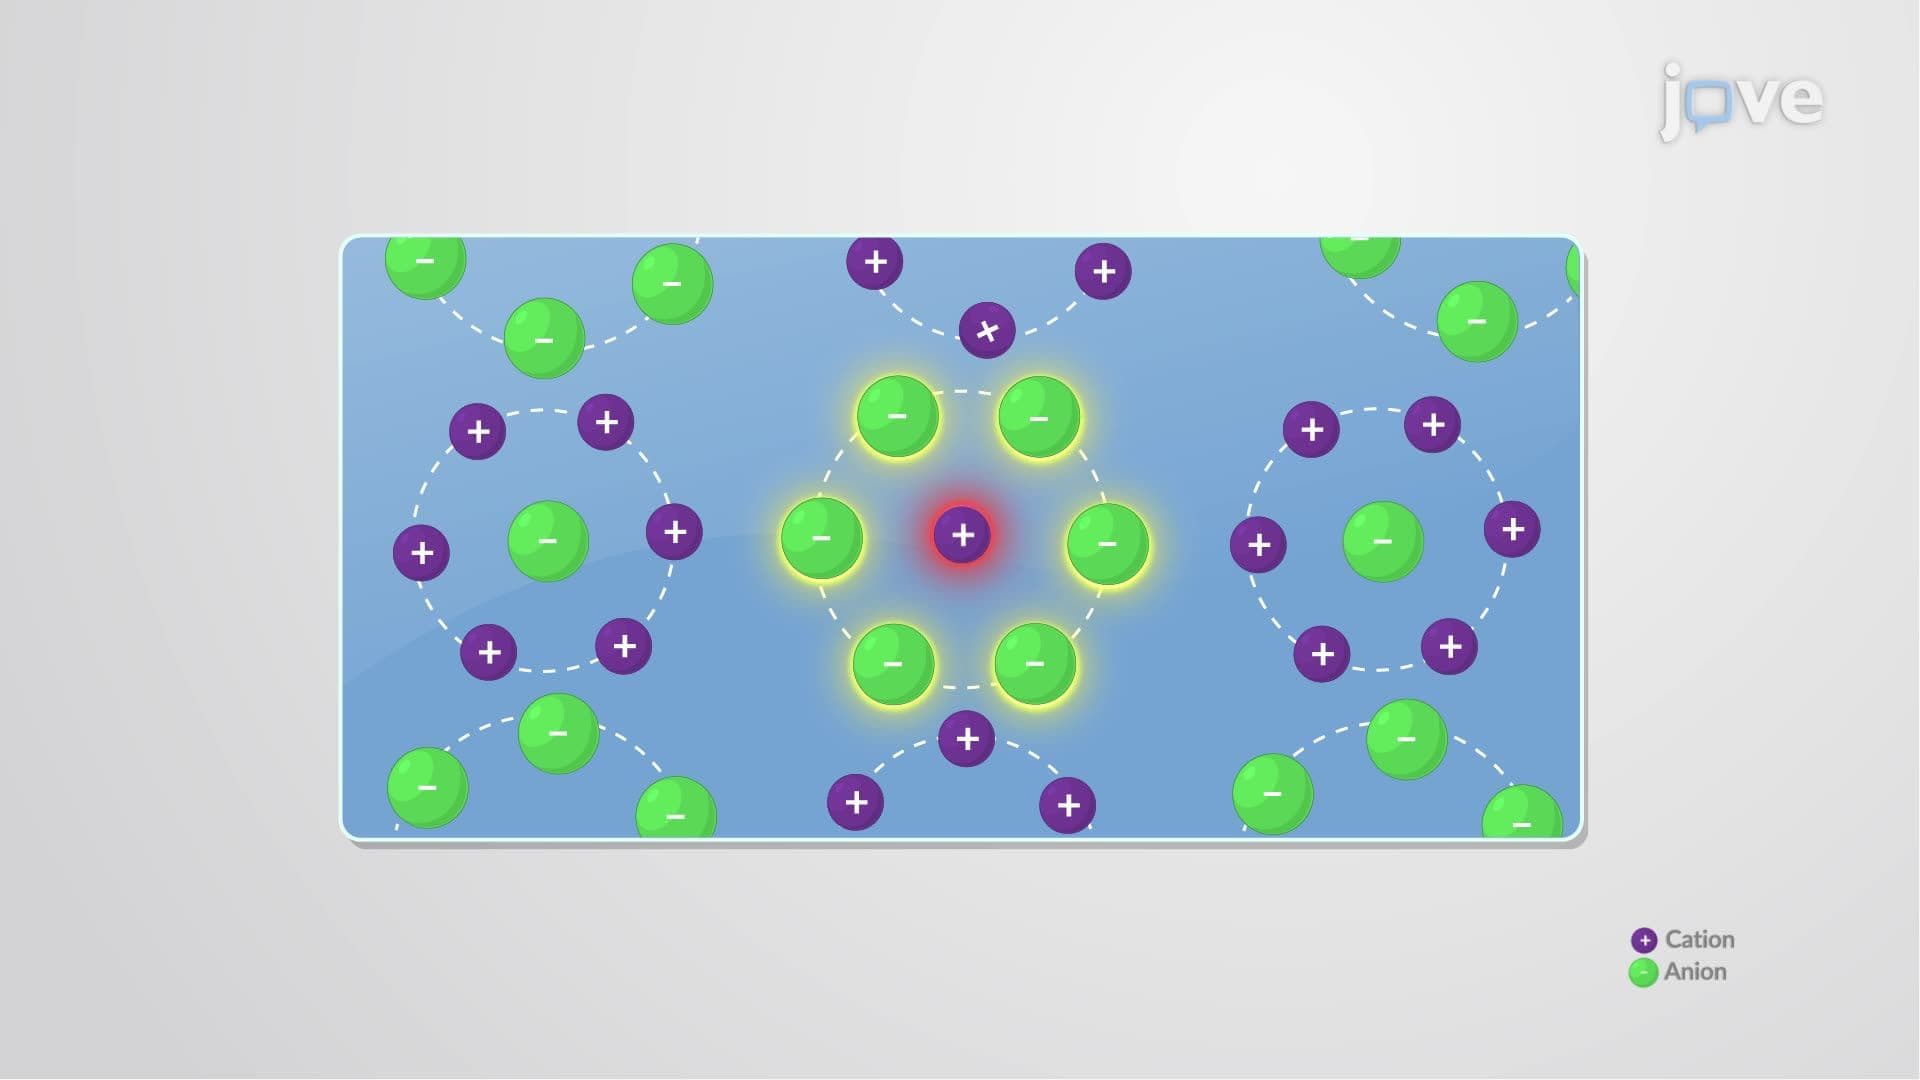
The activity coefficient value for an ion is close to one when the solution has almost zero ionic strength, i.e., when the solution shows close to ideal behavior. As the ionic strength of the solution increases from 0 to 0.1 mol/L, a decrease in the...
 01:27
The Debye–Hückel Theory of Electrolyte Solutions
 03:13
Intermolecular Forces
 01:14
Generation of Straight or Branched Actin Filaments
Arp2/3 Complex
Arp2/3 complex is a seven-subunit complex consisting of two proteins similar to actin- Arp2 and Arp3, and five other subunits that help keep Arp2 and Arp3 inactive. When required, the complex is...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...