Related Experiment Video
Updated: Aug 6, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Lessons Learned on Obtaining Reliable Conductivity Estimates From Molecular Dynamics Simulations
Paul Zaby1, Johannes Ingenmey1, Tuanan C Lourenço2
1Mulliken Center for Theoretical Chemistry, University of Bonn, Bonn, Germany.

Abstract:
The calculation of reliable ionic conductivities from molecular dynamics simulations is not a straightforward task, especially for strongly correlated systems, such as ionic liquids or highly concentrated electrolytes, where the Nernst-Einstein approach tends to fail. In this manuscript, we present the newly implemented conduct module for TRAVIS. It allows the calculation of the ionic conductivity using the Einstein-Helfand and Green-Kubo approaches, which explicitly include ionic correlations in their formalism. We provide a broad overview of accessible transport properties and compare methods and best practices for obtaining statistically reliable estimates of ionic conductivity and other physicochemical properties derived from electrolyte molecular dynamics simulations, including transport numbers and the inverse Haven ratio. To validate our implementation and demonstrate the conduct module's capabilities, we simulated the ionic liquid 1-ethyl-3-methylimidazolium dicyanamide ([EMIm][DCA]) as well as the ether-based electrolyte lithium bis(fluorosulfonyl)imide in ethylene glycol dimethyl ether (LiFSI/DME).

Related Concept Videos
 01:28
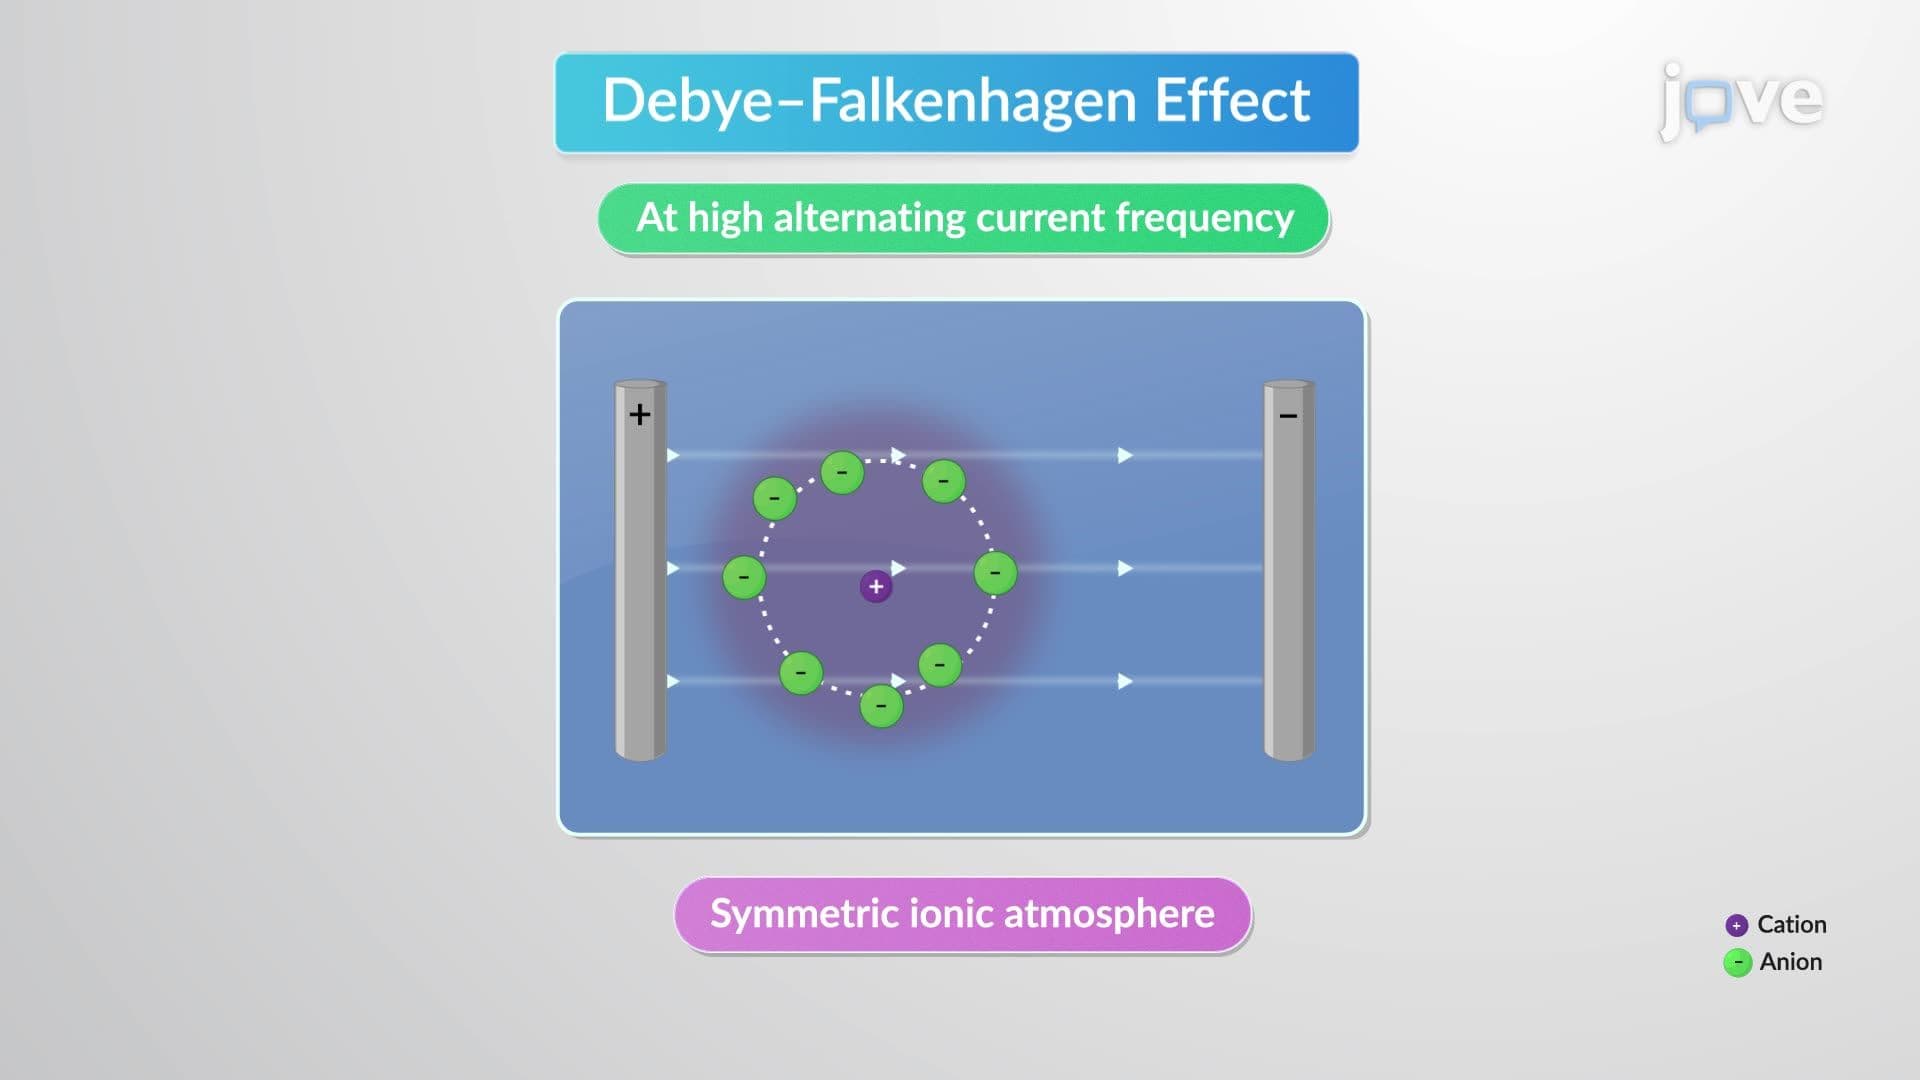
Debye–Huckel–Onsager Conductance Equation
 01:29
Electrical Transport
 01:29
Kohlraush’s Law and its Applications