Related Experiment Video
Updated: May 19, 2026

Real-Time Fluorescent Measurement of Synaptic Functions in Models of Amyotrophic Lateral Sclerosis
Published on: July 16, 2021
VAPB and C9orf72 mutations in 1 familial amyotrophic lateral sclerosis patient
Marka van Blitterswijk1, Michael A van Es, Max Koppers
1Department of Neurology, Rudolf Magnus Institute of Neuroscience, University Medical Center Utrecht, The Netherlands.

Abstract:
Previously, we have reported amyotrophic lateral sclerosis (ALS) families with multiple mutations in major ALS-associated genes. These findings provided evidence for an oligogenic basis of ALS. In our present study, we screened a cohort of 755 sporadic ALS patients, 111 familial ALS patients (97 families), and 765 control subjects of Dutch descent for mutations in vesicle-associated membrane protein B (VAPB). We have identified 1 novel VAPB mutation (p.V234I) in a familial ALS patient known to have a chromosome 9 open reading frame 72 (C9orf72) repeat expansion. This p.V234I mutation was absent in control subjects, located in a region with high evolutionary conservation, and predicted to have damaging effects. Taken together, these findings provide additional evidence for an oligogenic basis of ALS.
Related Concept Videos
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
03:03Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
 01:22
Lysosomal Hydrolases
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:21
Huntington Disease l: Introduction
 01:23
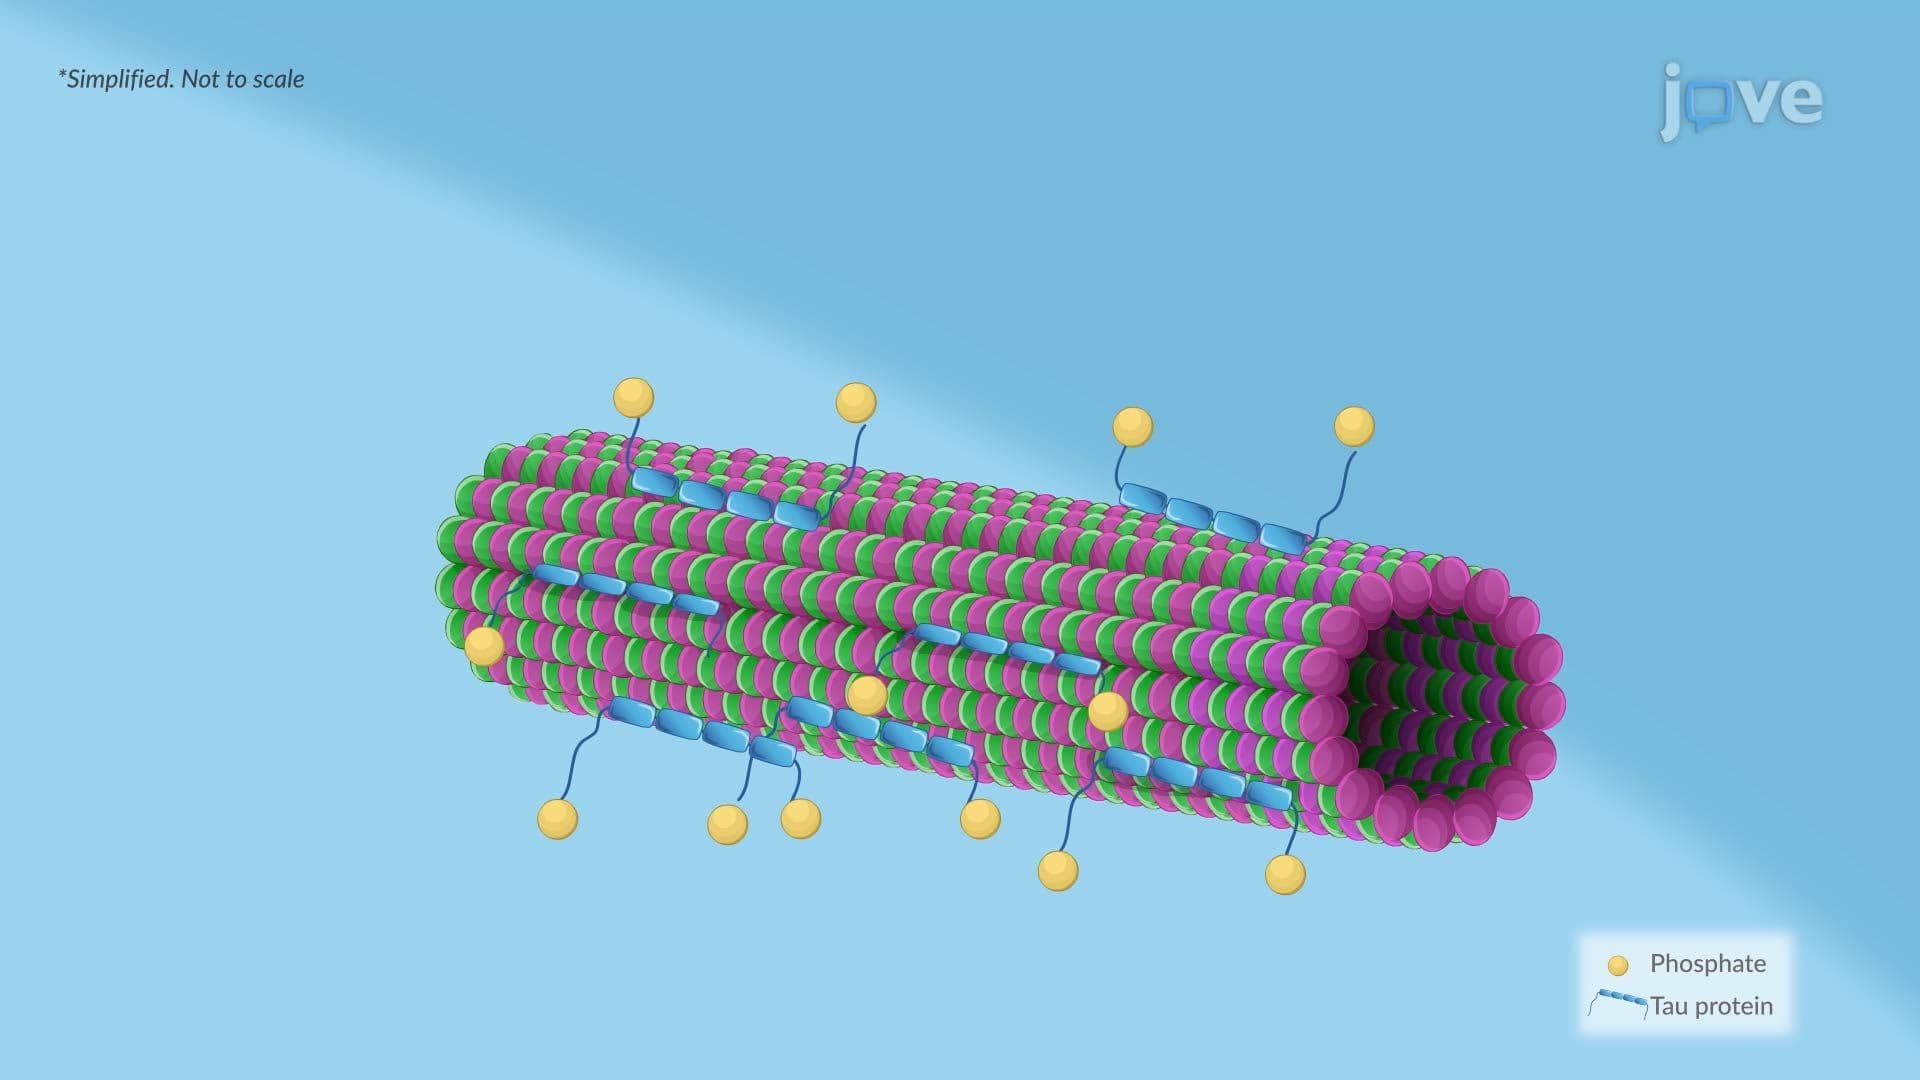
Alzheimer Disease ll: Pathophysiology