Specification, annotation, visualization and simulation of a large rule-based model for ERBB receptor signaling
Related Experiment Video
Updated: May 19, 2026
A Combined 3D Tissue Engineered In Vitro/In Silico Lung Tumor Model for Predicting Drug Effectiveness in Specific Mutational Backgrounds
Published on: April 6, 2016
Matthew S Creamer1, Edward C Stites, Meraj Aziz
1Clinical Translational Research Division, Translational Genomics Research Institute, Phoenix, AZ 85004, USA.

Background:
Mathematical/computational models are needed to understand cell signaling networks, which are complex. Signaling proteins contain multiple functional components and multiple sites of post-translational modification. The multiplicity of components and sites of modification ensures that interactions among signaling proteins have the potential to generate myriad protein complexes and post-translational modification states. As a result, the number of chemical species that can be populated in a cell signaling network, and hence the number of equations in an ordinary differential equation model required to capture the dynamics of these species, is prohibitively large. To overcome this problem, the rule-based modeling approach has been developed for representing interactions within signaling networks efficiently and compactly through coarse-graining of the chemical kinetics of molecular interactions.
Results:
Here, we provide a demonstration that the rule-based modeling approach can be used to specify and simulate a large model for ERBB receptor signaling that accounts for site-specific details of protein-protein interactions. The model is considered large because it corresponds to a reaction network containing more reactions than can be practically enumerated. The model encompasses activation of ERK and Akt, and it can be simulated using a network-free simulator, such as NFsim, to generate time courses of phosphorylation for 55 individual serine, threonine, and tyrosine residues. The model is annotated and visualized in the form of an extended contact map.
Conclusions:
With the development of software that implements novel computational methods for calculating the dynamics of large-scale rule-based representations of cellular signaling networks, it is now possible to build and analyze models that include a significant fraction of the protein interactions that comprise a signaling network, with incorporation of the site-specific details of the interactions. Modeling at this level of detail is important for understanding cellular signaling.
Related Concept Videos
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with one...
 01:15
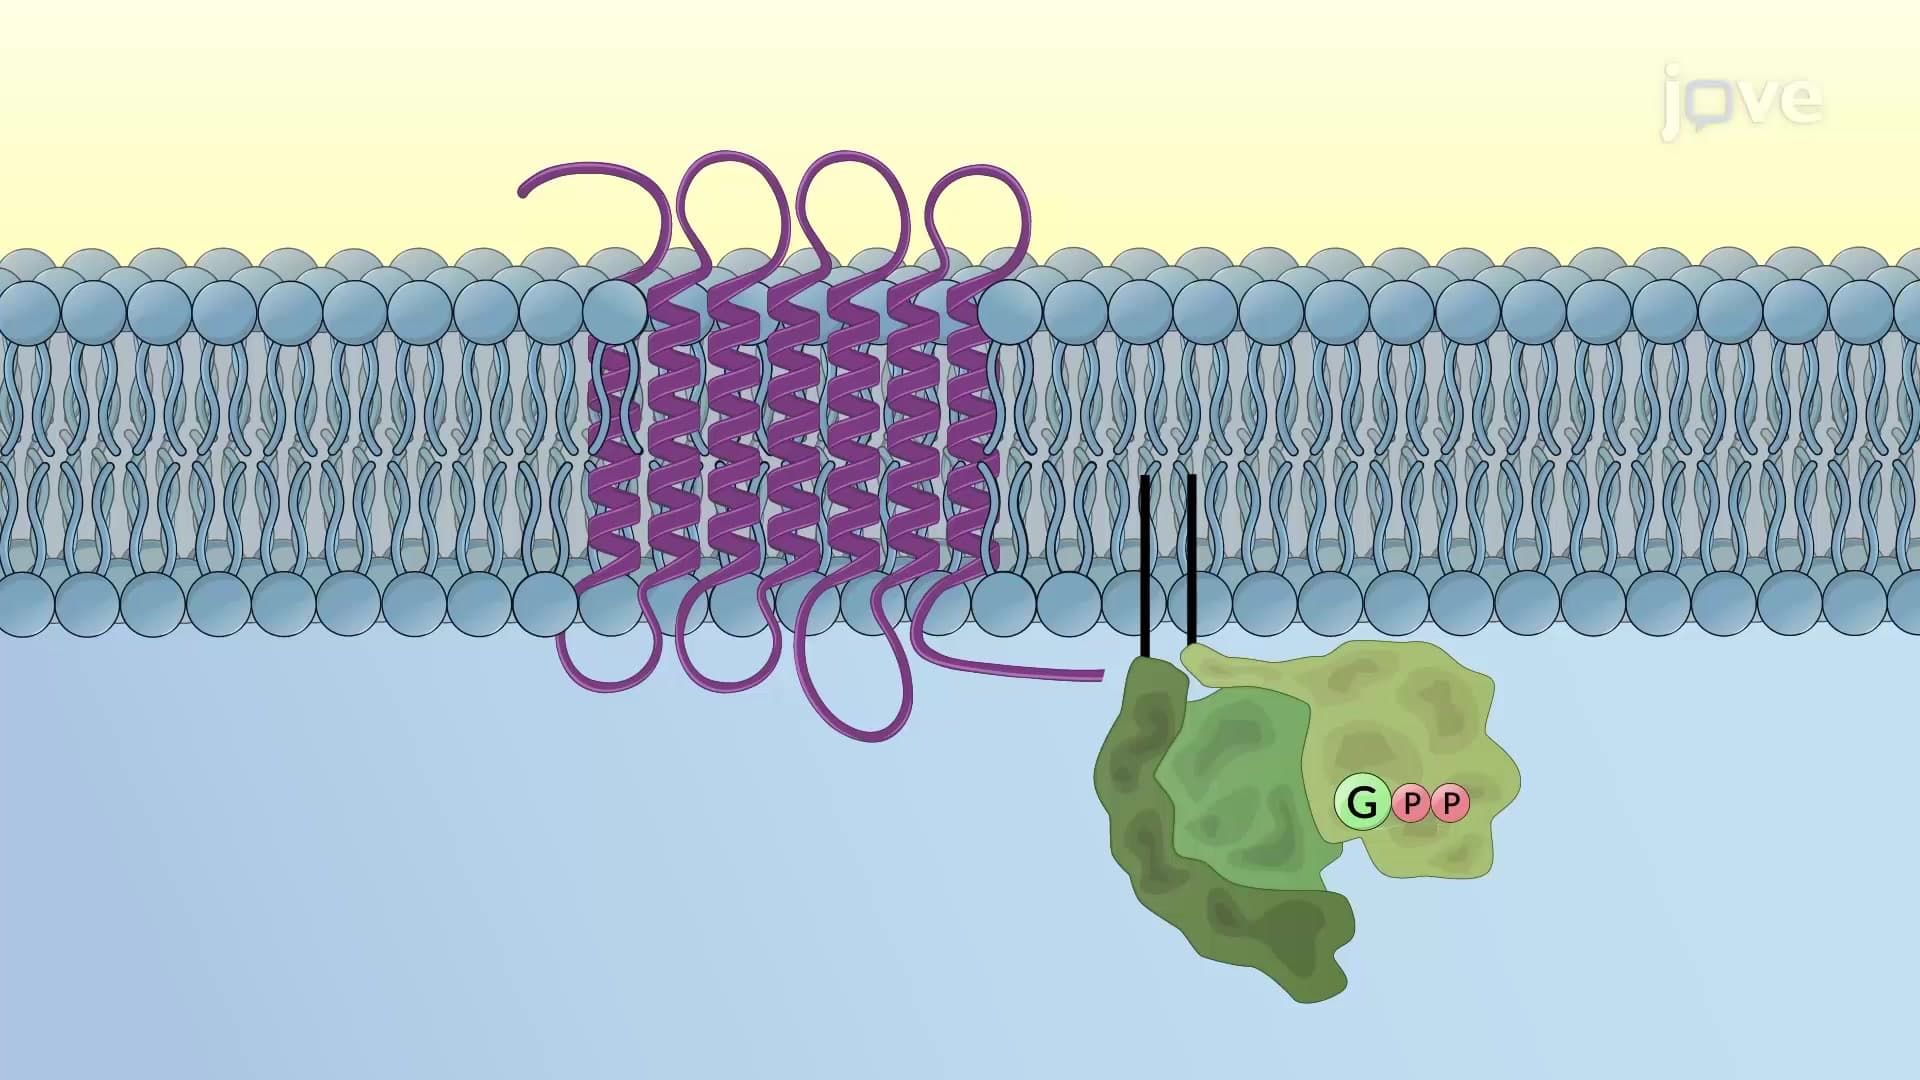
G Protein-coupled Receptors
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
 01:30
Assembly of Signaling Complexes
Interaction domains in cell signaling
Interaction domains recognize exposed features of their binding partners containing post-translationally modified sequences,...
 01:25
Cell Signaling in Plants
 01:26
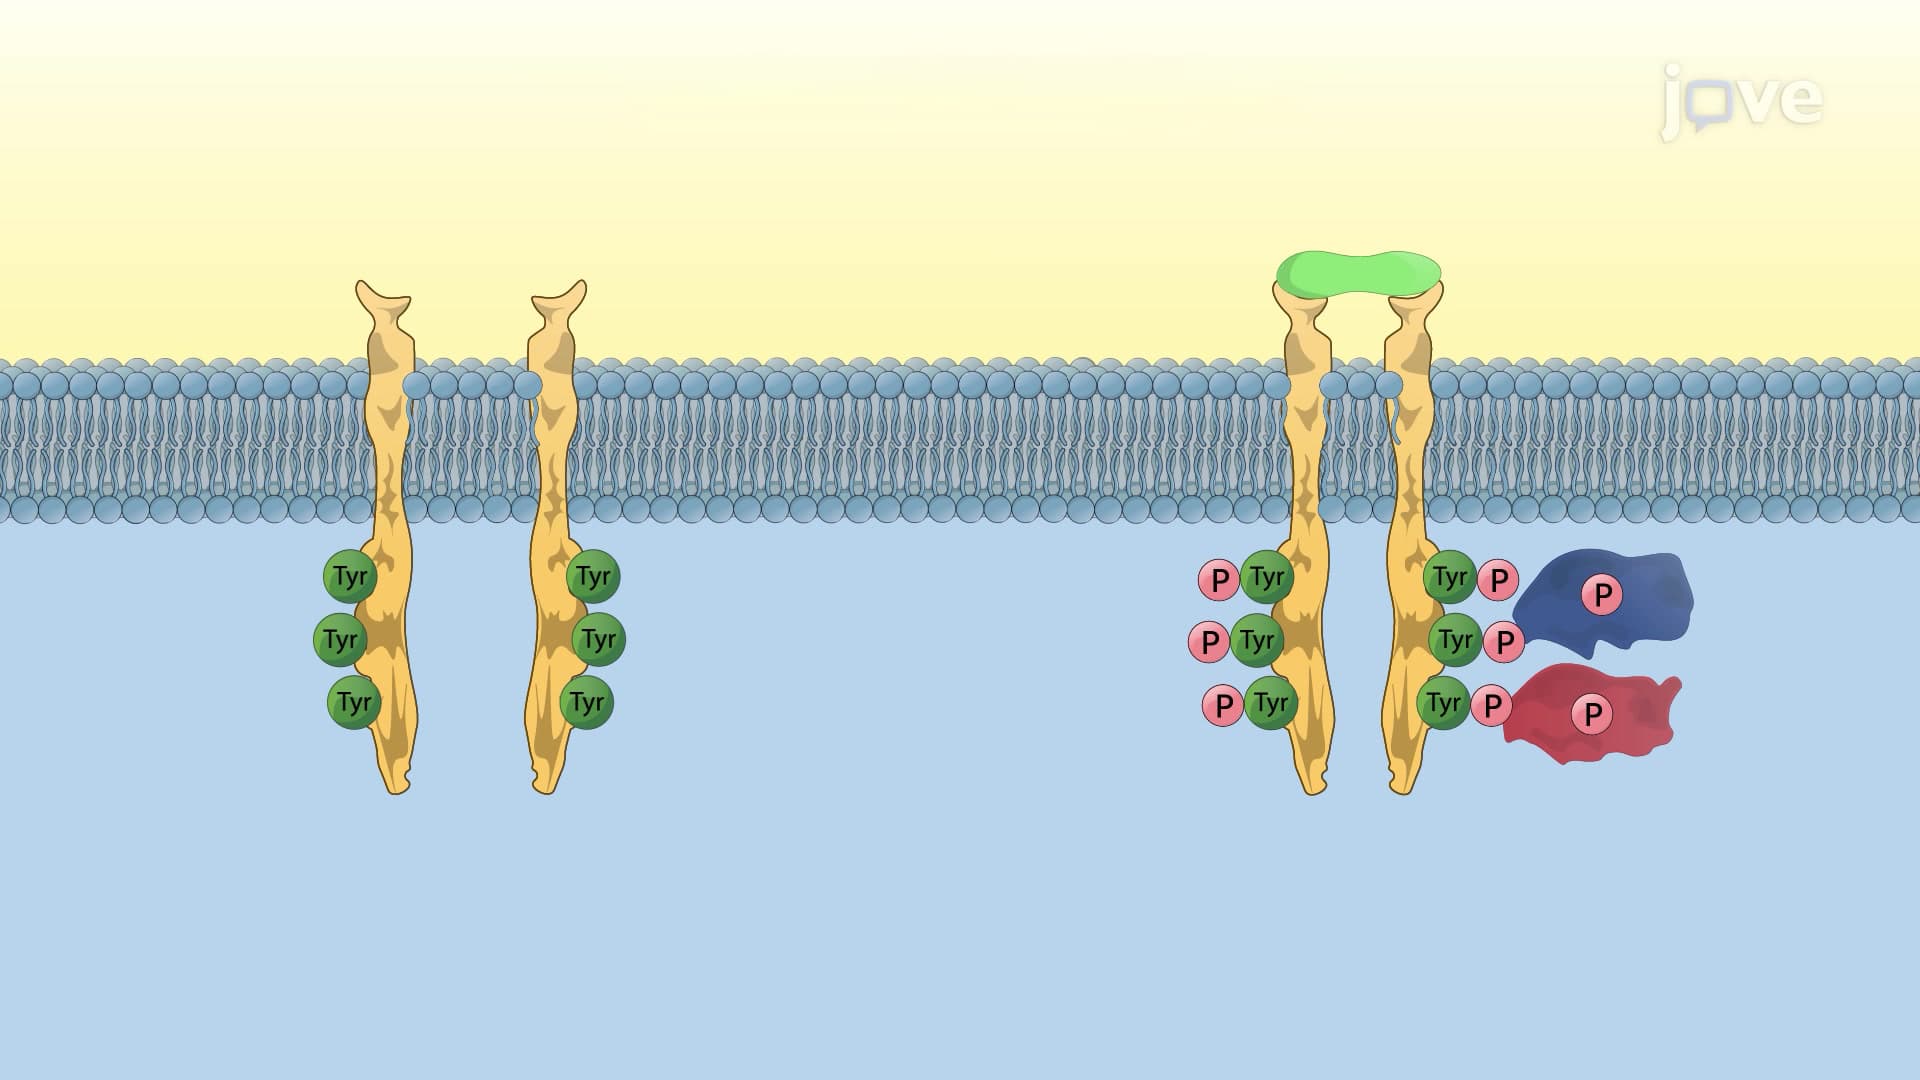
Receptor Tyrosine Kinases