Related Experiment Video
Updated: May 19, 2026


Generation of Human Nasal Epithelial Cell Spheroids for Individualized Cystic Fibrosis Transmembrane Conductance Regulator Study
Published on: April 11, 2018
CFTR regulates early pathogenesis of chronic obstructive lung disease in βENaC-overexpressing mice
Bjarki Johannesson1, Stephanie Hirtz, Jolanthe Schatterny
1Department of Translational Pulmonology, Translational Lung Research Center Heidelberg, Member of the German Center for Lung Research, University of Heidelberg, Heidelberg, Germany.

Background:
Factors determining the onset and severity of chronic obstructive pulmonary disease remain poorly understood. Previous studies demonstrated that airway surface dehydration in βENaC-overexpressing (βENaC-Tg) mice on a mixed genetic background caused either neonatal mortality or chronic obstructive lung disease suggesting that the onset of lung disease was modulated by the genetic background.
Methods:
To test this hypothesis, we backcrossed βENaC-Tg mice onto two inbred strains (C57BL/6 and BALB/c) and studied effects of the genetic background on neonatal mortality, airway ion transport and airway morphology. Further, we crossed βENaC-Tg mice with CFTR-deficient mice to validate the role of CFTR in early lung disease.
Results:
We demonstrate that the C57BL/6 background conferred increased CFTR-mediated Cl(-) secretion, which was associated with decreased mucus plugging and mortality in neonatal βENaC-Tg C57BL/6 compared to βENaC-Tg BALB/c mice. Conversely, genetic deletion of CFTR increased early mucus obstruction and mortality in βENaC-Tg mice.
Conclusions:
We conclude that a decrease or absence of CFTR function in airway epithelia aggravates the severity of early airway mucus obstruction and related mortality in βENaC-Tg mice. These results suggest that genetic or environmental factors that reduce CFTR activity may contribute to the onset and severity of chronic obstructive pulmonary disease and that CFTR may serve as a novel therapeutic target.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:24
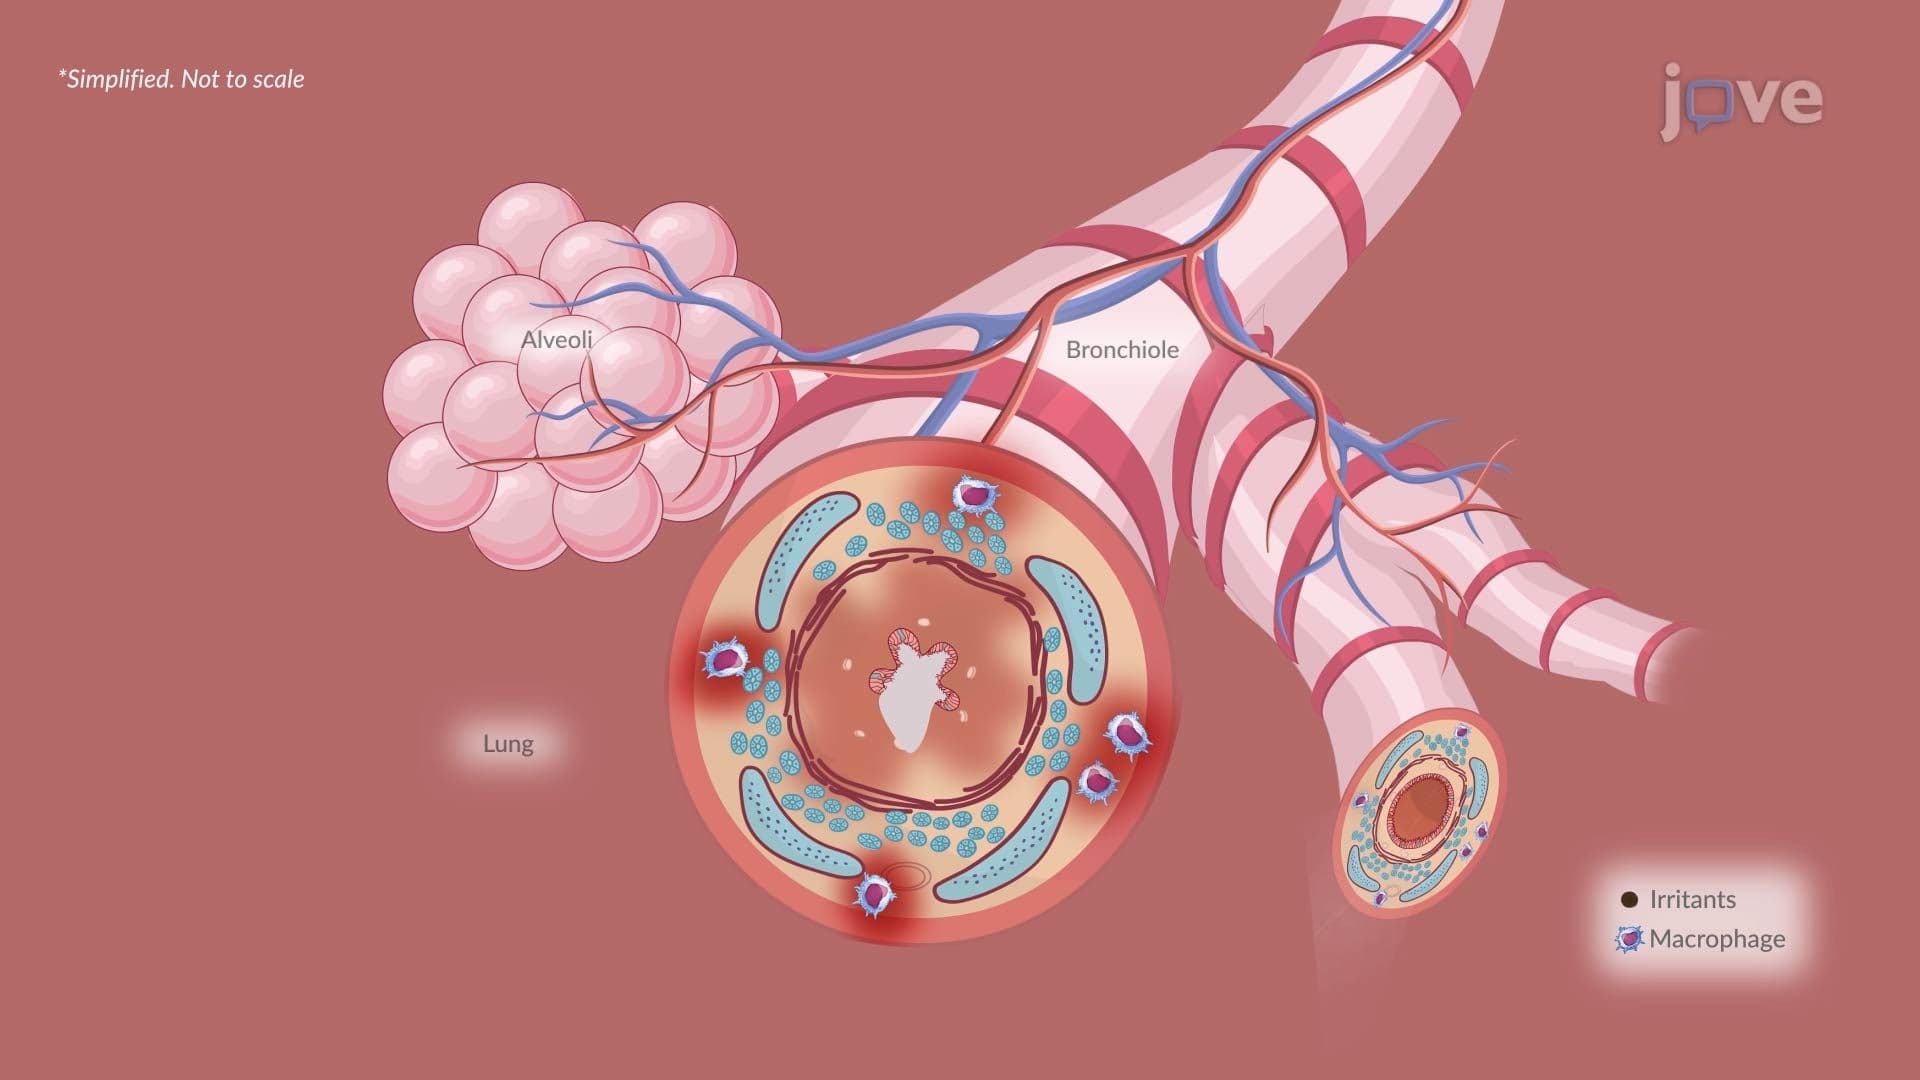
Chronic Obstructive Pulmonary Disease III: Chronic Bronchitis Features
 01:20
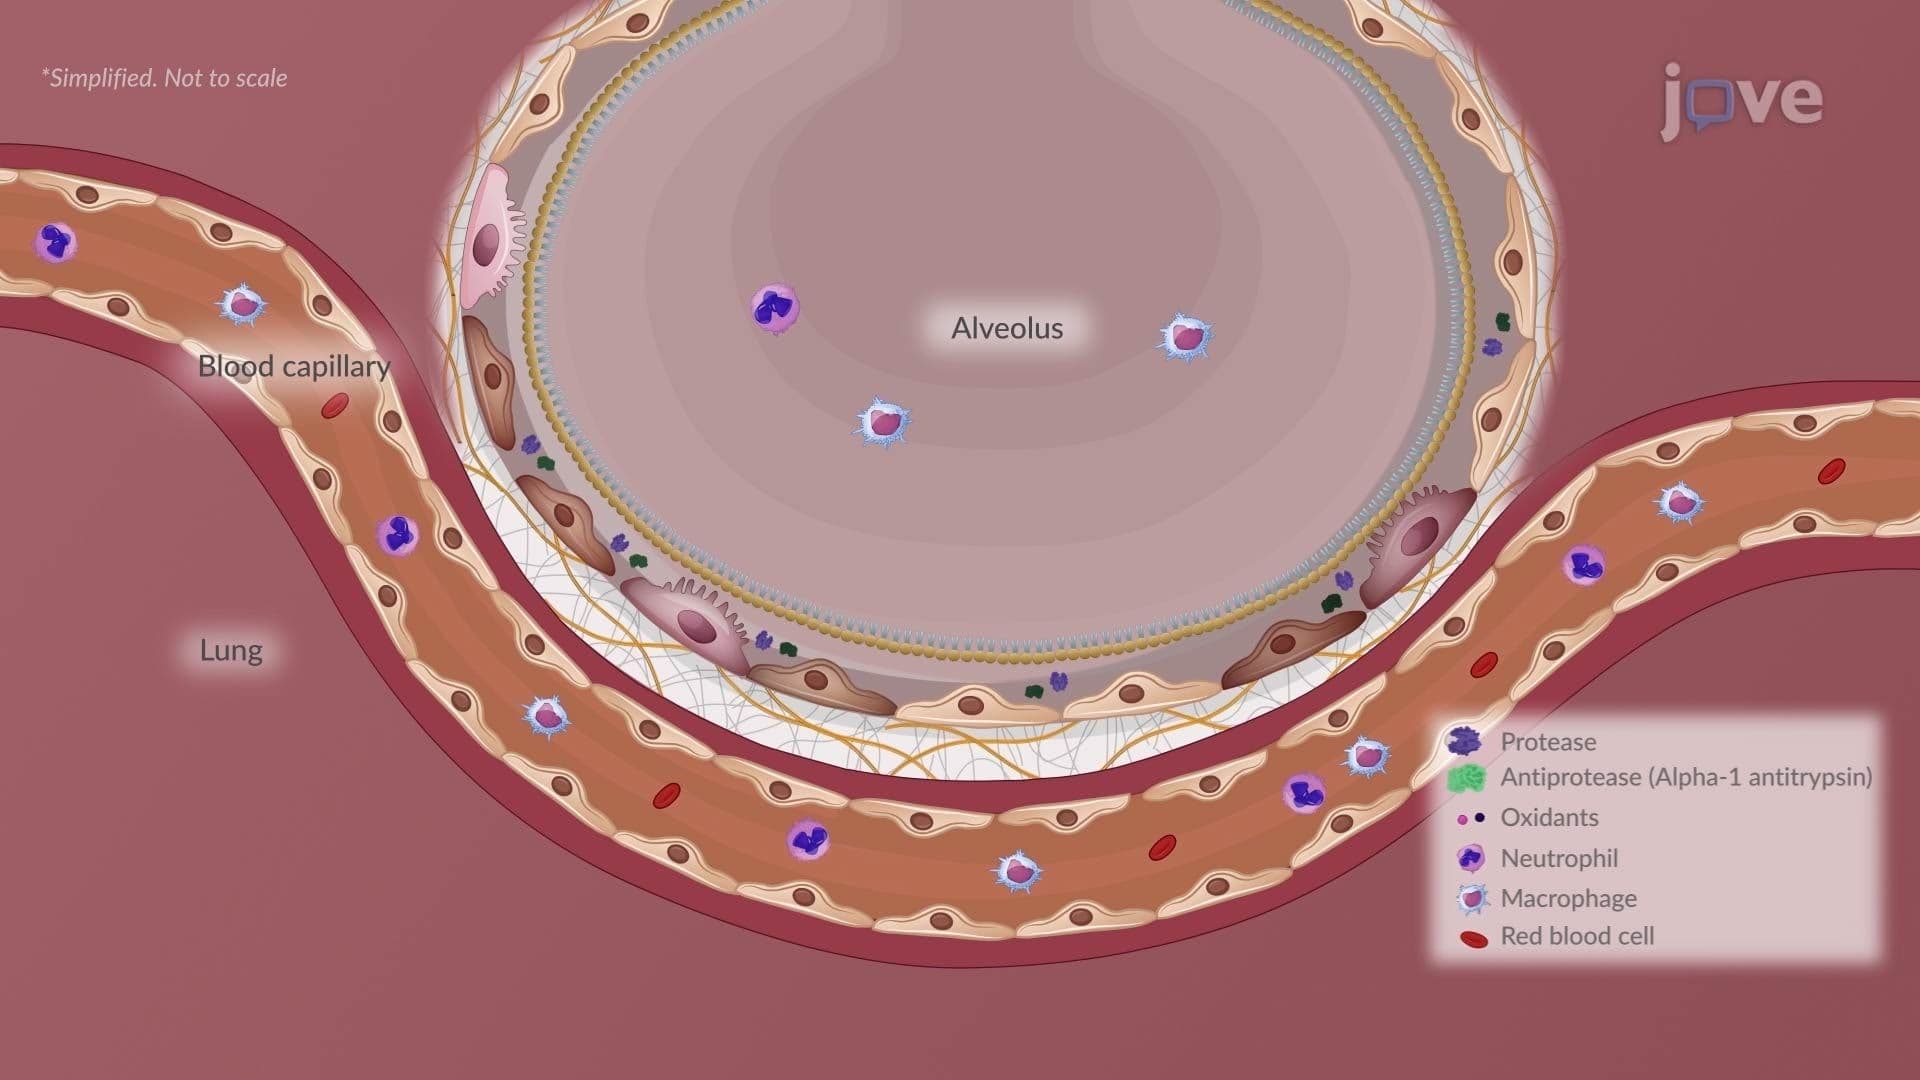
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
Chronic Inflammation
 01:23
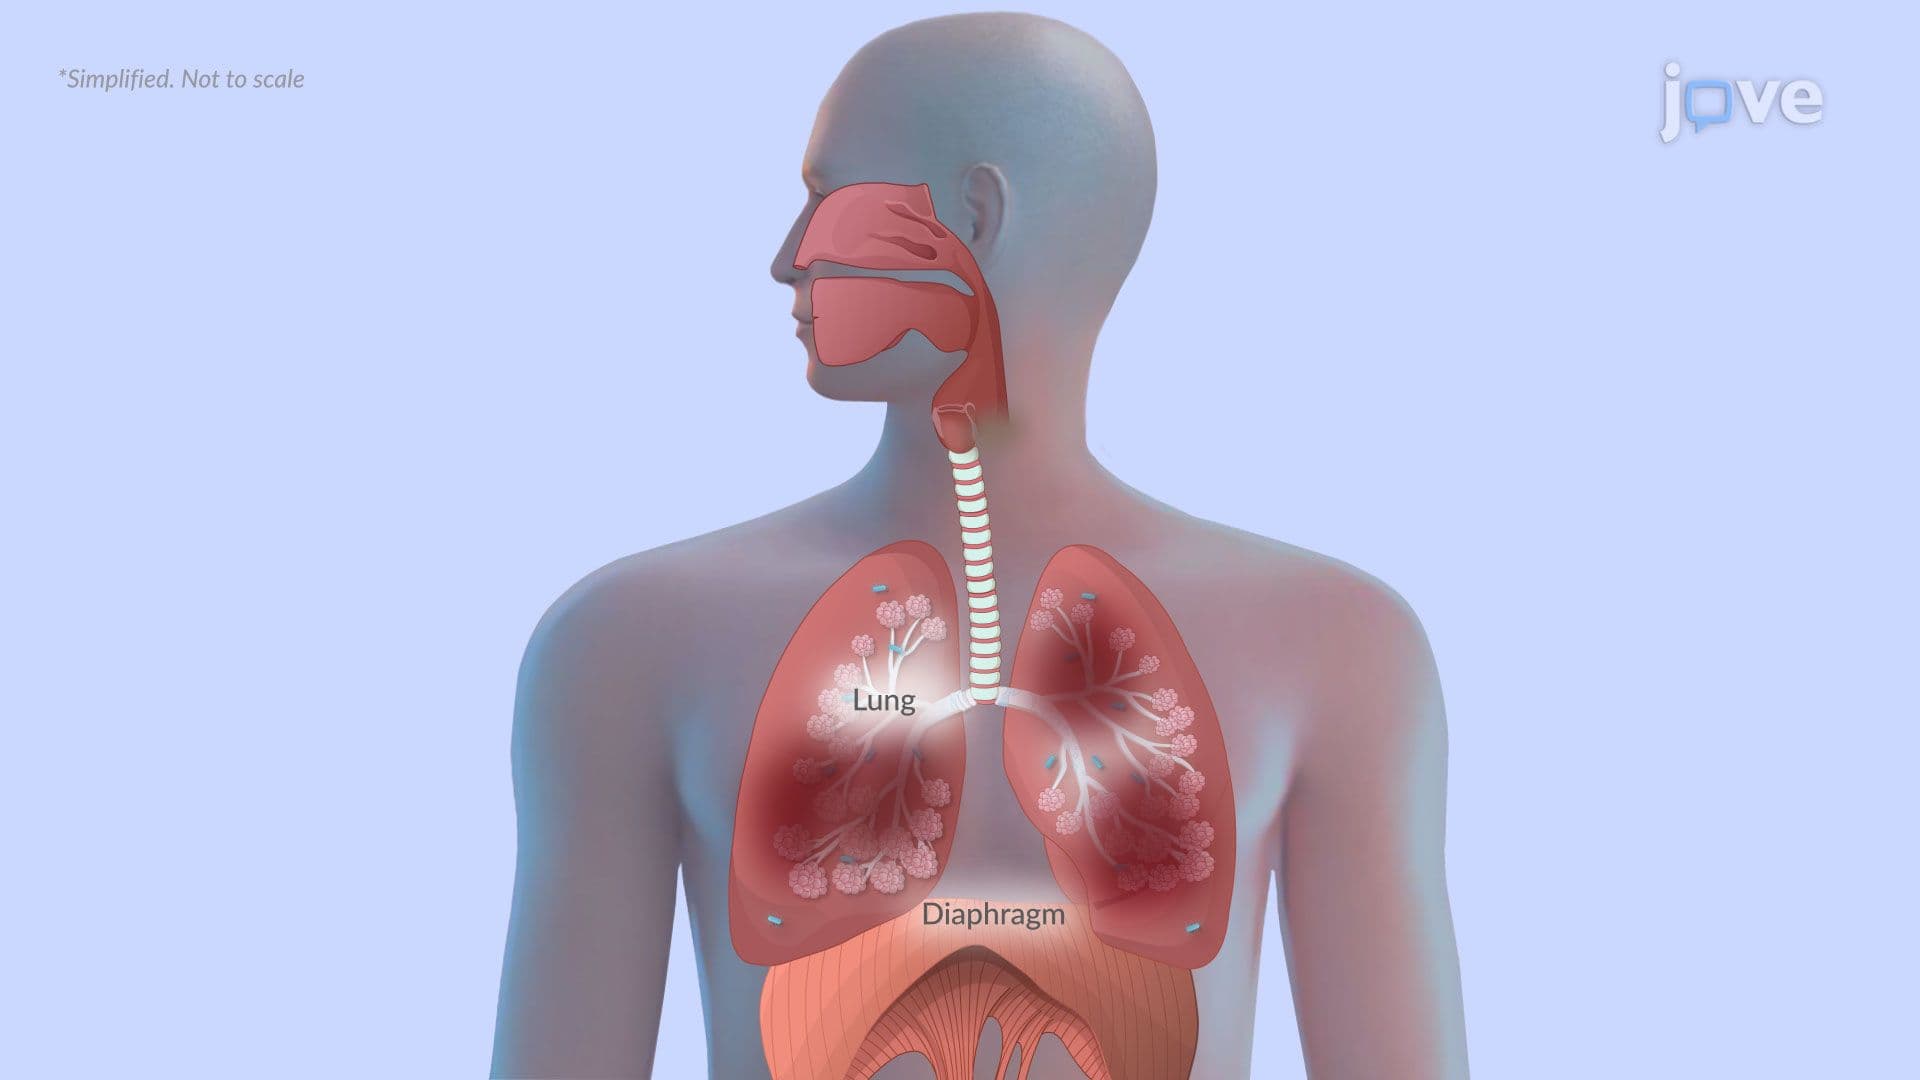
Chronic Obstructive Pulmonary Disease II: Emphysema
 01:23
Chronic Obstructive Pulmonary Disease I: Introduction