Related Experiment Video
Updated: May 17, 2026
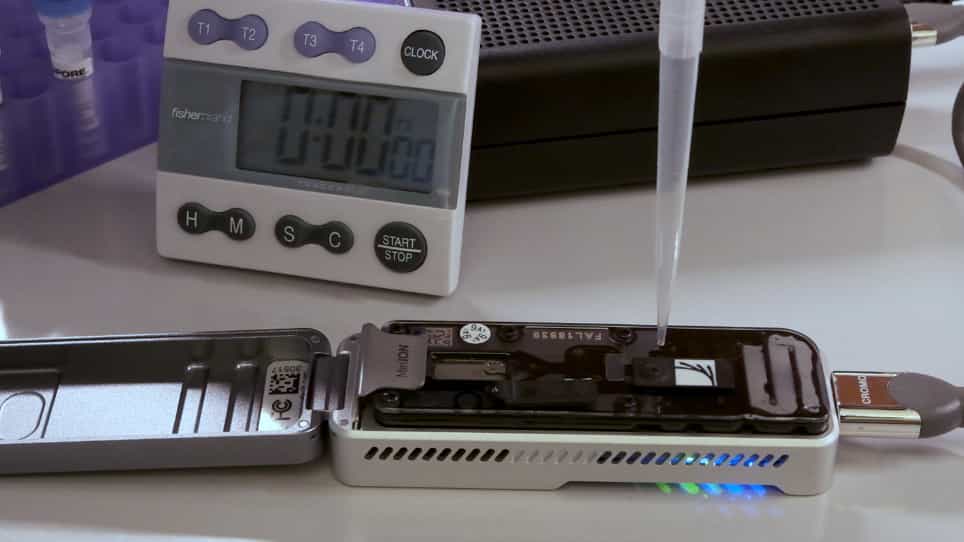

Hybrid De Novo Genome Assembly for the Generation of Complete Genomes of Urinary Bacteria using Short- and Long-read Sequencing Technologies
Published on: August 20, 2021
GapFiller: a de novo assembly approach to fill the gap within paired reads
Francesca Nadalin1, Francesco Vezzi, Alberto Policriti
1Department of Mathematics and Computer Science, University of Udine, Udine 33100, Italy. fnadalin@appliedgenomics.org

Background:
Next Generation Sequencing technologies are able to provide high genome coverages at a relatively low cost. However, due to limited reads' length (from 30 bp up to 200 bp), specific bioinformatics problems have become even more difficult to solve. De novo assembly with short reads, for example, is more complicated at least for two reasons: first, the overall amount of "noisy" data to cope with increased and, second, as the reads' length decreases the number of unsolvable repeats grows. Our work's aim is to go at the root of the problem by providing a pre-processing tool capable to produce (in-silico) longer and highly accurate sequences from a collection of Next Generation Sequencing reads.
Results:
In this paper a seed-and-extend local assembler is presented. The kernel algorithm is a loop that, starting from a read used as seed, keeps extending it using heuristics whose main goal is to produce a collection of error-free and longer sequences. In particular, GapFiller carefully detects reliable overlaps and operates clustering similar reads in order to reconstruct the missing part between the two ends of the same insert. Our tool's output has been validated on 24 experiments using both simulated and real paired reads datasets. The output sequences are declared correct when the seed-mate is found. In the experiments performed, GapFiller was able to extend high percentages of the processed seeds and find their mates, with a false positives rate that turned out to be nearly negligible.
Conclusions:
GapFiller, starting from a sufficiently high short reads coverage, is able to produce high coverages of accurate longer sequences (from 300 bp up to 3500 bp). The procedure to perform safe extensions, together with the mate-found check, turned out to be a powerful criterion to guarantee contigs' correctness. GapFiller has further potential, as it could be applied in a number of different scenarios, including the post-processing validation of insertions/deletions detection pipelines, pre-processing routines on datasets for de novo assembly pipelines, or in any hierarchical approach designed to assemble, analyse or validate pools of sequences.
Related Concept Videos
 03:36
Genome Annotation and Assembly
 02:35
RACE - Rapid Amplification of cDNA Ends
Since the...
 01:57
Sanger Sequencing
 03:21
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while microarray-based...
 03:00
Next-generation Sequencing
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features.
 01:02
Long-patch Base Excision Repair