Related Experiment Video
Updated: May 14, 2026

Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Discovery and optimization of triazine derivatives as ROCK1 inhibitors: molecular docking, molecular dynamics
Mingyun Shen1, Shunye Zhou, Youyong Li
1Institute of Functional Nano & Soft Materials (FUNSOM) and Jiangsu Key Laboratory for Carbon-Based Functional Materials & Devices, Soochow University, Suzhou, Jiangsu 215123, China.

Abstract:
Rho-associated protein kinases (ROCK1 and ROCK2) are promising targets for a number of diseases, including cardiovascular disorders, nervous system diseases, cancers, etc. Recently, we have successfully identified a ROCK1 inhibitor (1) with the triazine core. In order to gain a deeper insight into the microscopic binding of this inhibitor with ROCK1 and design derivatives with improved potency, the interactions between ROCK1 and a series of triazine/pyrimidine-based inhibitors were studied by using an integrated computational protocol that combines molecular docking, molecular dynamics (MD) simulations, binding free energy calculations, and binding energy decomposition analysis. First, three docking protocols, rigid receptor docking, induced fit docking, QM-polarized ligand docking, were used to determine the binding modes of the studied inhibitors in the active site of ROCK1. The results illustrate that rigid receptor docking achieves the best performance to rank the binding affinities of the studied inhibitors. Then, based on the predicted structures from molecular docking, MD simulations and MM/GBSA free energy calculations were employed to determine the dynamic binding process and compare the binding modes of the inhibitors with different activities. The binding free energies predicted by MM/GBSA are in good agreement with the experimental bioactivities, and the analysis of the individual energy terms suggests that the van der Waals interaction is the major driving force for ligand binding. In addition, the residue-inhibitor interaction spectra were obtained by the MM/GBSA free energy decomposition analysis, and the important residues for achieving strong binding were highlighted, which affords important guidance for the rational design of novel ROCK inhibitors. Finally, a variety of derivatives of inhibitor 1 were designed and four of them showed promising potency according to the predictions. We expect that our study can provide significant insight into the development of improved inhibitors of ROCK1.
Related Concept Videos
 01:26
Drug Discovery: Overview
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:16
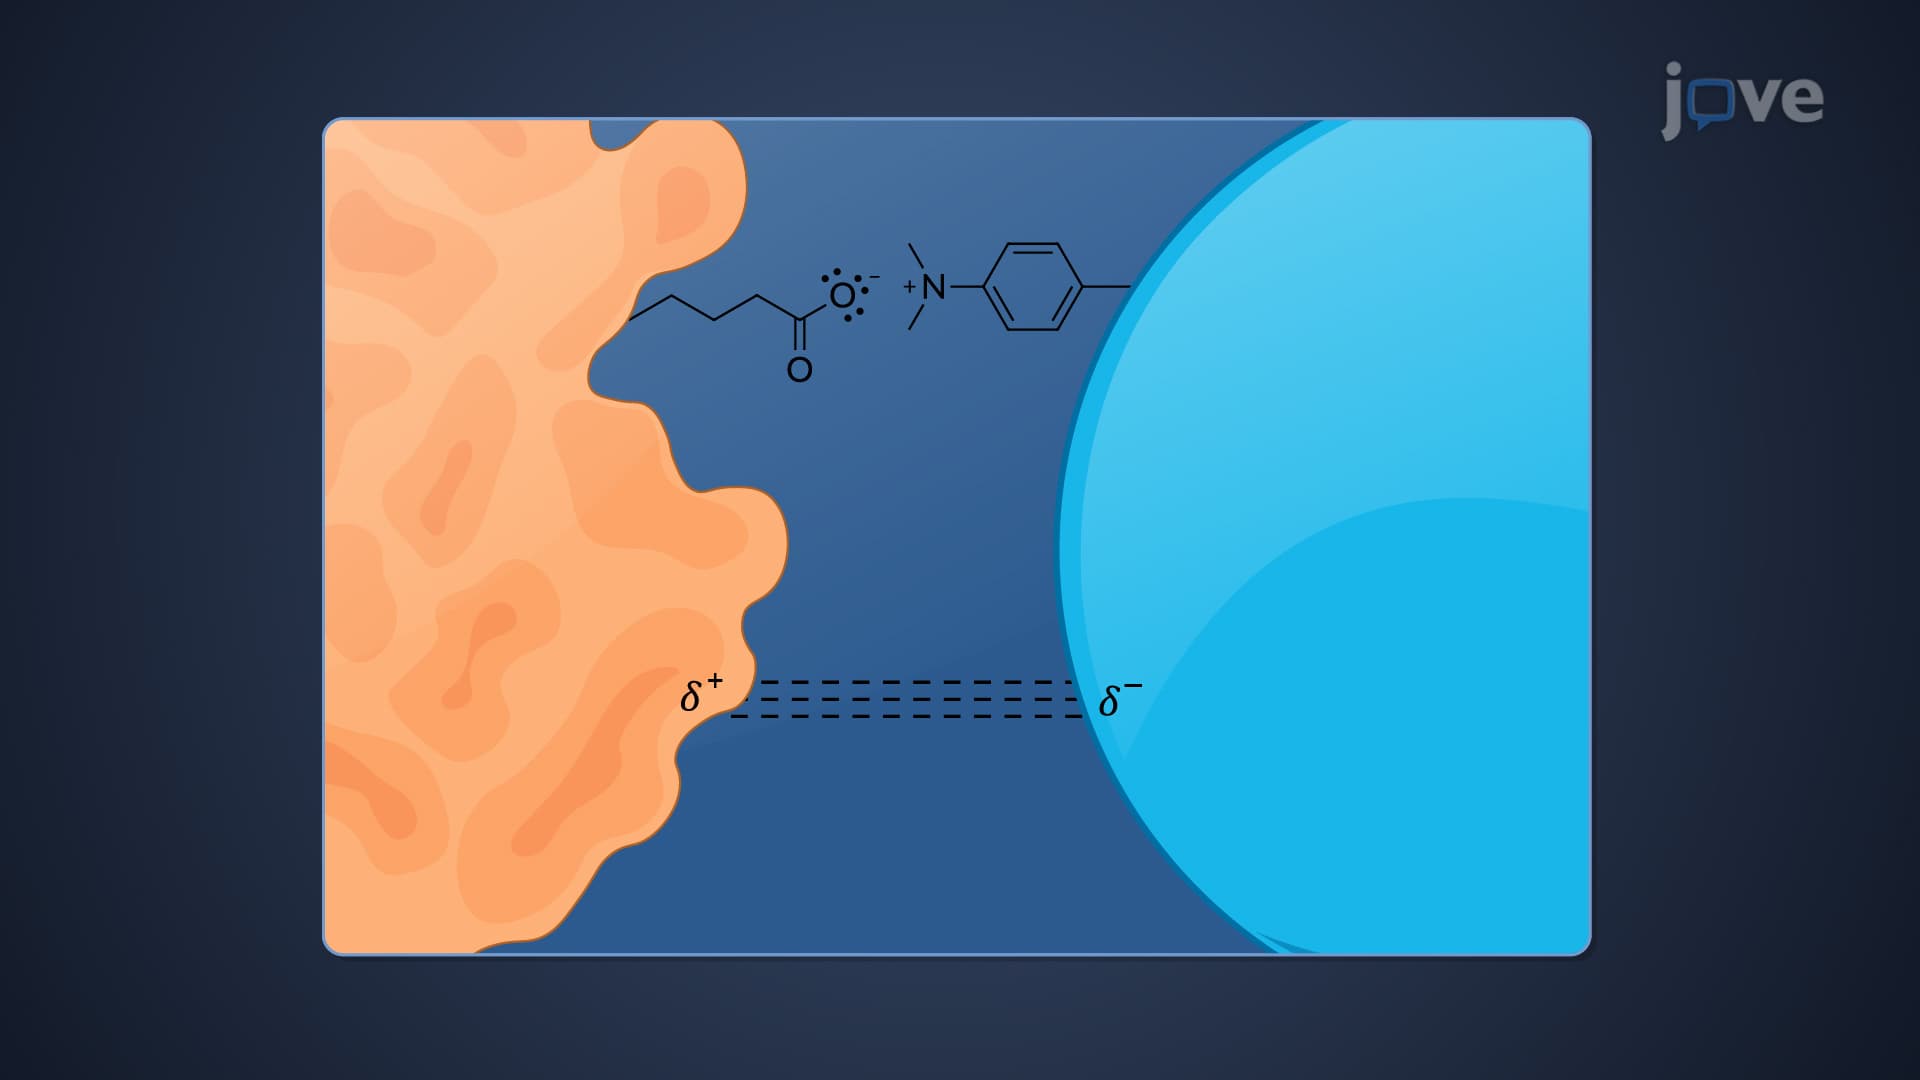
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...