Resonance Raman spectra of ortho-nitrophenol calculated by real-time time-dependent density functional theory

Martin Thomas1, Federico Latorre, Philipp Marquetand

  • 1Institute of Physical Chemistry, Friedrich-Schiller-University Jena, Helmholtzweg 4, 07743 Jena, Germany.

Related Concept Videos

Raman Spectroscopy: Overview01:20

Raman Spectroscopy: Overview

The underlying principle of Raman spectroscopy is based on the interaction between light and matter, specifically molecules' inelastic scattering of photons. When a monochromatic beam of light, typically from a laser source, interacts with a sample, most scattered light has the same frequency as the incident light. This is known as Rayleigh scattering.
However, a small fraction of the scattered light exhibits a frequency shift due to the exchange of energy between the incident photons and the...
IR Spectrum Peak Splitting: Symmetric vs Asymmetric Vibrations01:08

IR Spectrum Peak Splitting: Symmetric vs Asymmetric Vibrations
Identical bonds within a polyatomic group can stretch symmetrically (in-phase) or asymmetrically (out-of-phase). Similar to hydrogen bonding, these vibrations also influence the shape of the IR peak. Generally, asymmetric stretching frequencies are higher than symmetric stretching frequencies. For example, primary amines exhibit two distinct IR peaks between 3300–3500 cm−1 corresponding to the symmetric and asymmetric N-H stretching, while secondary amines exhibit a single stretching vibration...
NMR Spectroscopy of Aromatic Compounds01:14

NMR Spectroscopy of Aromatic Compounds

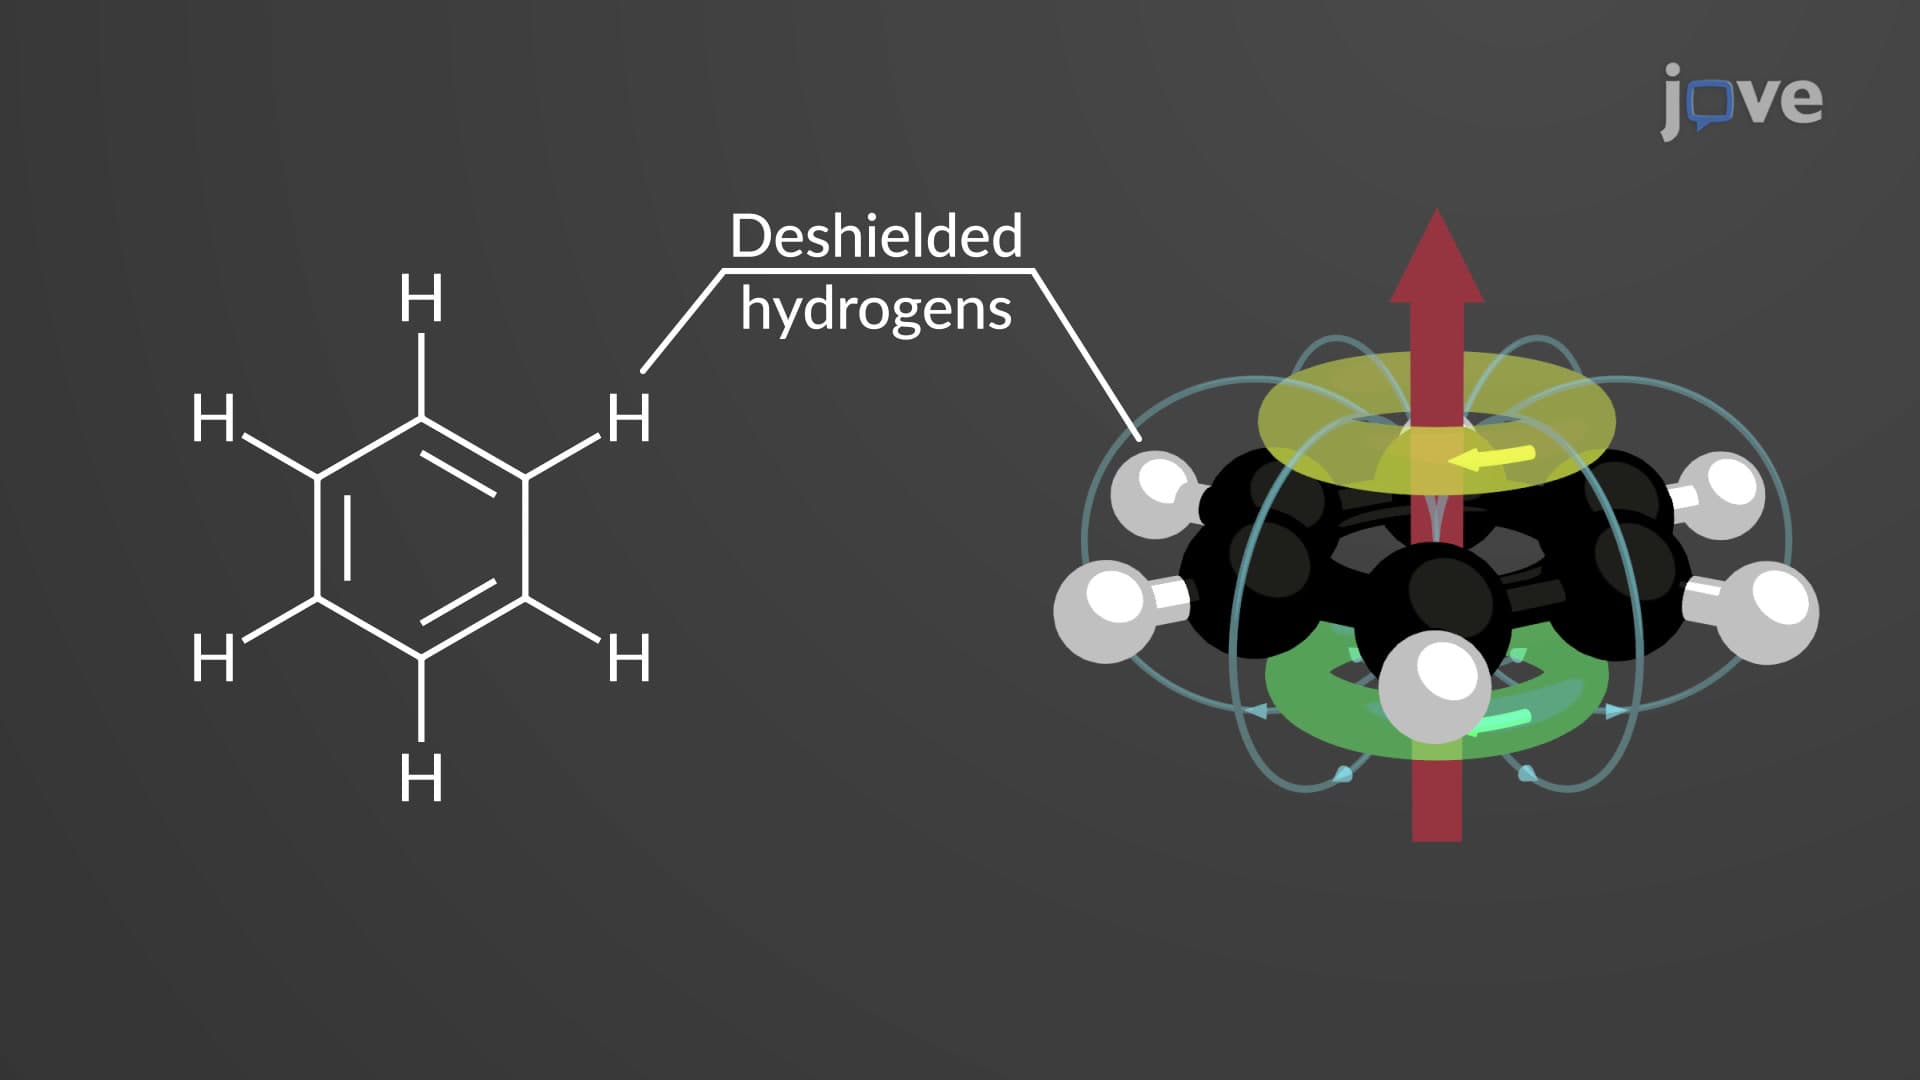
Aromatic compounds can be identified or analyzed using proton NMR and carbon‐13 NMR. Typically, aromatic hydrogens or hydrogens directly bonded to the aromatic rings are strongly deshielded by the aromatic ring current. Therefore, they absorb in the range of 6.5–8.0 ppm in proton NMR spectra. For instance, aromatic hydrogens directly bonded to the benzene ring absorb at 7.3 ppm. However, aromatic hydrogens of larger rings absorb farther upfield or downfield than the ideal range. Consider...
ortho–para-Directing Activators: –CH3, –OH, –⁠NH2, –OCH301:11

ortho–para-Directing Activators: –CH3, –OH, –⁠NH2, –OCH3

All ortho–para directors, excluding halogens, are activating groups. These groups donate electrons to the ring, making the ring carbons electron-rich. Consequently, the reactivity of the aromatic ring towards electrophilic substitution increases. For instance, the nitration of anisole is about 10,000 times faster than the nitration of benzene. The electron-donating effect of the methoxy group in anisole activates the ortho and para positions on the ring and stabilizes the corresponding...
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution00:52

¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
At room temperature, the chair conformer of cyclohexane undergoes rapid ring flipping between two equivalent chair conformers at a rate of approximately 105 times per second. These two chair conformers are in equilibrium. The rapid ring flipping results in the interconversion of the axial proton to an equatorial proton and an equatorial to the axial proton. Such interconversions are too rapid and cannot be detected on the NMR timescale. Hence, the NMR spectrometer cannot distinguish between the...
¹H NMR of Labile Protons: Temporal Resolution01:10

¹H NMR of Labile Protons: Temporal Resolution

Protons bonded to heteroatoms such as nitrogen and oxygen exhibit a range of chemical shift values. This is due to the varying degree of hydrogen bonding between the proton and the heteroatom in other molecules. The extent of hydrogen bonding affects the electron density around the proton, thereby giving different chemical shift values for the protons in the proton NMR spectrum.
The –OH proton in alcohols typically appears in the range of δ 2 to 5 ppm but can vary depending on the specific...