Related Experiment Video
Updated: May 14, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Ligand-optimized homology models of D₁ and D₂ dopamine receptors: application for virtual screening
Marcin Kołaczkowski1, Adam Bucki, Marcin Feder
1Department of Pharmaceutical Chemistry, Jagiellonian University Collegium Medicum , 9 Medyczna Street, 30-688 Kraków, Poland.

Abstract:
Recent breakthroughs in crystallographic studies of G protein-coupled receptors (GPCRs), together with continuous progress in molecular modeling methods, have opened new perspectives for structure-based drug discovery. A crucial enhancement in this area was development of induced fit docking procedures that allow optimization of binding pocket conformation guided by the features of its active ligands. In the course of our research program aimed at discovery of novel antipsychotic agents, our attention focused on dopaminergic D2 and D1 receptors (D2R and D1R). Thus, we decided to investigate whether the availability of a novel structure of the closely related D3 receptor and application of induced fit docking procedures for binding pocket refinement would permit the building of models of D2R and D1R that facilitate a successful virtual screening (VS). Here, we provide an in-depth description of the modeling procedure and the discussion of the results of a VS benchmark we performed to compare efficiency of the ligand-optimized receptors in comparison with the regular homology models. We observed that application of the ligand-optimized models significantly improved the VS performance both in terms of BEDROC (0.325 vs 0.182 for D1R and 0.383 vs 0.301 for D2R) as well as EF1% (20 vs 11 for D1R and 18 vs 10 for D2R). In contrast, no improvement was observed for the performance of a D2R model built on the D3R template, when compared with that derived from the structure of the previously published and more evolutionary distant β2 adrenergic receptor. The comparison of results for receptors built according to various protocols and templates revealed that the most significant factor for the receptor performance was a proper selection of "tool ligand" used in induced fit docking procedure. Taken together, our results suggest that the described homology modeling procedure could be a viable tool for structure-based GPCR ligand design, even for the targets for which only a relatively distant structural template is available.
Related Concept Videos
 01:15
G Protein-coupled Receptors
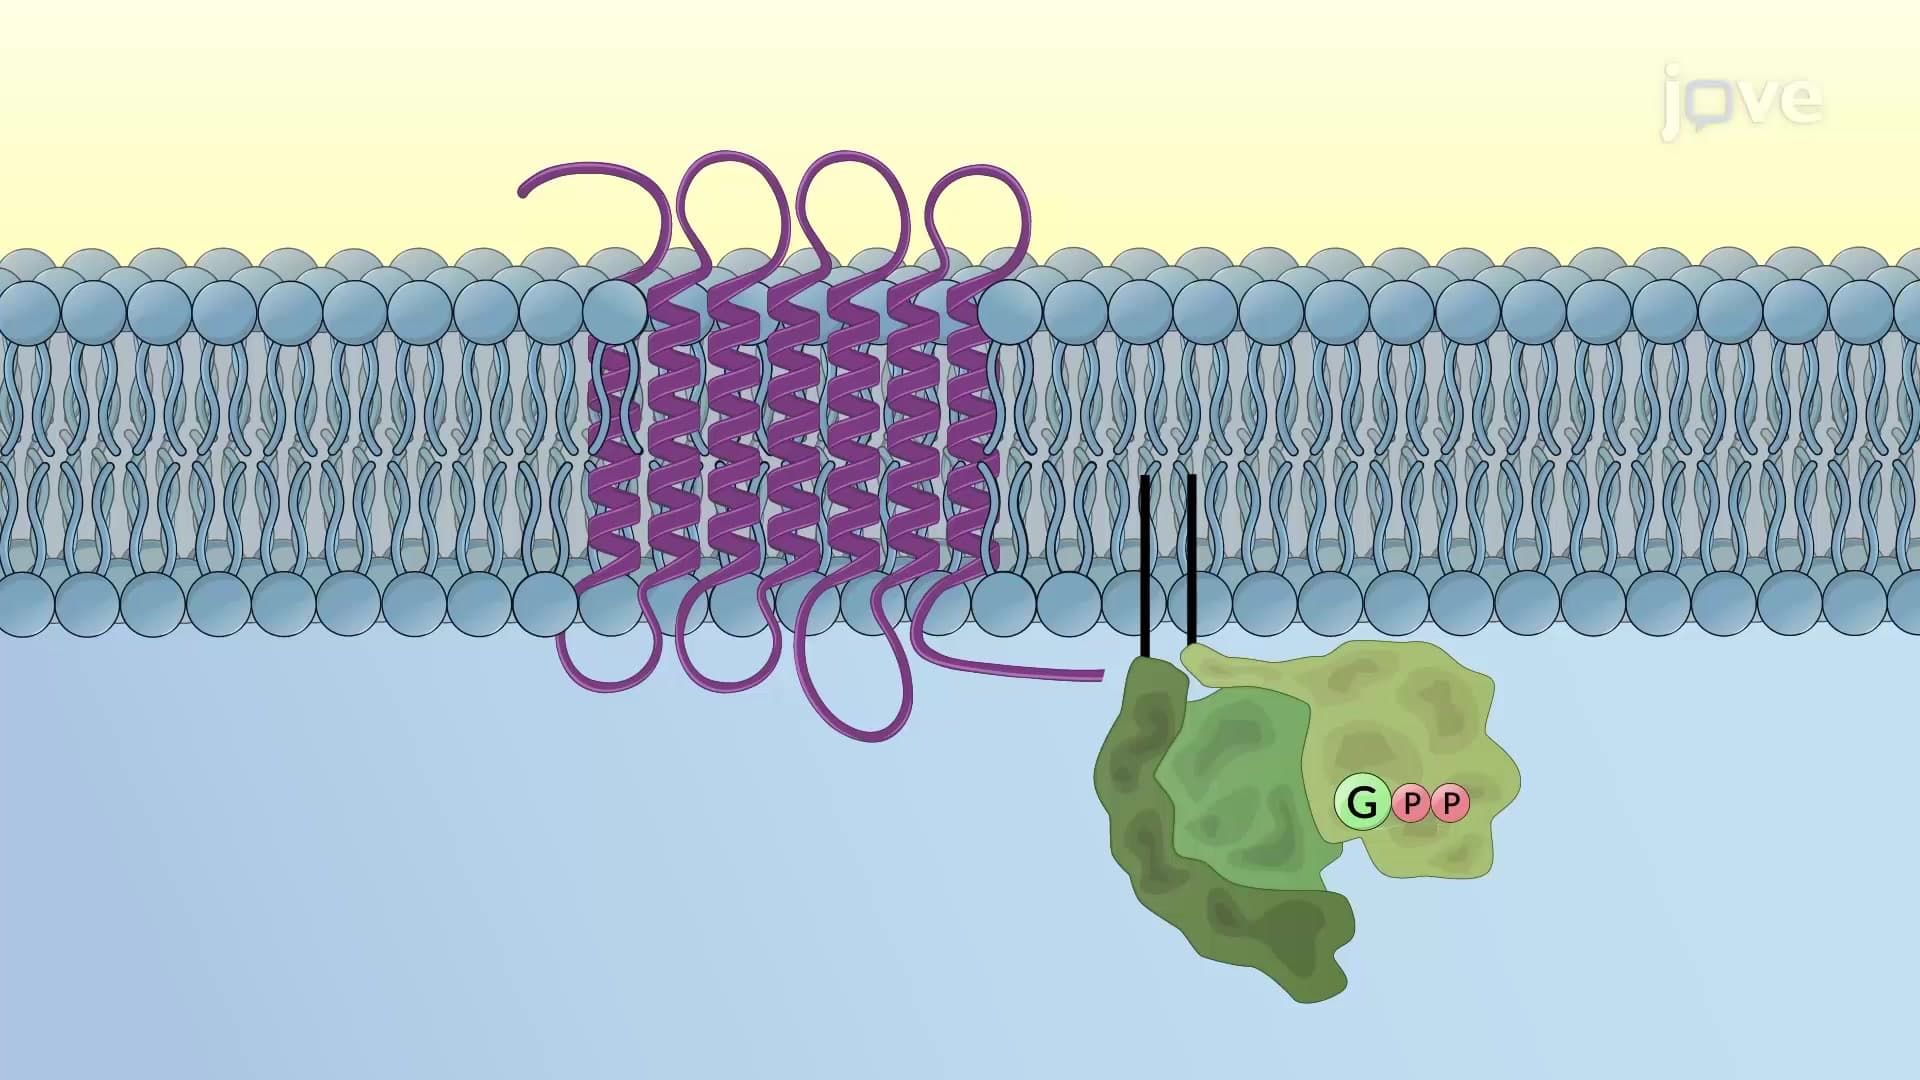
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:26
Drug Discovery: Overview
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...