Related Experiment Video
Updated: May 14, 2026

Metabolic Glycoengineering of Sialic Acid Using N-acyl-modified Mannosamines
Published on: November 25, 2017
Mucopolysacccharidoses: From understanding to treatment, a century of discoveries
1Departamento de Genética and Programa de Pós-Graduação em Genética e Biologia Molecular, Universidade Federal de Rio Grande do Sul, Porto Alegre, RS, Brazil. ; Medical Genetics Service, Hospital de Clínicas de Porto Alegre and WHO Collaborating Center for the Development of Medical Genetic Services in Latin America, Porto Alegre, RS, Brazil. ; Instituto Nacional de Genética Médica Populacional, Porto Alegre, RS, Brazil.

Abstract:
After the first description of a patient recognized as a MPS case was made in 1917, several similar cases were described and identified. Observations reported in the middle of the twentieth century concerning the presence of acid mucopolysaccharides (later called glycosaminoglycans, or GAGs) in tissues and especially in urine of patients were instrumental in providing an identity for these diseases, which became referred as "mucopolysaccharidoses" (MPS). In the late 1960's it was demonstrated that MPS were caused by defects in the breakdown of GAGs, and the specific enzyme deficiencies for the 11 types and subtypes of MPS were identified thereafter. Genes involved in the MPS were subsequently identified, and a large number of disease-causing mutations were identified in each one. Although individually rare, MPS are relatively frequent as a group, with an overall incidence estimated as 1:22,000. The increased excretion of urinary GAGs observed in the vast majority of MPS patients provides a simple screening method, the diagnosis usually being confirmed by the identification of the specific enzyme deficiency. Molecular analysis also plays a role, being helpful for phenotype prediction, prenatal diagnosis and especially for the identification of carriers. As the diseases are rare and diagnosis requires sophisticated methods, the establishment of reference laboratories for MPS identification is recommended. The successful experience of the MPS Brazil Network in providing access to information and diagnosis may be considered as an option for developing countries. The development of therapeutic strategies for MPS, including bone marrow/hematopoietic stem cell transplantation (BMT/HSCT) and enzyme replacement therapy (ERT), changed the natural history of many MPS types. However, some challenges still remain, including the prevention of cognitive decline which occurs in some MPS. Newer approaches, such as intratechal ERT, substrate reduction therapy, read-through, gene therapy and encapsulated modified cells may provide a better outcome for these diseases in the near future. As early diagnosis and early treatment seems to improve treatment outcomes, and as newborn screening is now technically feasible, pilot programs (including one in progress in an area with high-incidence of MPS VI in northeastern Brazil) should provide information about its potential impact in reducing the morbidity associated with MPS diseases.
Insights
Mucopolysaccharidoses (MPS) are rare genetic disorders caused by enzyme deficiencies. Early diagnosis and emerging therapies like gene therapy offer hope for improved outcomes in MPS patients.
Area of Science:
- Biochemistry
- Genetics
- Rare Diseases
Background:
- Mucopolysaccharidoses (MPS) are a group of inherited metabolic disorders characterized by the accumulation of glycosaminoglycans (GAGs).
- Historically, the identification of GAGs in patient tissues and urine led to the naming and understanding of MPS diseases.
- Specific enzyme deficiencies responsible for MPS were identified in the late 1960s, followed by the identification of involved genes and mutations.
Purpose of the Study:
- To provide a comprehensive overview of Mucopolysaccharidoses (MPS), from historical discovery to current therapeutic strategies.
- To highlight the diagnostic approaches, including screening methods and molecular analysis.
- To discuss the evolving landscape of MPS treatments and future therapeutic directions.
Main Methods:
- Review of historical case descriptions and scientific observations regarding MPS.
- Identification of specific enzyme deficiencies and genetic mutations causing MPS.
- Analysis of diagnostic methods, including urinary GAG excretion and molecular testing.
- Evaluation of current and emerging therapeutic interventions for MPS.
Main Results:
- MPS are a group of rare diseases with a combined incidence of approximately 1:22,000.
- Increased urinary GAG excretion serves as a screening tool, with enzyme deficiency confirmation.
- Bone marrow/hematopoietic stem cell transplantation (BMT/HSCT) and enzyme replacement therapy (ERT) have altered disease progression.
- Newer therapies like gene therapy show promise for improved outcomes.
Conclusions:
- Early diagnosis and treatment are crucial for improving outcomes in MPS.
- Reference laboratories and networks like MPS Brazil are vital for diagnosis and information dissemination.
- Ongoing research into novel therapies holds significant potential for managing MPS diseases.
- Newborn screening for MPS is technically feasible and pilot programs are evaluating its impact.
Related Concept Videos
 01:05
Proteoglycans
 01:23
Glycosaminoglycans
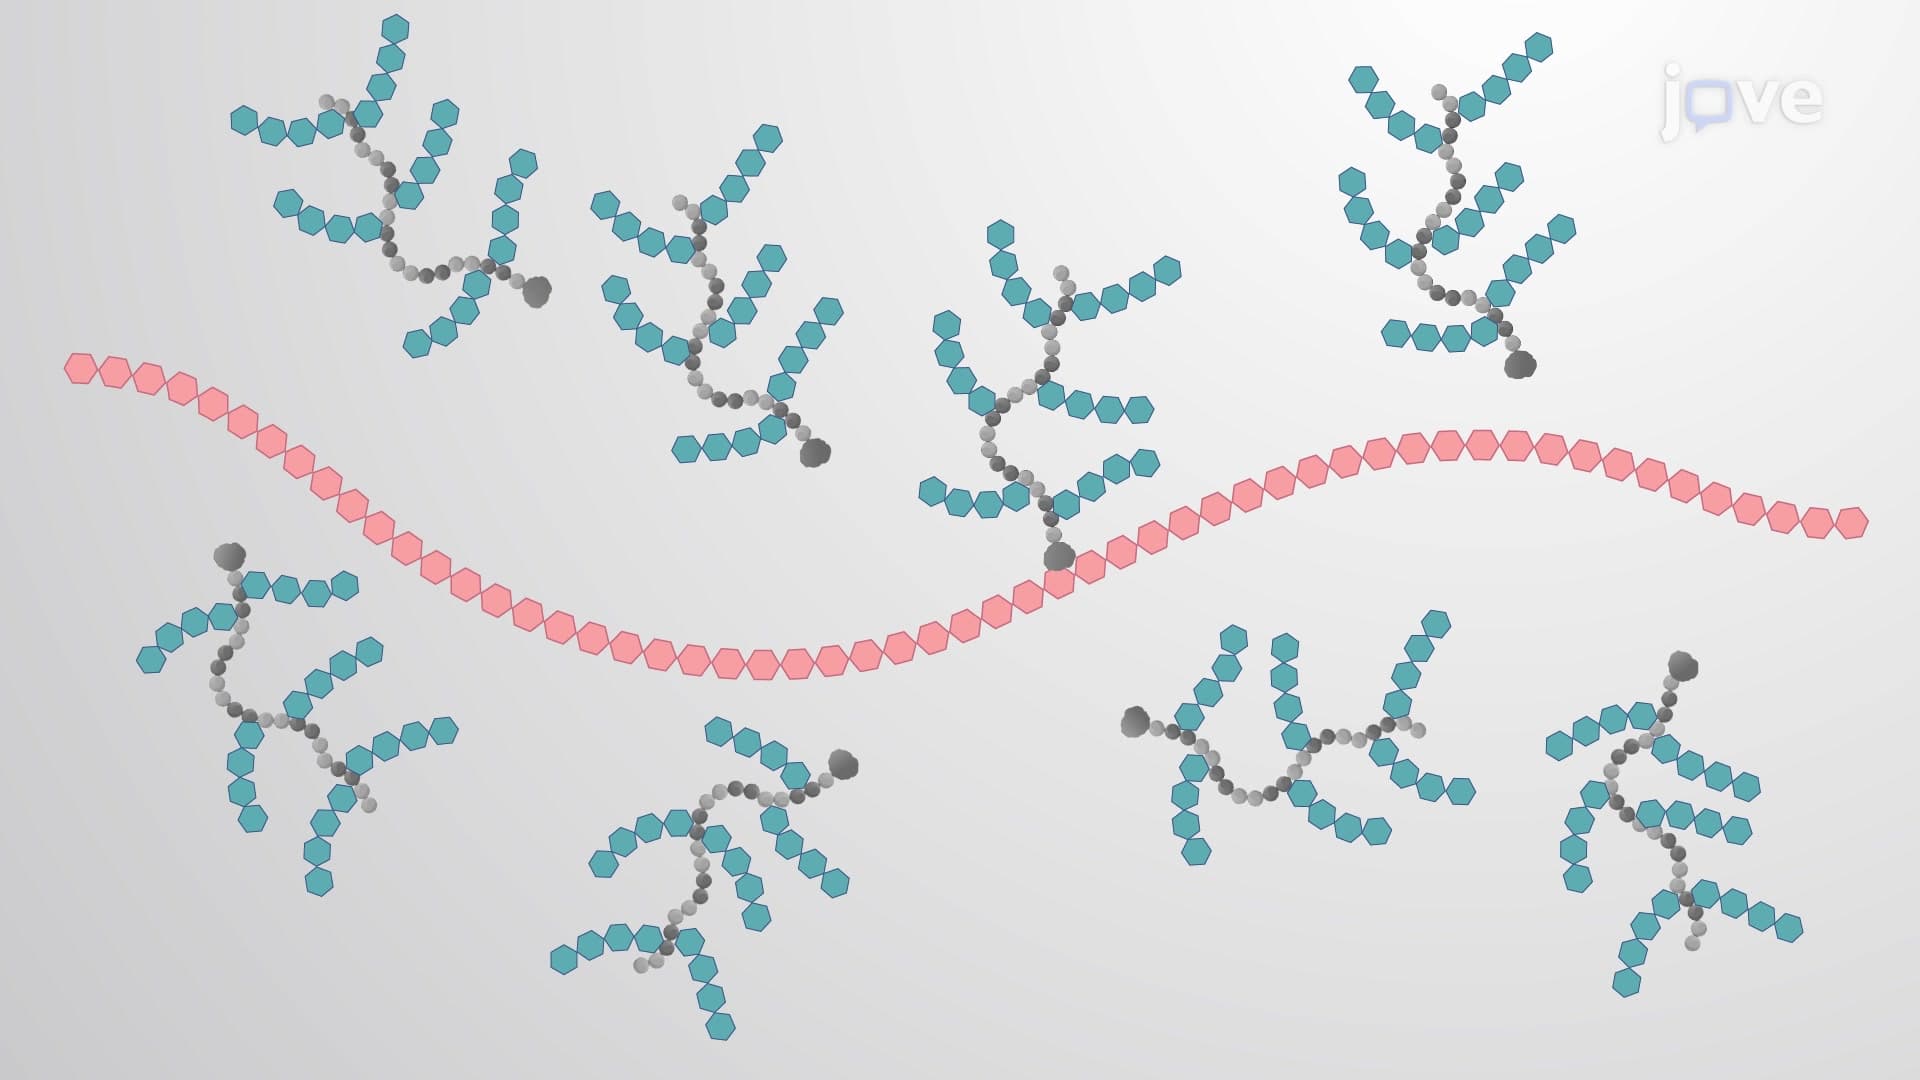
GAGS are found in the extracellular matrix of vertebrates, invertebrates, and bacteria. Due to their polar nature they attract water, and serve as excellent lubricants or shock absorbers in an animal body.
Hyaluronic...
 01:26
Biosynthesis of Polysaccharides
 01:22
Lysosomal Hydrolases
 01:29
Pharmacogenomics: Identification of New Drug Targets
 01:25
Protein Glycosylation
Glycosylation occurs in...