Related Experiment Video
Updated: Feb 20, 2026

High-Resolution Neutron Spectroscopy to Study Picosecond-Nanosecond Dynamics of Proteins and Hydration Water
Published on: April 28, 2022
Absolute hydration free energies of blocked amino acids: implications for protein solvation and stability
Gerhard König1, Stefan Bruckner, Stefan Boresch
1Laboratory of Computational Biology, National Heart, Lung, and Blood Institute, National Institutes of Health, Bethesda, Maryland, USA. gerhard.koenig@nih.gov

Abstract:
Most proteins perform their function in aqueous solution. The interactions with water determine the stability of proteins and the desolvation costs of ligand binding or membrane insertion. However, because of experimental restrictions, absolute solvation free energies of proteins or amino acids are not available. Instead, solvation free energies are estimated based on side chain analog data. This approach implies that the contributions to free energy differences are additive, and it has often been employed for estimating folding or binding free energies. However, it is not clear how much the additivity assumption affects the reliability of the resulting data. Here, we use molecular dynamics-based free energy simulations to calculate absolute hydration free energies for 15 N-acetyl-methylamide amino acids with neutral side chains. By comparing our results with solvation free energies for side chain analogs, we demonstrate that estimates of solvation free energies of full amino acids based on group-additive methods are systematically too negative and completely overestimate the hydrophobicity of glycine. The largest deviation of additive protocols using side chain analog data was 6.7 kcal/mol; on average, the deviation was 4 kcal/mol. We briefly discuss a simple way to alleviate the errors incurred by using side chain analog data and point out the implications of our findings for the field of biophysics and implicit solvent models. To support our results and conclusions, we calculate relative protein stabilities for selected point mutations, yielding a root-mean-square deviation from experimental results of 0.8 kcal/mol.
Related Concept Videos
 03:42
Amino acids
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 01:22
Protein Folding
 02:12
Solvating Effects
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:21
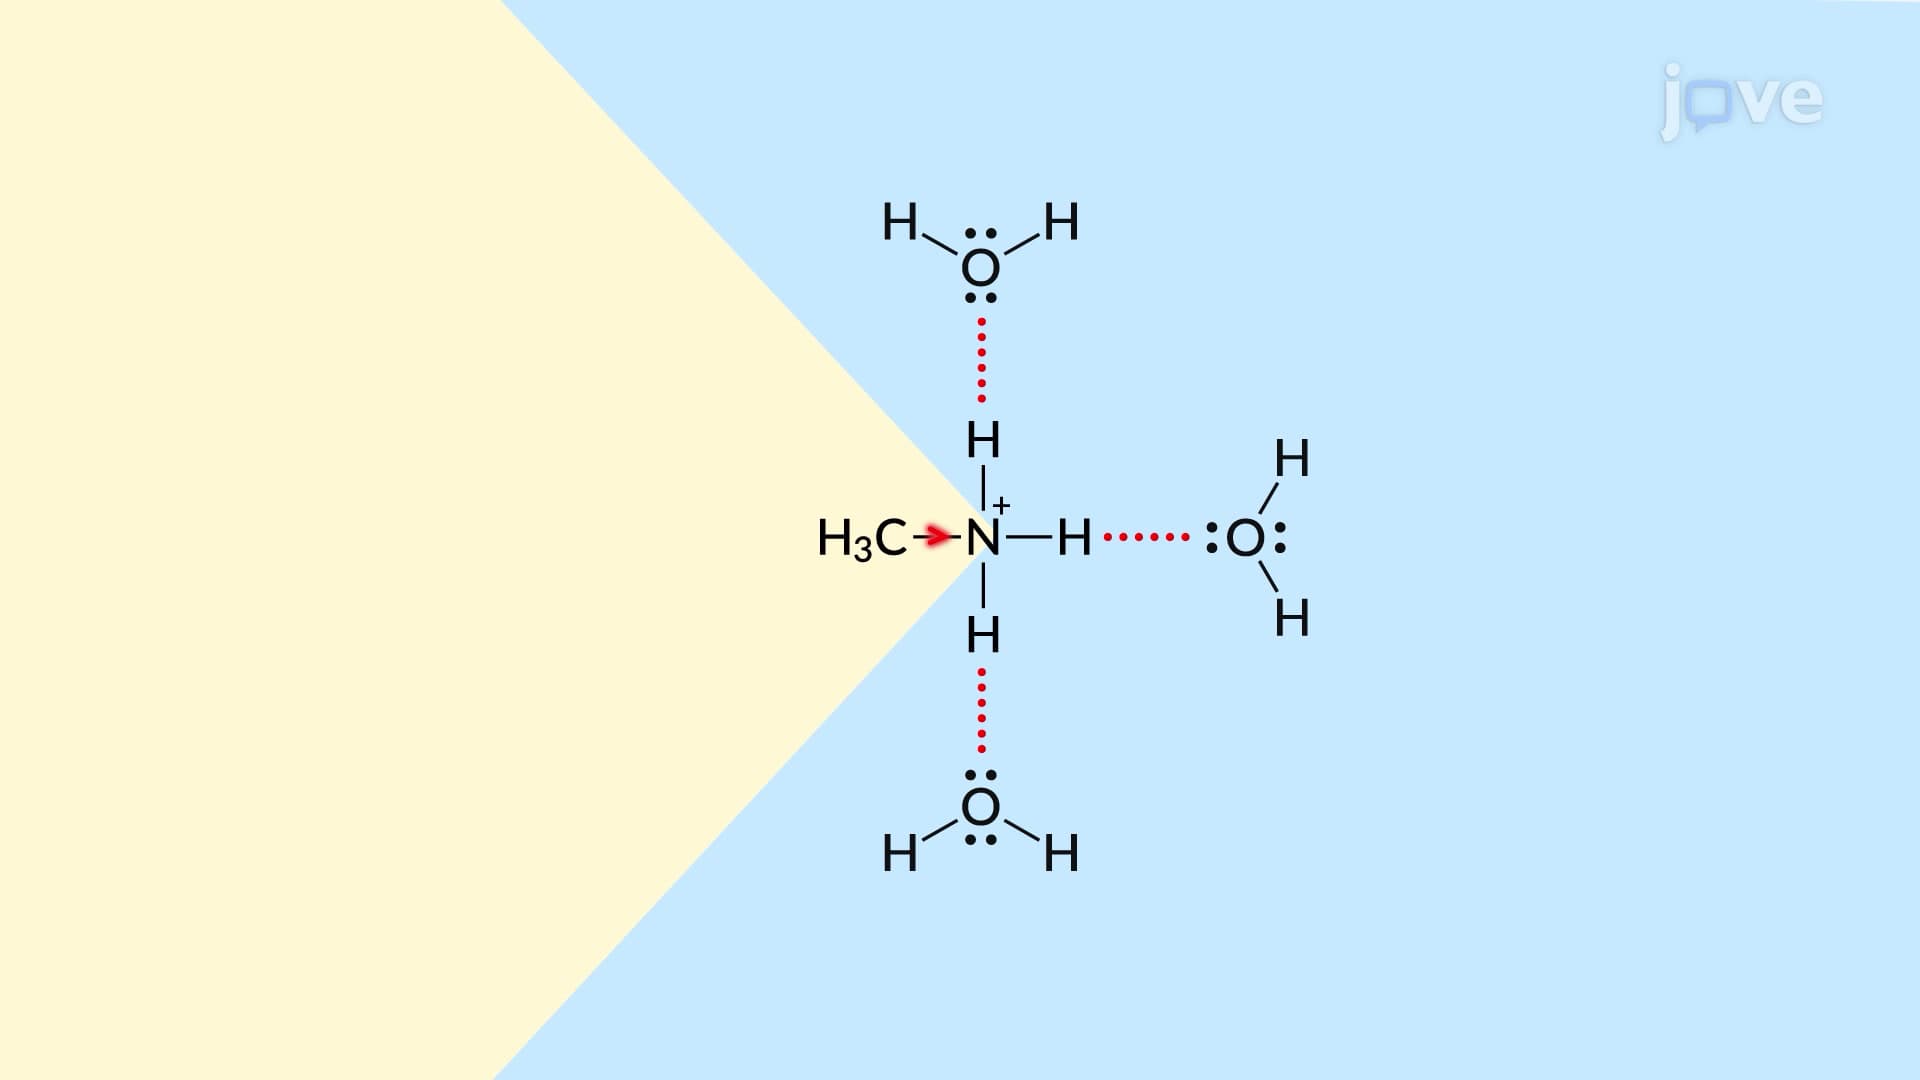
Basicity of Aliphatic Amines
To measure the basicity of amines, two conventions are generally used. The first defines Kb as the basicity constant for the deprotonation reaction of water by the amine, as presented in Figure 1. Conventionally, lower Kb indicates higher...