Related Experiment Video
Updated: May 13, 2026
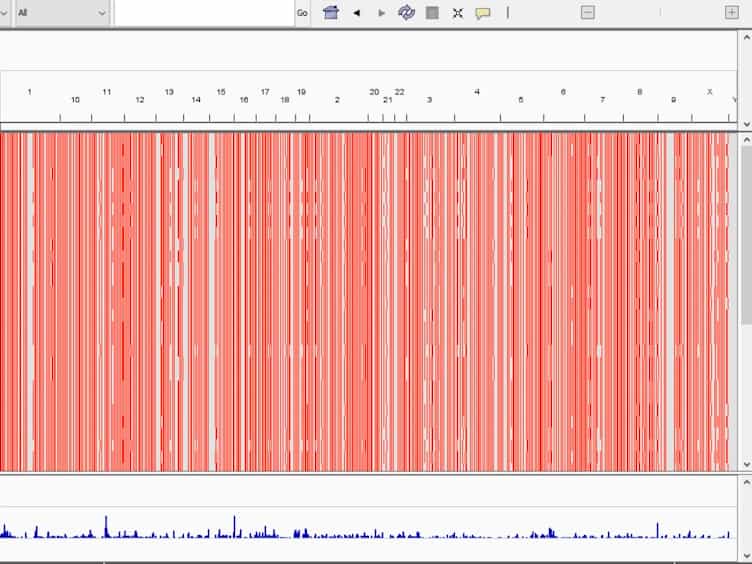

A Fast and Quantitative Method for Post-translational Modification and Variant Enabled Mapping of Peptides to Genomes
Published on: May 22, 2018
Whole human genome proteogenomic mapping for ENCODE cell line data: identifying protein-coding regions
Jainab Khatun1, Yanbao Yu, John A Wrobel
1College of Arts and Sciences, Boise State University, Boise, ID, USA. jainabkhatun@boisestate.edu

Background:
Proteogenomic mapping is an approach that uses mass spectrometry data from proteins to directly map protein-coding genes and could aid in locating translational regions in the human genome. In concert with the ENcyclopedia of DNA Elements (ENCODE) project, we applied proteogenomic mapping to produce proteogenomic tracks for the UCSC Genome Browser, to explore which putative translational regions may be missing from the human genome.
Results:
We generated ~1 million high-resolution tandem mass (MS/MS) spectra for Tier 1 ENCODE cell lines K562 and GM12878 and mapped them against the UCSC hg19 human genome, and the GENCODE V7 annotated protein and transcript sets. We then compared the results from the three searches to identify the best-matching peptide for each MS/MS spectrum, thereby increasing the confidence of the putative new protein-coding regions found via the whole genome search. At a 1% false discovery rate, we identified 26,472, 24,406, and 13,128 peptides from the protein, transcript, and whole genome searches, respectively; of these, 481 were found solely via the whole genome search. The proteogenomic mapping data are available on the UCSC Genome Browser at http://genome.ucsc.edu/cgi-bin/hgTrackUi?db=hg19&g=wgEncodeUncBsuProt.
Conclusions:
The whole genome search revealed that ~4% of the uniquely mapping identified peptides were located outside GENCODE V7 annotated exons. The comparison of the results from the disparate searches also identified 15% more spectra than would have been found solely from a protein database search. Therefore, whole genome proteogenomic mapping is a complementary method for genome annotation when performed in conjunction with other searches.
Related Concept Videos
 02:24
Ribosome Profiling
Applications of ribosome profiling
Ribosome profiling has many applications, including in vivo monitoring of translation inside a particular organ or tissue type and quantifying new protein synthesis levels.
The technique helps...
 01:33
Proteomics
Proteomics is the study of proteomes' function. It involves the large-scale systematic study of the proteome to denote the protein complement expressed by a genome. Scientist Mark Wilkins coined the term proteomics...
 00:58
Genomic DNA in Eukaryotes
 03:36
Genome Annotation and Assembly
 02:02
Genomics
 00:46
Genomic DNA in Prokaryotes
Genomic Diversity in Bacteria
Although bacterial genomes are much...