Related Experiment Video
Updated: May 12, 2026

Genotyping Single Nucleotide Polymorphisms in the Mitochondrial Genome by Pyrosequencing
Published on: February 10, 2023
Clinical and molecular features of POLG-related mitochondrial disease
Jeffrey D Stumpf1, Russell P Saneto, William C Copeland
1Laboratory of Molecular Genetics, National Institute of Environmental Health Sciences, National Institutes of Health, Research Triangle Park, NC 27709, USA.

Abstract:
The inability to replicate mitochondrial genomes (mtDNA) by the mitochondrial DNA polymerase (pol γ) leads to a subset of mitochondrial diseases. Many mutations in POLG, the gene that encodes pol γ, have been associated with mitochondrial diseases such as myocerebrohepatopathy spectrum (MCHS) disorders, Alpers-Huttenlocher syndrome, myoclonic epilepsy myopathy sensory ataxia (MEMSA), ataxia neuropathy spectrum (ANS), and progressive external ophthalmoplegia (PEO). This chapter explores five important topics in POLG-related disease: (1) clinical symptoms that identify and distinguish POLG-related diseases, (2) molecular characterization of defects in polymerase activity by POLG disease variants, (3) the importance of holoenzyme formation in disease presentation, (4) the role of pol γ exonuclease activity and mutagenesis in disease and aging, and (5) novel approaches to therapy and avoidance of toxicity based on primary research in pol γ replication.
Insights
Mitochondrial DNA polymerase (pol γ) defects cause POLG-related diseases. This chapter details clinical symptoms, molecular causes, and novel therapeutic strategies for these debilitating conditions.
Area of Science:
- Mitochondrial Biology
- Genetics
- Neurology
Background:
- Defects in mitochondrial DNA replication, specifically by the mitochondrial DNA polymerase (pol γ), are linked to various inherited diseases.
- Mutations in the POLG gene, encoding pol γ, are associated with a spectrum of severe conditions including Alpers-Huttenlocher syndrome and progressive external ophthalmoplegia (PEO).
Purpose of the Study:
- To provide a comprehensive overview of POLG-related mitochondrial diseases.
- To elucidate the clinical presentations, molecular mechanisms, and therapeutic avenues for these disorders.
Main Methods:
- Review of clinical data and genetic analyses of POLG mutations.
- Molecular characterization of polymerase activity and holoenzyme formation.
- Exploration of exonuclease activity, mutagenesis, and novel therapeutic research.
Main Results:
- Identified distinct clinical symptoms differentiating various POLG-related diseases.
- Characterized molecular defects in pol γ activity and the impact of holoenzyme formation.
- Highlighted the role of pol γ exonuclease activity in disease pathogenesis and aging.
Conclusions:
- Understanding POLG mutations and their functional consequences is crucial for diagnosing and managing mitochondrial diseases.
- Novel therapeutic strategies targeting pol γ replication are under development to address toxicity and improve patient outcomes.
More Related Videos
Related Concept Videos
 01:24
Parkinson Disease ll: Pathophysiology
 02:59
Animal Mitochondrial Genetics
 01:22
Lysosomal Hydrolases
 01:39
Mitochondrial Precursor Proteins
Most of the mitochondrial precursors...
 01:24
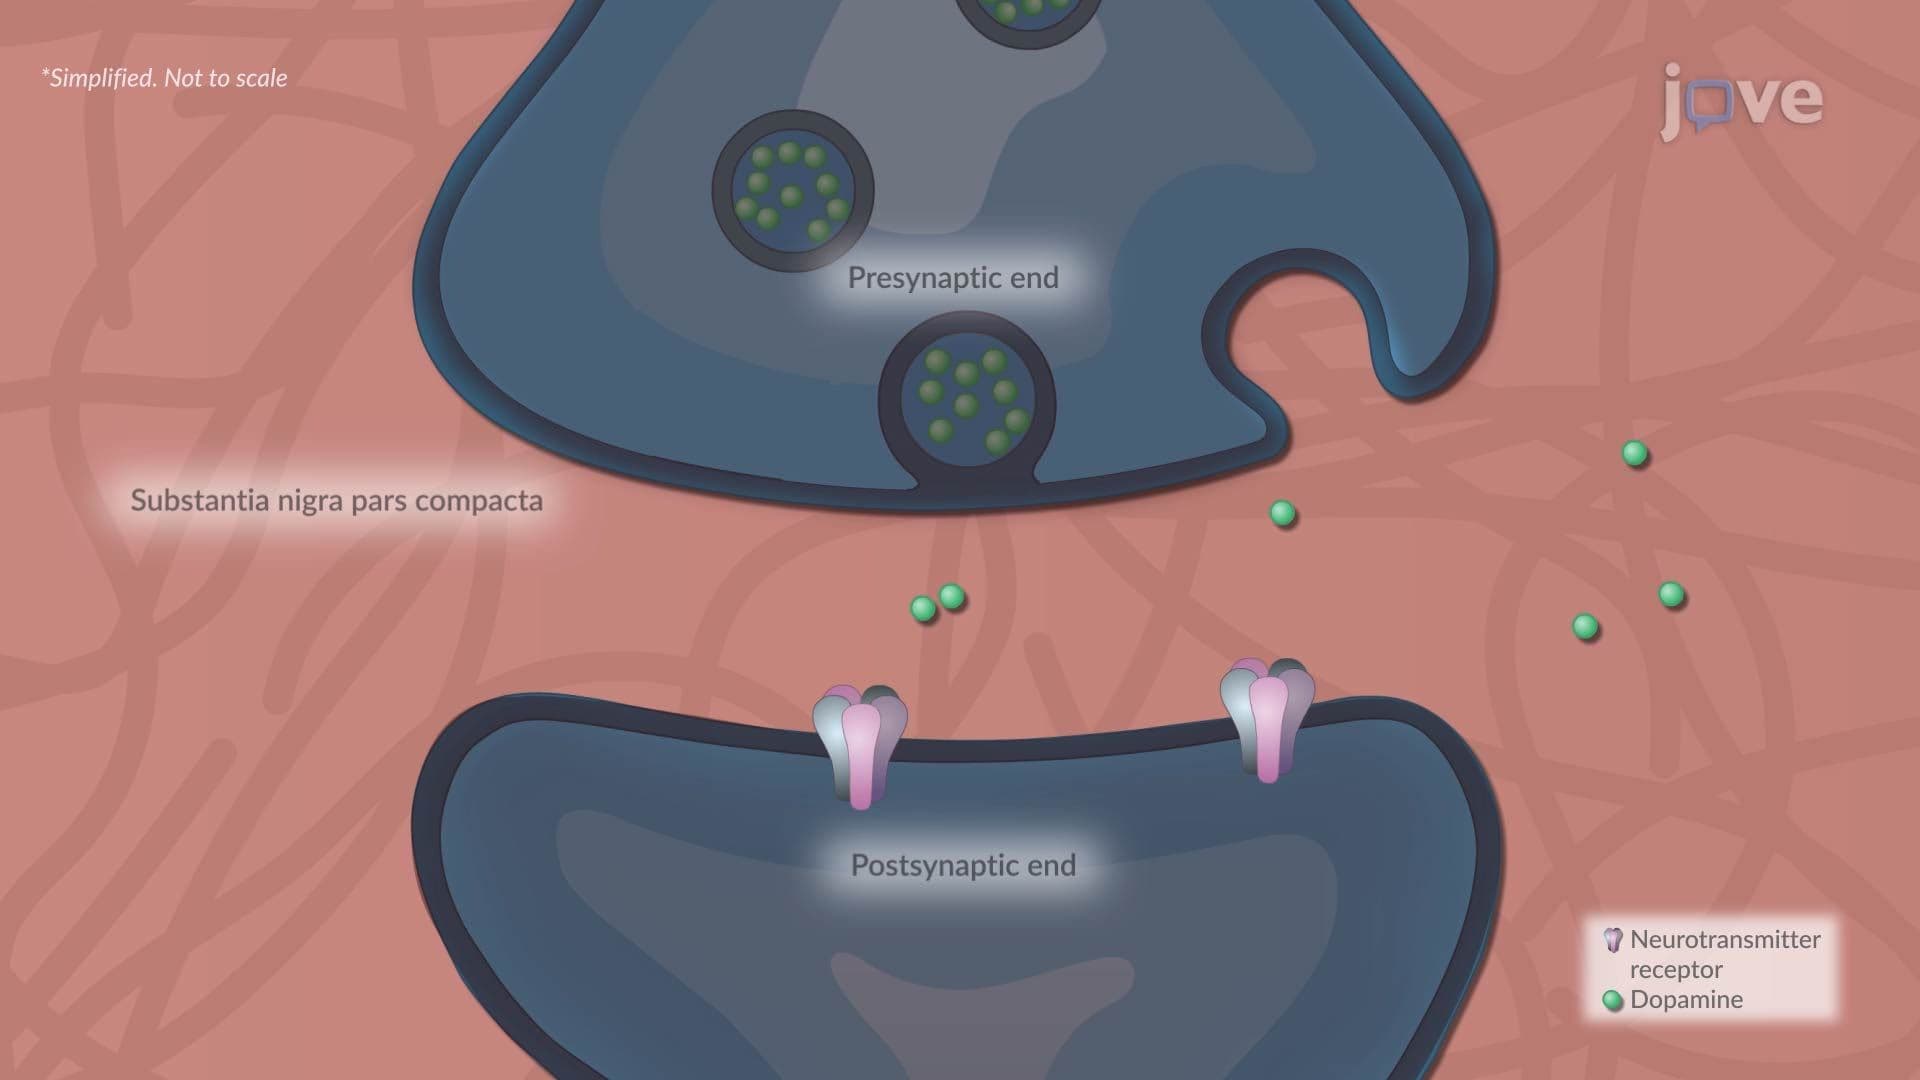
Parkinson Disease l: Introduction
 01:39
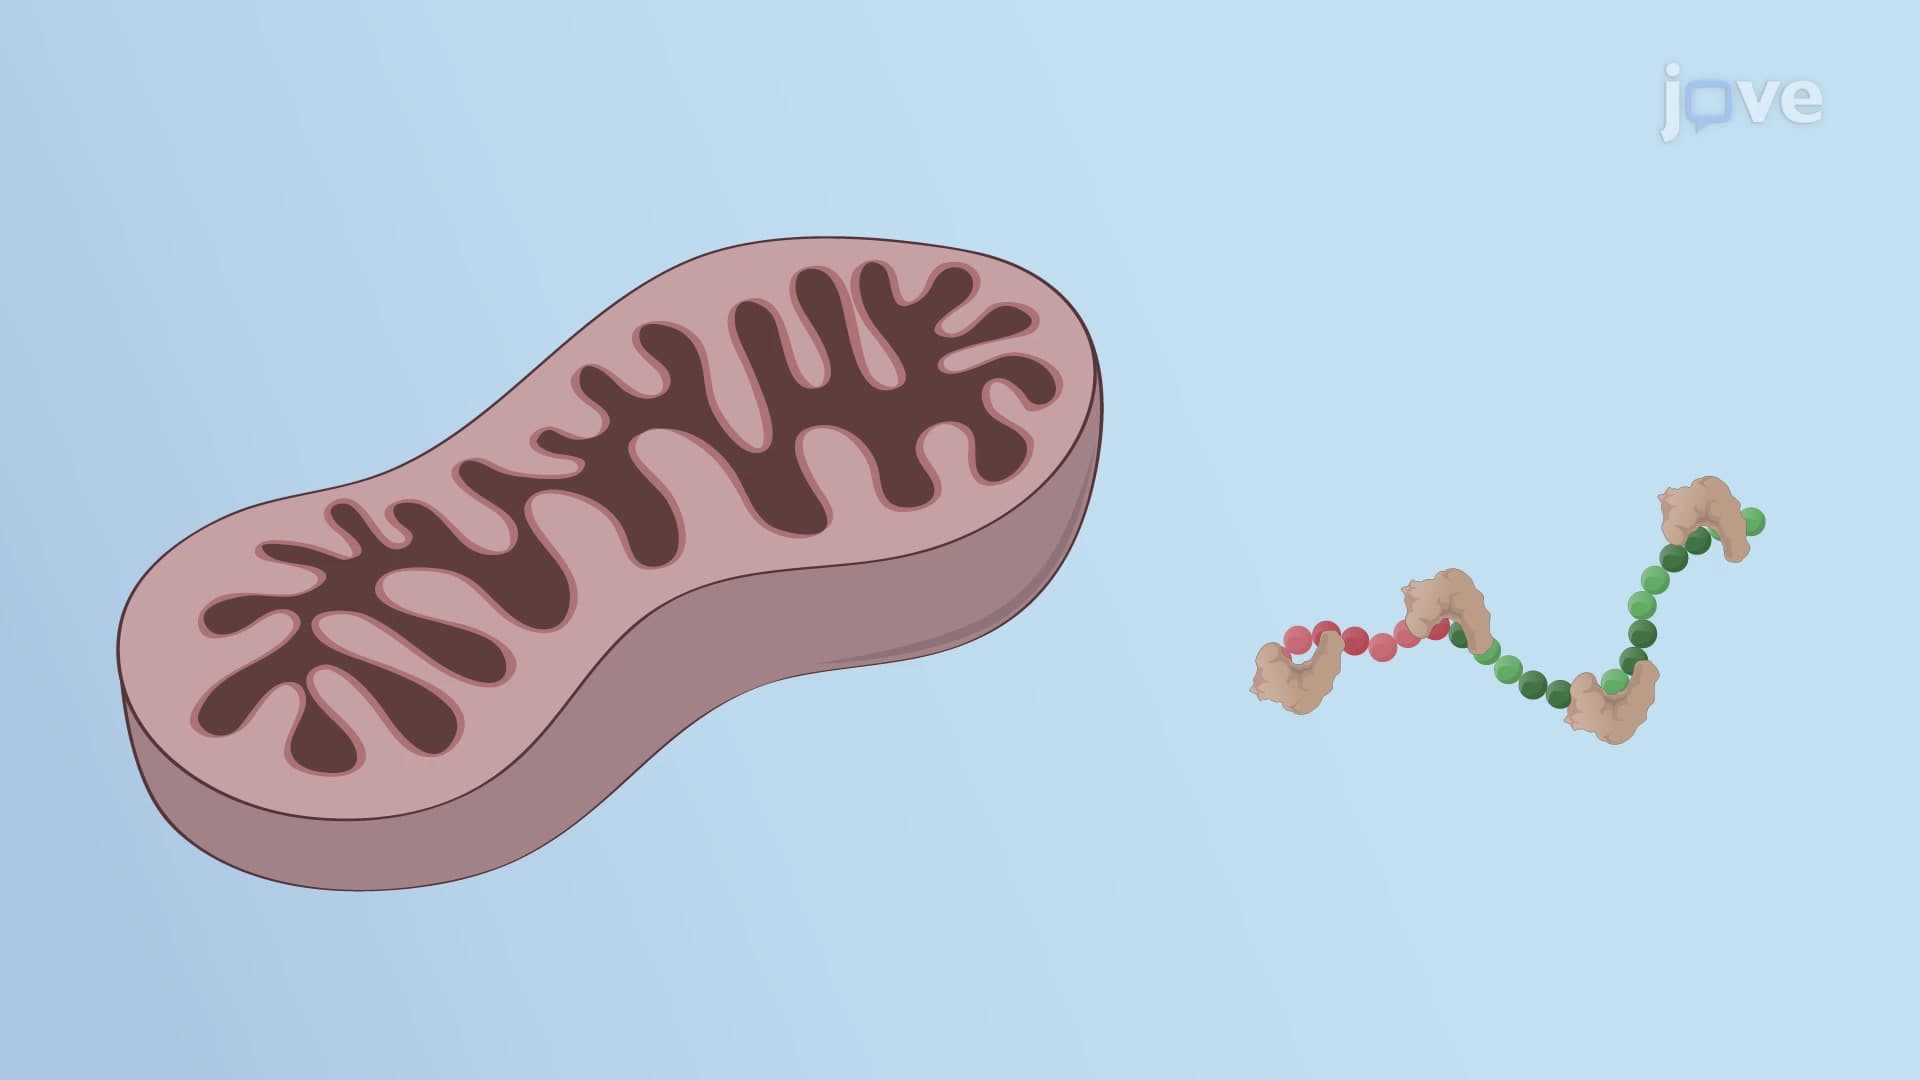
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...