

Characterization of biaryl torsional energetics and its treatment in OPLS all-atom force fields
Markus K Dahlgren1, Patric Schyman, Julian Tirado-Rives
1Department of Chemistry, Yale University, New Haven, Connecticut 06520-8107, United States.

Abstract:
The frequency of biaryl substructures in a database of approved oral drugs has been analyzed. This led to designation of 20 prototypical biaryls plus 10 arylpyridinones for parametrization in the OPLS all-atom force fields. Bond stretching, angle-bending, and torsional parameters were developed to reproduce the MP2 geometries and torsional energy profiles. The transferability of the new parameters was tested through their application to three additional biaryls. The torsional energetics for the 33 biaryl molecules are analyzed and factors leading to preferences for planar and nonplanar geometries are identified. For liquid biphenyl, the computed density and heat of vaporization at the boiling point (255 °C) are also reported.

Related Concept Videos
 03:51
Molecular Orbital Theory II
 01:15
Thin-Walled Hollow Shafts
 01:18
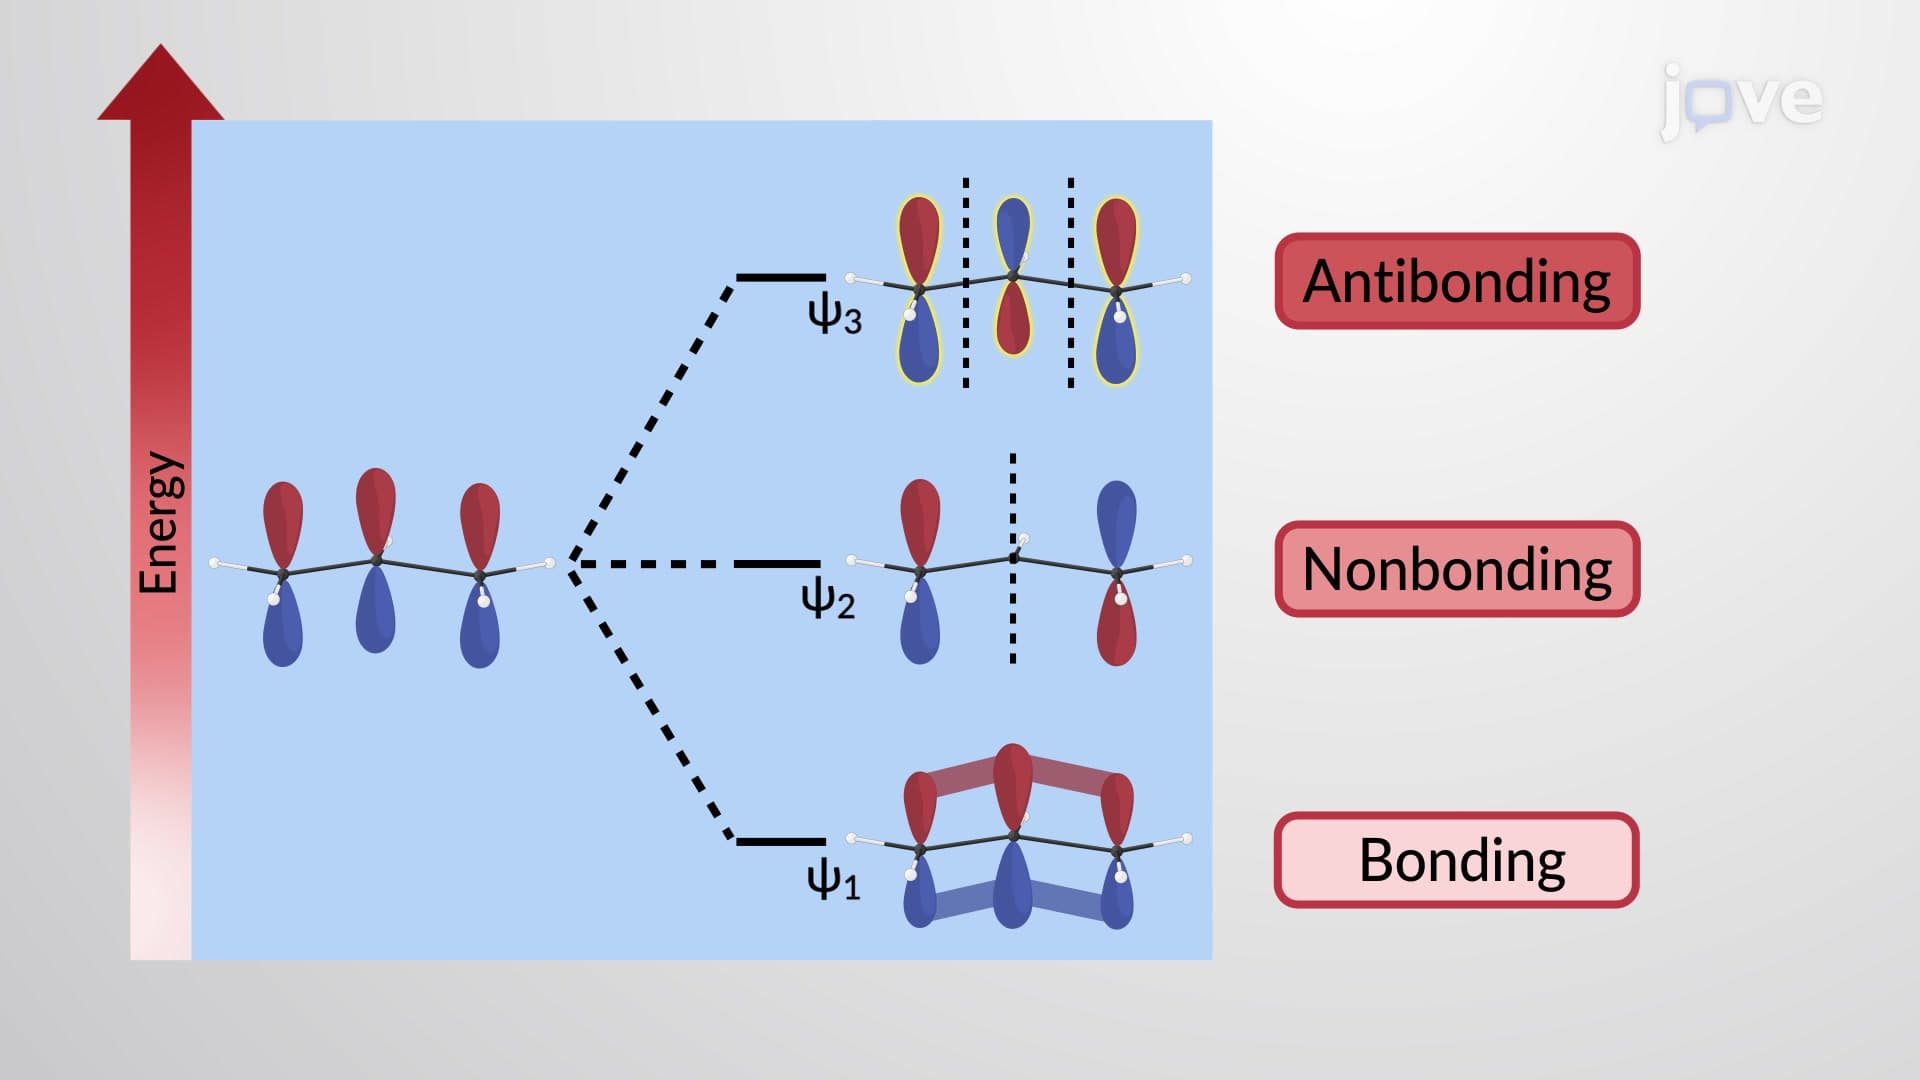
π Molecular Orbitals of the Allyl Cation and Anion
 01:09
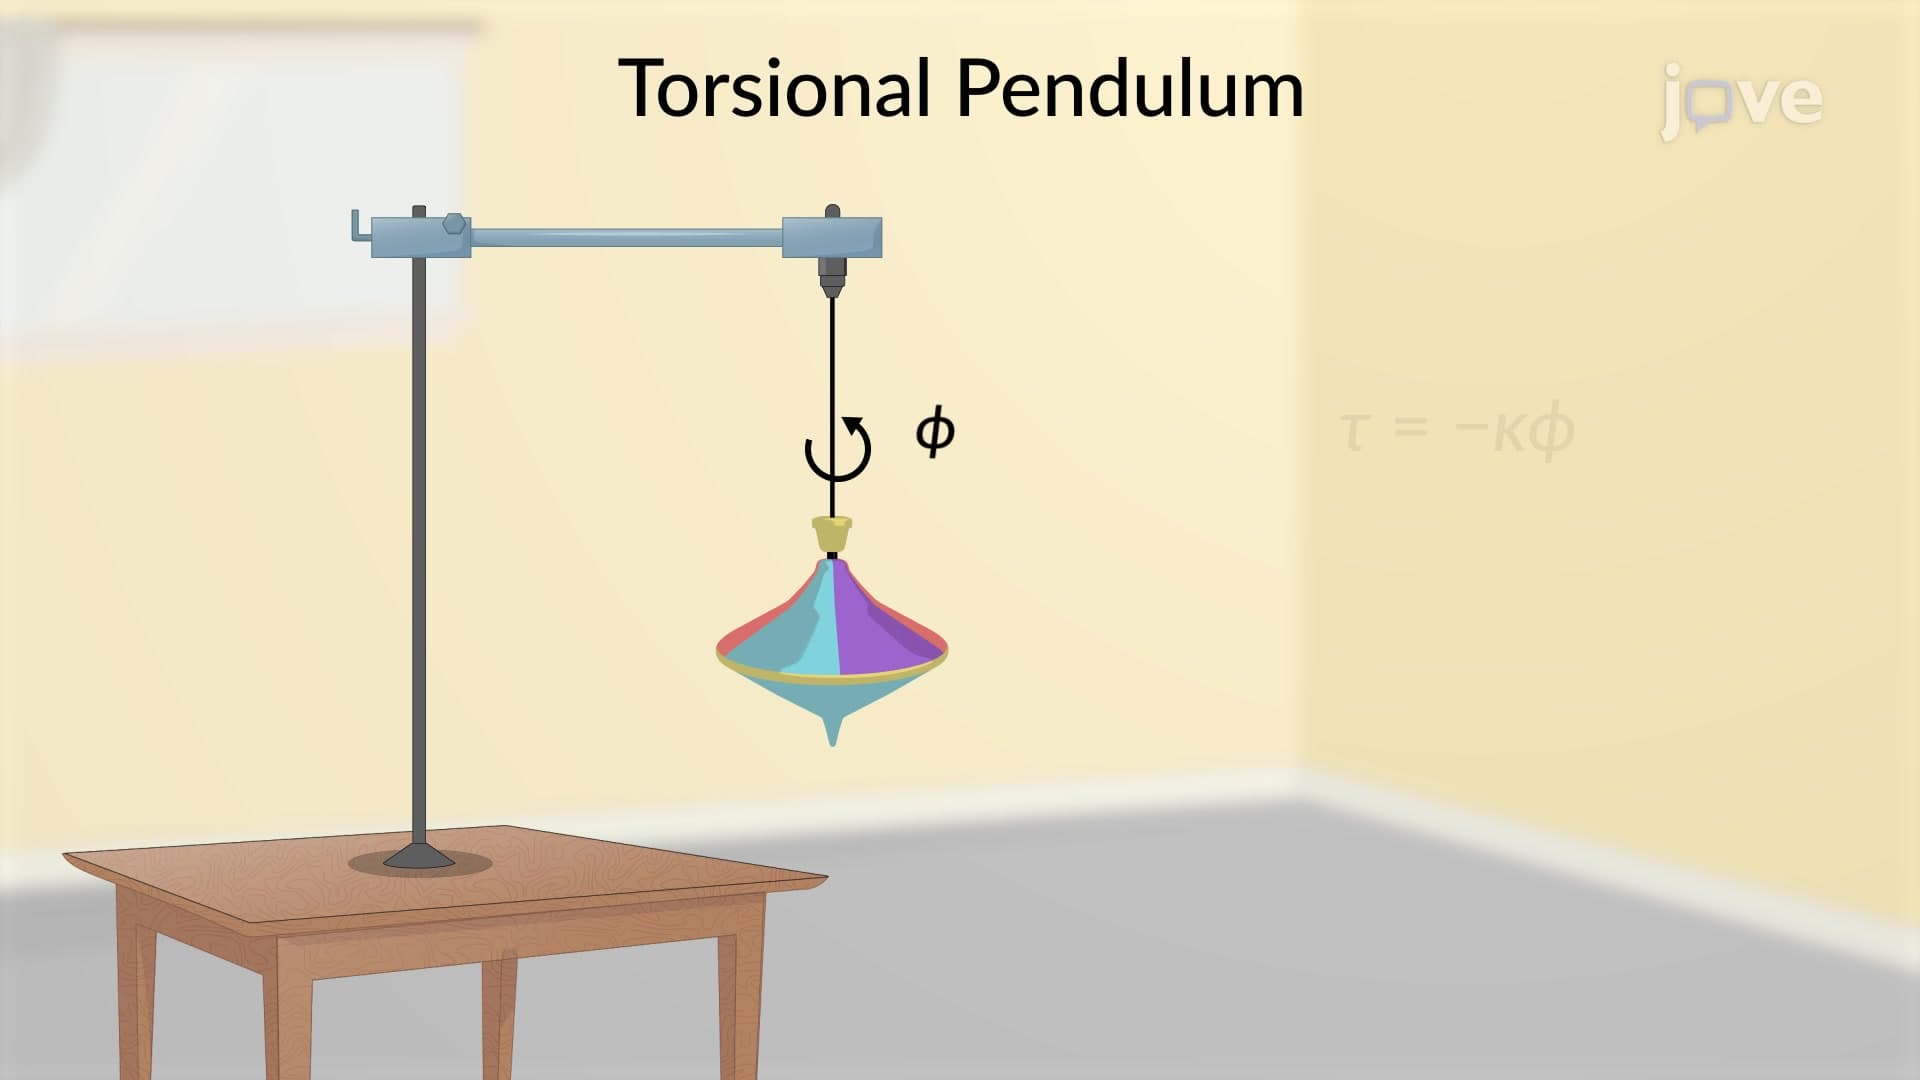
Torsional Pendulum
As long as the rigid body's angular displacement is small, its oscillation can be modeled as a linear angular oscillation. The amplitude of the oscillation is an angle. The role of mass is played by the...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 02:35
Molecular Orbital Theory I