Corrected small basis set Hartree-Fock method for large systems
Related Experiment Video
Updated: May 11, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
1Mulliken Center for Theoretical Chemistry, Institut für Physikalische und Theoretische Chemie der Universität Bonn, Beringstr. 4, D-53115, Bonn, Germany.

Abstract:
A quantum chemical method based on a Hartree-Fock calculation with a small Gaussian AO basis set is presented. Its main area of application is the computation of structures, vibrational frequencies, and noncovalent interaction energies in huge molecular systems. The method is suggested as a partial replacement of semiempirical approaches or density functional theory (DFT) in particular when self-interaction errors are acute. In order to get accurate results three physically plausible atom pair-wise correction terms are applied for London dispersion interactions (D3 scheme), basis set superposition error (gCP scheme), and short-ranged basis set incompleteness effects. In total nine global empirical parameters are used. This so-called Hartee-Fock-3c (HF-3c) method is tested for geometries of small organic molecules, interaction energies and geometries of noncovalently bound complexes, for supramolecular systems, and protein structures. In the majority of realistic test cases good results approaching large basis set DFT quality are obtained at a tiny fraction of computational cost.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 03:35
Hybridization of Atomic Orbitals II
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum numbers: n, l, ml, and...
 03:24
Hybridization of Atomic Orbitals I
 01:26
The Van der Waals Equation
 03:51
Molecular Orbital Theory II
 01:10
Van der Waals Equation
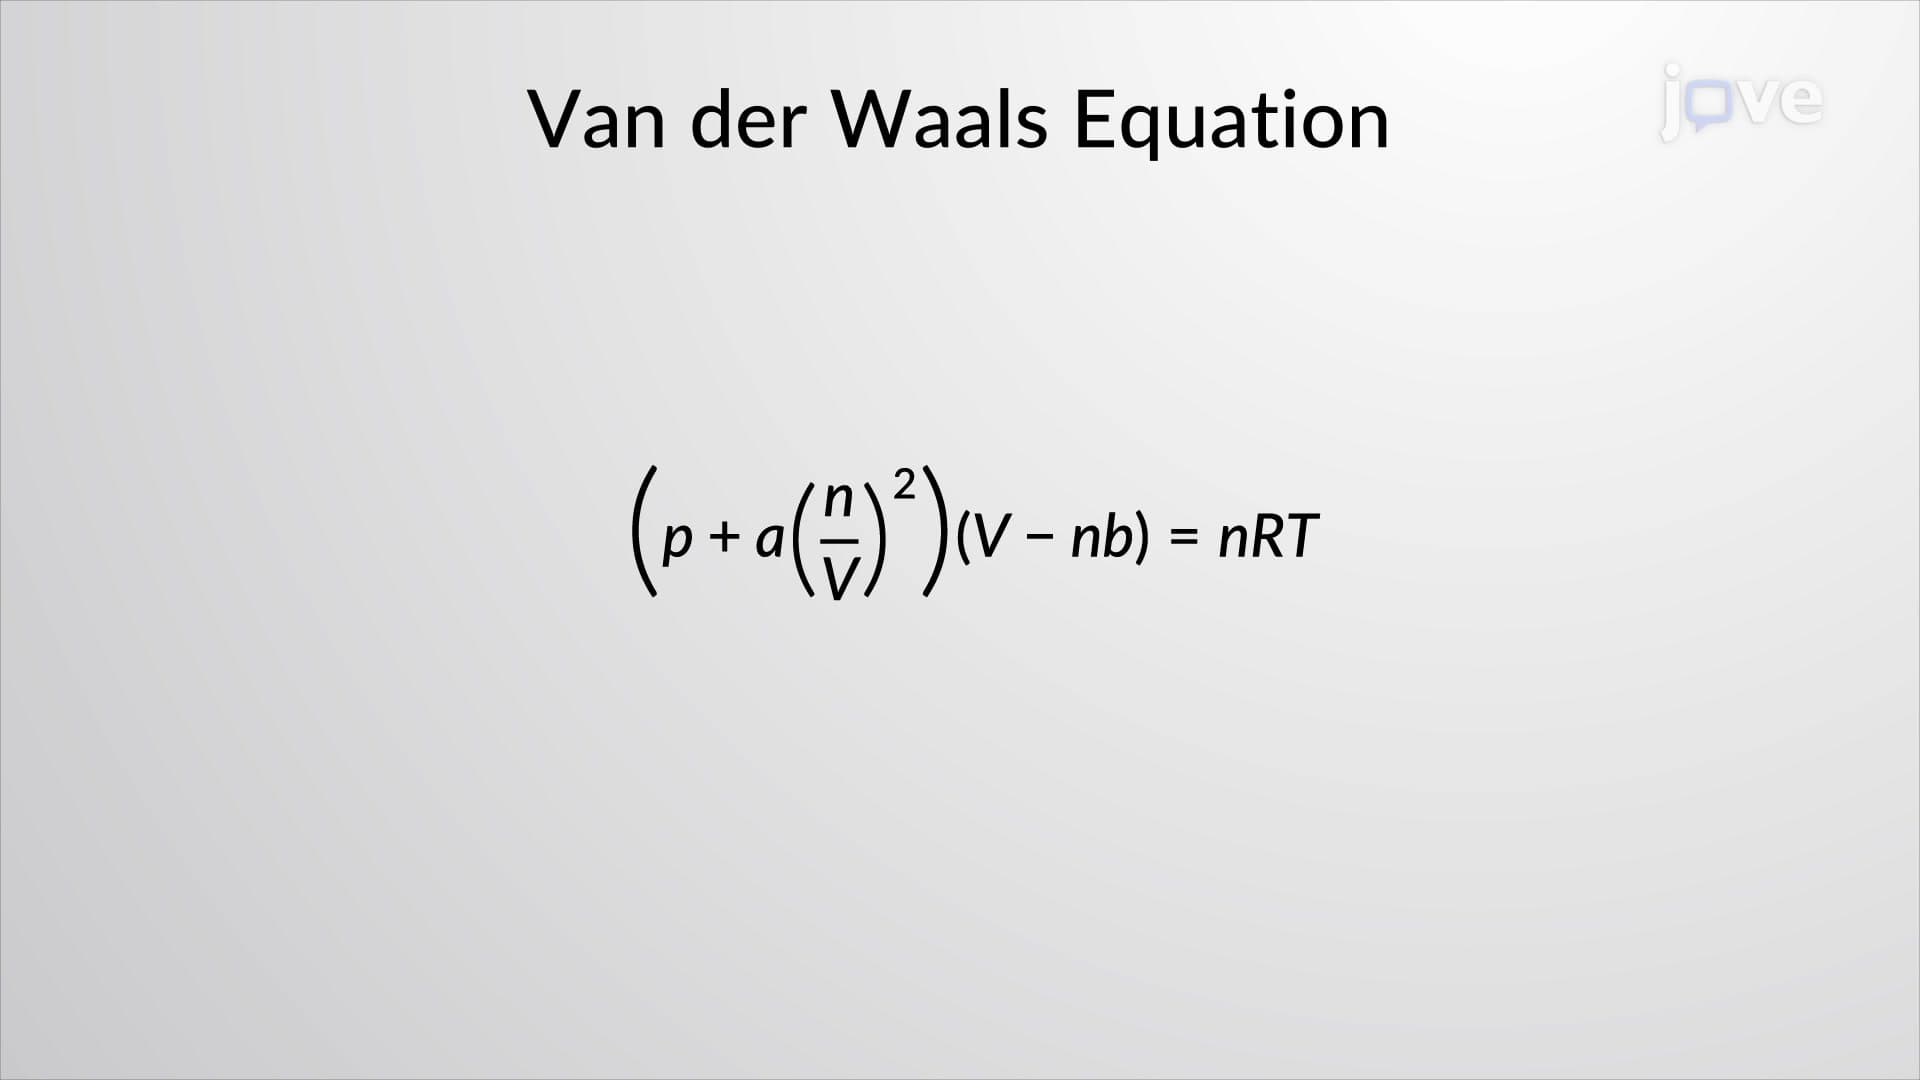
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...