Related Experiment Video
Updated: May 11, 2026
Catalytic Reactions at Amine-Stabilized and Ligand-Free Platinum Nanoparticles Supported on Titania During Hydrogenation of Alkenes and Aldehydes
Published on: June 24, 2022
Energetics of C-N coupling reactions on Pt(111) and Ni(111) surfaces from application of density-functional theory
Wei-Jia Chen1, Chen-Hao Yeh, Chun-Chih Chang
1Department of Chemistry, National Taiwan Normal University, 88, Section 4, Tingchow Road, Taipei 116, Taiwan.

Abstract:
We applied density-functional theory (DFT) with the projector-augmented-wave method (PAW) to investigate systematically the energetics of C-N coupling reactions on Pt(111) and Ni(111)surfaces. Our approach includes several steps: the adsorption of reactants and products (CHx, NHy and CHxNHy, x = 0-3, y = 0-2), movement of molecular fragments on the surface, and then C-N coupling. According to our calculations, the energies (ignoring the conventional negative sign) of adsorption of CHx and NHy on Pt(111)/Ni(111) surfaces decrease in the order C > CH > CH2 > CH3 and N > NH > NH2, with values 7.41/6.91, 6.97/6.52, 4.58/4.39, 2.19/2.01 eV and 5.10/5.49, 4.12/4.79, 2.75/2.87 eV, respectively. Regarding the adsorption energies among CHxNHy, the adsorption energy of CNH2 species is the highest on the Pt(111) surface, whereas on the Ni(111) surface CH3N is the most stable. The C-N coupling barriers differ on the two metallic surfaces despite the structures of initial, transition and final states being similar. On the Pt(111) surface, the coupling reaction of CH2 + NH2 has the smallest barrier, whereas CH + NH2 is the most favorable on the Ni(111) surface. The detailed local density of states (LDOS) and electron-localization functions (ELF) were investigated to rationalize the calculated outcomes.
Related Concept Videos
 02:50
Catalysis
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene π orbitals.
 01:17
Spin–Spin Coupling: One-Bond Coupling
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:22
Heterogeneous Catalysis
 01:22
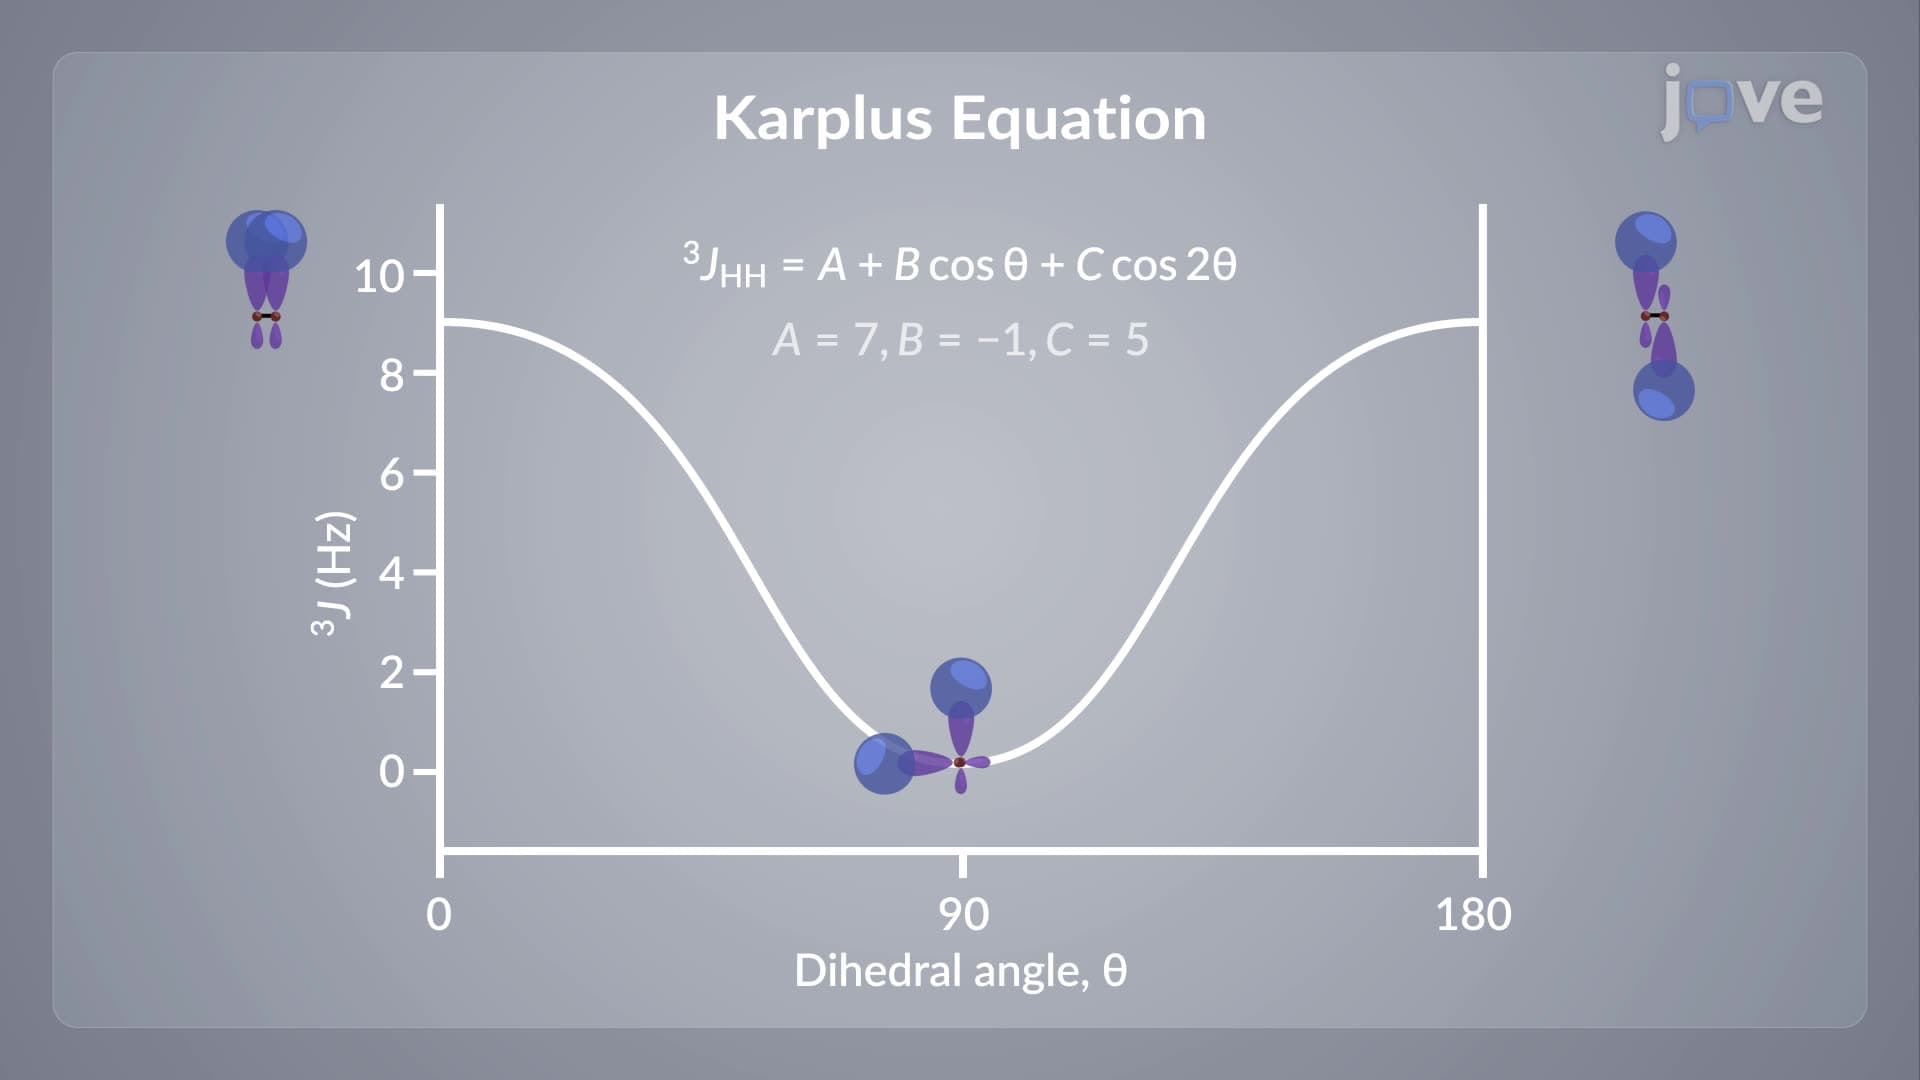
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...