

Ab initio and DFT studies of the structure and vibrational spectra of anhydrous caffeine
Santosh K Srivastava1, Vipin B Singh
1Department of Physics, Udai Pratap Autonomous College, Varanasi 221 002, India.

Abstract:
Vibrational spectra and molecular structure of anhydrous caffeine have been systematically investigated by second order Moller-Plesset (MP2) perturbation theory and density functional theory (DFT) calculations. Vibrational assignments have been made and many previous ambiguous assignments in IR and Raman spectra are amended. The calculated DFT frequencies and intensities at B3LYP/6-311++G(2d,2p) level, were found to be in better agreement with the experimental values. It was found that DFT with B3LYP functional predicts harmonic vibrational wave numbers more close to experimentally observed value when it was performed on MP2 optimized geometry rather than DFT geometry. The calculated TD-DFT vertical excitation electronic energies of the valence excited states of anhydrous caffeine are found to be in consonance to the experimental absorption peaks.
More Related Videos
 12:08
Catalytic Reactions at Amine-Stabilized and Ligand-Free Platinum Nanoparticles Supported on Titania During Hydrogenation of Alkenes and Aldehydes
Published on: June 24, 2022
 09:11
Caffeine Extraction, Enzymatic Activity and Gene Expression of Caffeine Synthase from Plant Cell Suspensions
Published on: October 2, 2018
Related Concept Videos
 01:29
IR and UV–Vis Spectroscopy of Aldehydes and Ketones
 01:26
Spectroscopy of Carboxylic Acid Derivatives
In the...
 01:18
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous overlap of p...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 01:29
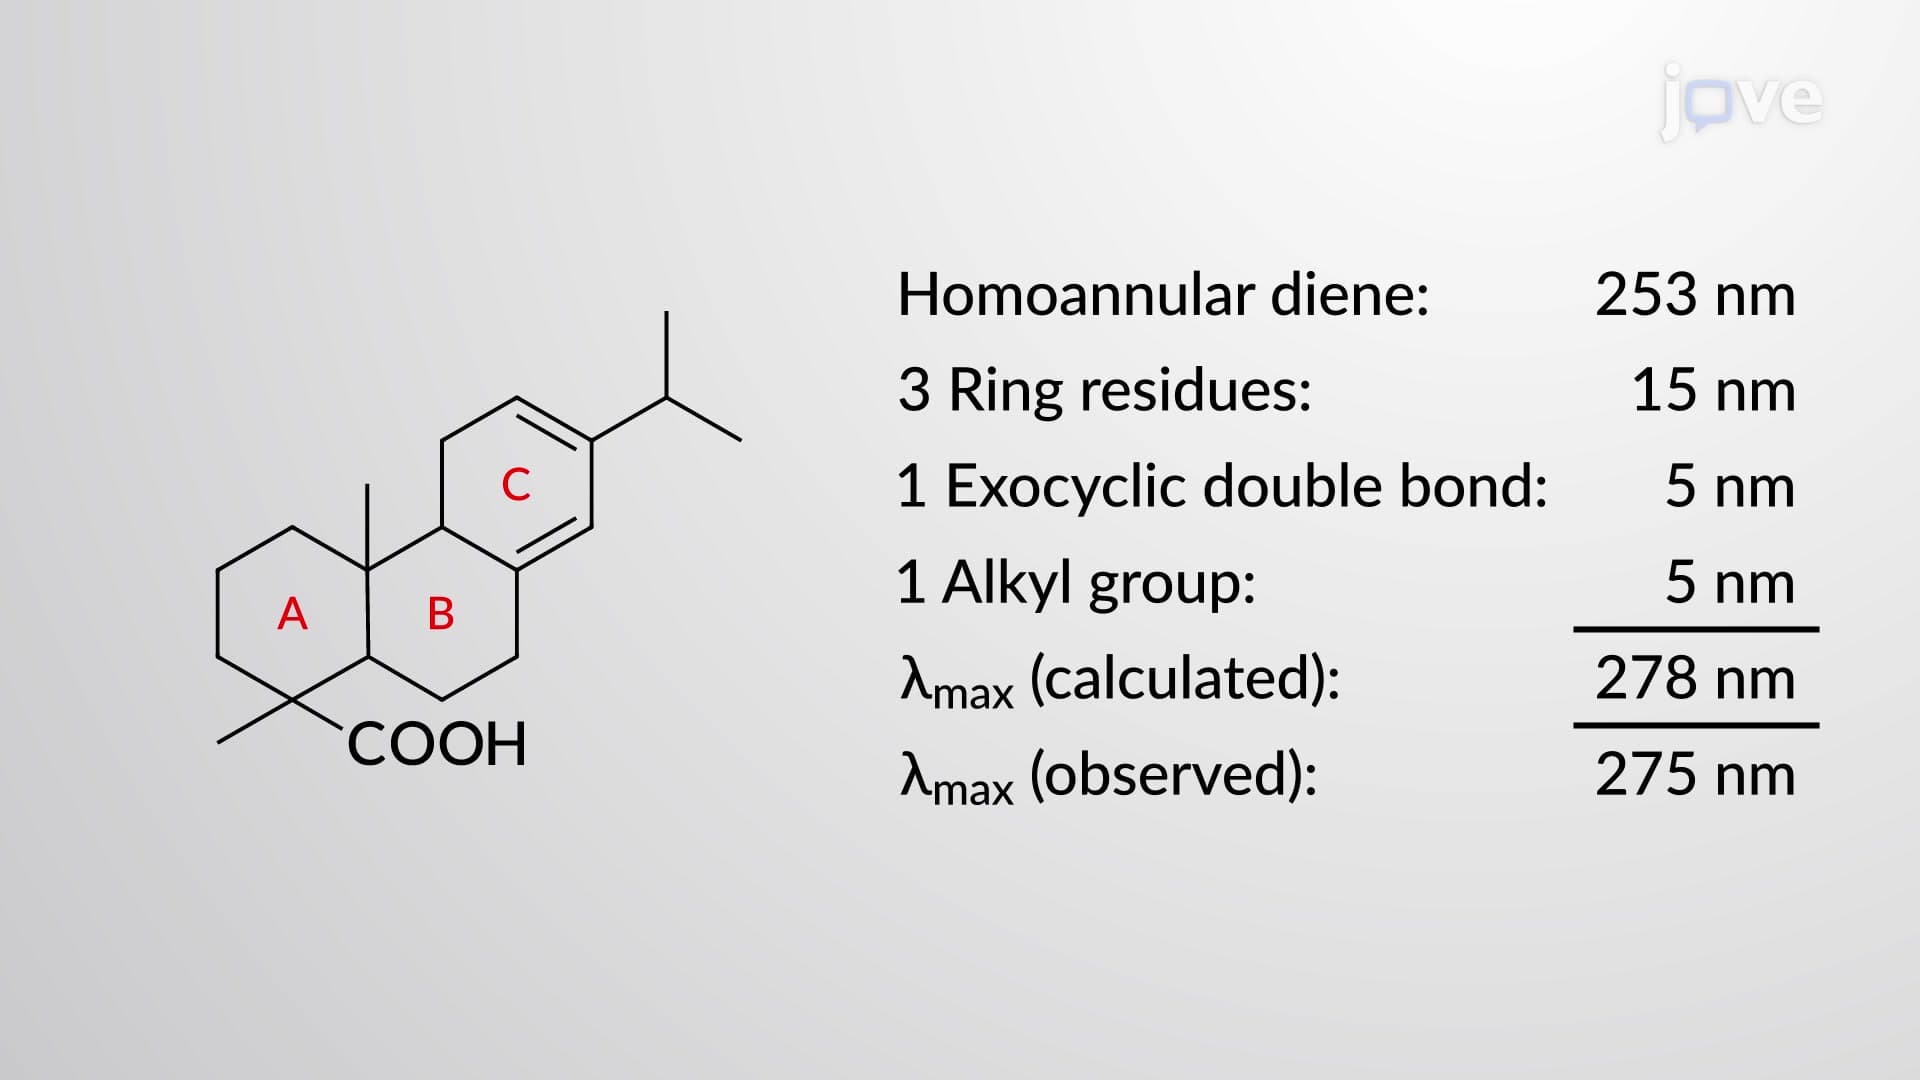
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:15
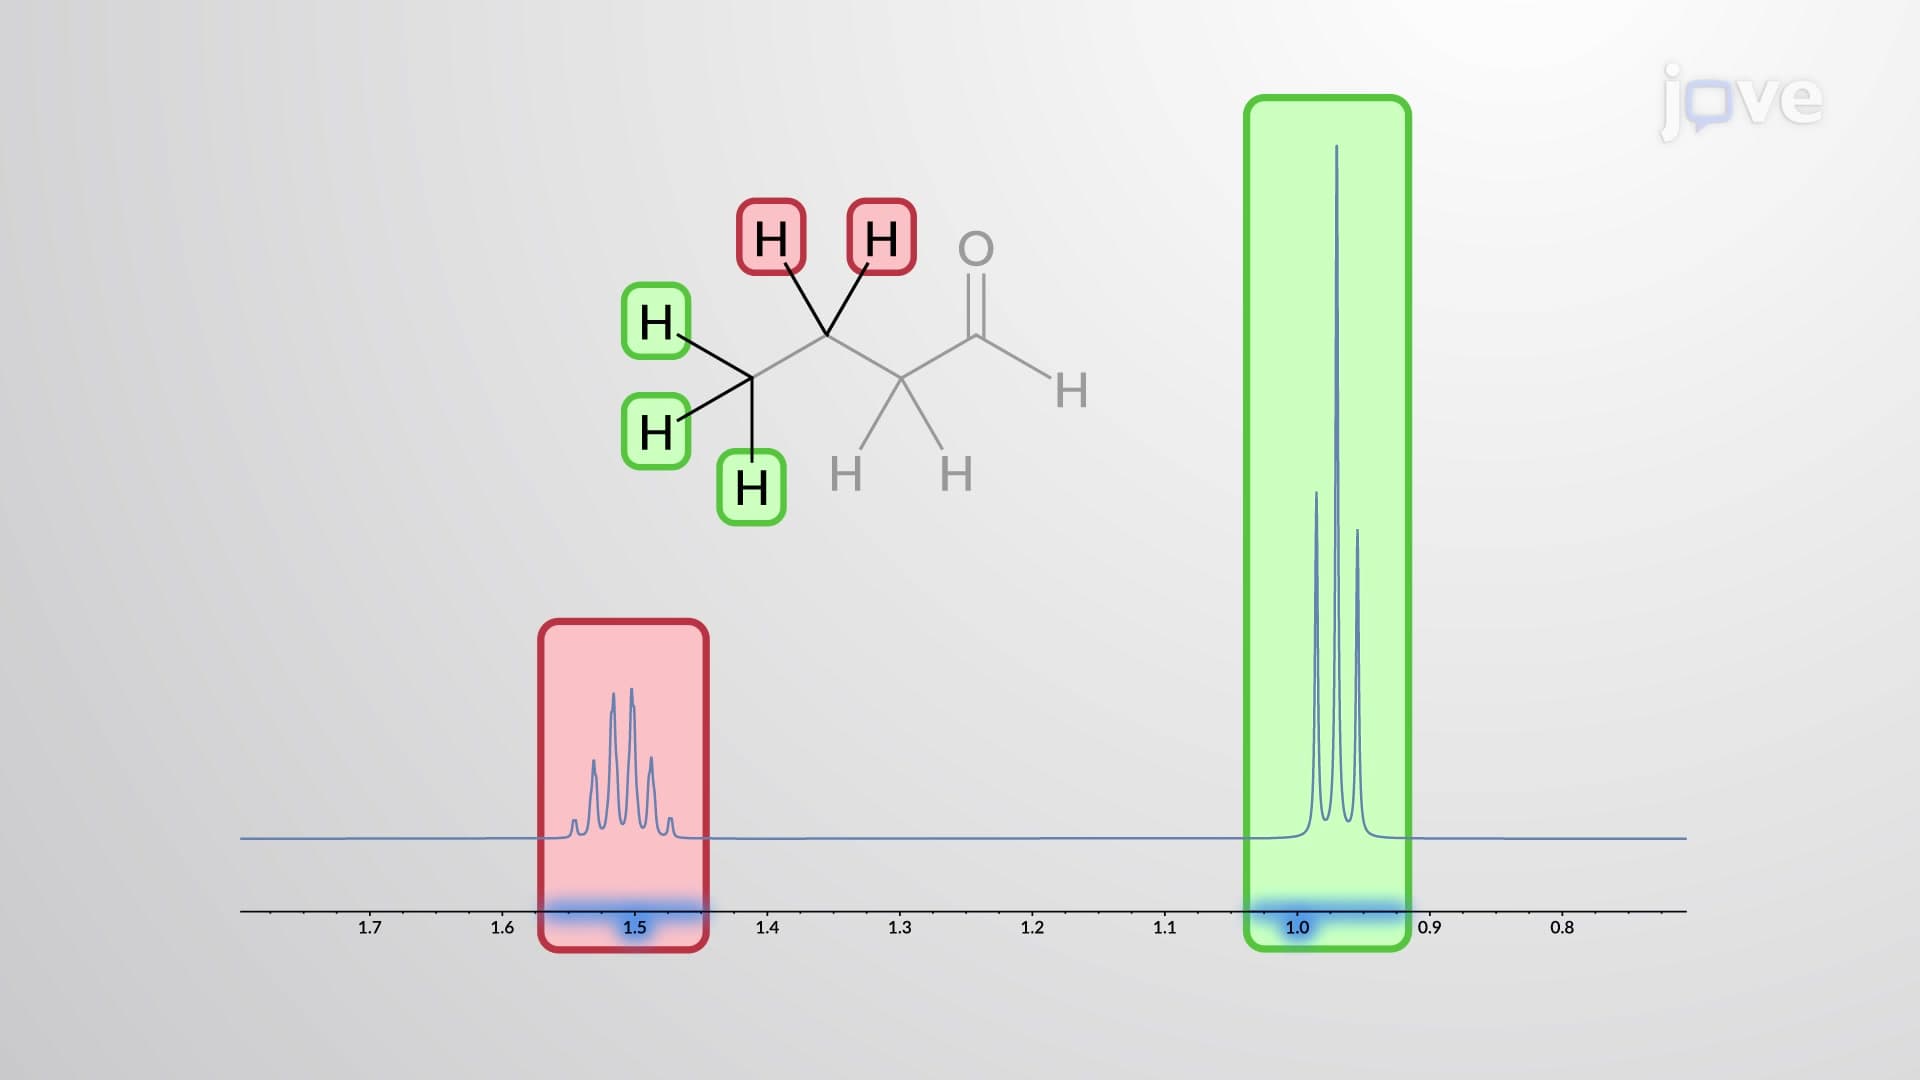
NMR Spectroscopy and Mass Spectrometry of Aldehydes and Ketones