Related Experiment Video
Updated: May 9, 2026
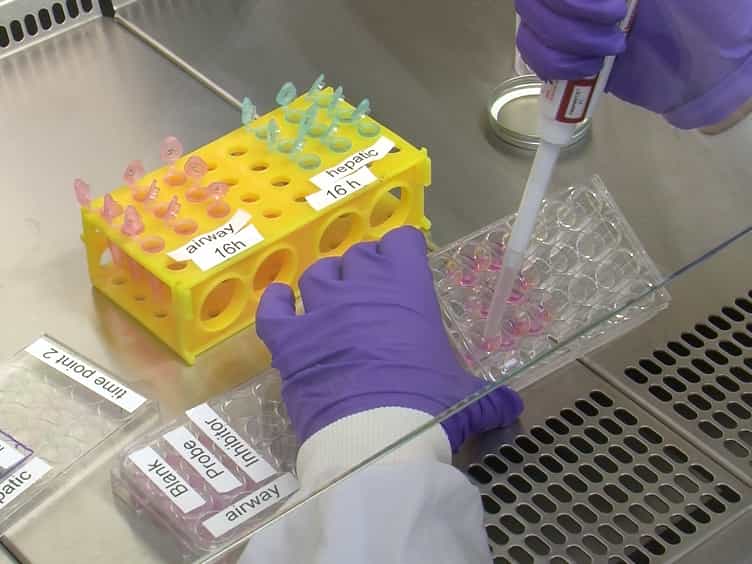
Mass Spectrometry and Luminogenic-based Approaches to Characterize Phase I Metabolic Competency of In Vitro Cell Cultures
Published on: March 28, 2017
CYP2B6 pharmacogenetics-based in vitro-in vivo extrapolation of efavirenz clearance by physiologically based
Cong Xu1, Sara K Quinney, Yingying Guo
1Division of Clinical Pharmacology (C.X., Z.D.), Department of Obstetrics and Gynecology (S.K.Q.) and Center for Computational Biology and Bioinformatics (L.L.), Indiana University School of Medicine, Indianapolis, Indiana; and Department of Drug Disposition, Lilly Research Laboratories, Eli Lilly and Co., Indianapolis, Indiana (Y.G., S.D.H.).

Abstract:
Efavirenz is mainly cleared by CYP2B6. The CYP2B6*6 allele is associated with lower efavirenz clearance. Efavirenz clearance was predictable using in vitro data for carriers of the CYP2B6*1/*1 genotype, but the prediction in carriers of the CYP2B6*6 allele was poor. To test the hypothesis that incorporation of mechanism of reduced efavirenz metabolism by the CYP2B6*6 allele can predict the genetic effect on efavirenz pharmacokinetics, in vitro-in vivo extrapolation of efavirenz clearance was performed by physiologically based pharmacokinetic modeling (Simcyp Simulator; Simcyp Ltd., Sheffield, UK) using data obtained from expressed CYP2B6.1 and CYP2B6.6 as well as human liver microsomes (HLMs) with CYP2B6*1/*1, *1/*6, and *6/*6 genotypes. Simulated pharmacokinetics of a single 600-mg oral dose of efavirenz for individuals with each genotype was compared with data observed in healthy subjects genotyped for the CYP2B6*6 allele (n = 20). Efavirenz clearance for carriers of the CYP2B6*1/*1 genotype was predicted reasonably well using HLM data, but the clearance in carriers of the CYP2B6*6 allele was underpredicted using both expressed and HLM systems. Improved prediction of efavirenz clearance was obtained from expressed CYP2B6 after recalculating intersystem extrapolation factors for CYP2B6.1 and CYP2B6.6 based on in vitro intrinsic clearance of bupropion 4-hydroxylation. These findings suggest that genetic effect on both CYP2B6 protein expression and catalytic efficiency needs to be taken into account for the prediction of pharmacokinetics in individuals carrying the CYP2B6*6/*6 genotype. Expressed CYP2B6 proteins may be a reliable in vitro system to predict effect of the CYP2B6*6 allele on the metabolism of CYP2B6 substrates.

Related Concept Videos
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion, mediated...
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:15
Impact of Pharmacokinetic–Pharmacodynamic Models: Regulatory Decisions
 01:15
Pharmacodynamic Models: Linear Concentration–Effect Model
 01:27
Pharmacogenetics of Drug Metabolism: Overview
 01:18
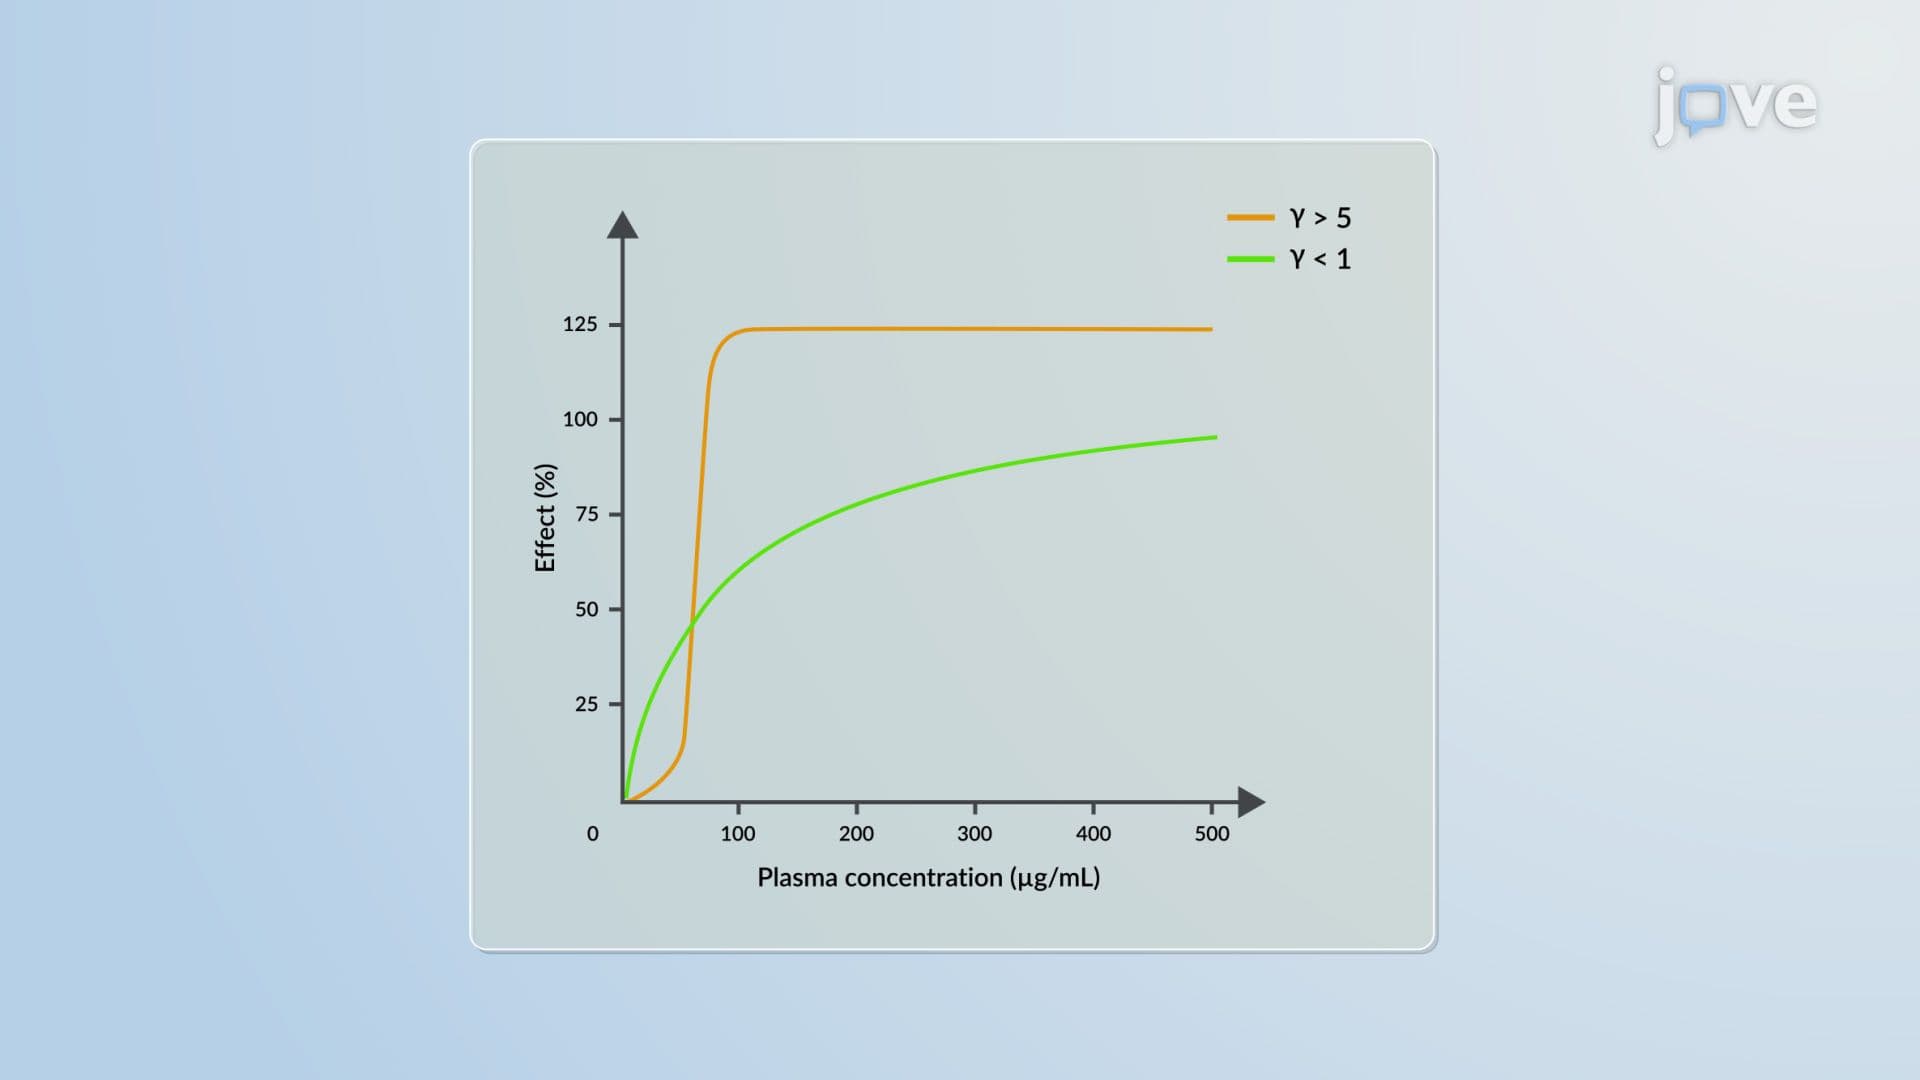
Pharmacodynamic Models: Emax Drug–Concentration Effect Model