Related Experiment Video
Updated: May 7, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Response to "Comment on 'Going beyond the frozen core approximation: development of coordinate-dependent
Argyris Kahros1, Benjamin J Schwartz
1Department of Chemistry and Biochemistry, UCLA, Los Angeles, California 90095-1569, USA.

Abstract:
Stoll, Fuentealba, and Szentpály (SFS) argue that the coordinate-dependent pseudopotential we developed for the sodium dimer cation molecule is inferior to other potentials that have been presented in the literature for this molecule. The goal of our work, however, was to present a novel method for the development of rigorous coordinate-dependent pseudopotentials. Our method is designed to reproduce all-electron Hartree-Fock calculations without the inclusion of adjustable parameters. Moreover, our method starts from the superposition of unoptimized, non-norm-conserved atomic potentials, so that when complete, the resulting norm-conserving potential can reproduce an all-electron Hartree-Fock calculation without the inclusion of adjustable parameters. We chose the sodium dimer cation system as a proof of principle for our method, and showed that our method does indeed allow a one-electron calculation to correctly reproduce the all-electron Hartree-Fock calculation from bonding to the dissociation limit. Our purpose in developing this method is to use such potentials in condensed-phase mixed quantum/classical molecular dynamics simulations, where inclusion of valence polarization effects is unimportant or can be added on after the fact. Thus we do not claim that our method provides a potential that is superior to potentials that have been specifically constructed to go beyond the static exchange approximation and/or include valence polarization effects-such potentials are beyond the scope of our work. We also note that although we made a numerical error in the application of our method to Na2(+) in our original work [A. Kahros and B. J. Schwartz, J. Chem. Phys. 138, 054110 (2013)] that led to an overestimation of the magnitude of core polarization effects for this particular molecule, out method does work as derived for this molecule and the error does not affect the significance of our method or its general applicability.
More Related Videos


Related Concept Videos
 01:26
Thermodynamic Potentials
 01:10
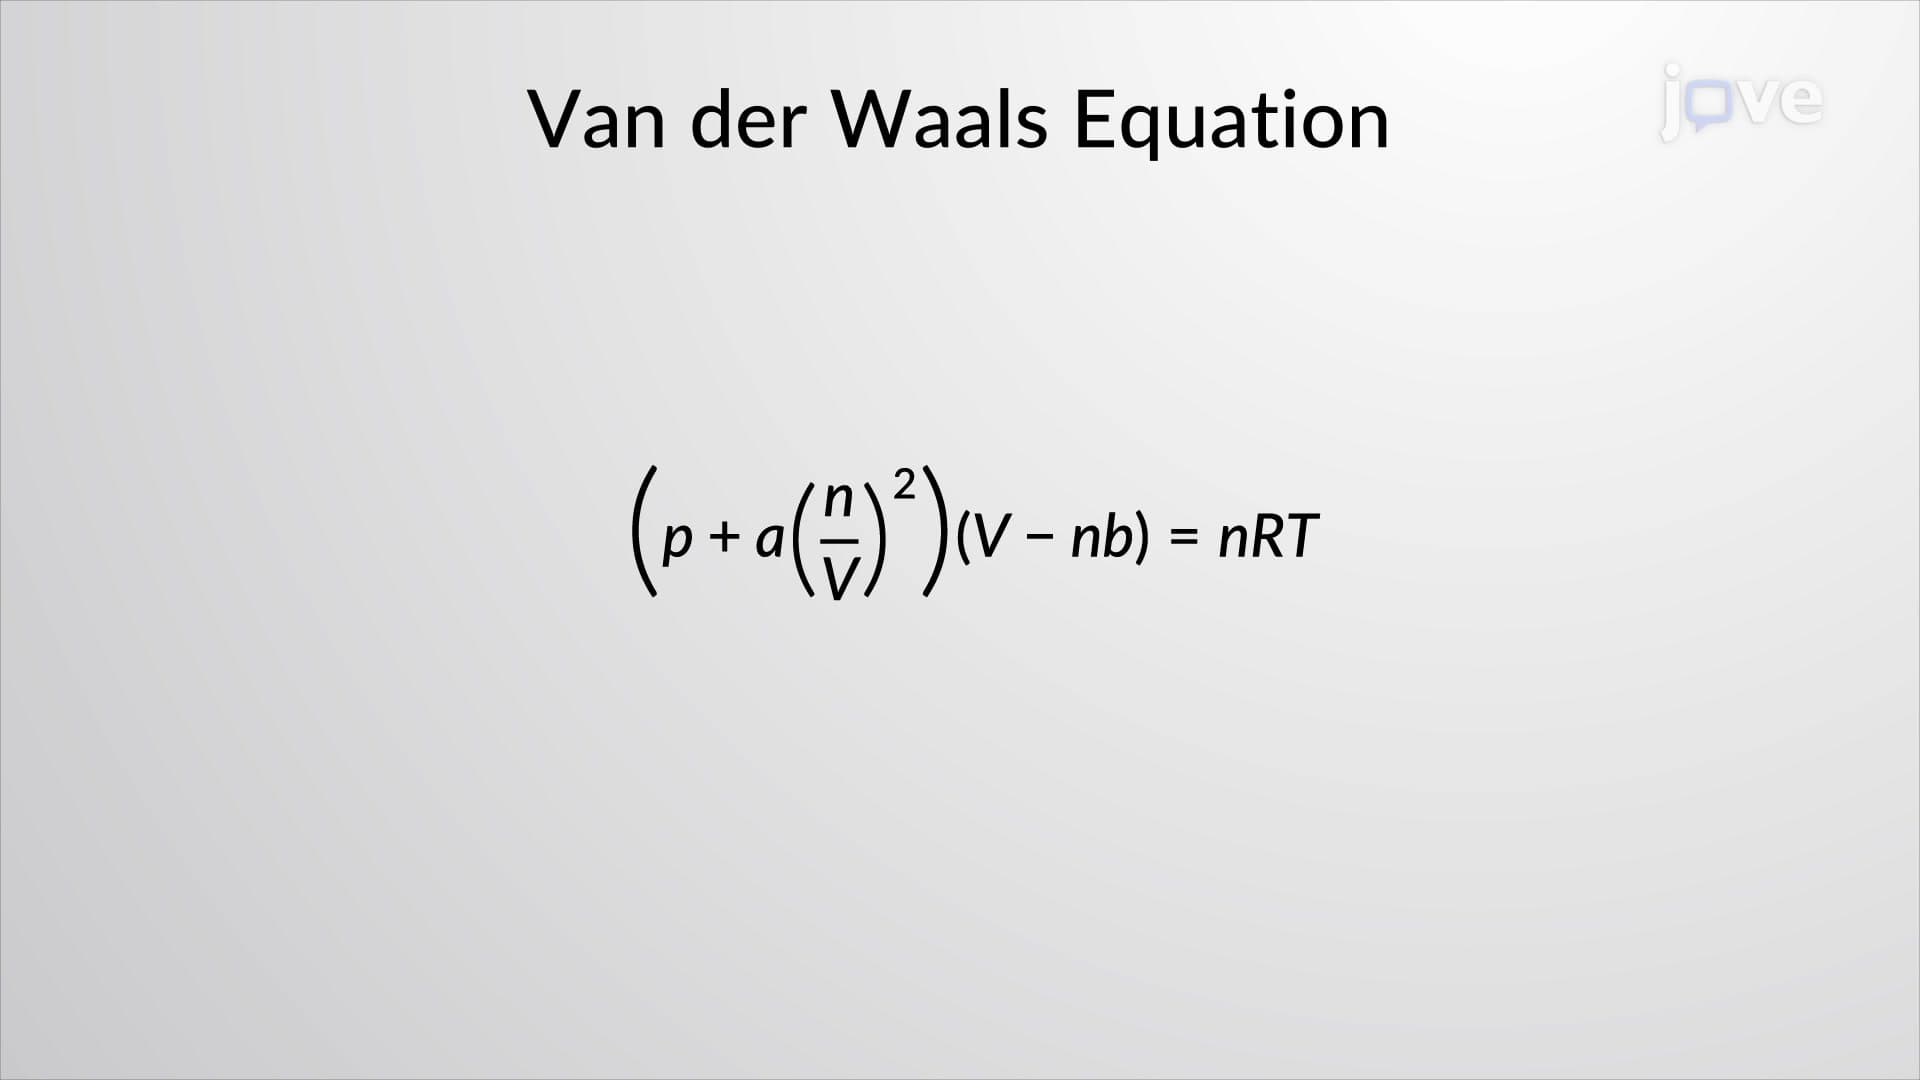
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the...
 01:29
Potential Due to a Polarized Object
 01:26
The Van der Waals Equation
 03:35
Hybridization of Atomic Orbitals II
 03:24
Hybridization of Atomic Orbitals I