Related Experiment Video
Updated: May 6, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Accurate ab initio-based adiabatic global potential energy surface for the 2(2)A" state of NH2 by extrapolation to

Abstract:
A full three-dimensional global potential energy surface is reported first time for the title system, which is important for the photodissociation processes. It is obtained using double many-body expansion theory and an extensive set of accurate ab initio energies extrapolated to the complete basis set limit. Such a work can be recommended for dynamics studies of the N((2)D) + H2 reaction, a reliable theoretical treatment of the photodissociation dynamics and as building blocks for constructing the double many-body expansion potential energy surface of larger nitrogen/hydrogen containing systems. In turn, a preliminary theoretical study of the reaction N((2)D)+H2(X(1)Σg (+))(ν=0,j=0)→NH(a(1)Δ)+H((2)S) has been carried out with the method of quasi-classical trajectory on the new potential energy surface. Integral cross sections and thermal rate constants have been calculated, providing perhaps the most reliable estimate of the integral cross sections and the rate constants known thus far for such a reaction.
More Related Videos

Related Concept Videos
 01:25
Adsorption Isotherms II
 01:18
Adiabatic Processes for an Ideal Gas
 02:34
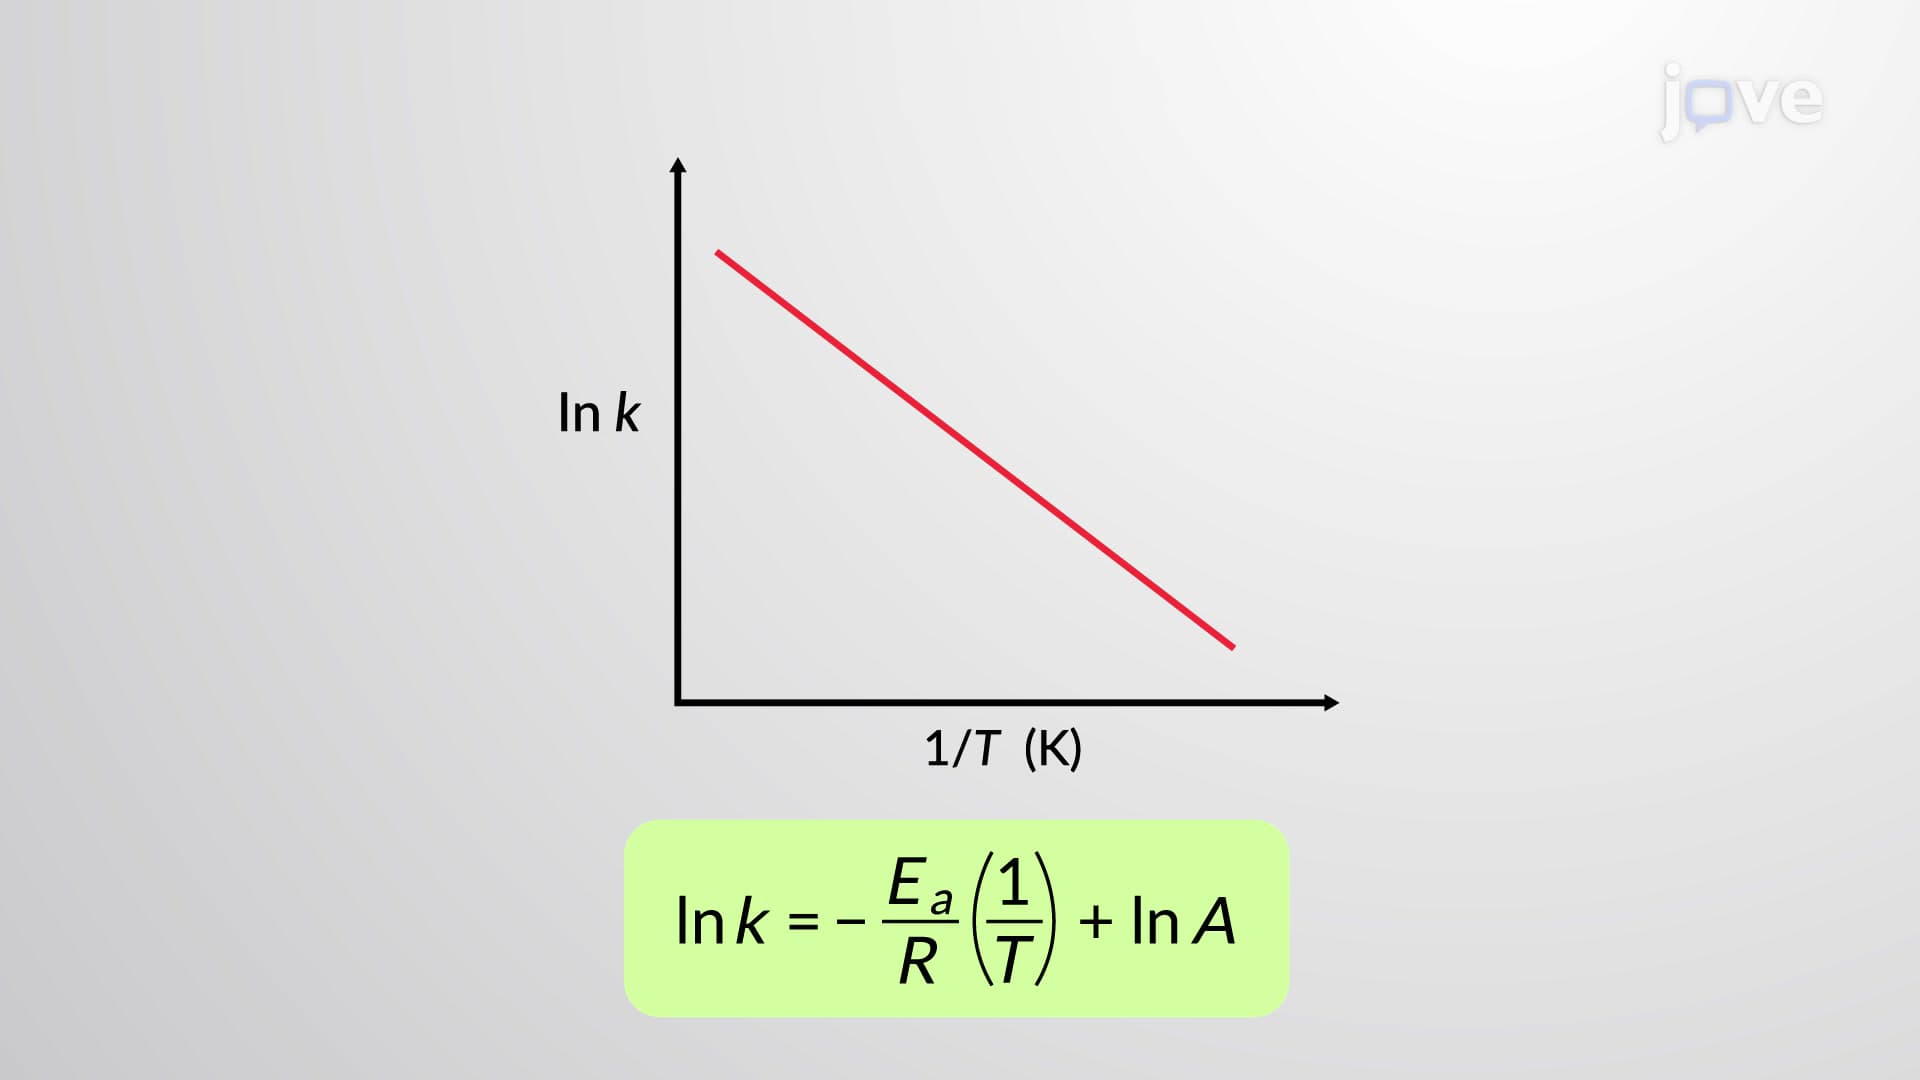
Arrhenius Plots
The Arrhenius equation can...
 01:26
Thermodynamic Potentials
 01:13
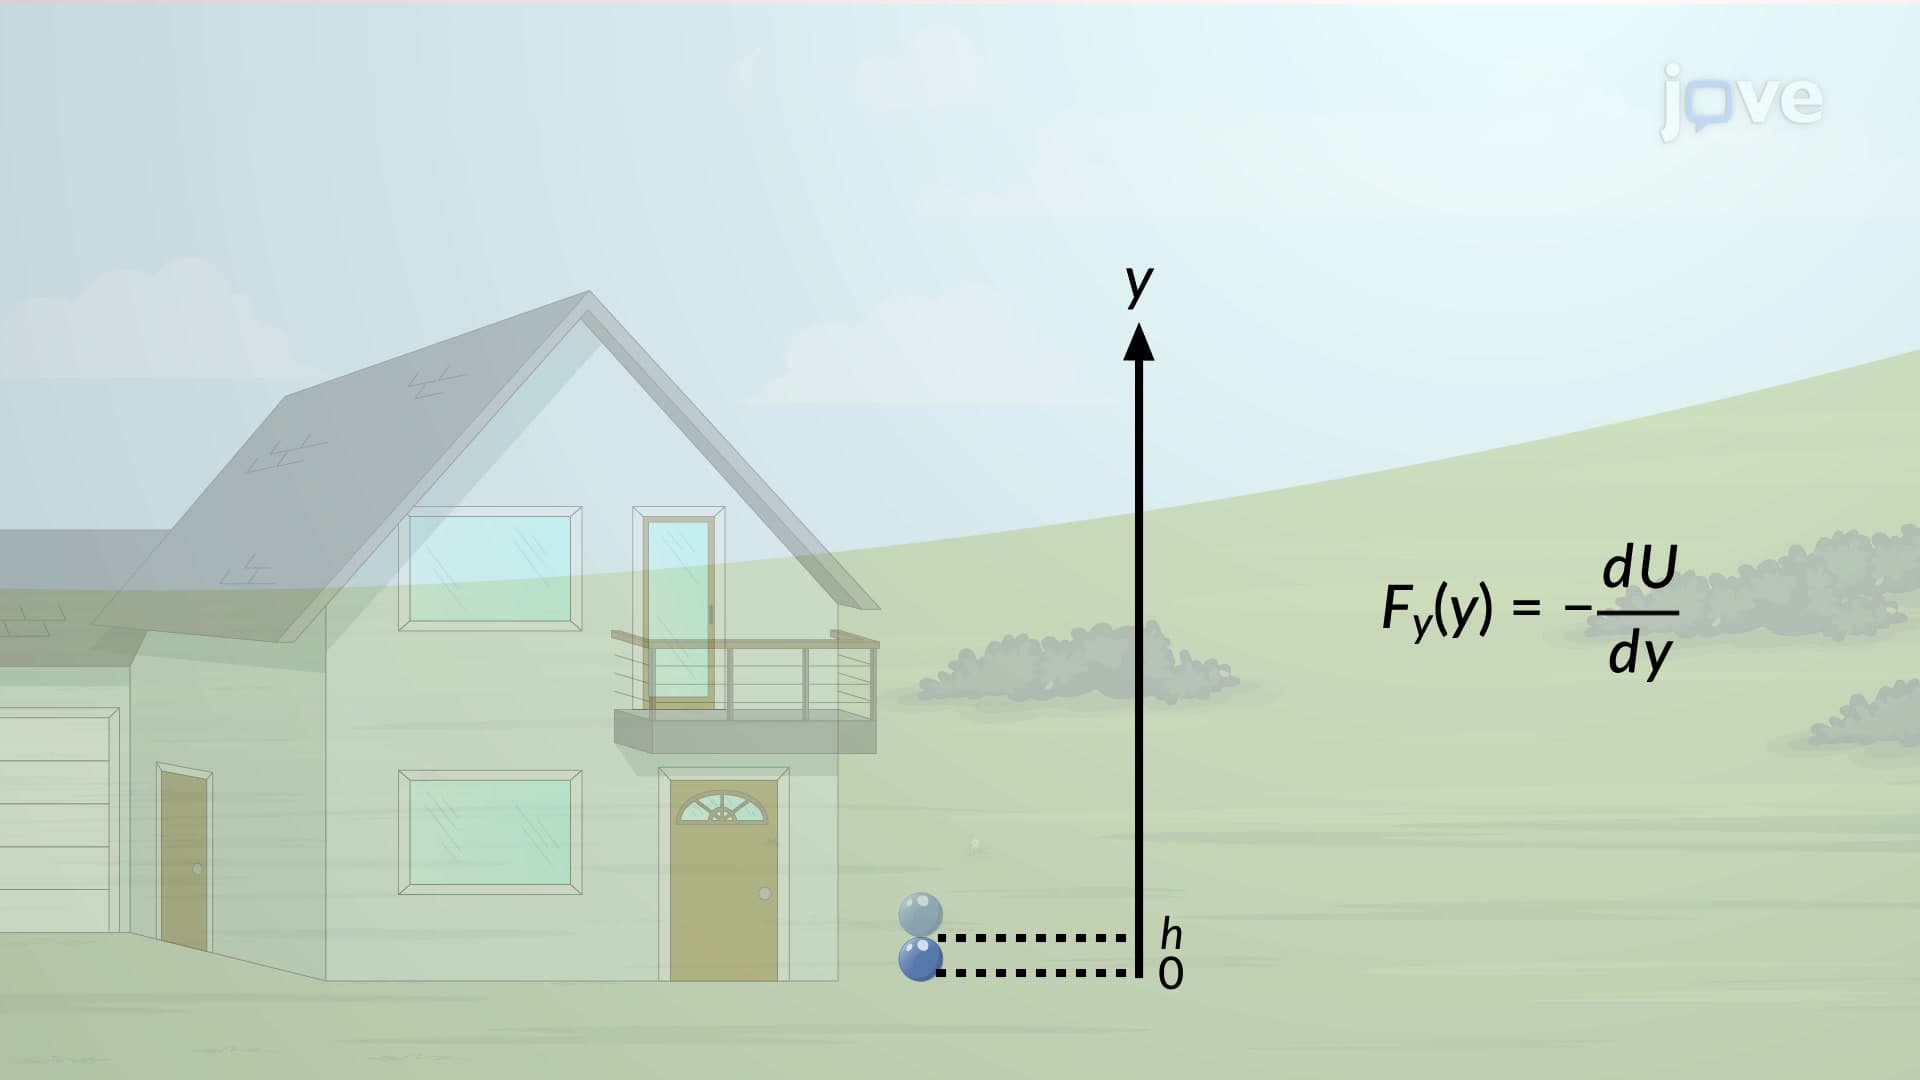
Force and Potential Energy in One Dimension
 03:25
Free Energy Changes for Nonstandard States