Related Experiment Video
Updated: May 6, 2026

Synthesis of Cyclic Polymers and Characterization of Their Diffusive Motion in the Melt State at the Single Molecule Level
Published on: September 26, 2016
Applicability of mode-coupling theory to polyisobutylene: a molecular dynamics simulation study
1Centro de Física de Materiales (CSIC-UPV/EHU) - Materials Physics Center (MPC), Paseo Manuel de Lardizabal 5, 20018 San Sebastián, Spain.

Abstract:
The applicability of Mode Coupling Theory (MCT) to the glass-forming polymer polyisobutylene (PIB) has been explored by using fully atomistic molecular dynamics simulations. MCT predictions for the so-called asymptotic regime have been successfully tested on the dynamic structure factor and the self-correlation function of PIB main-chain carbons calculated from the simulated cell. The factorization theorem and the time-temperature superposition principle are satisfied. A consistent fitting procedure of the simulation data to the MCT asymptotic power-laws predicted for the α-relaxation regime has delivered the dynamic exponents of the theory-in particular, the exponent parameter λ-the critical non-ergodicity parameters, and the critical temperature T(c). The obtained values of λ and T(c) agree, within the uncertainties involved in both studies, with those deduced from depolarized light scattering experiments [A. Kisliuk et al., J. Polym. Sci. Part B: Polym. Phys. 38, 2785 (2000)]. Both, λ and T(c)/T(g) values found for PIB are unusually large with respect to those commonly obtained in low molecular weight systems. Moreover, the high T(c)/T(g) value is compatible with a certain correlation of this parameter with the fragility in Angell's classification. Conversely, the value of λ is close to that reported for real polymers, simulated "realistic" polymers and simple polymer models with intramolecular barriers. In the framework of the MCT, such finding should be the signature of two different mechanisms for the glass-transition in real polymers: intermolecular packing and intramolecular barriers combined with chain connectivity.
Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the...
 02:40
MO Theory and Covalent Bonding
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene...
 01:28
Stability of Conjugated Dienes
A comparison of the enthalpies of hydrogenation of dienes reveals that conjugated dienes release less heat on hydrogenation, rendering them more stable than their nonconjugated analogs.
 01:22
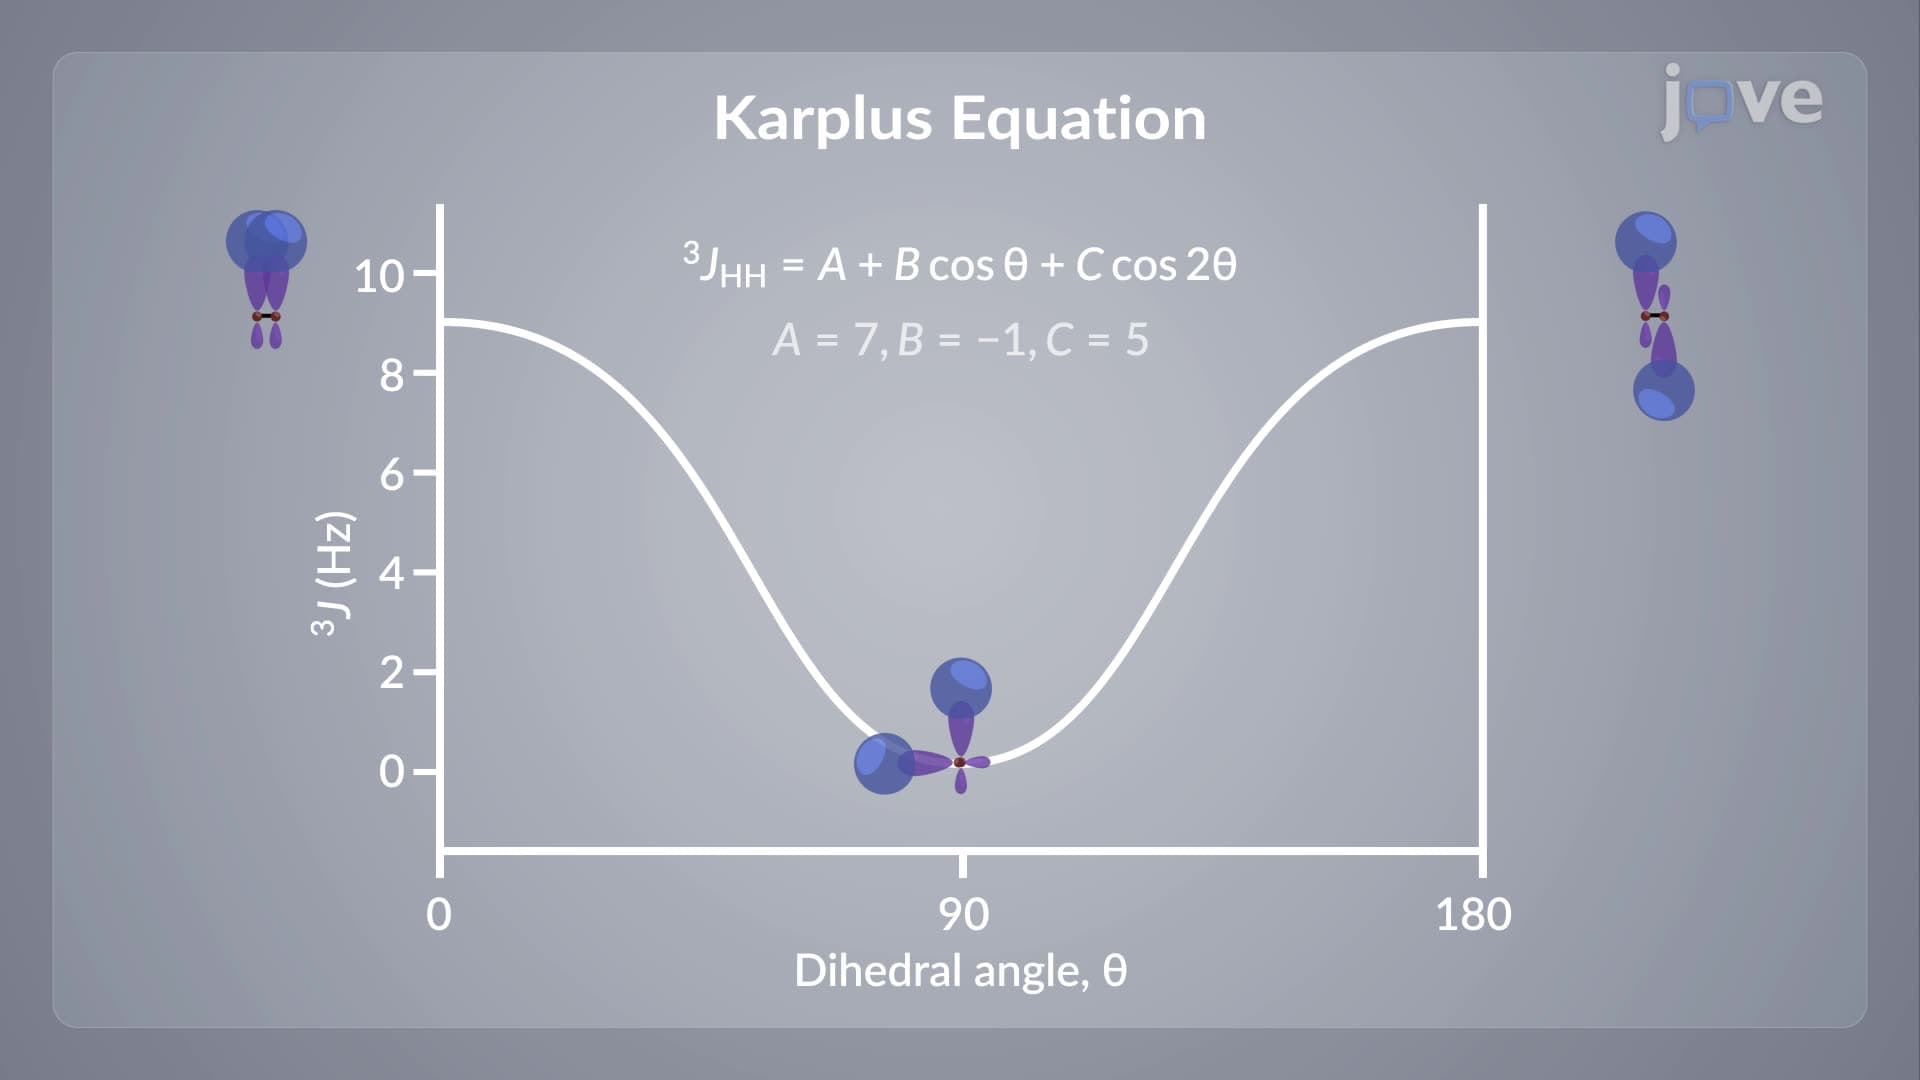
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
 01:08
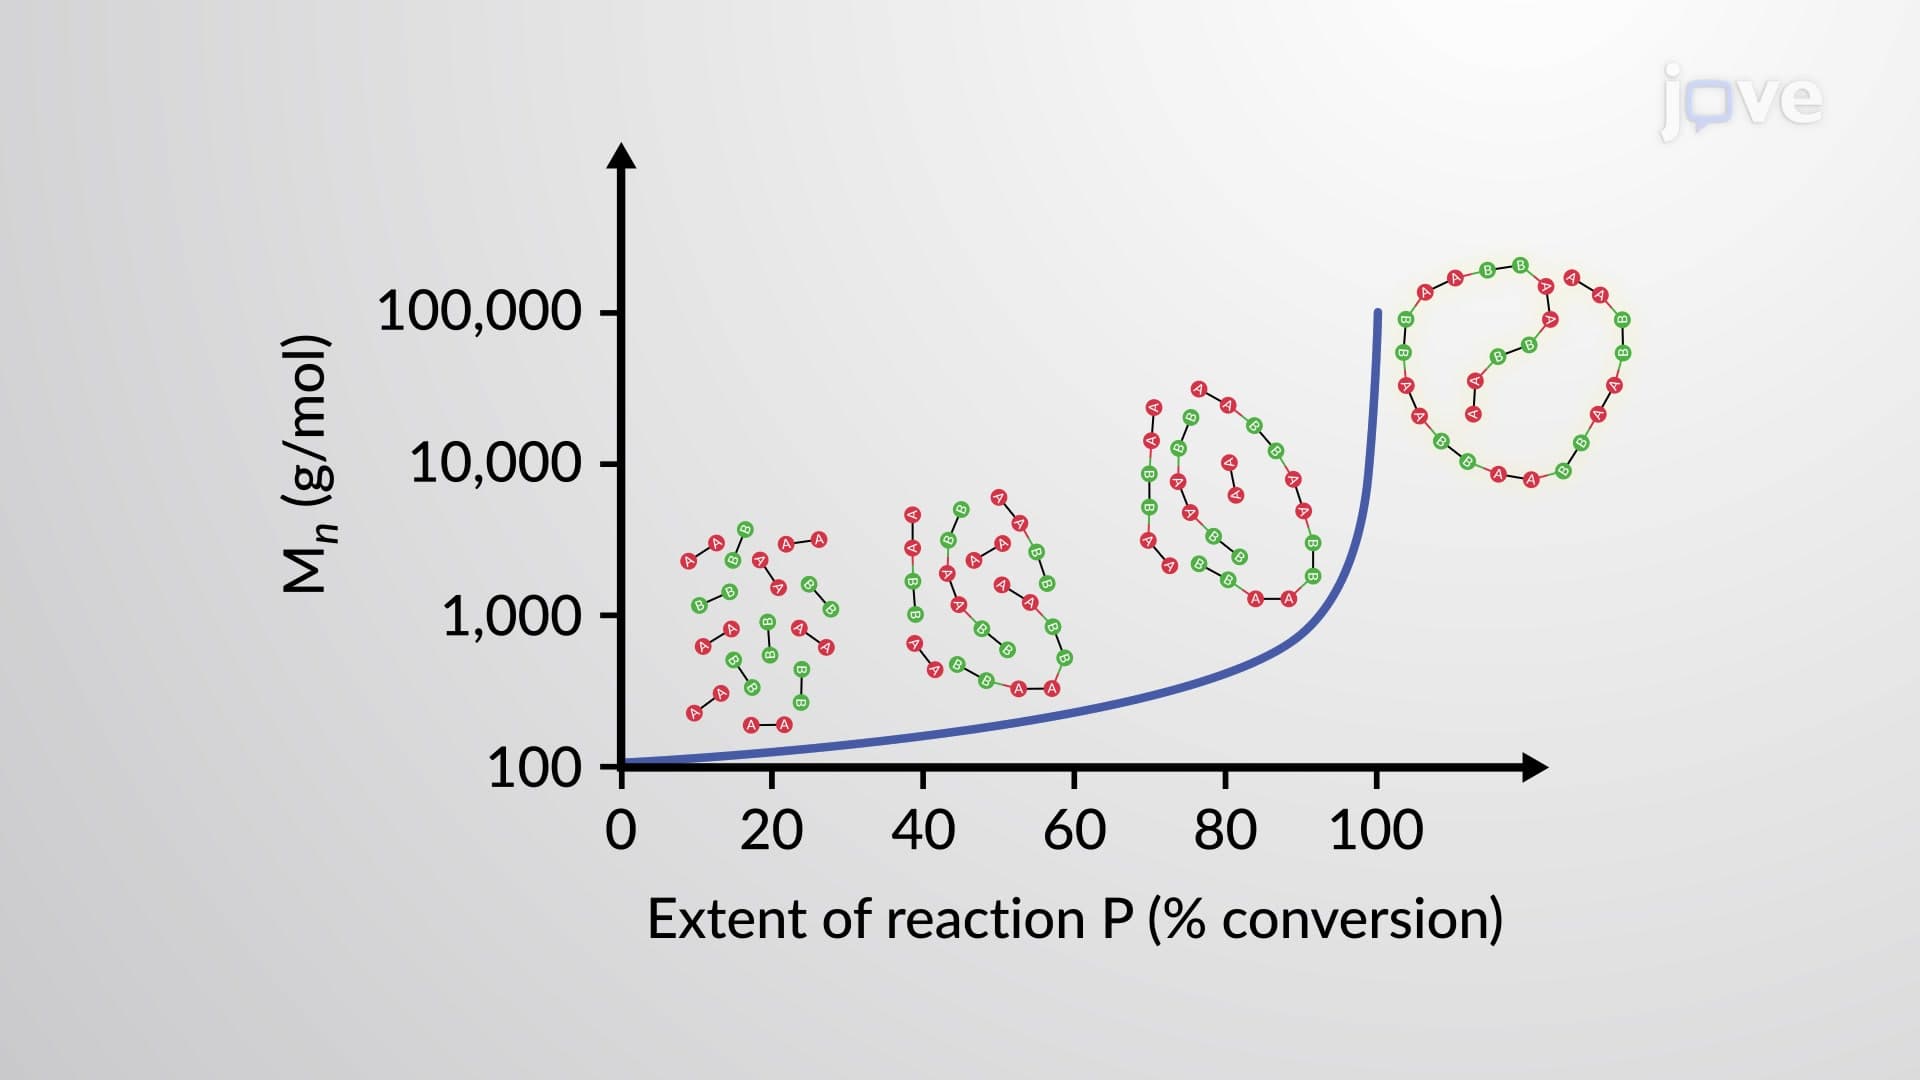
Molecular Weight of Step-Growth Polymers
As the step-growth polymerization involves step-wise condensation of monomers, the molecular weight also builds up eventually. Consequently, high molecular weight polymers are obtained at the late stages of the polymerization, where 99% of monomers have been consumed.
The extent of the...